Related Experiment Video
Updated: Aug 1, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
A double-hybrid density functional based on good local physics with outstanding performance on the GMTKN55 database
Axel D Becke1, Golokesh Santra2, Jan M L Martin2
1Department of Chemistry, Dalhousie University, 6274 Coburg Road, P.O. Box 15000, Halifax, Nova Scotia B3H 4R2, Canada.

Abstract:
In two recent papers [A. D. Becke, J. Chem. Phys. 156, 214101 (2022) and A. D. Becke, J. Chem. Phys. 157, 234102 (2022)], we compared two Kohn-Sham density functionals based on physical modeling and theory with the best density-functional power-series fits in the literature. The best error statistics reported to date for a hybrid functional on the general main-group thermochemistry, kinetics, and noncovalent interactions (GMTKN55) chemical database of Goerigk et al. [Phys. Chem. Chem. Phys. 19, 32184 (2017)] were obtained. In the present work, additional second-order perturbation-theory terms are considered. The result is a 12-parameter double-hybrid density functional with the lowest GMTKN55 WTMAD2 "weighted total mean absolute deviation" error (1.76 kcal/mol) yet seen for any hybrid or double-hybrid density-functional approximation. We call it "DH23."
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Related Concept Videos
 03:35
Hybridization of Atomic Orbitals II
 03:24
Hybridization of Atomic Orbitals I
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 02:36
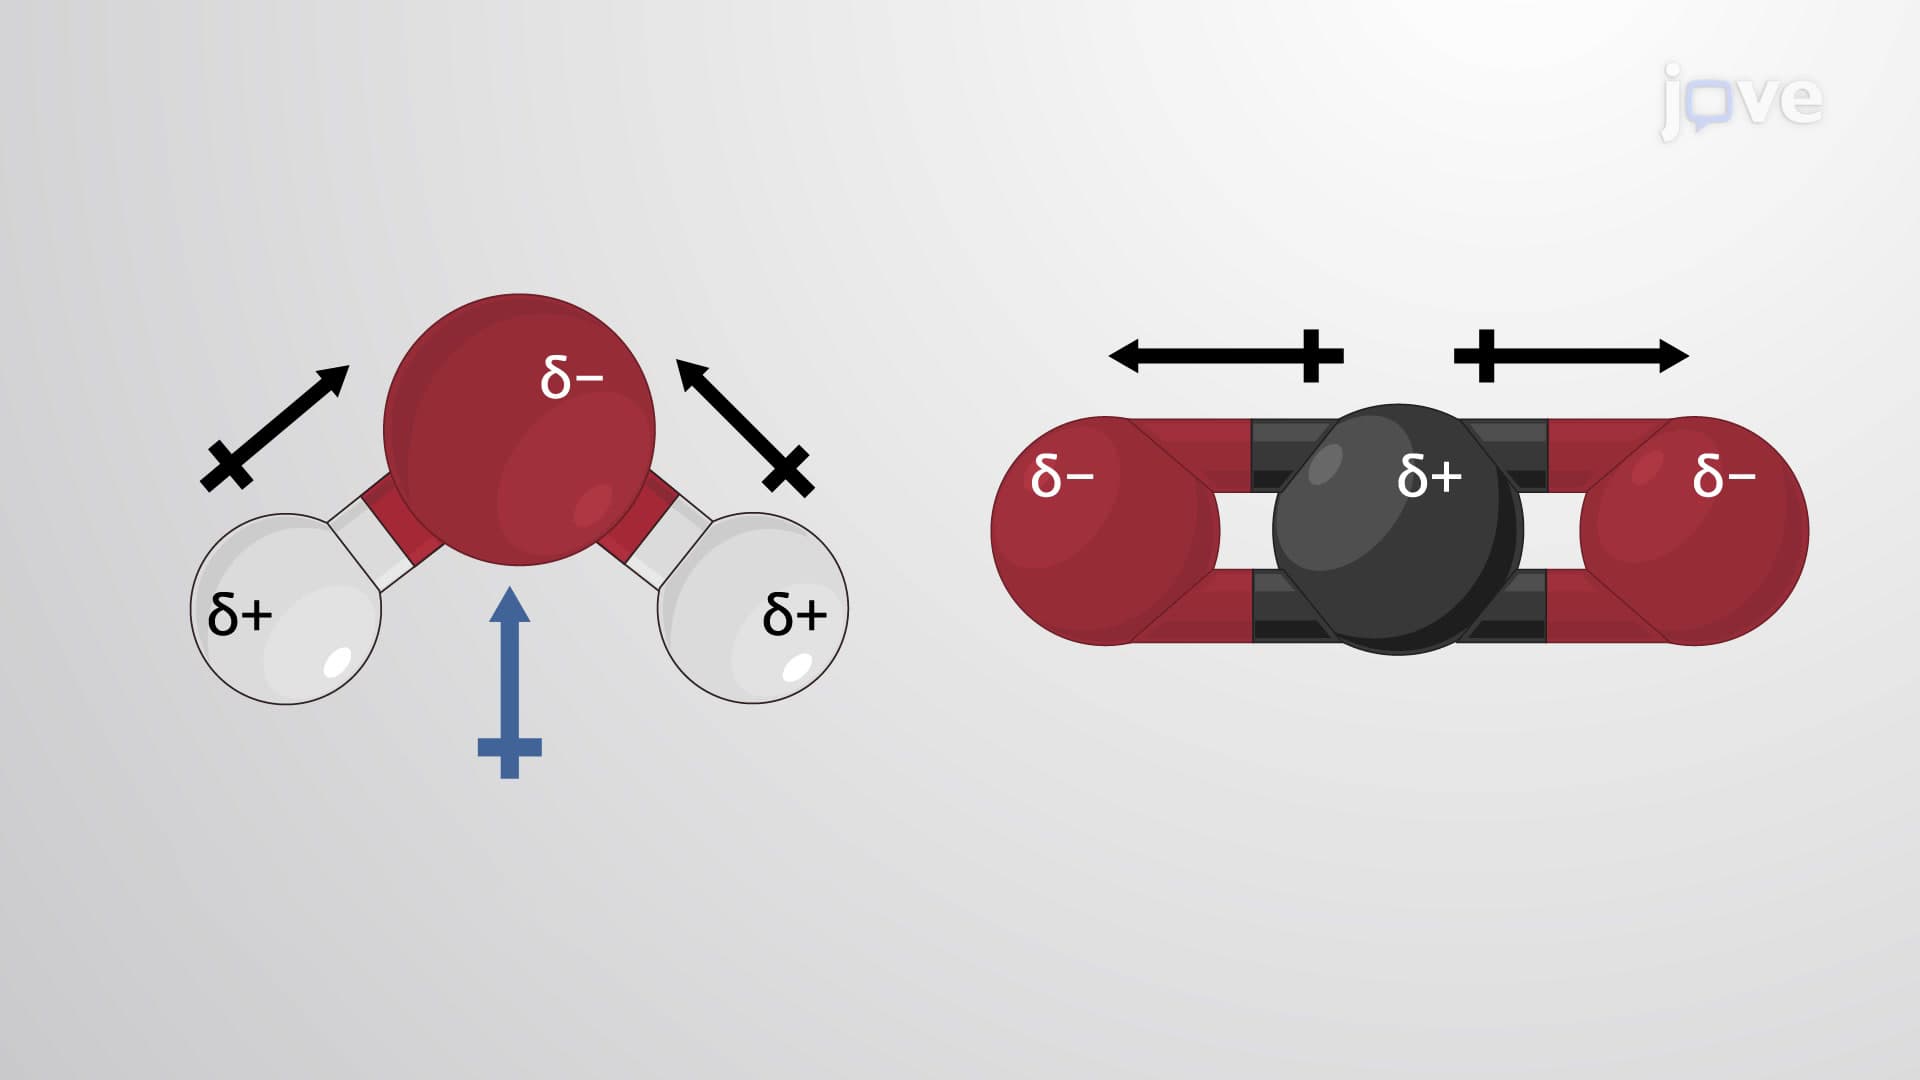
Molecular Geometry and Dipole Moments
 03:51
Molecular Orbital Theory II
 02:27
Predicting Molecular Geometry