Related Experiment Video
Updated: Aug 1, 2025

Methods for the Discovery of Novel Compounds Modulating a Gamma-Aminobutyric Acid Receptor Type A Neurotransmission
Published on: August 16, 2018
Conformationally Selective 2-Aminotetralin Ligands Targeting the alpha2A- and alpha2C-Adrenergic Receptors

Abstract:
Many important physiological processes are mediated by alpha2A- and alpha2C-adrenergic receptors (α2Rs), a subtype of class A G protein-coupled receptors (GPCRs). However, α2R signaling is poorly understood, and there are few approved medications targeting these receptors. Drug discovery aimed at α2Rs is complicated by the high degree of binding pocket homology between α2AR and α2CR, which confounds ligand-mediated selective activation or inactivation of signaling associated with a particular subtype. Meanwhile, α2R signaling is complex and it is reported that activating α2AR is beneficial in many clinical contexts, while activating α2CR signaling may be detrimental to these positive effects. Here, we report on a novel 5-substituted-2-aminotetralin (5-SAT) chemotype that, depending on substitution, has diverse pharmacological activities at α2Rs. Certain lead 5-SAT analogues act as partial agonists at α2ARs, while functioning as inverse agonists at α2CRs, a novel pharmacological profile. Leads demonstrate high potency (e.g., EC50 < 2 nM) at the α2AR and α2CRs regarding Gαi-mediated inhibition of adenylyl cyclase and production of cyclic adenosine monophosphate (cAMP). To help understand the molecular basis of 5-SAT α2R multifaceted functional activity, α2AR and α2CR molecular models were built from the crystal structures and 1 μs molecular dynamics (MD) simulations and molecular docking experiments were performed for a lead 5-SAT with α2AR agonist and α2CR inverse agonist activity, i.e., (2S)-5-(2'-fluorophenyl)-N,N-dimethyl-1,2,3,4-tetrahydronaphthalen-2-amine (FPT), in comparison to the FDA-approved (for opioid withdrawal symptoms) α2AR/α2CR agonist lofexidine. Results reveal several interactions between FPT and α2AR and α2CR amino acids that may impact the functional activity. The computational data in conjunction with experimental in vitro affinity and function results provide information to understand ligand stabilization of functionally distinct GPCR conformations regarding α2AR and α2CRs.
More Related Videos
Related Concept Videos
 01:17
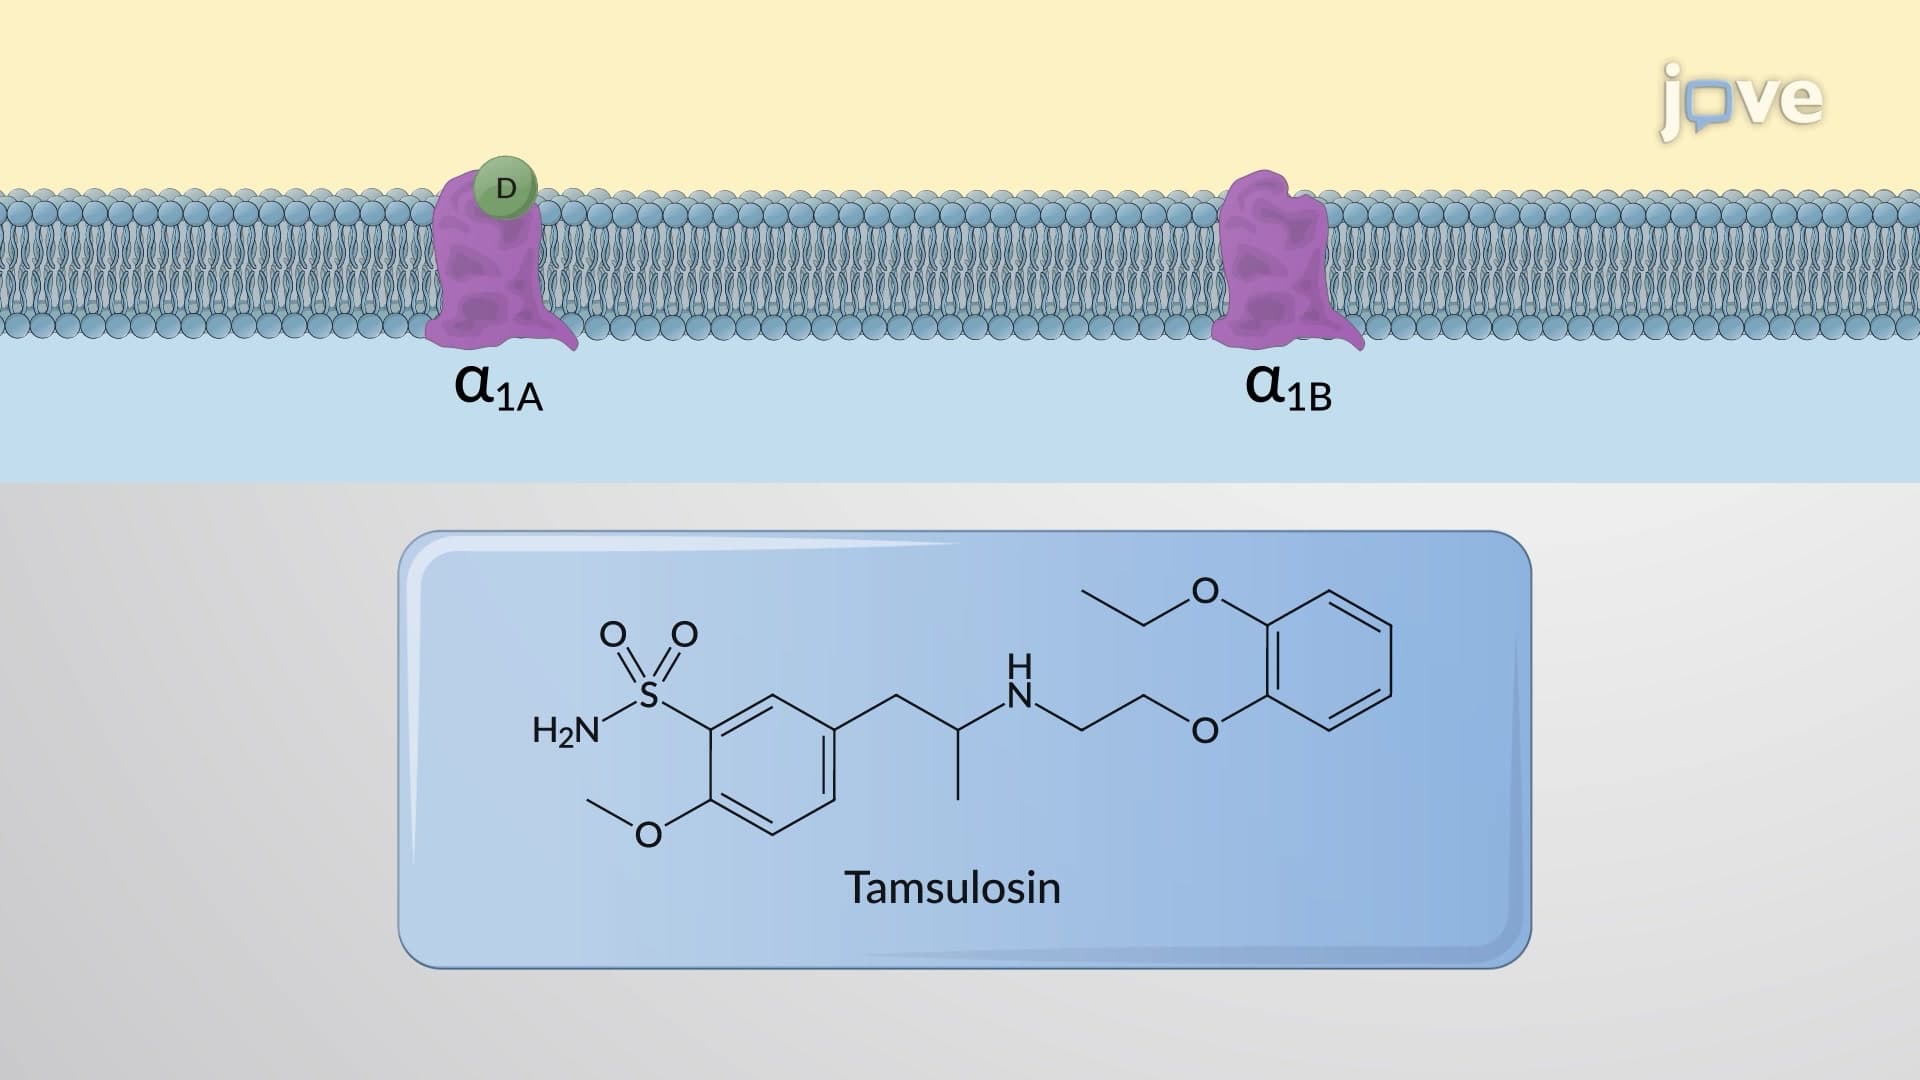
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline...
 01:31
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of...
 01:30
Adrenergic Agonists: Direct-Acting Agents
These agents can be classified...
 01:27
Adrenergic Receptors (Adrenoceptors): Classification
α-Adrenoceptors
α-Adrenoceptors are classified into two main subtypes: α1 and α2. The α1 adrenoceptors,...
 01:22
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally,...