Related Experiment Video
Updated: Jun 22, 2026
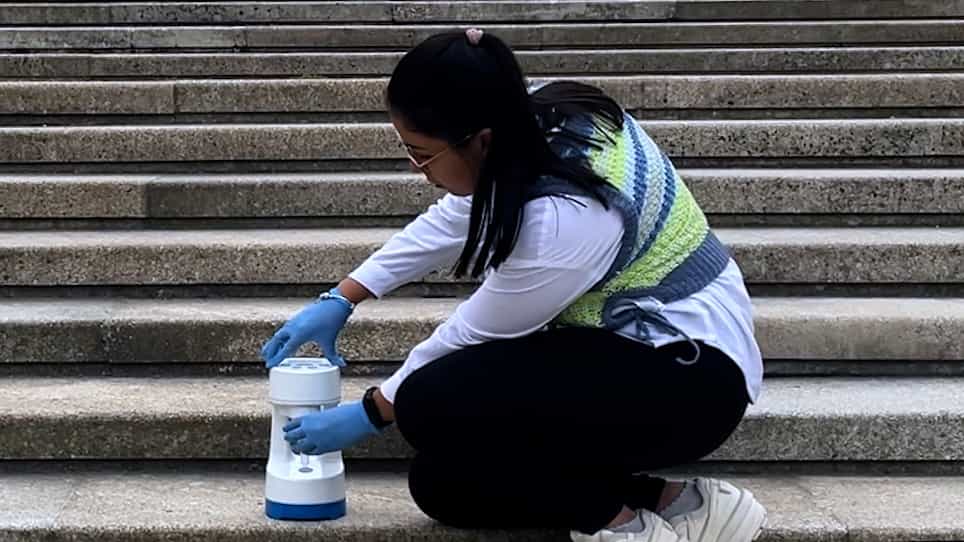

Quantification and Whole Genome Characterization of SARS-CoV-2 RNA in Wastewater and Air Samples
Published on: June 30, 2023
SARS-CoV-2 Variants by Whole-Genome Sequencing in a University Hospital in Bangkok: First to Third COVID-19 Waves
Chayanee Setthapramote1, Thanwa Wongsuk1, Chuphong Thongnak1
1Department of Clinical Pathology, Faculty of Medicine Vajira Hospital, Navamindradhiraj University, Bangkok 10300, Thailand.

Background:
Multiple severe acute respiratory syndrome coronavirus 2 (SARS-CoV-2) variants emerged globally during the recent coronavirus disease (COVID-19) pandemic. From April 2020 to April 2021, Thailand experienced three COVID-19 waves, and each wave was driven by different variants. Therefore, we aimed to analyze the genetic diversity of circulating SARS-CoV-2 using whole-genome sequencing analysis.
Methods:
A total of 33 SARS-CoV-2 positive samples from three consecutive COVID-19 waves were collected and sequenced by whole-genome sequencing, of which, 8, 10, and 15 samples were derived from the first, second, and third waves, respectively. The genetic diversity of variants in each wave and the correlation between mutations and disease severity were explored.
Results:
During the first wave, A.6, B, B.1, and B.1.375 were found to be predominant. The occurrence of mutations in these lineages was associated with low asymptomatic and mild symptoms, providing no transmission advantage and resulting in extinction after a few months of circulation. B.1.36.16, the predominant lineage of the second wave, caused more symptomatic COVID-19 cases and contained a small number of key mutations. This variant was replaced by the VOC alpha variant, which later became dominant in the third wave. We found that B.1.1.7 lineage-specific mutations were crucial for increasing transmissibility and infectivity, but not likely associated with disease severity. There were six additional mutations found only in severe COVID-19 patients, which might have altered the virus phenotype with an inclination toward more highly pathogenic SARS-CoV-2.
Conclusion:
The findings of this study highlighted the importance of whole-genome analysis in tracking newly emerging variants, exploring the genetic determinants essential for transmissibility, infectivity, and pathogenicity, and helping better understand the evolutionary process in the adaptation of viruses in humans.
Related Concept Videos
 00:36
Viral Mutations
 01:05
Single Nucleotide Polymorphisms-SNPs