Related Experiment Video
Updated: Jul 31, 2025
Paramagnetic Relaxation Enhancement for Detecting and Characterizing Self-Associations of Intrinsically Disordered Proteins
Published on: September 23, 2021
Critical Assessment of Self-Consistency Checks in the All-Atom Molecular Dynamics Simulation of Intrinsically
Kari Gaalswyk1, Austin Haider2, Kingshuk Ghosh1,2
1Department of Physics and Astronomy, University of Denver, Denver, Colorado 80208, United States.

Abstract:
All atom simulations can be used to quantify conformational properties of Intrinsically Disordered Proteins (IDP). However, simulations must satisfy convergence checks to ensure observables computed from simulation are reliable and reproducible. While absolute convergence is purely a theoretical concept requiring infinitely long simulation, a more practical, yet rigorous, approach is to impose Self Consistency Checks (SCCs) to gain confidence in the simulated data. Currently there is no study of SCCs in IDPs, unlike their folded counterparts. In this paper, we introduce different criteria for self-consistency checks for IDPs. Next, we impose these SCCs to critically assess the performance of different simulation protocols using the N terminal domain of HIV Integrase and the linker region of SARS-CoV-2 Nucleoprotein as two model IDPs. All simulation protocols begin with all-atom implicit solvent Monte Carlo (MC) simulation and subsequent clustering of MC generated conformations to create the representative structures of the IDPs. These representative structures serve as the initial structure for subsequent molecular dynamics (MD) runs with explicit solvent. We conclude that generating multiple short (∼3 μs) MD simulation trajectories─all starting from the most representative MC generated conformation─and merging them is the protocol of choice due to (i) its ability to satisfy multiple SCCs, (ii) consistently reproducing experimental data, and (iii) the efficiency of running independent trajectories in parallel by harnessing multiple cores available in modern GPU clusters. Running one long trajectory (greater than 20 μs) can also satisfy the first two criteria but is less desirable due to prohibitive computation time. These findings help resolve the challenge of identifying a usable starting configuration, provide an objective measure of SCC, and establish rigorous criteria to determine the minimum length (for one long simulation) or number of trajectories needed in all-atom simulation of IDPs.
Related Concept Videos
 02:18
Intrinsically Disordered Proteins
 01:22
Protein Folding
 01:29
Protein Folding Quality Check in the RER
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:24
Protein Organization
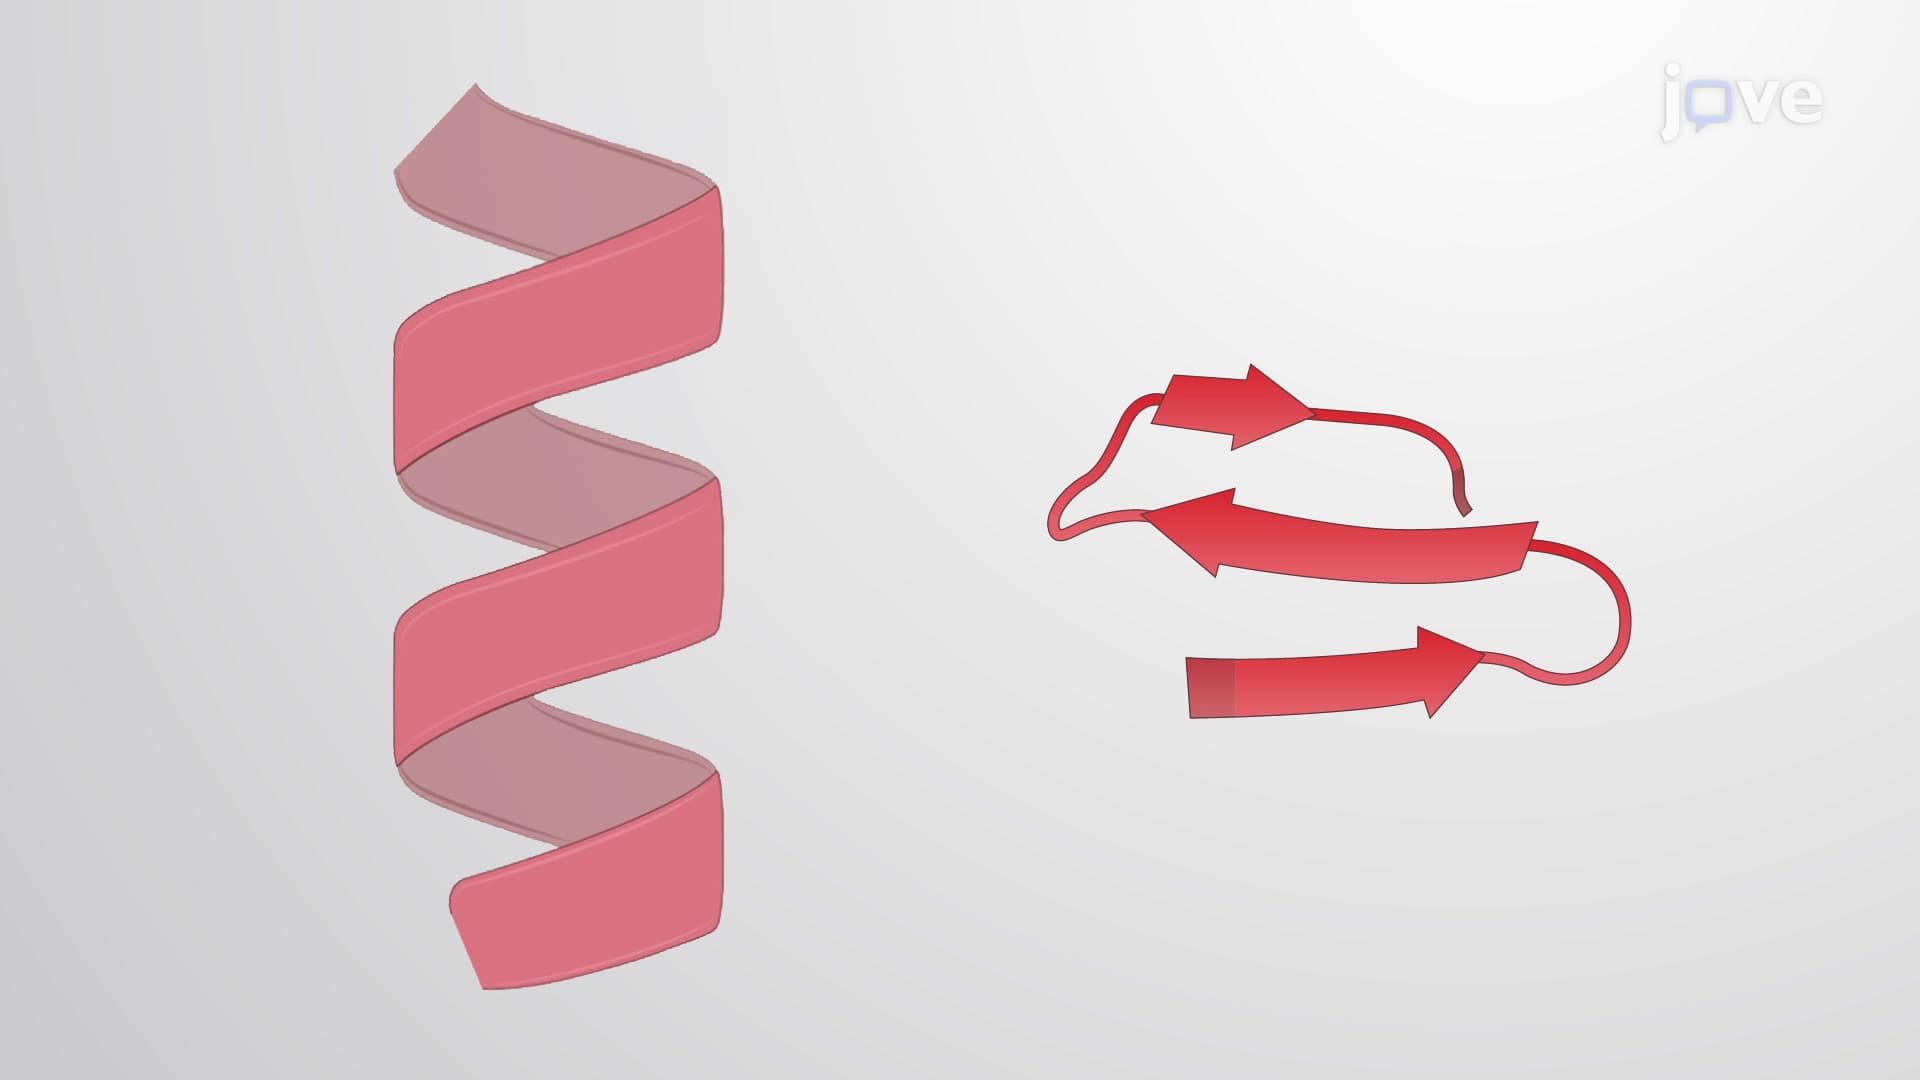
The primary structure of a protein is its amino acid sequence....
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to...