Related Experiment Video
Updated: Jul 29, 2025


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Discovery of andrographolide hit analog as a potent cyclooxygenase-2 inhibitor through consensus MD-simulation,
Priyanka Jain1, Jitendra Satija2, C Sudandiradoss3
1School of Biosciences and Technology, Vellore Institute of Technology, Vellore, Tamil Nadu, 632014, India.

Abstract:
Cyclooxygenase-2 (COX-2) is the key enzyme responsible for the conversion of arachidonic acid to prostaglandins that display pro-inflammatory properties and thus, it is a potential target protein to develop anti-inflammatory drugs. In this study, chemical and bio-informatics approaches have been employed to find a novel potent andrographolide (AGP) analog as a COX-2 inhibitor having better pharmacological properties than aspirin and rofecoxib (controls). The full amino acid sequenced human Alpha fold (AF) COX-2 protein (604AA) was selected and validated for its accuracy against the reported COX-2 protein structures (PDB ID: 5F19, 5KIR, 5F1A, 5IKQ and 1V0X) followed by multiple sequence alignment analysis to establish the sequence conservation. The systematic virtual screening of 237 AGP analogs against AF-COX-2 protein yielded 22 lead compounds based on the binding energy score (< - 8.0 kcal/mol). These were further screened out to 7 analogs by molecular docking analysis and investigated further for ADMET prediction, ligand efficiency metrics calculations, quantum mechanical analysis, MD simulation, electrostatic potential energy (EPE) docking simulation, and MM/GBSA. In-depth analysis revealed that AGP analog A3 (3-[2-[(1R,4aR,5R,6R,8aR)-6-hydroxy-5,6,8a-trimethyl-2-methylidene-3,4,4a,5,7,8-hexahydro-1H-naphthalen-1-yl]ethylidene]-4-hydroxyoxolan-2-one) forms the most stable complex with the AF-COX-2 showing the least RMSD value (0.37 ± 0.03 nm), a good number of hydrogen bonds (protein-ligand H-bond = 11, and protein H-bond = 525), minimum EPE score (- 53.81 kcal/mol), and lowest MM-GBSA before and after simulation (- 55.37 and - 56.25 kcal/mol, respectively) value compared to other analogs and controls. Thus, we suggest that the identified A3 AGP analog could be developed as a promising plant-based anti-inflammatory drug by inhibiting COX-2.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 08:15
Author Spotlight: Network Pharmacology and Molecular Docking to Decipher the Action of Jiawei Shengjiang San Against Diabetic Kidney Disease
Published on: May 10, 2024
Related Concept Videos
 01:26
Drug Discovery: Overview
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:15
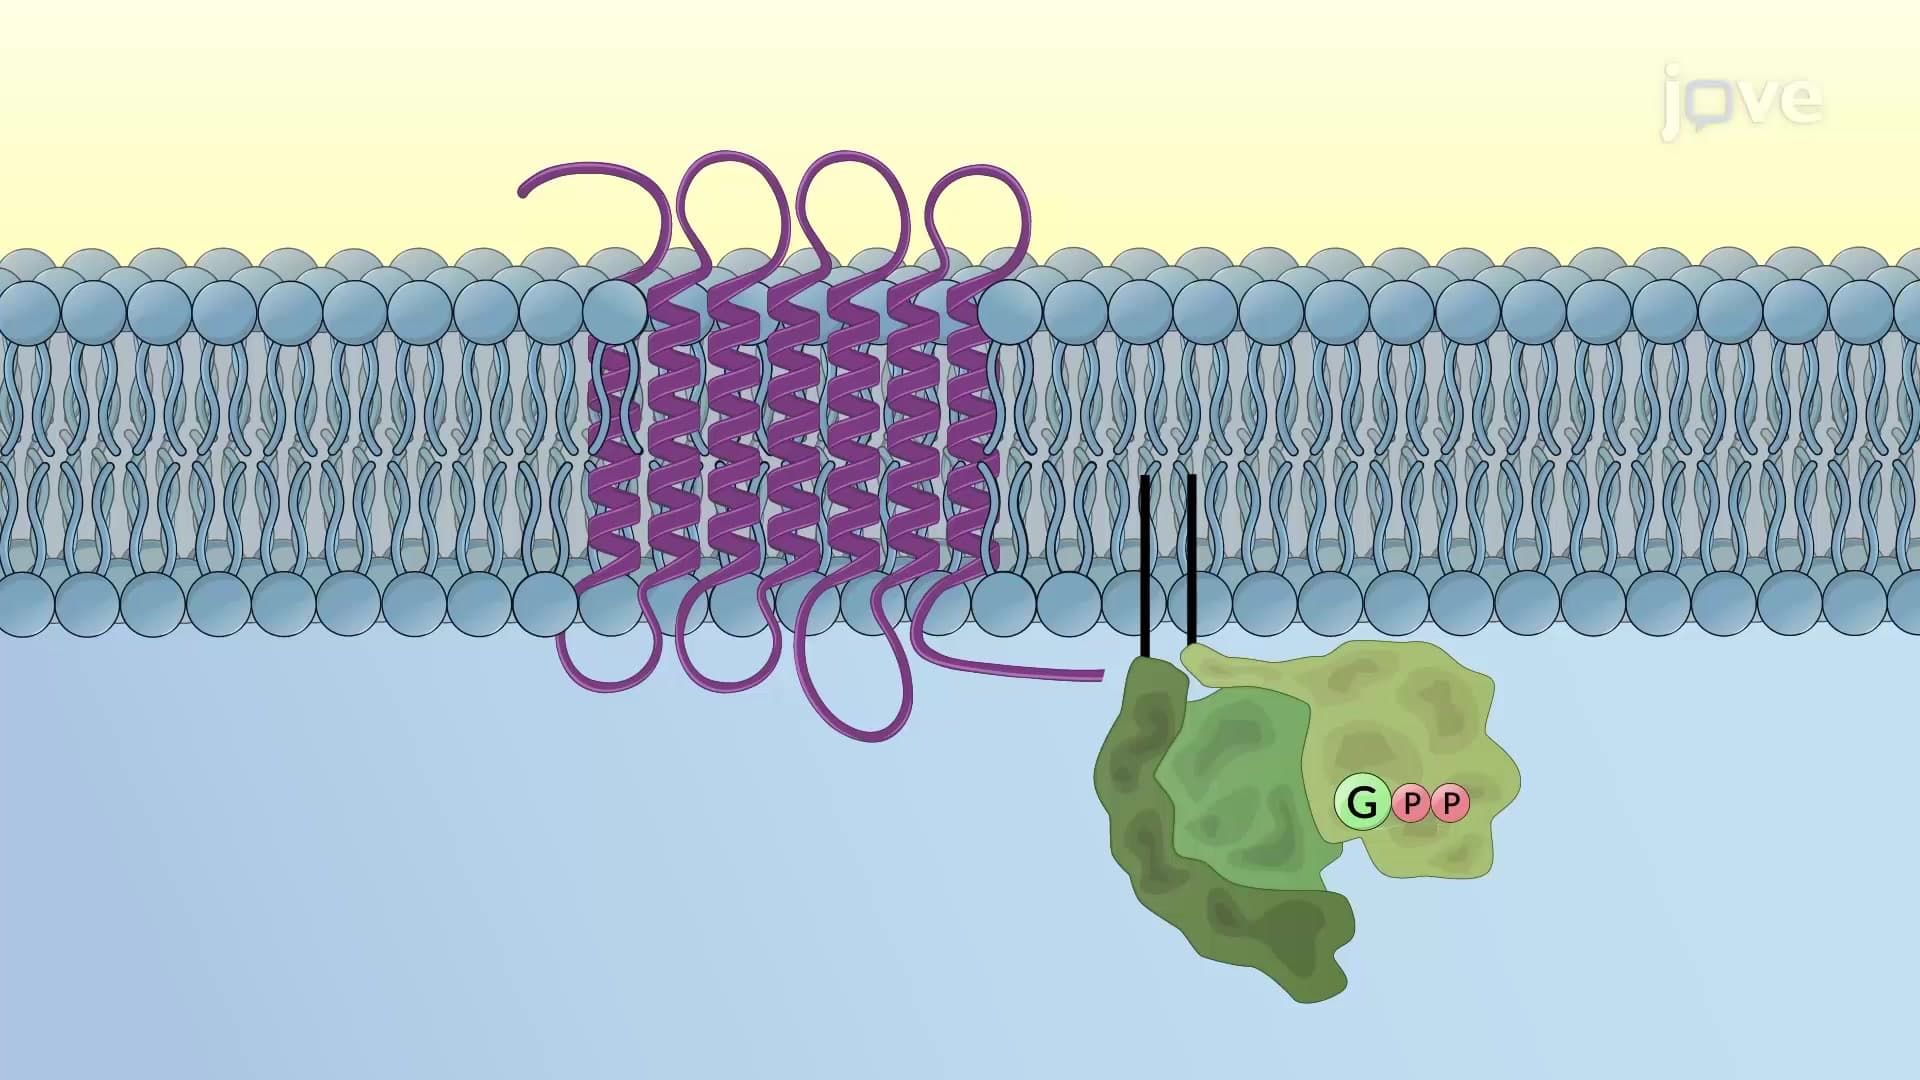
G Protein-coupled Receptors
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
 01:25
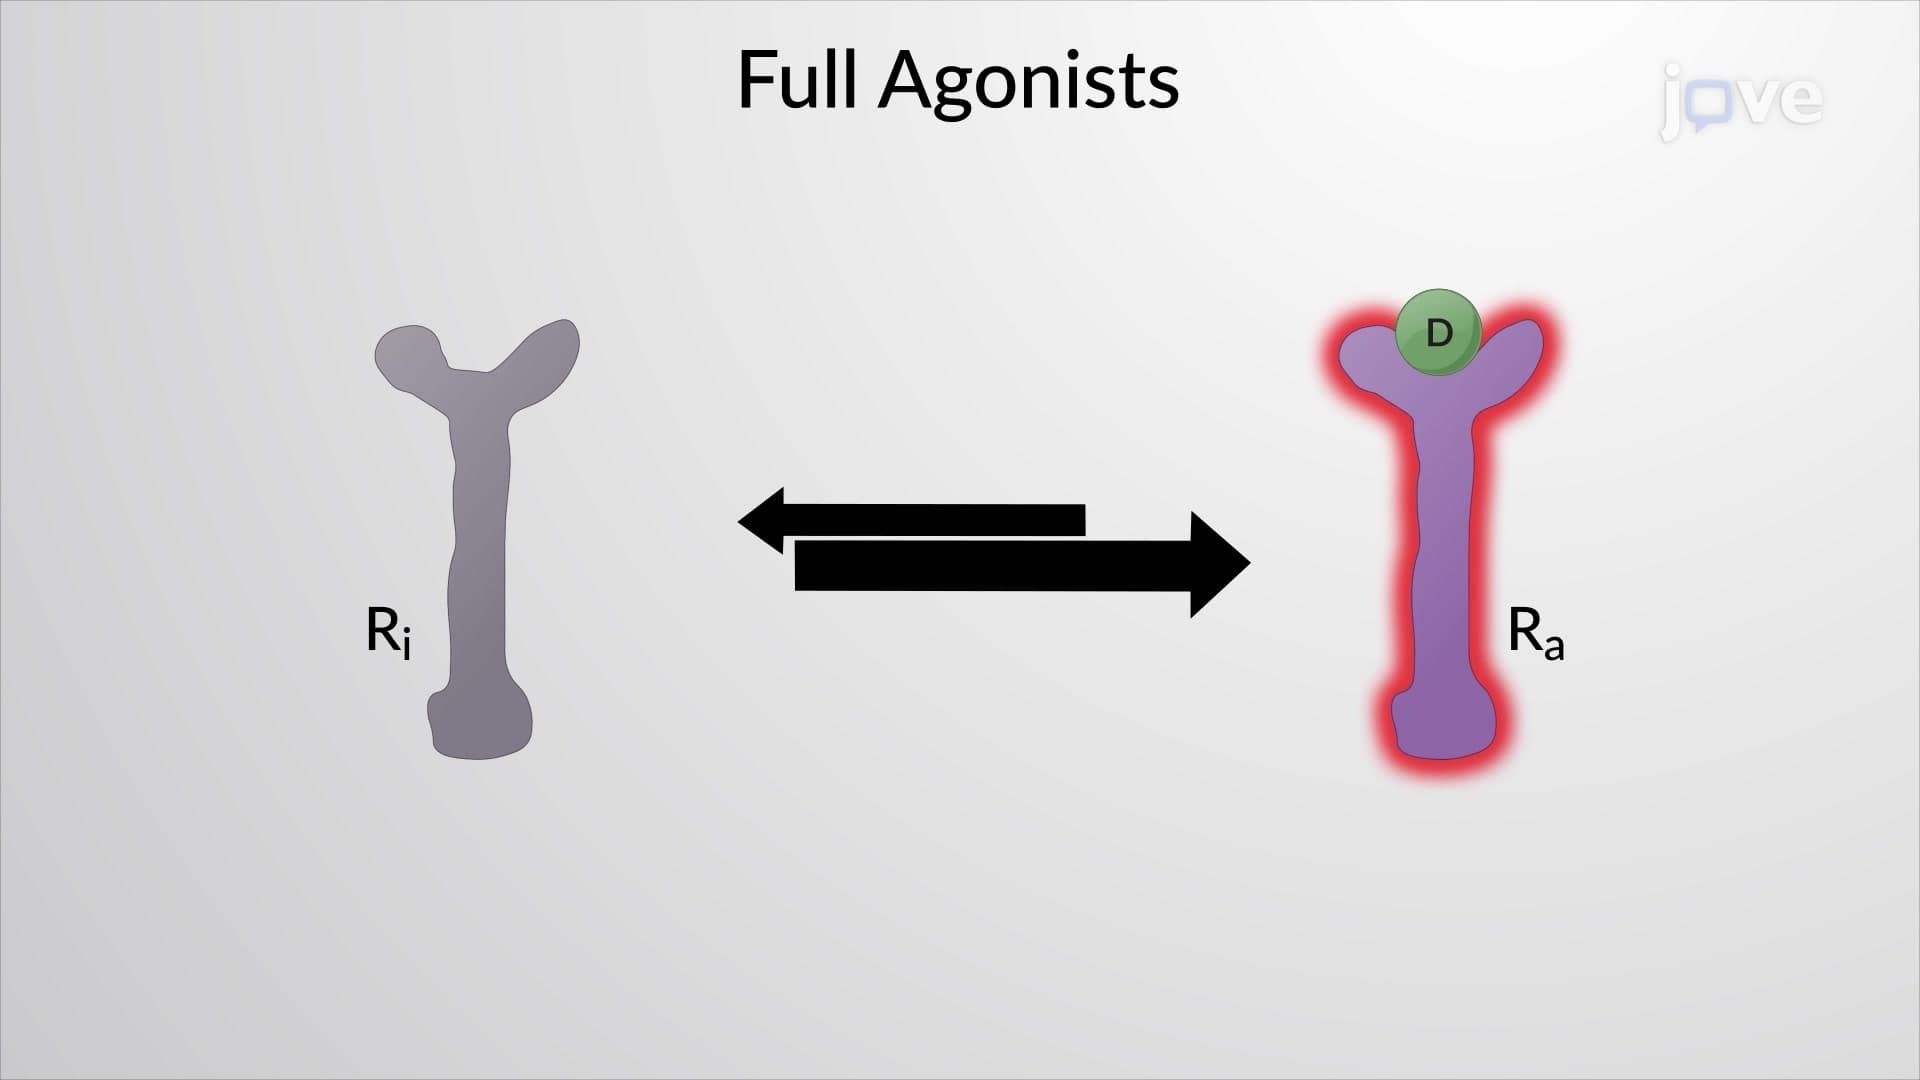
Drug-Receptor Interaction: Agonist
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous...
 01:22
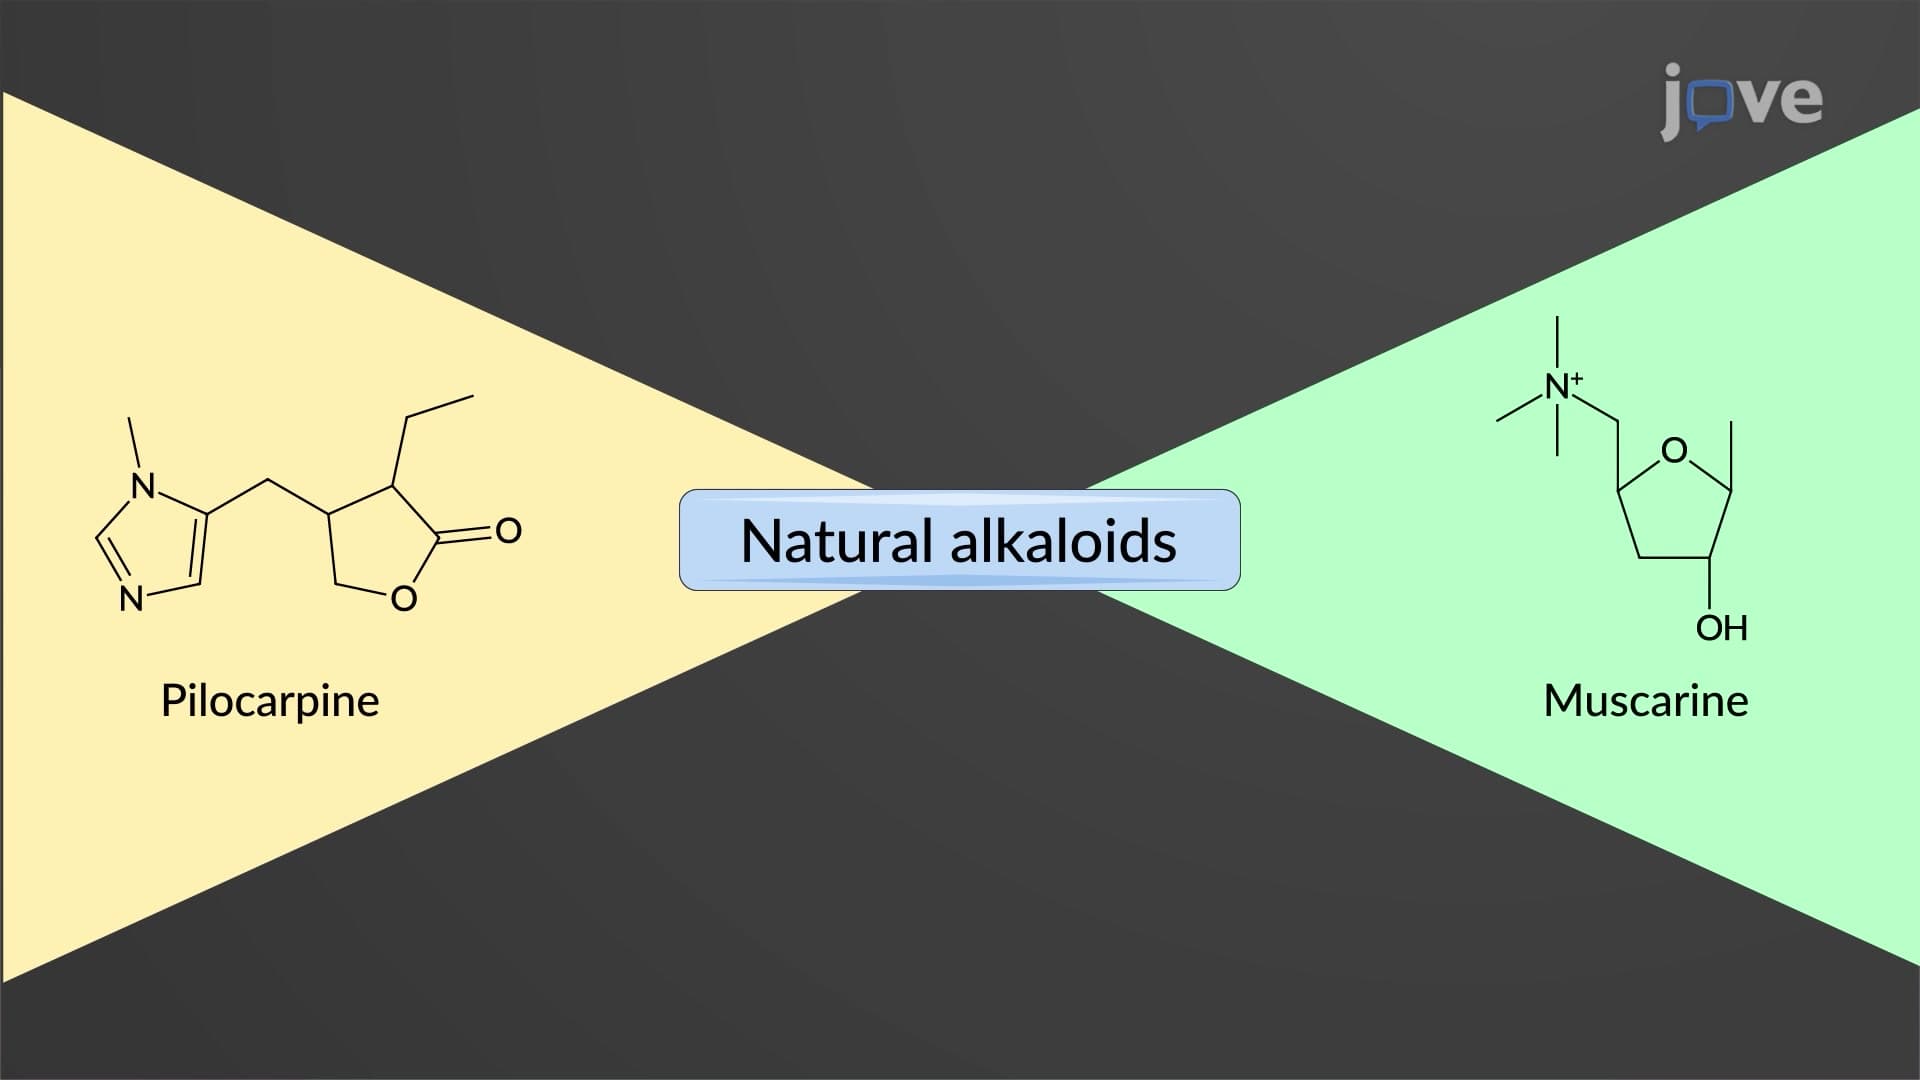
Direct-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
The direct-acting...