Related Experiment Video
Updated: Jul 28, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Computing vibrational energy levels using a canonical polyadic tensor method with a fixed rank and a contraction tree
Sangeeth Das Kallullathil1, Tucker Carrington1
1Chemistry Department, Queen's University, Kingston, Ontario K7L 3N6, Canada.

Abstract:
In this paper, we use the previously introduced Canonical Polyadic (CP)-Multiple Shift Block Inverse Iteration (MSBII) eigensolver [S. D. Kallullathil and T. Carrington, J. Chem. Phys. 155, 234105 (2021)] in conjunction with a contraction tree to compute vibrational spectra. The CP-MSBII eigensolver uses the CP format. The memory cost scales linearly with the number of coordinates. A tensor in CP format represents a wavefunction constrained to be a sum of products (SOP). An SOP wavefunction can be made more accurate by increasing the number of terms, the rank. When the required rank is large, the runtime of a calculation in CP format is long, although the memory cost is small. To make the method more efficient, we break the full problem into pieces using a contraction tree. The required rank for each of the sub-problems is small. To demonstrate the effectiveness of the ideas, we computed vibrational energy levels of acetonitrile (12-D) and ethylene oxide (15-D).
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 01:08
IR Spectrum Peak Splitting: Symmetric vs Asymmetric Vibrations
 01:09
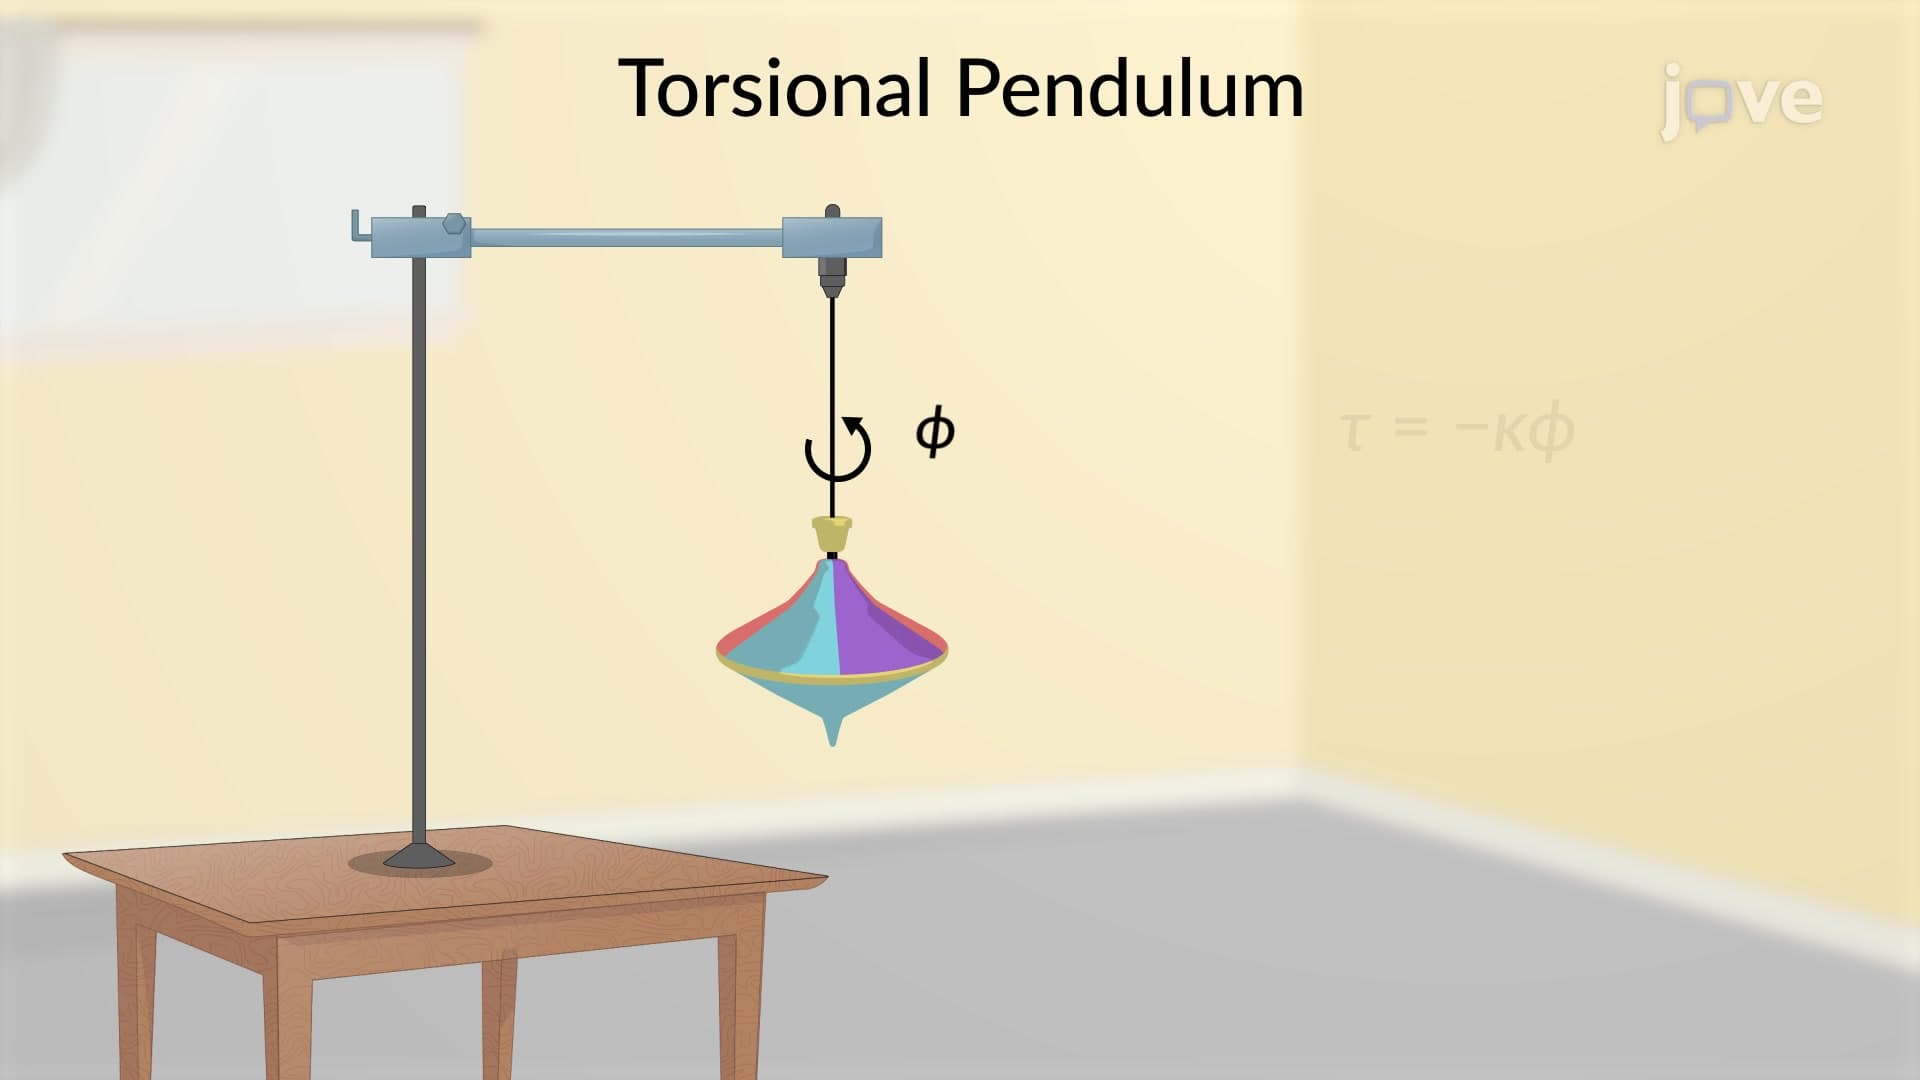
Torsional Pendulum
As long as the rigid body's angular displacement is small, its oscillation can be modeled as a linear angular oscillation. The amplitude of the oscillation is an angle. The role of mass is played...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 03:35
Hybridization of Atomic Orbitals II
 01:08
Vector Algebra: Method of Components
In many applications, the magnitudes and directions of...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...