Related Experiment Video
Updated: Jul 27, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Riemannian Trust Region Method for Minimization of the Fourth Central Moment for Localized Molecular Orbitals
Aliakbar Sepehri1, Run R Li2, Mark R Hoffmann1
1Chemistry Department, University of North Dakota, Grand Forks, North Dakota 58202-9024, United States.

Abstract:
The importance of localized molecular orbitals (MOs) in correlation treatments beyond mean-field calculation and in the illustration of chemical bonding (and antibonding) can hardly be overstated. However, the generation of orthonormal localized occupied MOs is significantly more straightforward than obtaining orthonormal localized virtual MOs. Orthonormal MOs allow facile use of highly efficient group theoretical methods (e.g., graphical unitary group approach) for calculation of Hamiltonian matrix elements in multireference configuration interaction calculations (such as MRCISD) and in quasi-degenerate perturbation treatments, such as the Generalized Van Vleck Perturbation Theory. Moreover, localized MOs can elucidate qualitative understanding of bonding in molecules, in addition to high-accuracy quantitative descriptions. We adopt the powers of the fourth moment cost function introduced by Jørgensen and coworkers. Because the fourth moment cost functions are prone to having multiple negative Hessian eigenvalues when starting from easily available canonical (or near-canonical) MOs, standard optimization algorithms can fail to obtain the orbitals of the virtual or partially occupied spaces. To overcome this drawback, we applied a trust region algorithm on an orthonormal Riemannian manifold with an approximate retraction from the tangent space built into the first and second derivatives of the cost function. Moreover, the Riemannian trust region outer iterations were coupled to truncated Conjugate Gradient inner loops, which avoided any costly solutions of simultaneous linear equations or eigenvector/eigenvalue solutions. Numerical examples are provided on model systems, including the high-connectivity H10 set in 1-, 2-, and 3-dimensional arrangements, and on a chemically realistic description of cyclobutadiene (c-C4H4) and the propargyl radical (C3H3). In addition to demonstrating the algorithm on occupied and virtual blocks of orbitals, the method is also shown to work on the active space at the MCSCF level of theory.
More Related Videos
Related Concept Videos
 02:35
Molecular Orbital Theory I
 02:40
MO Theory and Covalent Bonding
 03:51
Molecular Orbital Theory II
 01:08
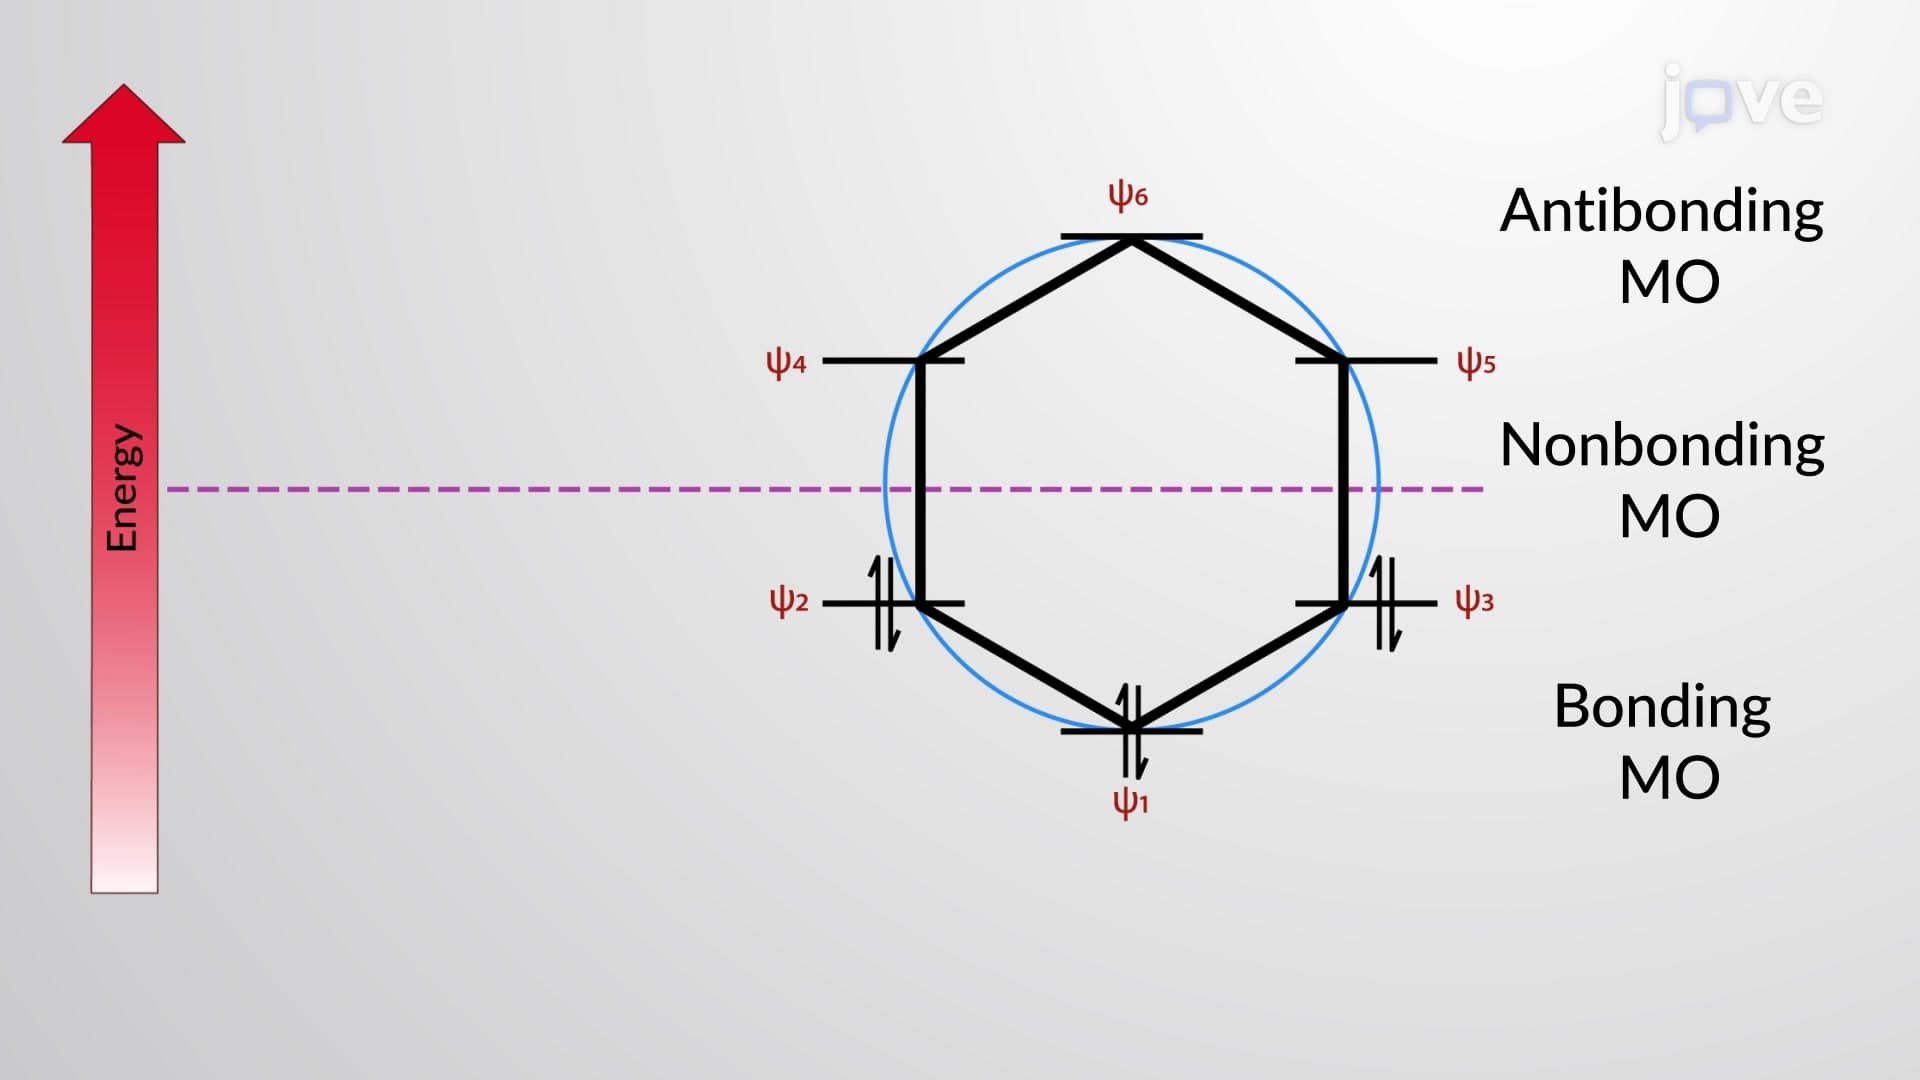
Hückel's Rule Diagram of π MOs: Frost Circle
A Frost circle is constructed by drawing a polygon whose number of edges is equal to the number of carbons of the given cyclic system, with one of the vertices pointing down. Then, a circle is drawn enclosing the polygon so...
 02:44
Atomic Orbitals
 02:36
Molecular Geometry and Dipole Moments