Related Experiment Video
Updated: Jul 26, 2025
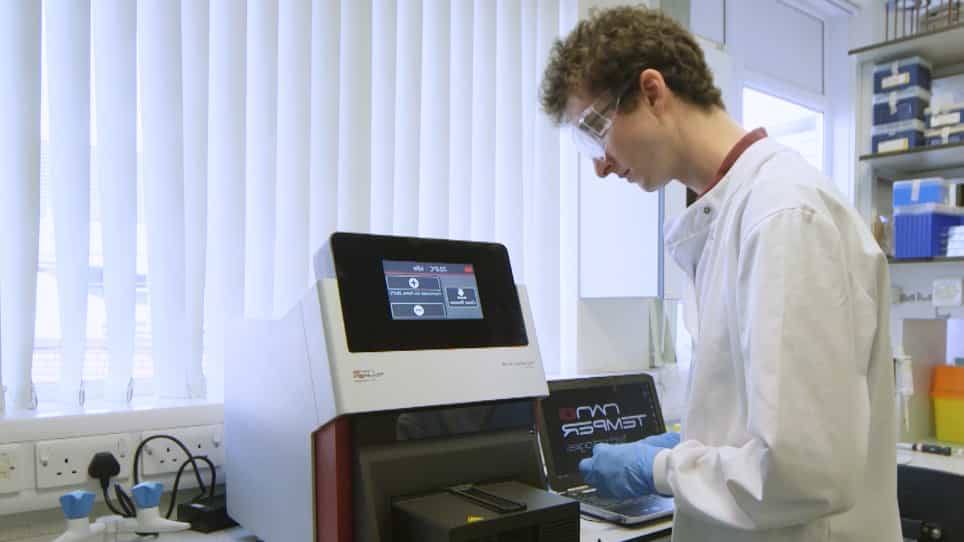

How to Stabilize Protein: Stability Screens for Thermal Shift Assays and Nano Differential Scanning Fluorimetry in the Virus-X Project
Published on: February 11, 2019
Probing protein stability: towards a computational atomistic, reliable, affordable, and improvable model
Germano Nobili1,2, Simone Botticelli1,2, Giovanni La Penna3,2
1Dipartimento di Fisica, Universitá di Roma Tor Vergata, Roma, Italy.

Abstract:
We present an improved application of a recently proposed computational method designed to evaluate the change of free energy as a function of the average value of a suitably chosen collective variable in proteins. The method is based on a full atomistic description of the protein and its environment. The goal is to understand how the protein melting temperature changes upon single-point mutations, because the sign of the temperature variation will allow us to discriminate stabilizing vs. destabilizing mutations in protein sequences. In this refined application the method is based on altruistic well-tempered metadynamics, a variant of multiple-walkers metadynamics. The resulting metastatistics is then modulated by the maximal constrained entropy principle. The latter turns out to be especially helpful in free-energy calculations as it is able to alleviate the severe limitations of metadynamics in properly sampling folded and unfolded configurations. In this work we apply the computational strategy outlined above in the case of the bovine pancreatic trypsin inhibitor, a well-studied small protein, which is a reference for computer simulations since decades. We compute the variation of the melting temperature characterizing the folding-unfolding process between the wild-type protein and two of its single-point mutations that are seen to have opposite effect on the free energy changes. The same approach is used for free energy difference calculations between a truncated form of frataxin and a set of five of its variants. Simulation data are compared to in vitro experiments. In all cases the sign of the change of melting temperature is reproduced, under the further approximation of using an empirical effective mean-field to average out protein-solvent interactions.
More Related Videos
 07:33
Analyzing Protein Architectures and Protein-Ligand Complexes by Integrative Structural Mass Spectrometry
Published on: October 15, 2018
 07:08
Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
Related Concept Videos
 01:24
Protein Organization
The primary structure of a protein is its amino acid sequence....
 01:22
Protein Folding
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to...
 02:04
Covalently Linked Protein Regulators
These groups modify specific amino acids in a protein....
 01:53
RNA Stability
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...