Related Experiment Video
Updated: Jul 26, 2025

Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Nuclear-Electronic Orbital QM/MM Approach: Geometry Optimizations and Molecular Dynamics
Mathew Chow1, Eleftherios Lambros2, Xiaosong Li2
1Department of Chemistry, Yale University, New Haven, Connecticut 06520, United States.

Abstract:
Hybrid quantum mechanical/molecular mechanical (QM/MM) methods allow simulations of chemical reactions in atomistic solvent and heterogeneous environments such as proteins. Herein, the nuclear-electronic orbital (NEO) QM/MM approach is introduced to enable the quantization of specified nuclei, typically protons, in the QM region using a method such as NEO-density functional theory (NEO-DFT). This approach includes proton delocalization, polarization, anharmonicity, and zero-point energy in geometry optimizations and dynamics. Expressions for the energies and analytical gradients associated with the NEO-QM/MM method, as well as the previously developed polarizable continuum model (NEO-PCM), are provided. Geometry optimizations of small organic molecules hydrogen bonded to water in either dielectric continuum solvent or explicit atomistic solvent illustrate that aqueous solvation can strengthen hydrogen-bonding interactions for the systems studied, as indicated by shorter intermolecular distances at the hydrogen-bond interface. We then performed a real-time direct dynamics simulation of a phenol molecule in explicit water using the NEO-QM/MM method. These developments and initial examples provide the foundation for future studies of nuclear-electronic quantum dynamics in complex chemical and biological environments.

Related Concept Videos
 02:35
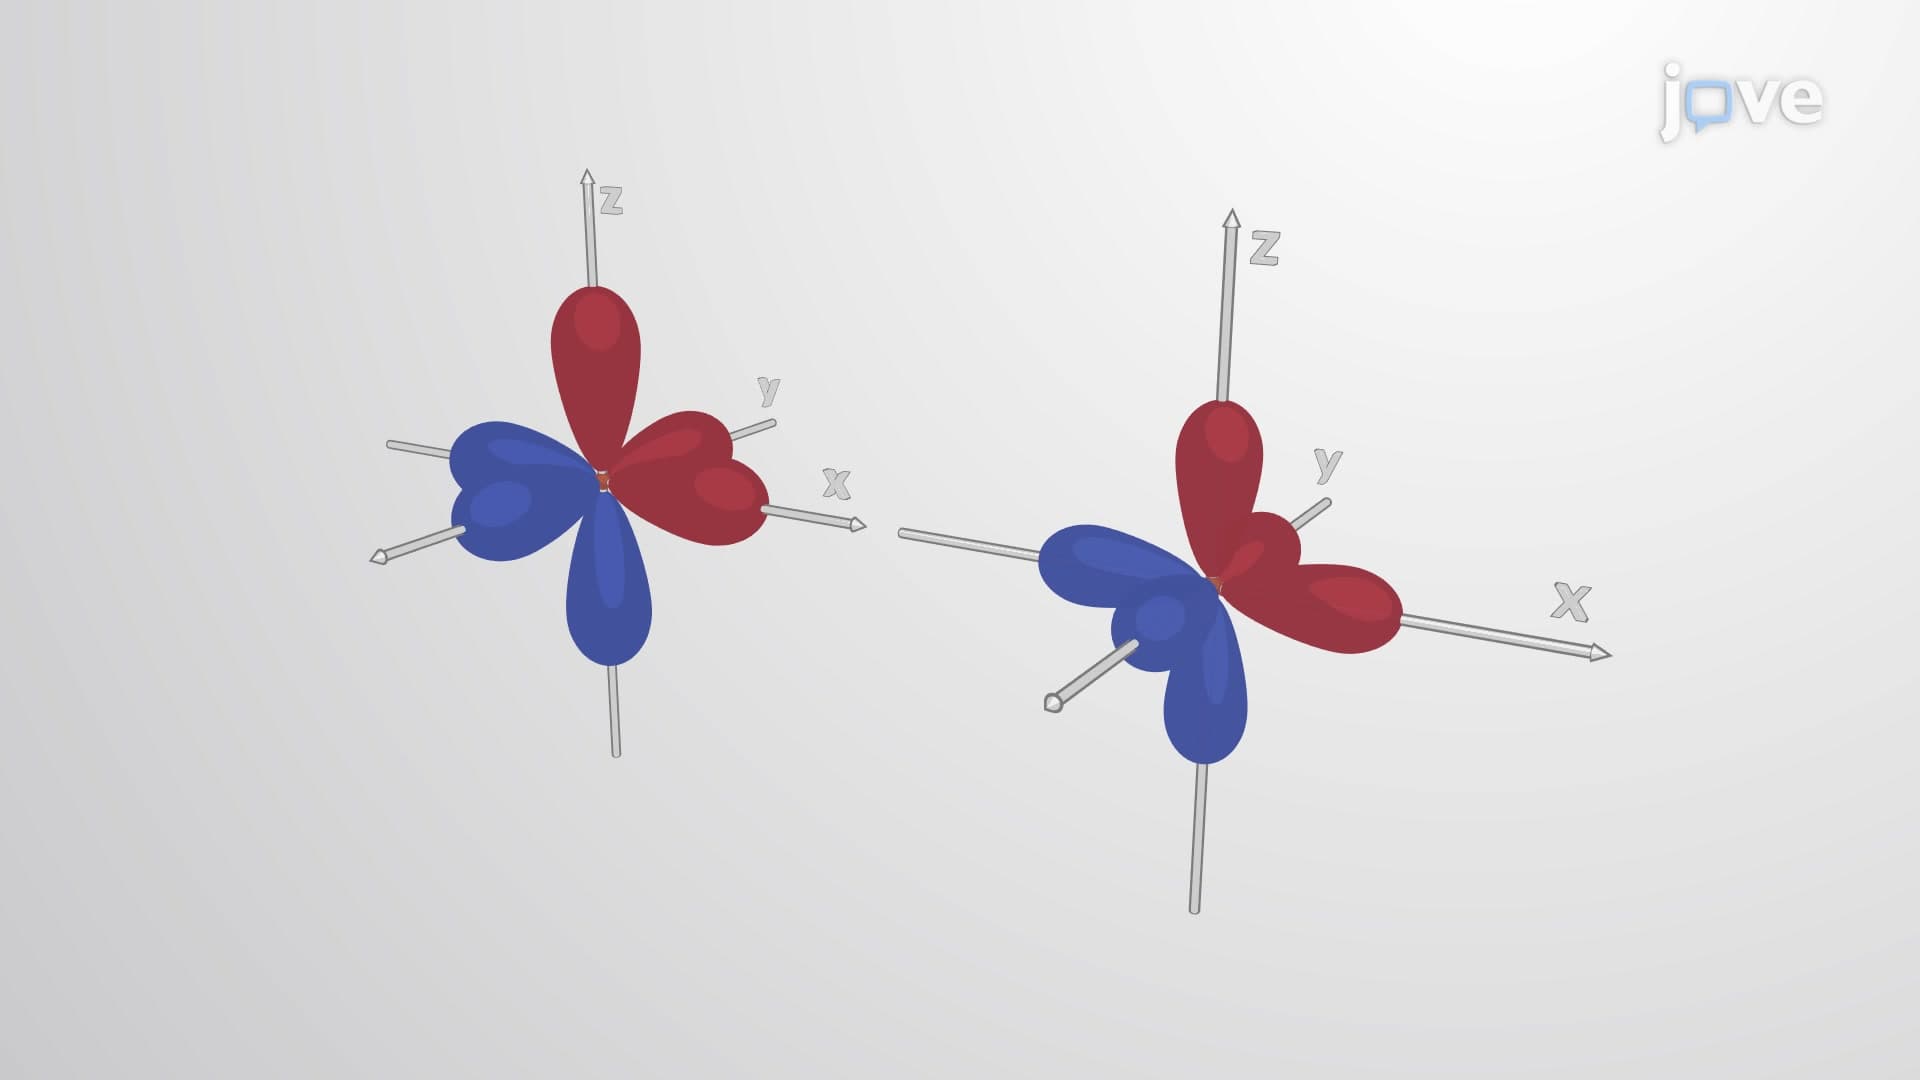
Molecular Orbital Theory I
 02:36
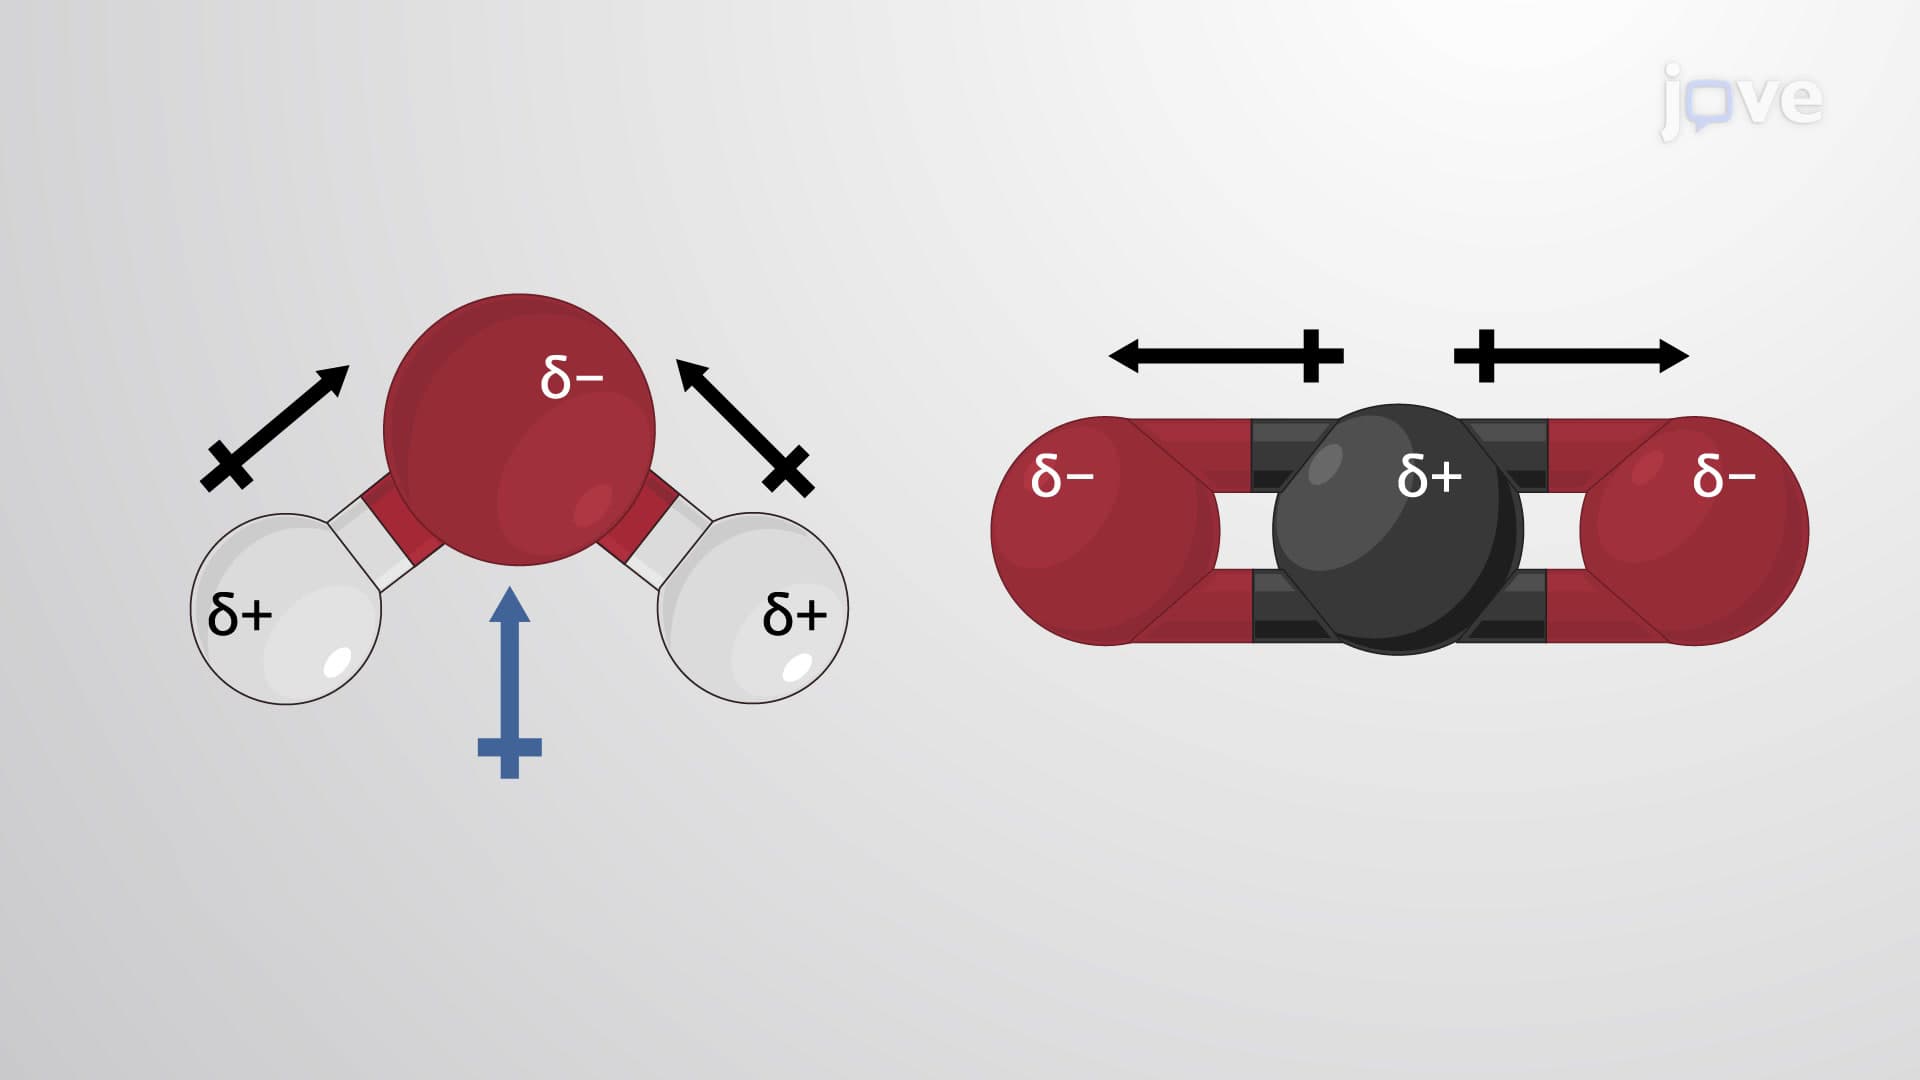
Molecular Geometry and Dipole Moments
 02:27
Predicting Molecular Geometry
 03:51
Molecular Orbital Theory II
 03:24
Hybridization of Atomic Orbitals I
 02:44
Atomic Orbitals