Related Experiment Video
Updated: Jul 26, 2025

Combining X-Ray Crystallography with Small Angle X-Ray Scattering to Model Unstructured Regions of Nsa1 from S. Cerevisiae
Published on: January 10, 2018
Validation of X-ray Crystal Structure Ensemble Representations of SARS-CoV-2 Main Protease by Solution NMR Residual
Yang Shen1, Angus J Robertson2, Ad Bax1
1Laboratory of Chemical Physics, National Institute of Diabetes and Digestive and Kidney Diseases, National Institutes of Health, Bethesda, MD 20892, USA.

Abstract:
Considerable debate has focused on whether sampling of molecular dynamics trajectories restrained by crystallographic data can be used to develop realistic ensemble models for proteins in their natural, solution state. For the SARS-CoV-2 main protease, Mpro, we evaluated agreement between solution residual dipolar couplings (RDCs) and various recently reported multi-conformer and dynamic-ensemble crystallographic models. Although Phenix-derived ensemble models showed only small improvements in crystallographic Rfree, substantially improved RDC agreement over fits to a conventionally refined 1.2-Å X-ray structure was observed, in particular for residues with above average disorder in the ensemble. For a set of six lower resolution (1.55-2.19 Å) Mpro X-ray ensembles, obtained at temperatures ranging from 100 to 310 K, no significant improvement over conventional two-conformer representations was found. At the residue level, large differences in motions were observed among these ensembles, suggesting high uncertainties in the X-ray derived dynamics. Indeed, combining the six ensembles from the temperature series with the two 1.2-Å X-ray ensembles into a single 381-member "super ensemble" averaged these uncertainties and substantially improved agreement with RDCs. However, all ensembles showed excursions that were too large for the most dynamic fraction of residues. Our results suggest that further improvements to X-ray ensemble refinement are feasible, and that RDCs provide a sensitive benchmark in such endeavors. Remarkably, a weighted ensemble of 350 PDB Mpro X-ray structures provided slightly better cross-validated agreement with RDCs than any individual ensemble refinement, implying that differences in lattice confinement also limit the fit of RDCs to X-ray coordinates.
More Related Videos
Related Concept Videos
 02:18
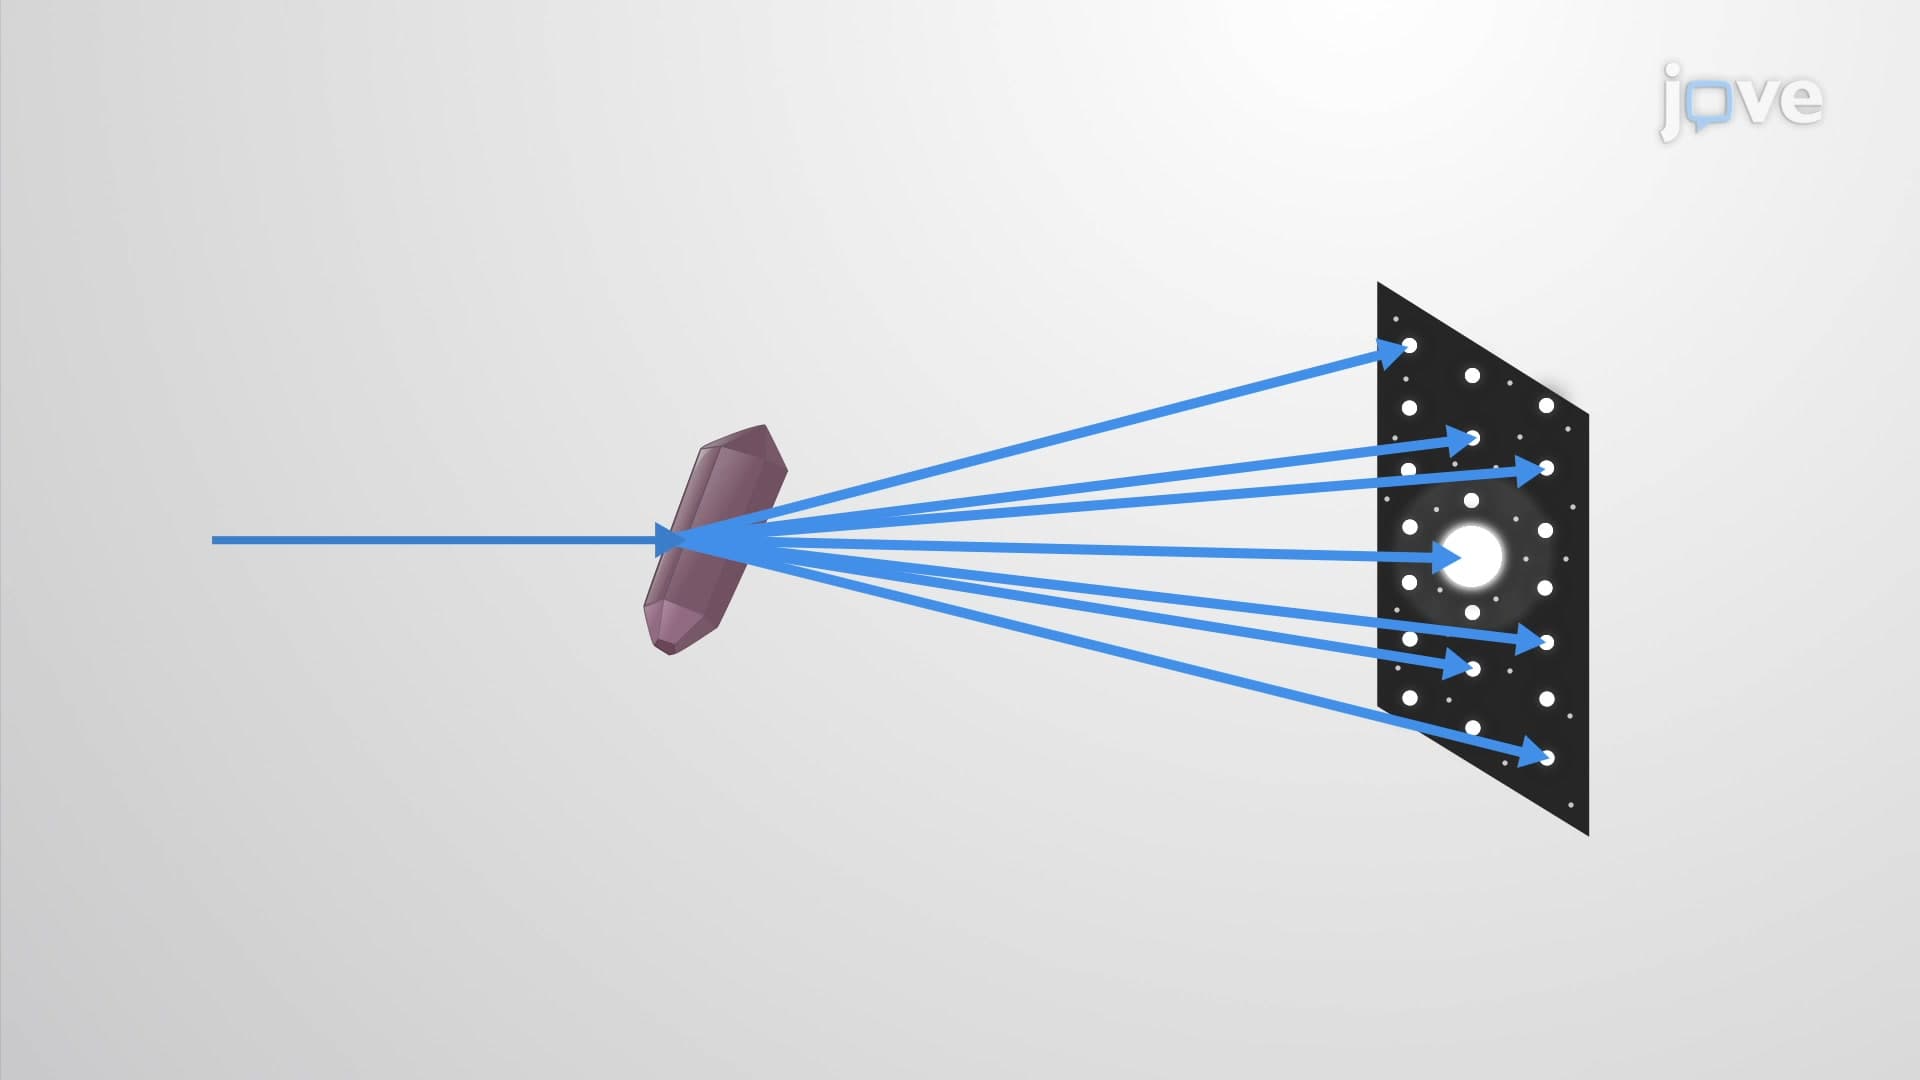
X-ray Crystallography
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 01:08
NMR Spectroscopy: Spin–Spin Coupling
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 01:16
2D NMR: Overview of Homonuclear Correlation Techniques
COSY90 is the standard two-dimensional (2D) COSY experiment that...
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:18
2D NMR: Overview of Heteronuclear Correlation Techniques