Related Experiment Video
Updated: Aug 17, 2026

A Phenotyping Regimen for Genetically Modified Mice Used to Study Genes Implicated in Human Diseases of Aging
Published on: July 14, 2016
Urinary hyaluronic acid elevation in Hutchinson-Gilford progeria syndrome

The basic genetic defect in the Hutchinson-Gilford Progeria Syndrome (progeria), a premature aging syndrome, is unknown. To investigate possible defects in hyaluronic acid (HA) metabolism in this disease, the urinary excretion of HA was studied. Urine specimens from 11 patients with this disorder were examined for HA by a novel high performance liquid chromatography (HPLC) technique. In patients with progeria, HA excretion ranged from 169 micrograms HA/g creatinine to 1440 micrograms HA/g creatinine. In normal age-matched controls, HA excreted ranged from 0 to 77 micrograms HA/g creatinine. In all, a mean 17-fold increase in HA excretion was observed in patients with progeria when compared with age-matched normal controls. Total glycosaminoglycan (GAG) excretion was not elevated. Amongst normal controls, a modest age-related increase in HA excretion was observed. These results suggest that urinary HA levels are abnormally elevated in progeria.
The basic genetic defect in the Hutchinson-Gilford Progeria Syndrome (progeria), a premature aging syndrome, is unknown. To investigate possible defects in hyaluronic acid (HA) metabolism in this disease, the urinary excretion of HA was studied. Urine specimens from 11 patients with this disorder were examined for HA by a novel high performance liquid chromatography (HPLC) technique. In patients with progeria, HA excretion ranged from 169 micrograms HA/g creatinine to 1440 micrograms HA/g creatinine. In normal age-matched controls, HA excreted ranged from 0 to 77 micrograms HA/g creatinine. In all, a mean 17-fold increase in HA excretion was observed in patients with progeria when compared with age-matched normal controls. Total glycosaminoglycan (GAG) excretion was not elevated. Amongst normal controls, a modest age-related increase in HA excretion was observed. These results suggest that urinary HA levels are abnormally elevated in progeria.
Related Concept Videos
 02:18
Alternative RNA Splicing
There are five types of alternative RNA splicing that vary in the ways the pre-mRNA segments are removed or retained in the mature mRNA. The first...
 01:22
Lysosomal Hydrolases
 01:23
Glycosaminoglycans
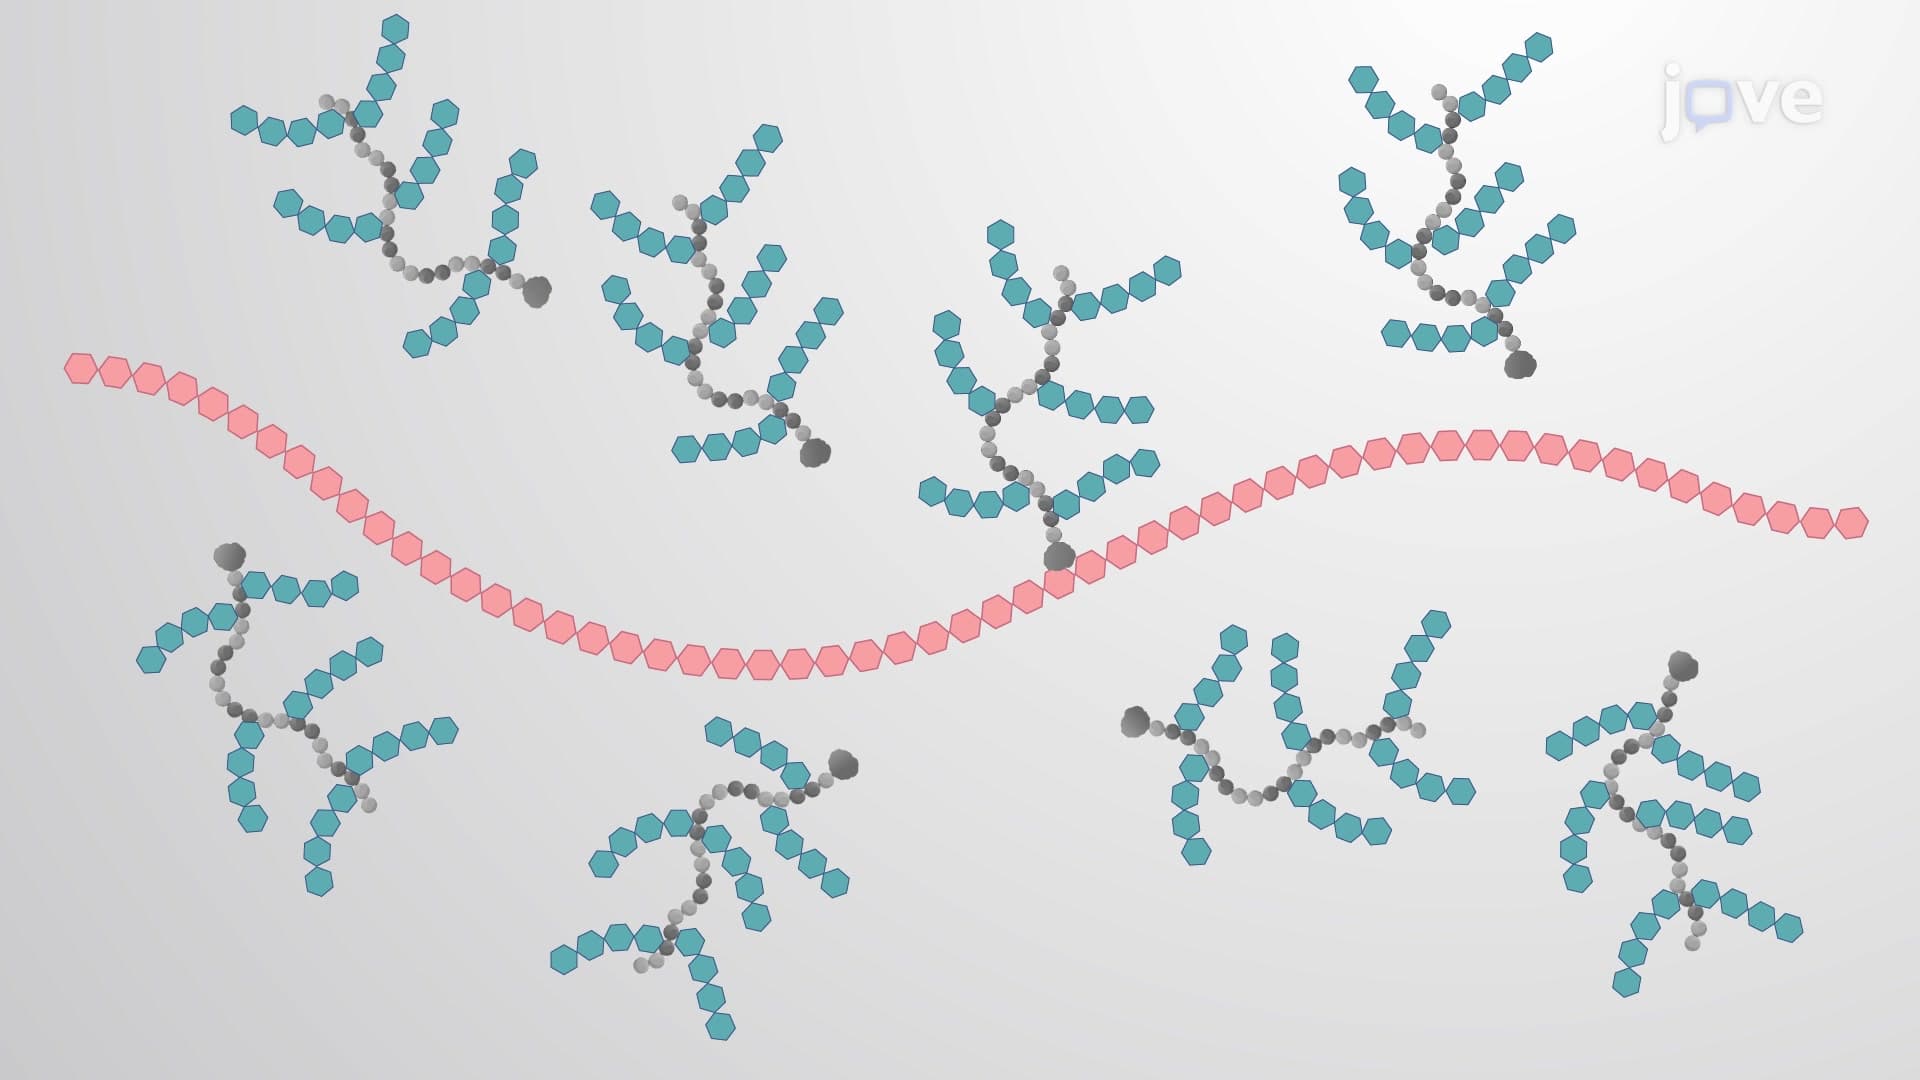
GAGS are found in the extracellular matrix of vertebrates, invertebrates, and bacteria. Due to their polar nature they attract water, and serve as excellent lubricants or shock absorbers in an animal body.
Hyaluronic...
 01:29
Cardiomyopathy III: Hypertrophic Cardiomyopathy
 01:21
Hyperosmolar Hyperglycemic State
 01:21
Huntington Disease l: Introduction