Related Experiment Video
Updated: Jul 24, 2025


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
charmm2gmx: An Automated Method to Port the CHARMM Additive Force Field to GROMACS
András F Wacha1, Justin A Lemkul2
1Institute of Materials and Environmental Chemistry, Research Centre for Natural Sciences, Eötvös Loránd Research Network, Magyar tudósok körútja 2, Budapest H-1117, Hungary.

Abstract:
CHARMM is one of the most widely used biomolecular force fields. Although developed in close connection with a dedicated molecular simulation engine of the same name, it is also usable with other codes. GROMACS is a well-established, highly optimized, and multipurpose software for molecular dynamics, versatile enough to accommodate many different force field potential functions and the associated algorithms. Due to conceptional differences related to software design and the large amount of numeric data inherent to residue topologies and parameter sets, conversion from one software format to another is not straightforward. Here, we present an automated and validated means to port the CHARMM force field to a format read by the GROMACS engine, harmonizing the different capabilities of the two codes in a self-documenting and reproducible way with a bare minimum of user interaction required. Being based entirely on the upstream data files, the presented approach does not involve any hard-coded data, in contrast with previous attempts to solve the same problem. The heuristic approach used for perceiving the local internal geometry is directly applicable for analogous transformations of other force fields.
More Related Videos
 05:00
Author Spotlight: Streamlining Visual Dynamics to Simplify Molecular Dynamics Simulations Using Gromacs
Published on: August 9, 2024
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:00
Molecular Models
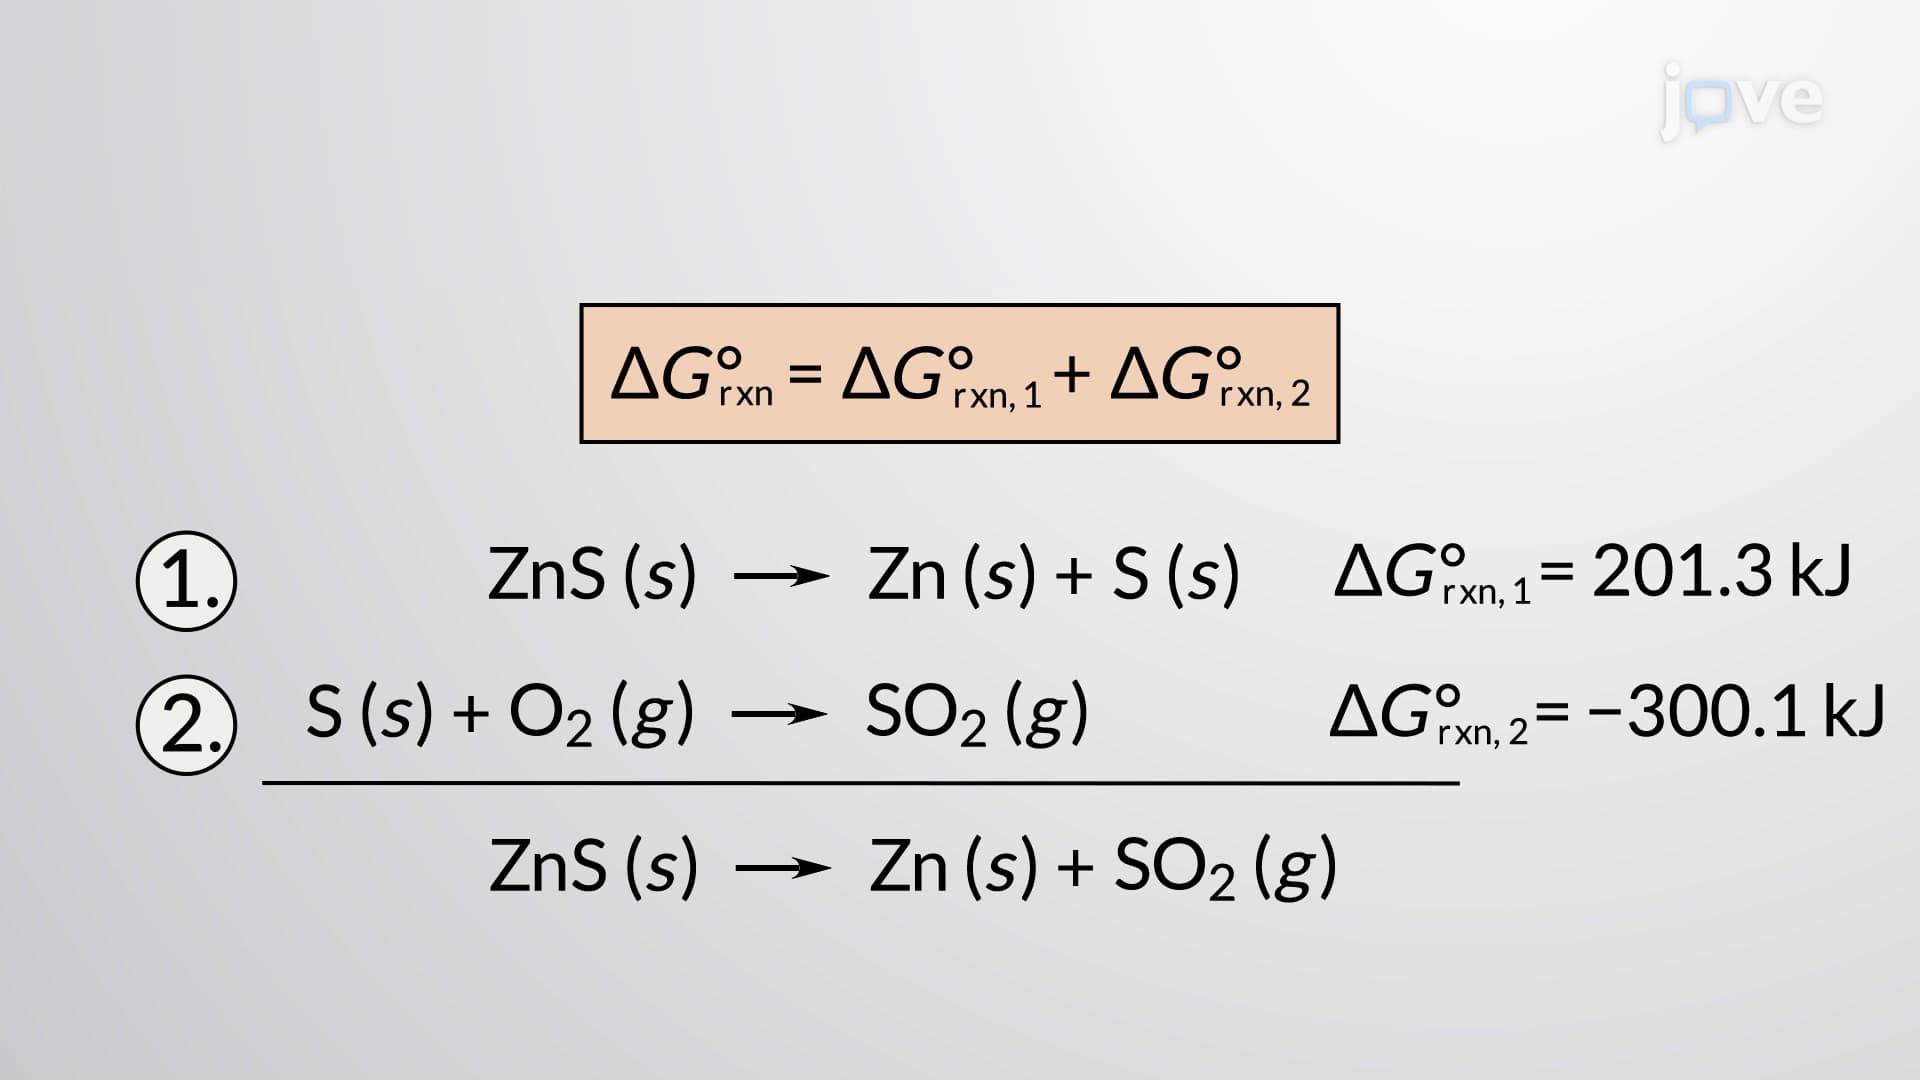 02:49
Calculating Standard Free Energy Changes
 01:06
Interfacial Electrochemical Methods: Overview
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 01:29
Mechanistic Models: Compartment Models in Algorithms for Numerical Problem Solving
In individual population analyses, different algorithms are employed, such as Cauchy's method, which uses a...