Related Experiment Video
Updated: Jul 23, 2025
Determining the Likelihood of Variant Pathogenicity Using Amino Acid-level Signal-to-Noise Analysis of Genetic Variation
Published on: January 16, 2019
Paired associated SARS-CoV-2 spike variable positions: a network analysis approach to emerging variants
Yiannis Manoussopoulos1,2, Cleo Anastassopoulou1, John P A Ioannidis3,4,5,6
1Department of Microbiology, Medical School, National and Kapodistrian University of Athens , Athens, Greece.

Abstract:
Amino acids in variable positions of proteins may be correlated, with potential structural and functional implications. Here, we apply exact tests of independence in R × C contingency tables to examine noise-free associations between variable positions of the SARS-CoV-2 spike protein, using as a paradigm sequences from Greece deposited in GISAID (N = 6,683/1,078 full length) for the period 29 February 2020 to 26 April 2021 that essentially covers the first three pandemic waves. We examine the fate and complexity of these associations by network analysis, using associated positions (exact P ≤ 0.001 and Average Product Correction ≥ 2) as links and the corresponding positions as nodes. We found a temporal linear increase of positional differences and a gradual expansion of the number of position associations over time, represented by a temporally evolving intricate web, resulting in a non-random complex network of 69 nodes and 252 links. Overconnected nodes corresponded to the most adapted variant positions in the population, suggesting a direct relation between network degree and position functional importance. Modular analysis revealed 25 k-cliques comprising 3 to 11 nodes. At different k-clique resolutions, one to four communities were formed, capturing epistatic associations of circulating variants (Alpha, Beta, B.1.1.318), but also Delta, which dominated the evolutionary landscape later in the pandemic. Cliques of aminoacidic positional associations tended to occur in single sequences, enabling the recognition of epistatic positions in real-world virus populations. Our findings provide a novel way of understanding epistatic relationships in viral proteins with potential applications in the design of virus control procedures. IMPORTANCE Paired positional associations of adapted amino acids in virus proteins may provide new insights for understanding virus evolution and variant formation. We investigated potential intramolecular relationships between variable SARS-CoV-2 spike positions by exact tests of independence in R × C contingency tables, having applied Average Product Correction (APC) to eliminate background noise. Associated positions (exact P ≤ 0.001 and APC ≥ 2) formed a non-random, epistatic network of 25 cliques and 1-4 communities at different clique resolutions, revealing evolutionary ties between variable positions of circulating variants and a predictive potential of previously unknown network positions. Cliques of different sizes represented theoretical combinations of changing residues in sequence space, allowing the identification of significant aminoacidic combinations in single sequences of real-world populations. Our analytic approach that links network structural aspects to mutational aminoacidic combinations in the spike sequence population offers a novel way to understand virus epidemiology and evolution.
More Related Videos
Related Concept Videos
 01:05
Single Nucleotide Polymorphisms-SNPs
 00:36
Viral Mutations
 01:26
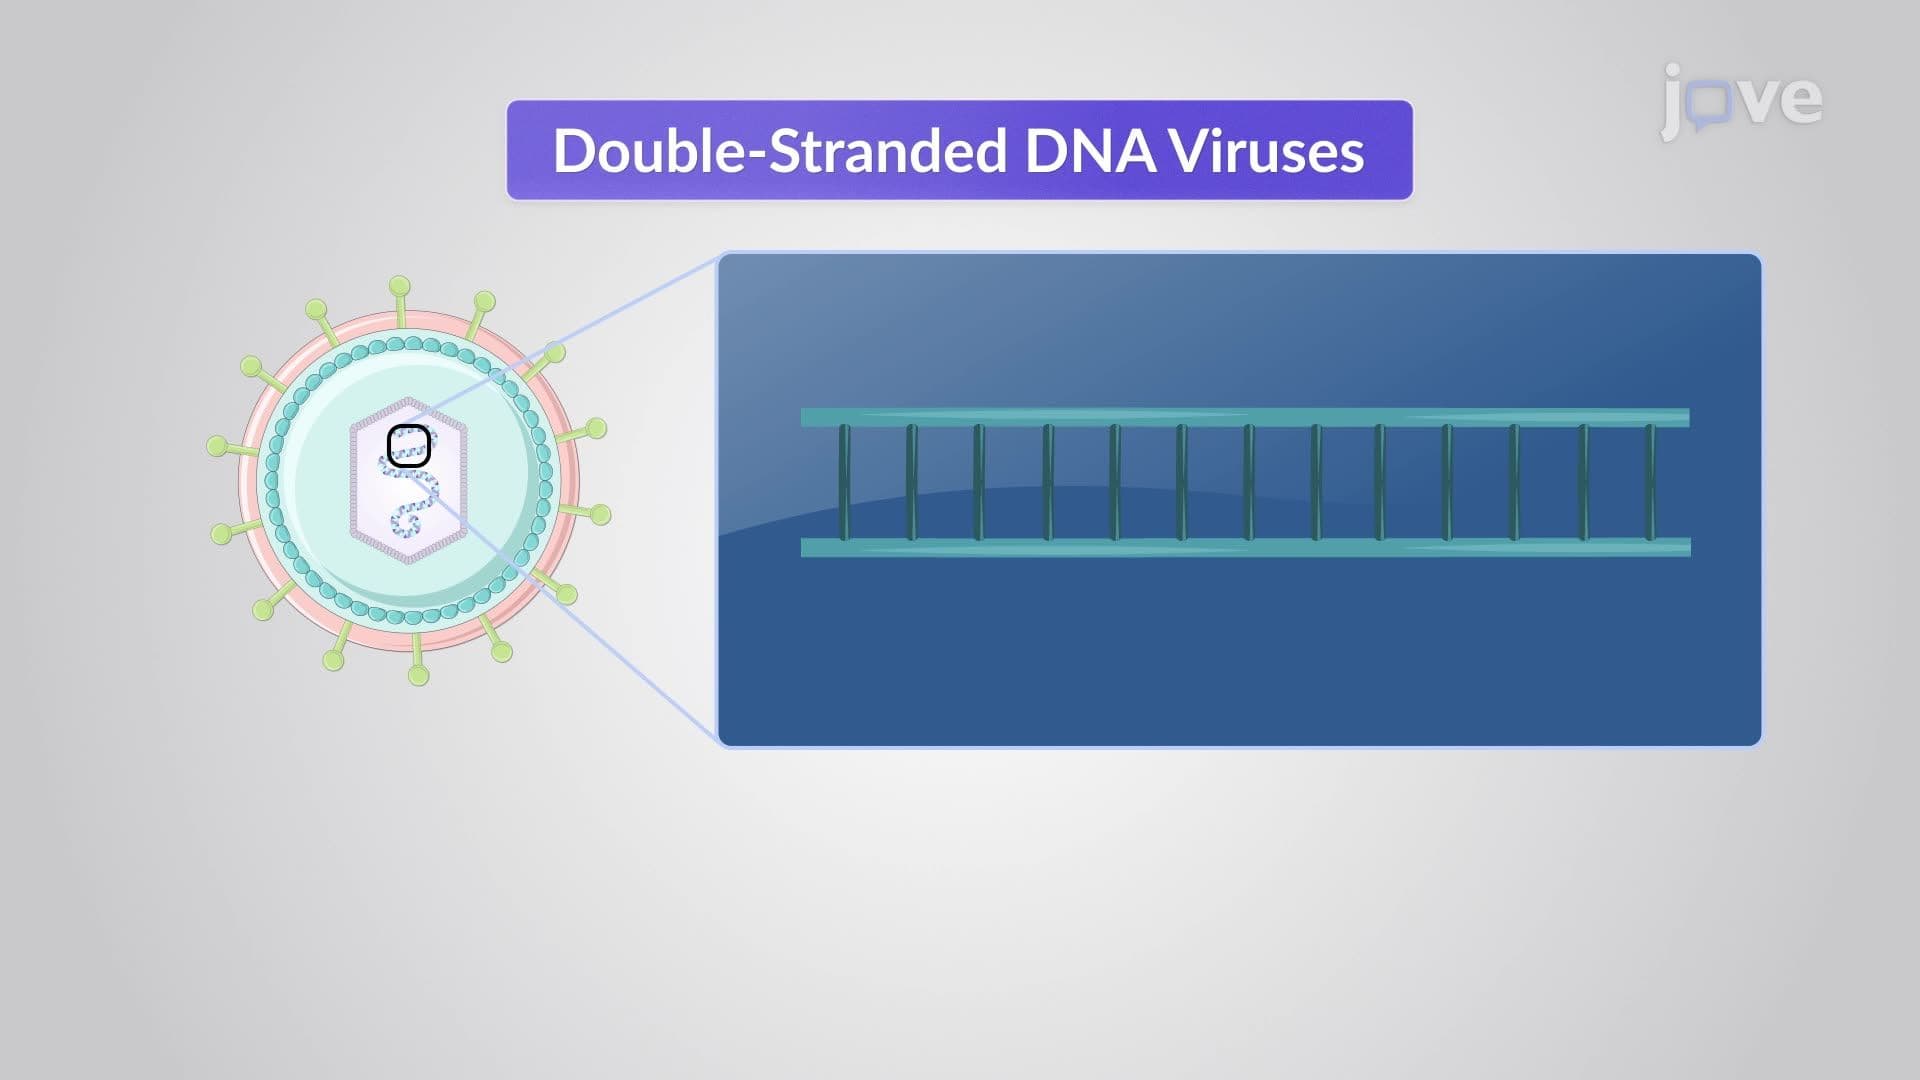
Size and Structure of Viral Genomes
 01:11
Variability: Analysis
The range is a simple measure of variability, indicating the difference between the highest and...
 02:26
Comparing Copy Number Variations and SNPs
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
 00:57
Viral Recombination