A quantitative sequencing method using synthetic internal standards including functional and phylogenetic marker genes
Related Experiment Video
Updated: Jul 23, 2025
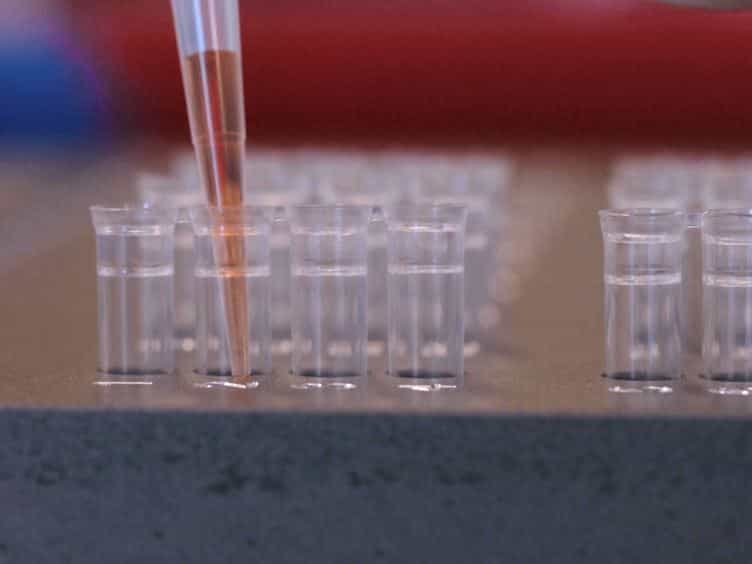
Pyrosequencing for Microbial Identification and Characterization
Published on: August 22, 2013
Kazuyoshi Koike1, Ryo Honda2, Masataka Aoki3
1Graduate School of Natural Science and Technology, Kanazawa University, Kanazawa, Japan.

Abstract:
The method of spiking synthetic internal standard genes (ISGs) to samples for amplicon sequencing, generating sequences and converting absolute gene numbers from read counts has been used only for phylogenetic markers and has not been applied to functional markers. In this study, we developed ISGs, including gene sequences of the 16S rRNA, pmoA, encoding a subunit of particulate methane monooxygenase and amoA, encoding a subunit of ammonia monooxygenase. We added ISGs to the samples, amplified the target genes and performed amplicon sequencing. For the mock community, the copy numbers converted from read counts using ISGs were equivalent to those obtained by the quantitative real-time polymerase chain reaction (4.0 × 10⁴ versus 4.1 × 10⁴ and 3.0 × 10³ versus 4.0 × 10³ copies μL-DNA-1 for 16S rRNA and pmoA genes, respectively), but we also identified underestimation, possibly due to primer coverage (7.8 × 10² versus 3.7 × 10³ μL-DNA-1 for amoA gene). We then applied this method to environmental samples and analysed phylogeny, functional diversity and absolute quantities. One Methylocystis population was most abundant in the sludge samples [16S rRNA gene (3.8 × 10⁹ copies g-1 ) and the pmoA gene (2.3 × 10⁹ copies g-1 )] and were potentially interrelated. This study demonstrates that ISG spiking is useful for evaluating sequencing data processing and quantifying functional markers.
Related Concept Videos
 03:00
Next-generation Sequencing
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
 03:21
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
 01:57
Sanger Sequencing
 01:29
Modern Molecular Taxonomy
 02:54
Evolutionary Relationships through Genome Comparisons