Related Experiment Video
Updated: Jul 22, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Comparison of Eight Classical Lennard-Jones-Based H2 Molecular Models in the Gas Phase at Temperatures and Pressures
Méryll Barraco1,2,3, Sylvie Neyertz1, Nieck E Benes2
1Université Savoie Mont Blanc, Université Grenoble Alpes, CNRS, Grenoble INP, LEPMI, 38000 Grenoble, France.

Abstract:
This work compares eight classical H2 molecular models in the gas phase taken from the existing literature. All models are based on Lennard-Jones (LJ) 12-6 terms for the van der Waals interactions and hence easier to transfer to multiphase molecular simulations than more sophisticated potentials. The H2 potentials tested include one-site, two-site, three-site, and five-site models, with the sites being either the H atoms, the center-of-mass of the H2 molecule, or massless sites. For the multisite models, high-frequency H-H stretching modes can lead to poor equipartition of the kinetic energy, and the timestep for molecular dynamics (MD) simulations should be reduced to maintain a stable numerical integration of the equations of motion. As such, only those models with rigid bonds are considered. In the present case, 600 MD simulations of H2 gas were carried out over a large range of temperatures (-50 to +90 °C) and at densities corresponding to a pressure range of 50 to 2000 bar, which include the operating conditions of on-board storage tanks in hydrogen-fueled vehicles. Most of the models under study were found to reproduce reasonably well the experimental pVT phase diagram as well as the solubility. Discrepancies only became significant at the highest densities tested, and these could be used to rank the different models. All model diffusion coefficients were essentially indistinguishable from experimental results, and as such, kinetically dominated dynamic properties could not be used as a criterion for the choice of model. Among the eight models tested, two of them, i.e., the two-site model of Yang and Zhong and the one-site model derived from Buch performed very well over the range of conditions tested. They represent a good compromise between realism, simplicity, and computational efficiency.
More Related Videos
 05:46
High-pressure Sapphire Cell for Phase Equilibria Measurements of CO2/Organic/Water Systems
Published on: January 24, 2014
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:26
Molecular Comparison of Gases, Liquids, and Solids
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 01:10
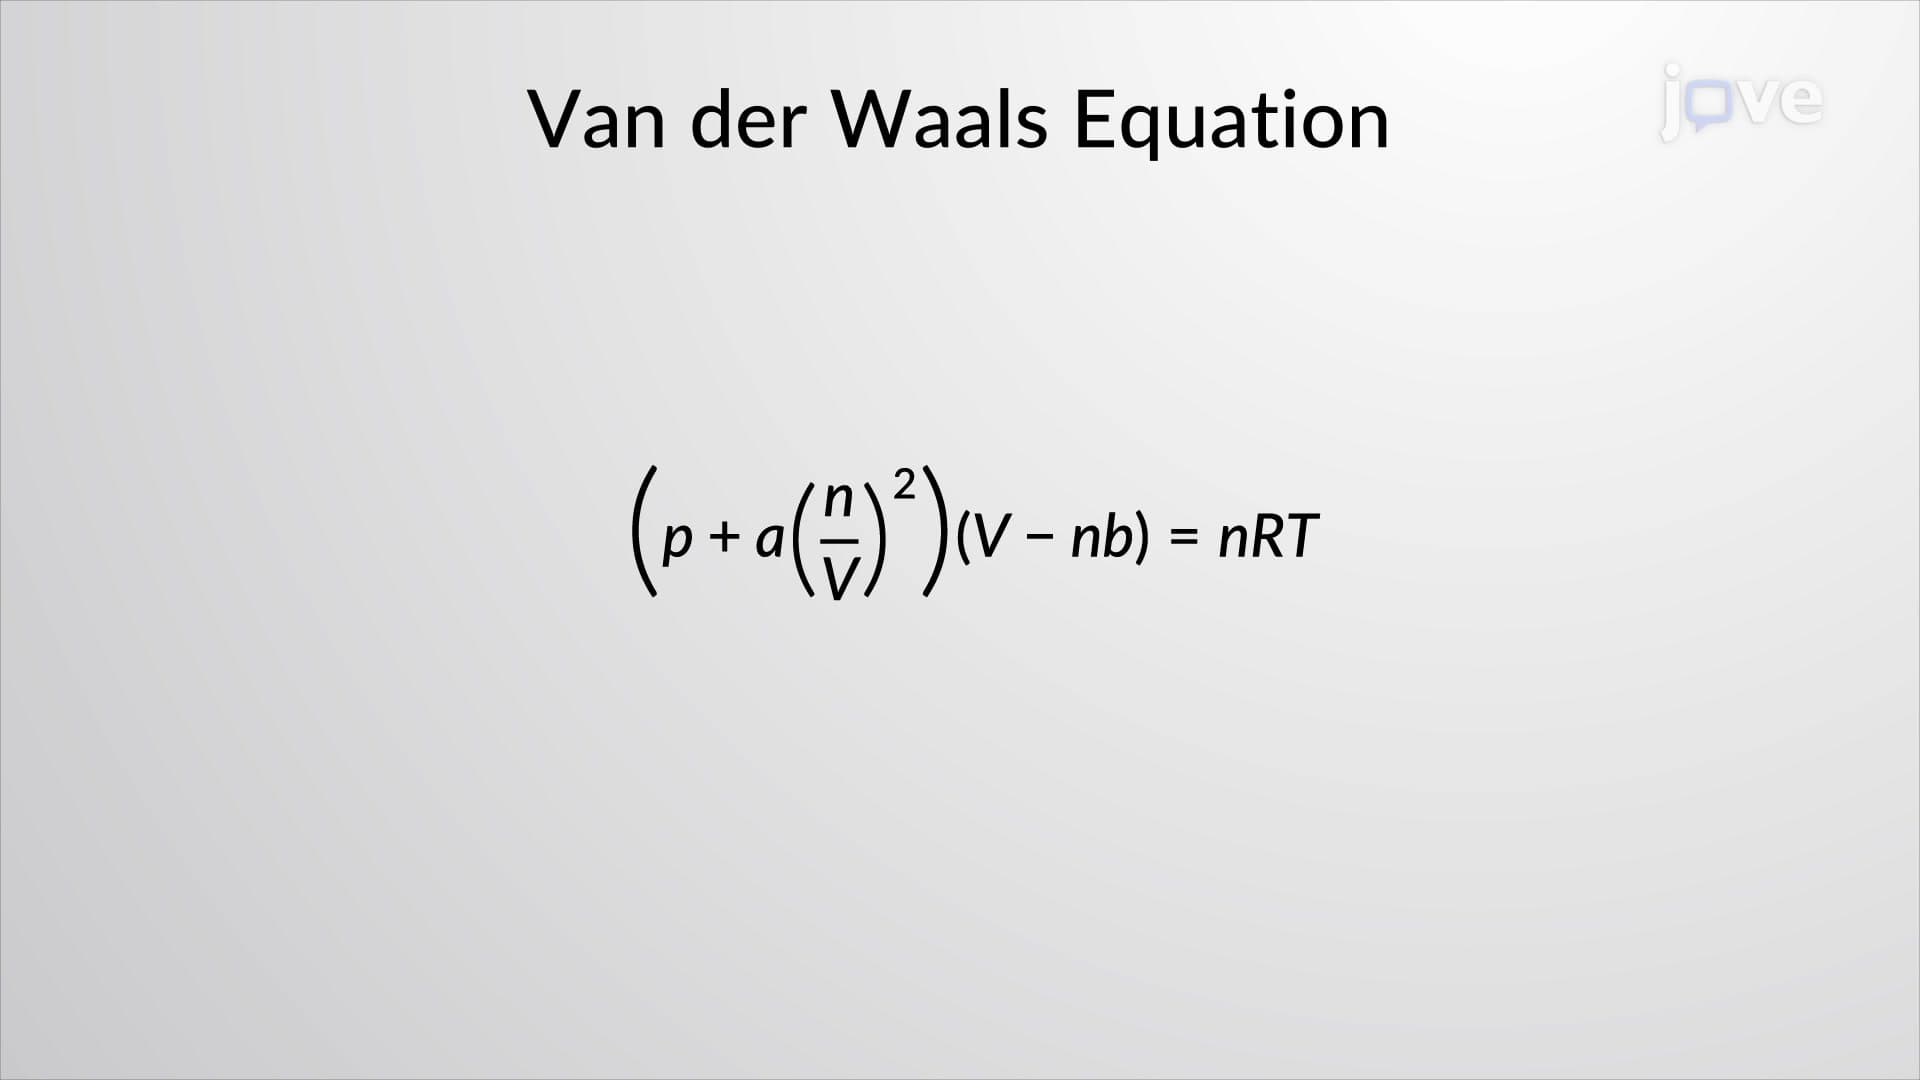
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 02:34
Comparing Intermolecular Forces: Melting Point, Boiling Point, and Miscibility
Temporary attractive forces like dispersion are present in all molecules, whether they are polar or nonpolar. They...
 03:19
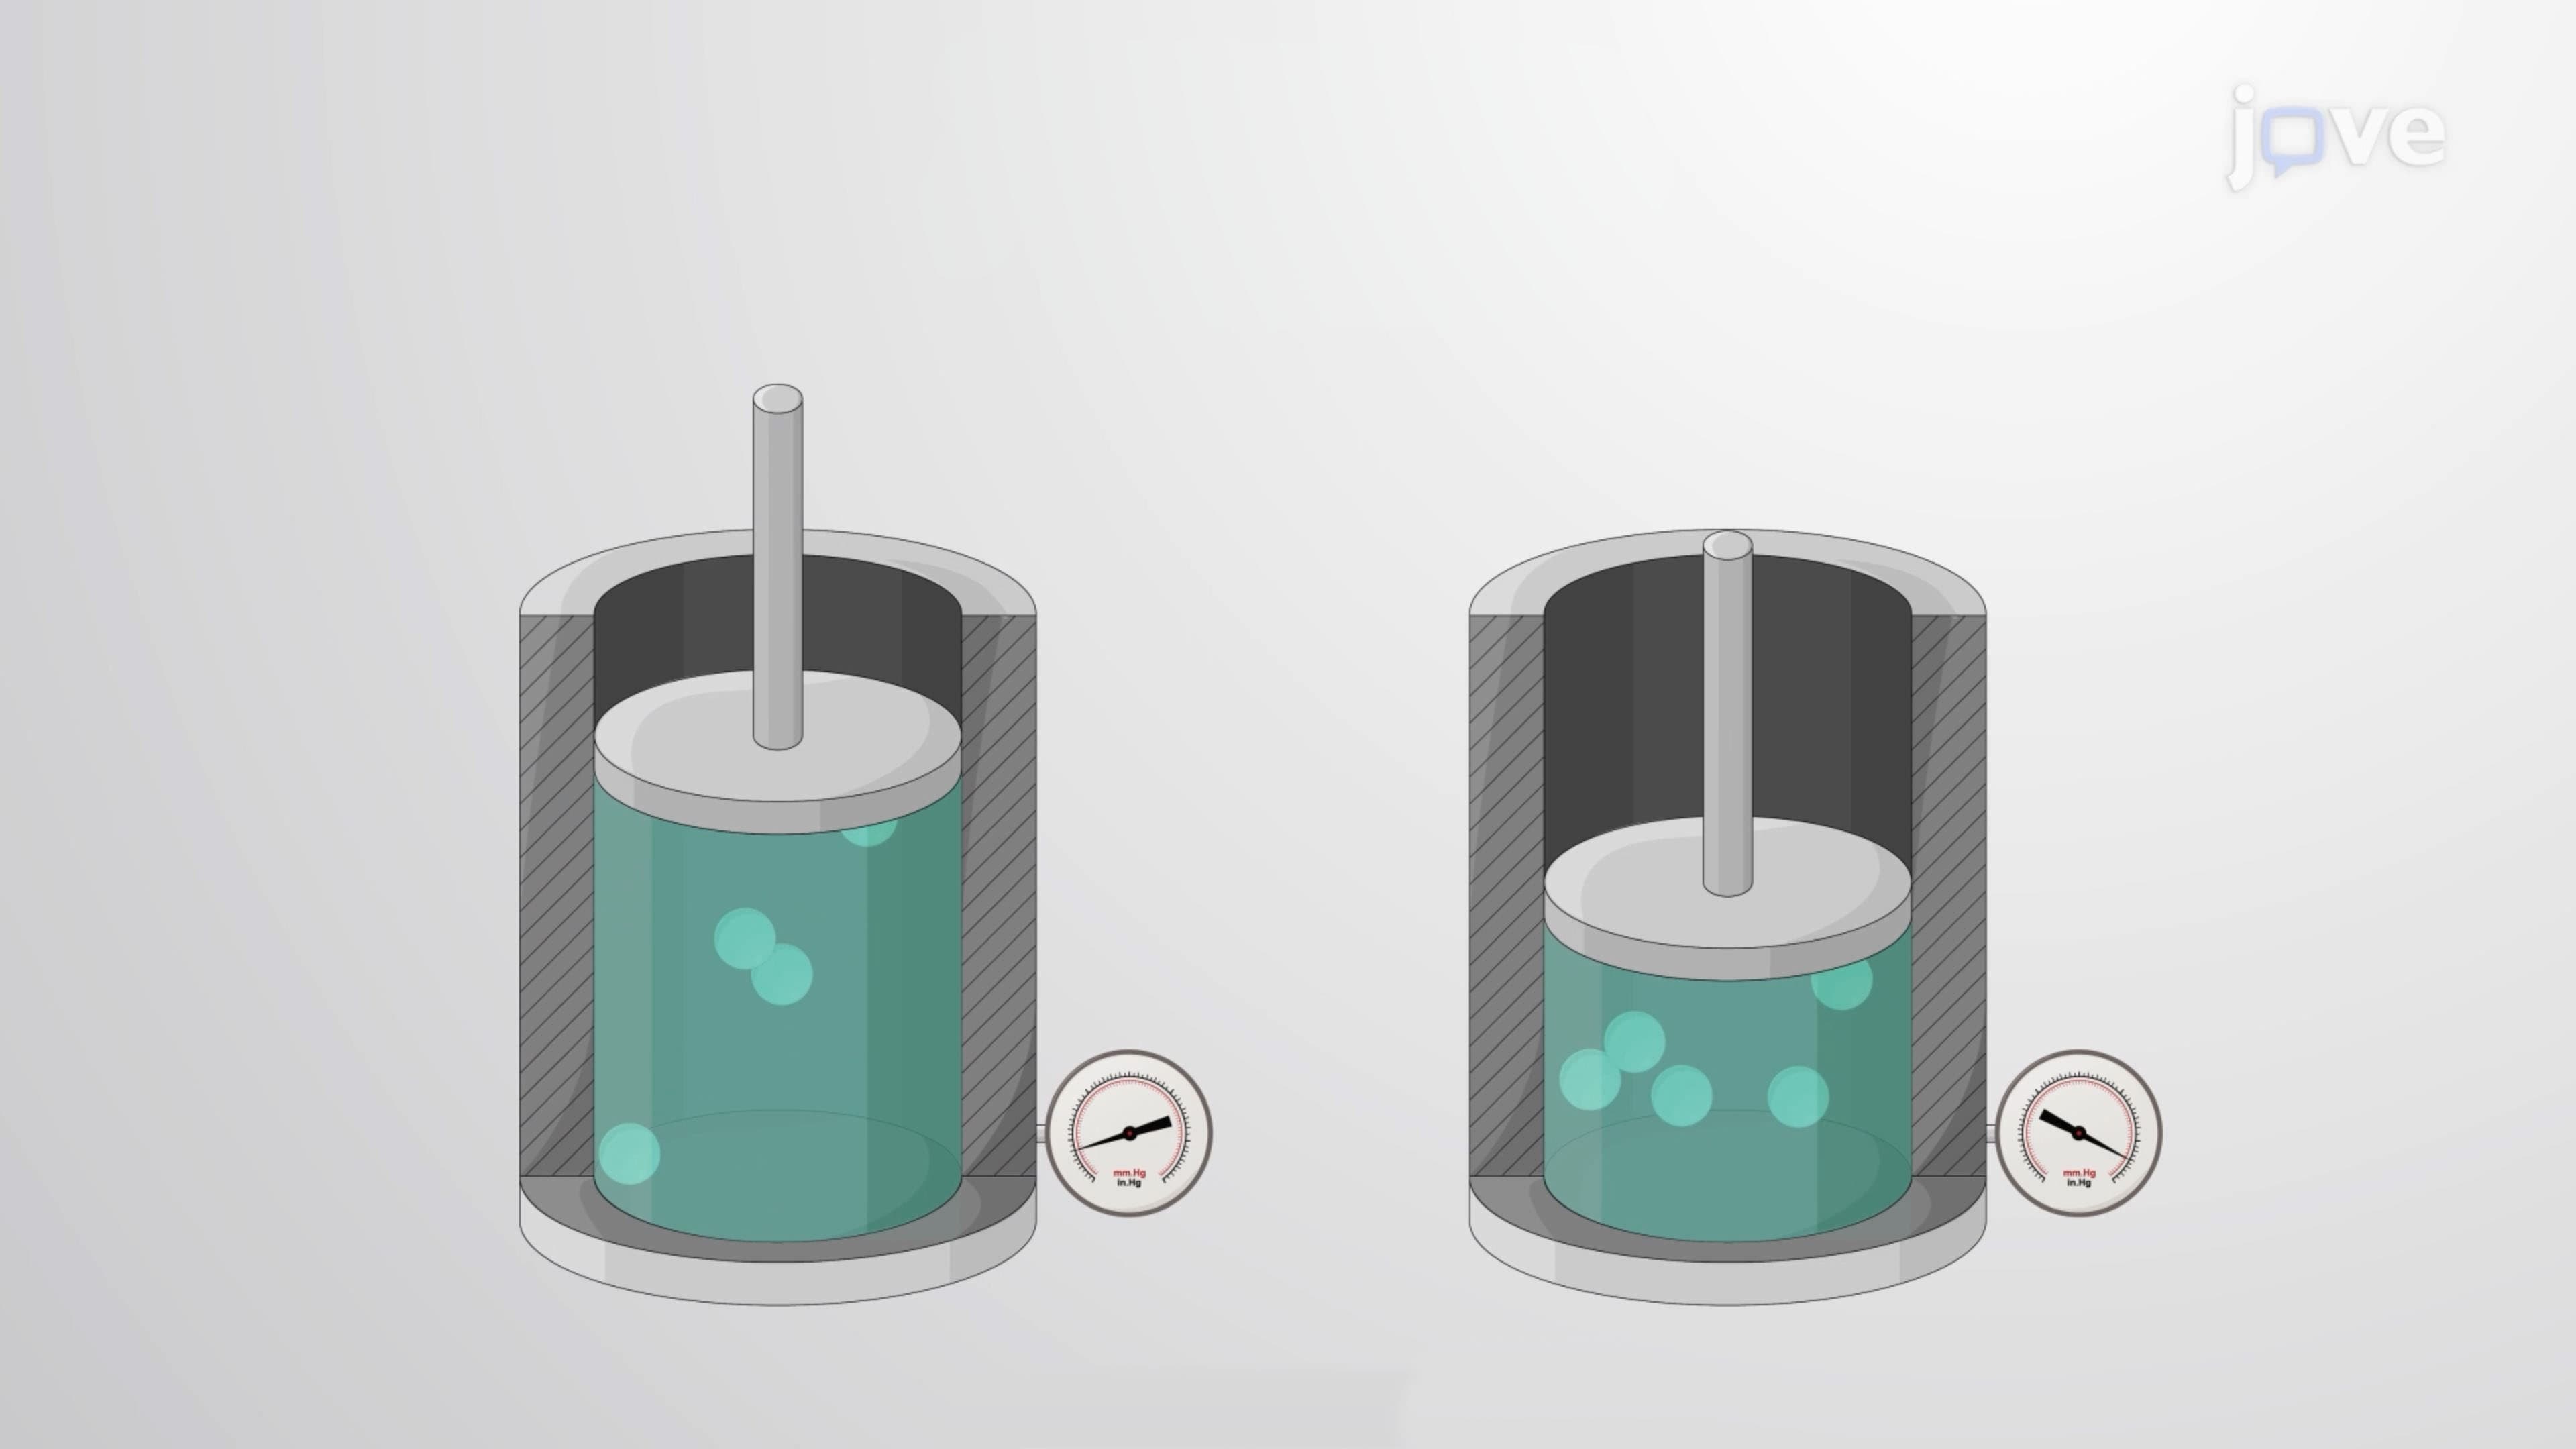
Gas Laws: Boyle's, Gay-Lussac, Charles', Avogadro's, and Ideal Gas Law
 01:25
Heat Capacities of an Ideal Gas III