Related Experiment Video
Updated: Jul 22, 2025


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Multi-combined QSAR, molecular docking, molecular dynamics simulation, and ADMET of Flavonoid derivatives as potent
Hatim Soufi1, Mohamed Moussaoui1, Soukayna Baammi2
1Laboratory of Physical Chemistry of Materials, Faculty of Sciences Ben M'Sick, Hassan II University of Casablanca, Benguerir, Morocco.

Abstract:
In searching for a new and efficient therapeutic agent against Alzheimer's disease, a Quantitative structure-activity relationship (QSAR) was derived for 45 Flavonoid derivatives recently synthesized and evaluated as cholinesterase inhibitors. The multiple linear regression method (MLR) was adopted to develop an adequate mathematical model that describes the relationship between a variety of molecular descriptors of the studied compounds and their biological activities (cholinesterase inhibitors). Golbraikh and Tropsha criteria were applied to verify the validity of the built model. The built MLR model was statistically reliable, robust, and predictive (R2 = 0.801, Q2cv = 0.876, R2test = 0.824). Dreiding energy and Molar Refractivity were the major factors that govern the Anti-cholinesterase activity. These results were further exploited to design a new series of Flavonoid derivatives with higher Anti-cholinesterase activities than the existing ones. Thereafter, molecular docking and molecular dynamic studies were performed to predict the binding types of the designed compounds and to investigate their stability at the active site of the Butyrylcholinestérase BuChE protein. The negative and low binding affinity calculated for all designed compounds shows that designed compound 1 has a favorable affinity for the 4TPK. Moreover, molecular dynamics simulation studies confirmed the stability of designed compound 1 in the active pocket of 4TPK over 100 ns. Finally, the ADMET analysis was incorporated to analyze the pharmacokinetics and toxicity parameters. The designed compounds were found to meet the ADMET descriptor criteria at an acceptable level having respectable intestinal permeability and water solubility and can reach the intended destinations.Communicated by Ramaswamy H. Sarma.
More Related Videos
 10:25
Screening Traditional Chinese Medicine Compounds for Inhibiting UCHL3 Activity Based on Molecular Docking and Deubiquitinating Enzyme Probe Technology
Published on: November 22, 2024
 00:05
In Silico Modeling Method for Computational Aquatic Toxicology of Endocrine Disruptors: A Software-Based Approach Using QSAR Toolbox
Published on: August 28, 2019
Related Concept Videos
 01:29
Indirect-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
Reversible inhibitors display short to medium durations of action. Short-acting agents include simple alcohols with...
 01:18
Indirect-Acting Cholinergic Agonists: Mechanism of Action
Reversible inhibitors like edrophonium bind to a specific part of the enzyme called the anionic catalytic site. They form noncovalent bonds, which means they are not strongly attached to the enzyme. This creates a temporary and less stable enzyme–inhibitor complex,...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:31
Direct-Acting Cholinergic Agonists: Pharmacokinetics
 01:22
Direct-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
The direct-acting...
 01:29
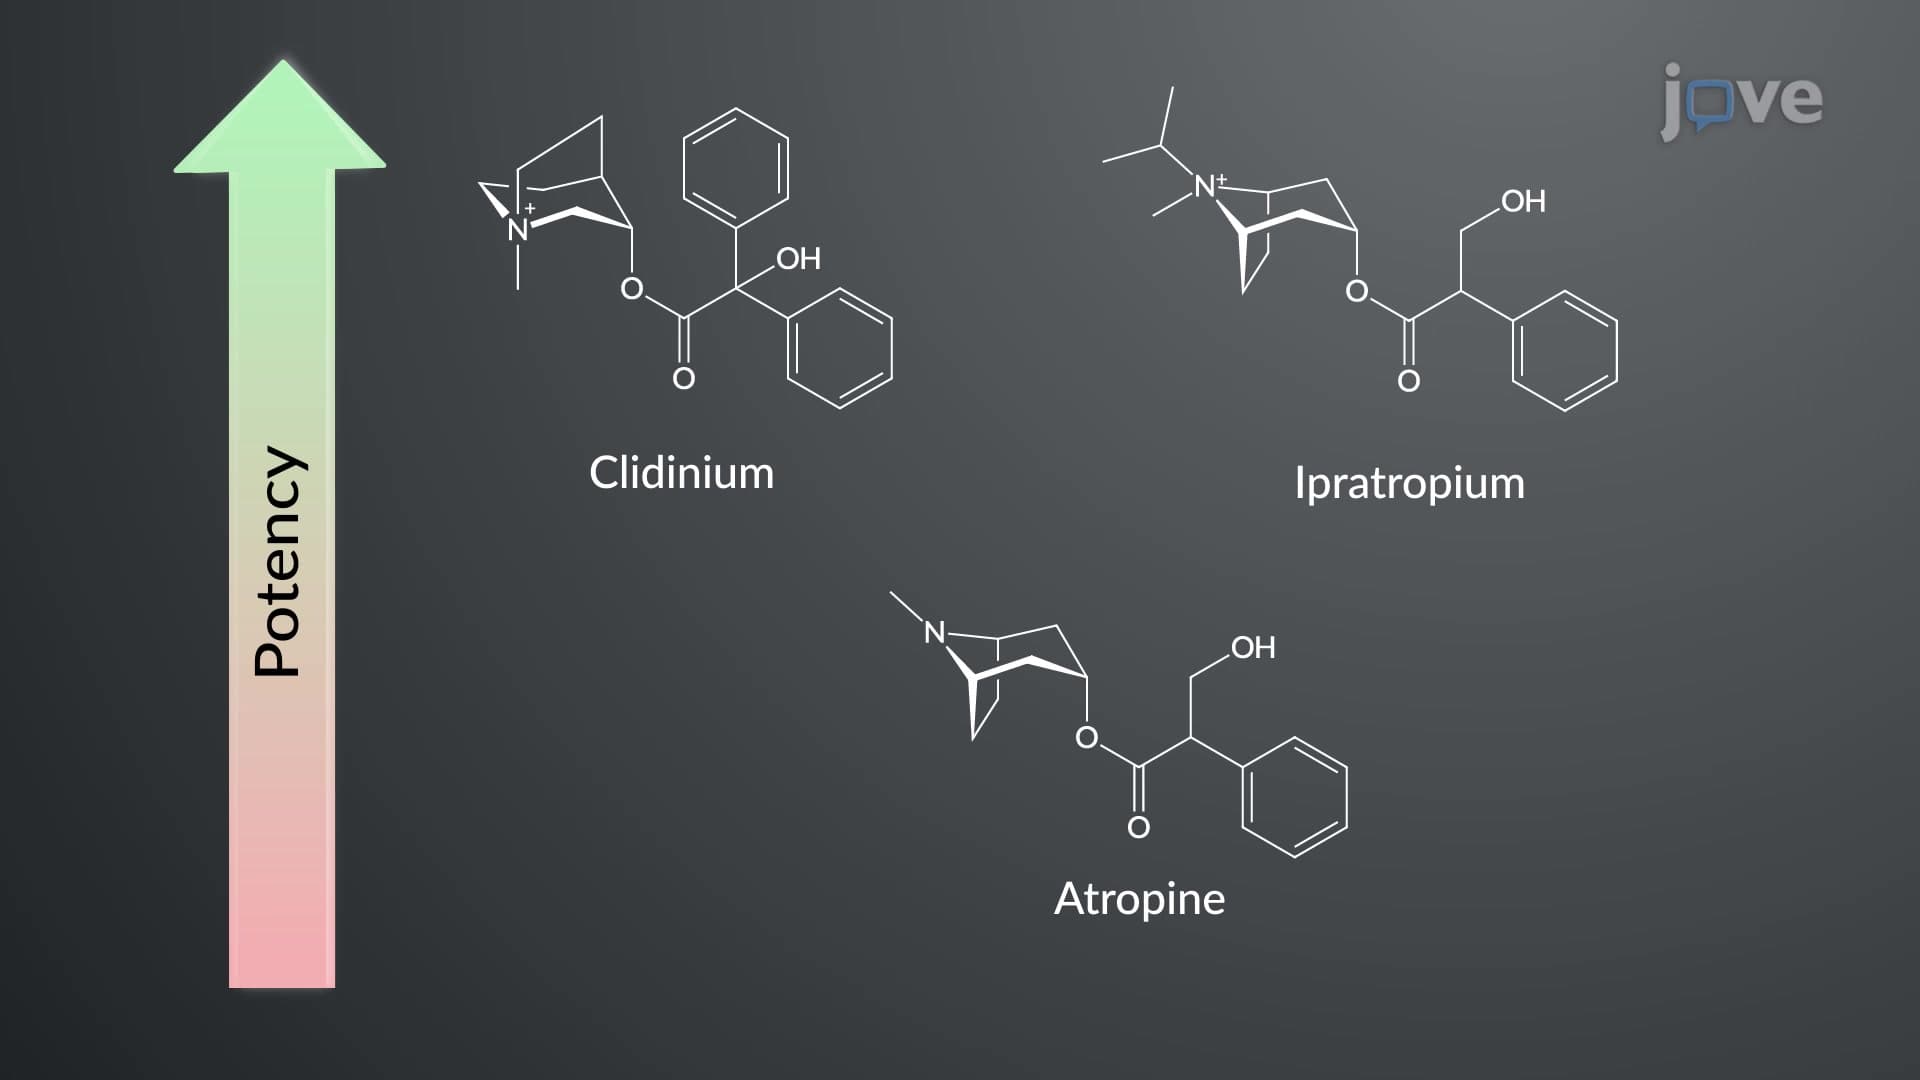
Cholinergic Antagonists: Chemistry and Structure-Activity Relationship