Related Experiment Video
Updated: Jul 21, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Partitioning Quantum Chemistry Simulations with Clifford Circuits
Philipp Schleich1,2,3, Joseph Boen3,4, Lukasz Cincio3
1Department of Computer Science, University of Toronto, Toronto M5S 1A1, Canada.

Abstract:
Current quantum computing hardware is restricted by the availability of only few, noisy qubits which limits the investigation of larger, more complex molecules in quantum chemistry calculations on quantum computers in the near term. In this work, we investigate the limits of their classical and near-classical treatment while staying within the framework of quantum circuits and the variational quantum eigensolver. To this end, we consider naive and physically motivated, classically efficient product ansatz for the parametrized wavefunction adapting the separable-pair ansatz form. We combine it with post-treatment to account for interactions between subsystems originating from this ansatz. The classical treatment is given by another quantum circuit that has support between the enforced subsystems and is folded into the Hamiltonian. To avoid an exponential increase in the number of Hamiltonian terms, the entangling operations are constructed from purely Clifford or near-Clifford circuits. While Clifford circuits can be simulated efficiently classically, they are not universal. In order to account for missing expressibility, near-Clifford circuits with only few, selected non-Clifford gates are employed. The exact circuit structure to achieve this objective is molecule-dependent and is constructed using simulated annealing and genetic algorithms. We demonstrate our approach on a set of molecules of interest and investigate the extent of our methodology's reach.
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:52
VSEPR Theory and the Basic Shapes
 01:29
Mechanistic Models: Compartment Models in Algorithms for Numerical Problem Solving
In individual population analyses, different algorithms are employed, such as Cauchy's method, which uses a...
 01:08
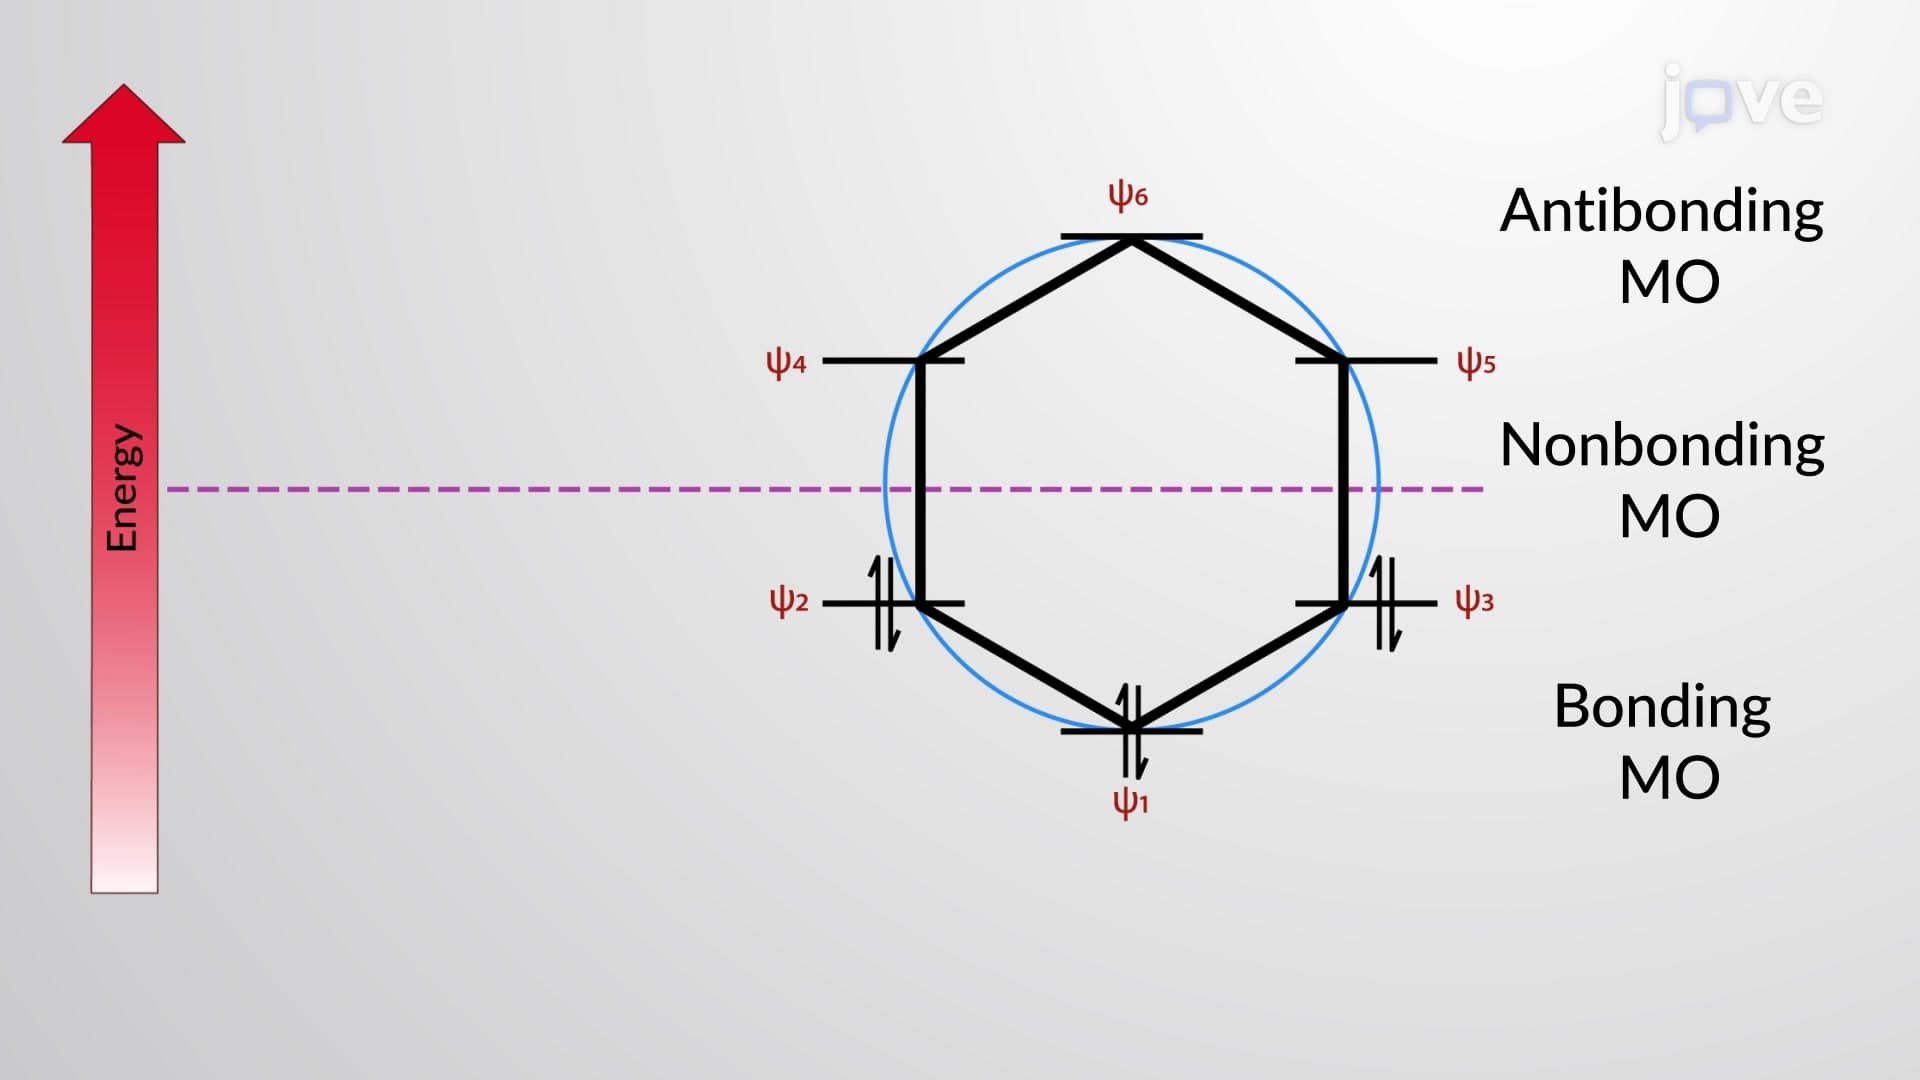
Hückel's Rule Diagram of π MOs: Frost Circle
A Frost circle is constructed by drawing a polygon whose number of edges is equal to the number of carbons of the given cyclic system, with one of the vertices pointing down. Then, a circle is drawn enclosing the polygon so...
 01:27
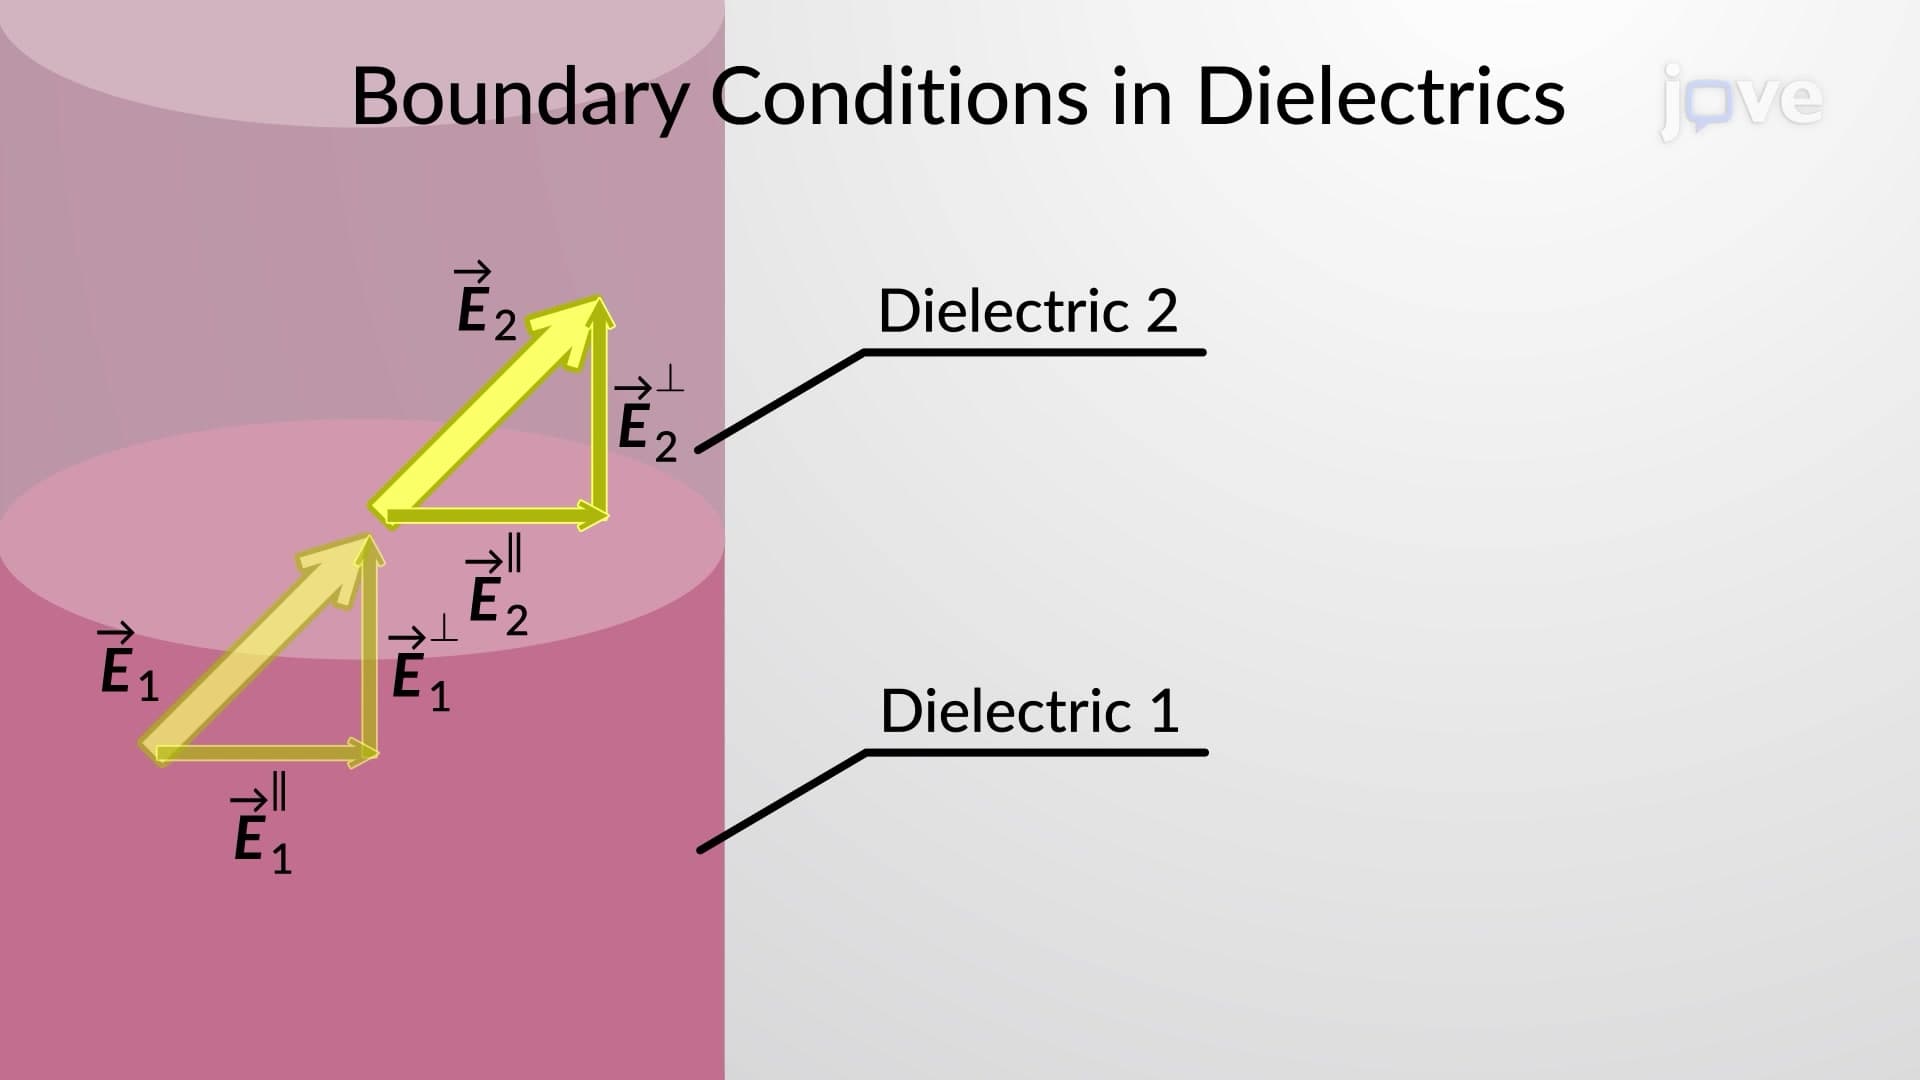
Electrostatic Boundary Conditions in Dielectrics
Consider a case where both the mediums across a boundary are two different dielectric materials. Recall that the electric field and electric displacement are proportional and related through the material's...