Related Experiment Video
Updated: Jul 21, 2025
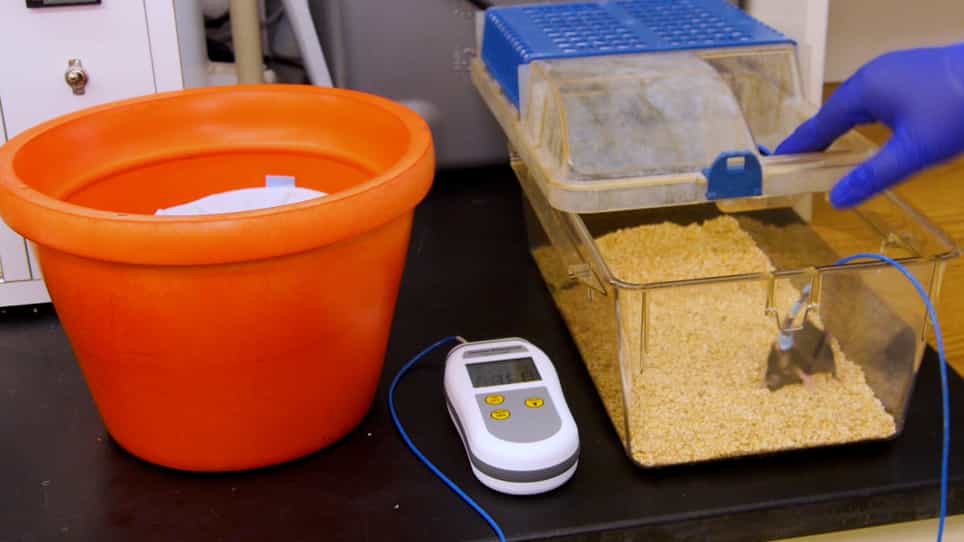

A Behavioral Screen for Heat-Induced Seizures in Mouse Models of Epilepsy
Published on: July 12, 2021
Patient-derived SLC6A1 variant S295L results in an epileptic phenotype similar to haploinsufficient mice
Britta E Lindquist1,2, Yuliya Voskobiynyk1,2, Kimberly Goodspeed3
1Gladstone Institute of Neurological Disease, Gladstone Institutes, San Francisco, California, USA.

Abstract:
The solute carrier family 6 member 1 (SLC6A1) gene encodes GAT-1, a γ-aminobutyric acid transporter expressed on astrocytes and inhibitory neurons. Mutations in SLC6A1 are associated with epilepsy and developmental disorders, including motor and social impairments, but variant-specific animal models are needed to elucidate mechanisms. Here, we report electrocorticographic (ECoG) recordings and clinical data from a patient with a variant in SLC6A1 that encodes GAT-1 with a serine-to-leucine substitution at amino acid 295 (S295L), who was diagnosed with childhood absence epilepsy. Next, we show that mice bearing the S295L mutation (GAT-1S295L/+ ) have spike-and-wave discharges with motor arrest consistent with absence-type seizures, similar to GAT-1+/- mice. GAT-1S295L/+ and GAT-1+/- mice follow the same pattern of pharmacosensitivity, being bidirectionally modulated by ethosuximide (200 mg/kg ip) and the GAT-1 antagonist NO-711 (10 mg/kg ip). By contrast, GAT-1-/- mice were insensitive to both ethosuximide and NO-711 at the doses tested. In conclusion, ECoG findings in GAT-1S295L/+ mice phenocopy GAT-1 haploinsufficiency and provide a useful preclinical model for drug screening and gene therapy investigations.
More Related Videos
 08:22
A Novel Strategy Combining Array-CGH, Whole-exome Sequencing and In Utero Electroporation in Rodents to Identify Causative Genes for Brain Malformations
Published on: December 1, 2017
 09:08
In Vivo Fiber-Coupled Pre-Clinical Confocal Laser-scanning Endomicroscopy pCLE of Hippocampal Capillaries in Awake Mice
Published on: April 21, 2023
