Related Experiment Video
Updated: Jul 21, 2025


Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
PyL3dMD: Python LAMMPS 3D molecular descriptors package
Pawan Panwar1, Quanpeng Yang2, Ashlie Martini3
1Department of Mechanical Engineering, University of California Merced, 5200 North Lake Road, Merced, CA, 95343, USA. ppanwar@ucmerced.edu.

Abstract:
Molecular descriptors characterize the biological, physical, and chemical properties of molecules and have long been used for understanding molecular interactions and facilitating materials design. Some of the most robust descriptors are derived from geometrical representations of molecules, called 3-dimensional (3D) descriptors. When calculated from molecular dynamics (MD) simulation trajectories, 3D descriptors can also capture the effects of operating conditions such as temperature or pressure. However, extracting 3D descriptors from MD trajectories is non-trivial, which hinders their wide use by researchers developing advanced quantitative-structure-property-relationship models using machine learning. Here, we describe a suite of open-source Python-based post-processing routines, called PyL3dMD, for calculating 3D descriptors from MD simulations. PyL3dMD is compatible with the popular simulation package LAMMPS and enables users to compute more than 2000 3D molecular descriptors from atomic trajectories generated by MD simulations. PyL3dMD is freely available via GitHub and can be easily installed and used as a highly flexible Python package on all major platforms (Windows, Linux, and macOS). A performance benchmark study used descriptors calculated by PyL3dMD to develop a neural network and the results showed that PyL3dMD is fast and efficient in calculating descriptors for large and complex molecular systems with long simulation durations. PyL3dMD facilitates the calculation of 3D molecular descriptors using MD simulations, making it a valuable tool for cheminformatics studies.
Related Concept Videos
 02:36
Molecular Geometry and Dipole Moments
 02:27
Predicting Molecular Geometry
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:08
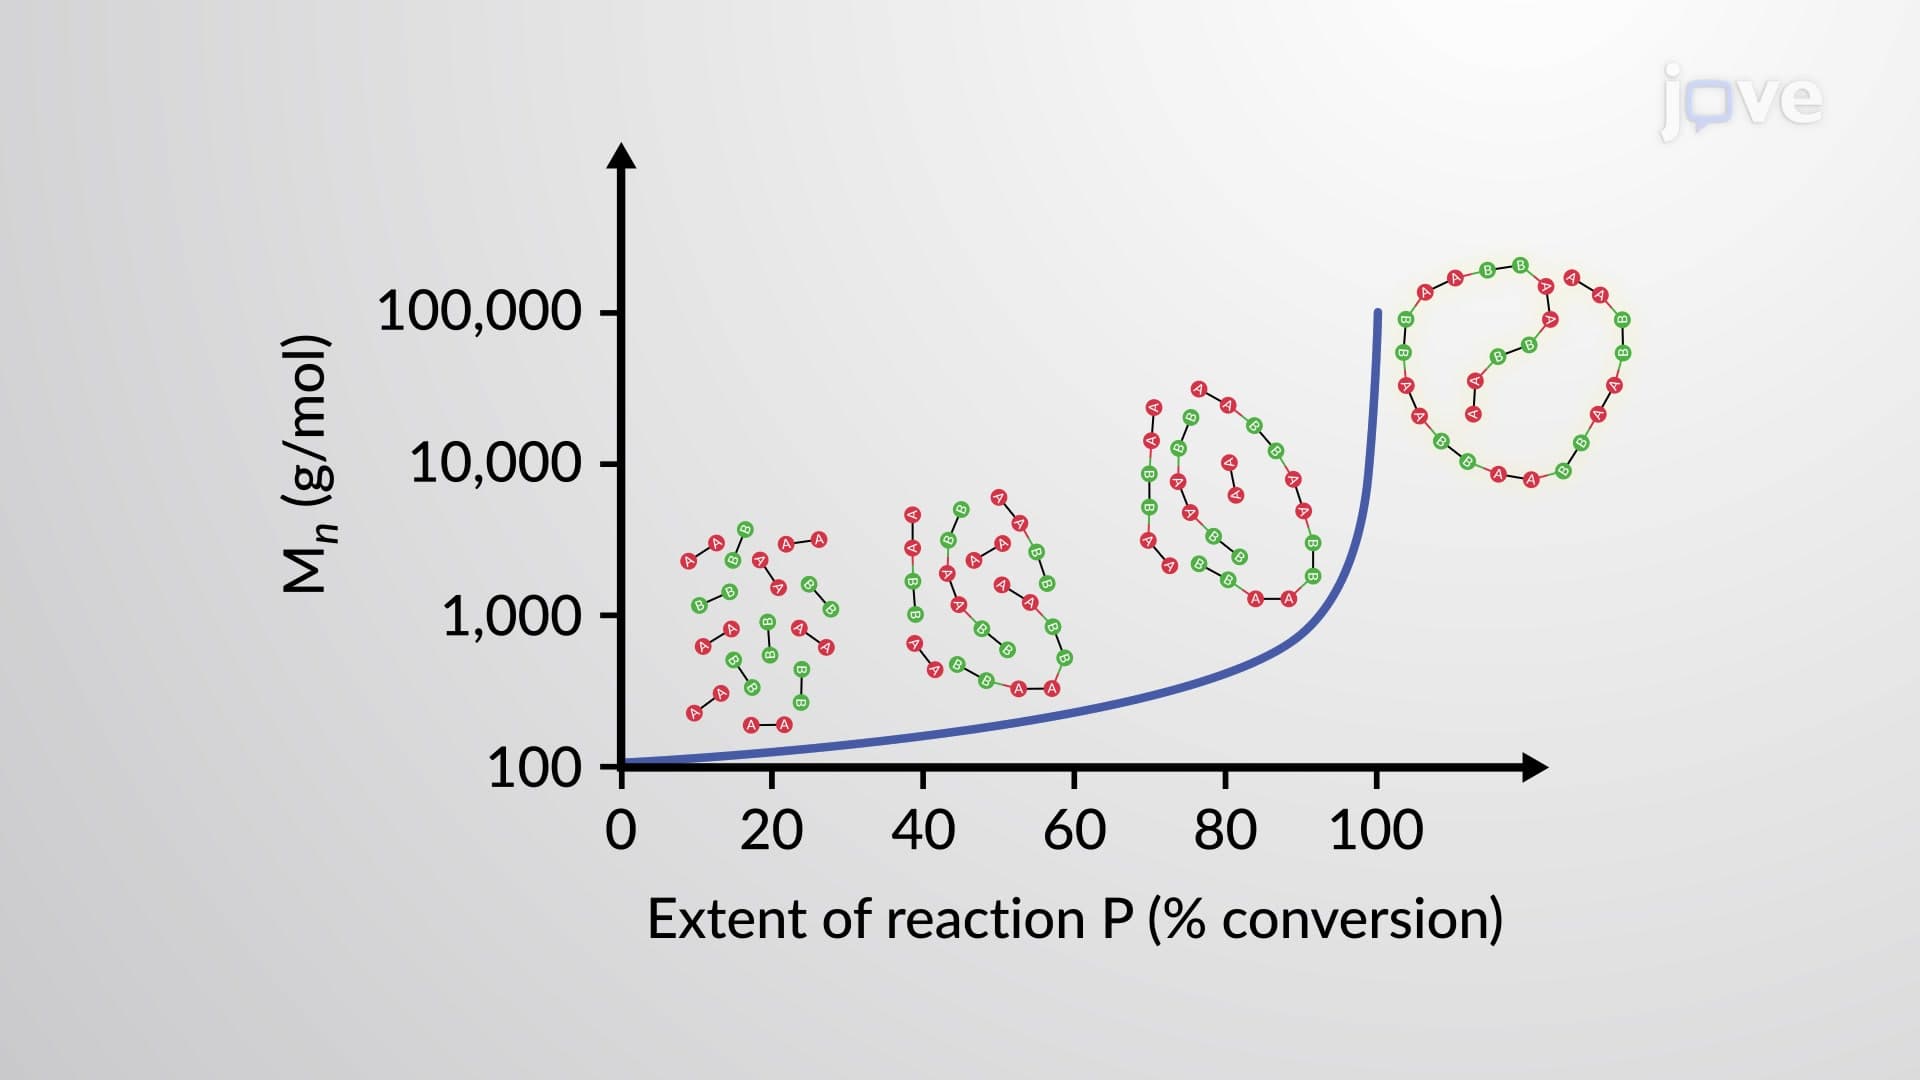
Molecular Weight of Step-Growth Polymers
As the step-growth polymerization involves step-wise condensation of monomers, the molecular weight also builds up eventually. Consequently, high molecular weight polymers are obtained at the late stages of the polymerization, where 99% of monomers have been consumed.
The extent of the...
 01:27
Distribution of Molecular Speeds
 01:10
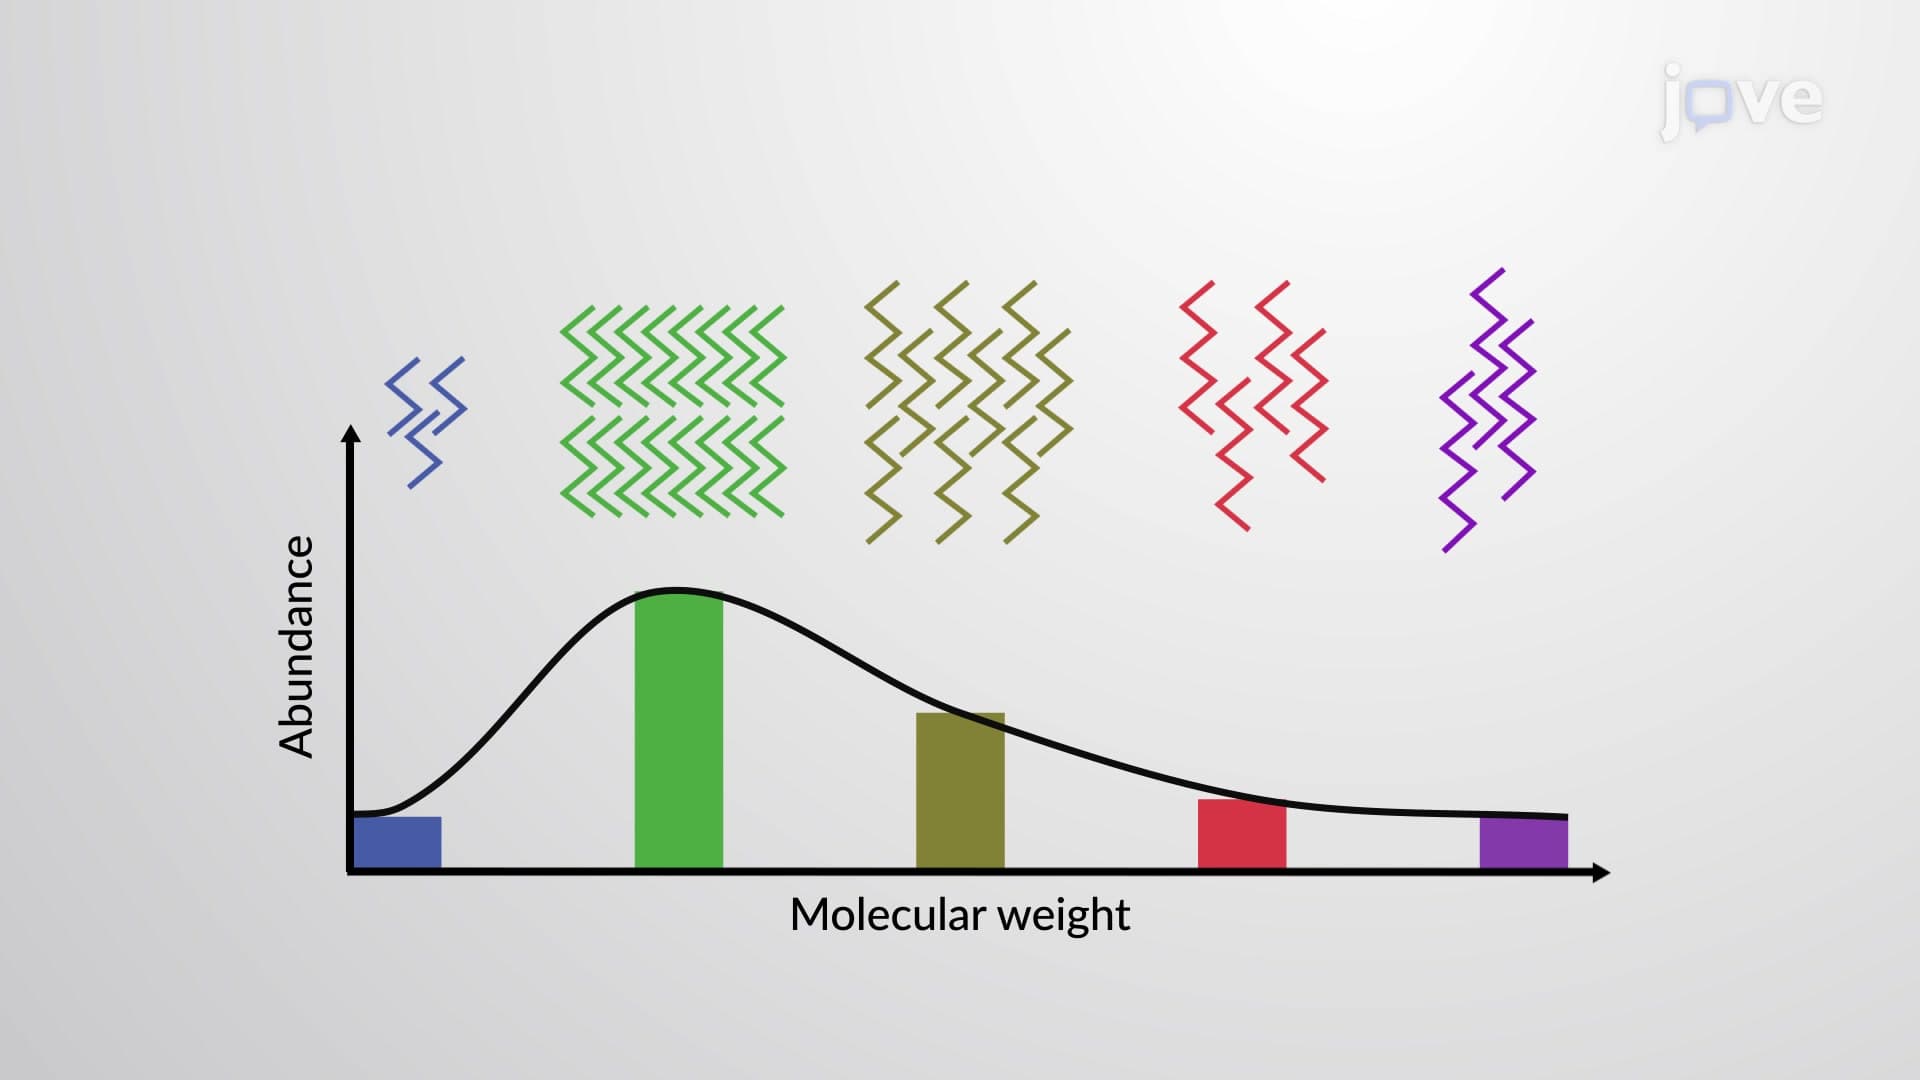
Polymers: Molecular Weight Distribution