Related Experiment Video
Updated: Aug 2, 2026
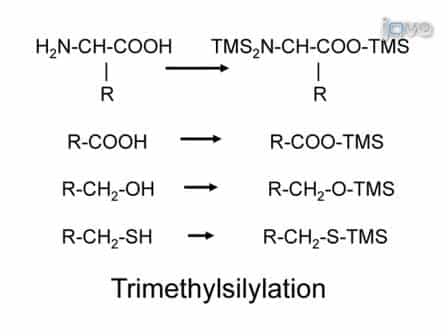

One-step Metabolomics: Carbohydrates, Organic and Amino Acids Quantified in a Single Procedure
Published on: June 25, 2010
Children with inborn errors of phenylalanine metabolism: prognosis and phenylalanine tolerance

Insights
Phenylketonuria (PKU) screening and early low phenylalanine diets support normal growth in children. Some children can normalize their diet later, suggesting regulatory mutations, reducing overly strict dietary needs.
Area of Science:
- Biochemistry
- Genetics
- Pediatrics
Background:
- Neonatal screening detects phenylketonuria (PKU), a metabolic disorder.
- Early dietary intervention is crucial for managing PKU and preventing neurological damage.
Purpose of the Study:
- To evaluate long-term growth and development in children with PKU treated with a low phenylalanine diet.
- To assess the feasibility of normalizing diet in PKU patients over time.
- To explore potential genetic factors influencing phenylalanine tolerance.
Main Methods:
- Follow-up of 23 children diagnosed with PKU via neonatal screening.
- Implementation of a low phenylalanine diet when blood phenylalanine levels exceeded 0.72 mmol/l.
- Monitoring growth, development, and blood phenylalanine levels over 8-18 years.
Main Results:
- Children's growth and development were comparable to the reference population, with two exceptions (mild intellectual disability, short attention span).
- Fourteen children remained on a strict phenylalanine-restricted diet.
- Nine children normalized their diet between 0.5 and 10 years of age, maintaining blood phenylalanine levels between 0.25-0.72 mmol/l.
Conclusions:
- Long-term low phenylalanine diets are effective in managing PKU.
- Increased phenylalanine tolerance in some children may indicate regulatory mutations in the phenylalanine hydroxylase system.
- Continuous reevaluation allows for less restrictive dietary management in PKU patients.
Abstract:
Twenty-three children, who were detected by neonatal PKU screening, were followed for 8-18 years in one paediatric centre. Dietary treatment was started if the blood phenylalanine level exceeded 0.72 mmol/l. All 23 infants were initially given a low phenylalanine diet. The growth and development rates of the children did not differ significantly from those in a reference population, although one child had mild mental retardation and another had a short attention span. Fourteen children were still on a strict phenylalanine-restricted diet on their last follow-up (at 8-18 years of age). In nine children who were initially put on a low phenylalanine diet, it was possible to normalize the diet between 1/2 and 10 years of age, while maintaining the blood phenylalanine levels between 0.25 and 0.72 mmol/l. It seems likely that those of our patients who markedly increased their phenylalanine tolerance during childhood had a regulatory mutation of the phenylalanine hydroxylase system. A continuous reevaluation of each child treated with a low phenylalanine diet reduces the use of unnecessarily restricted diets.

Related Concept Videos
 01:21
Overview of Protein Metabolism
Amino acids play various roles in the body once they are absorbed into cells. They are restructured...
 01:20
Inborn Errors of Metabolism
 01:23
Pharmacokinetics in Pediatric Patients: Overview and Drug Absorption
 01:17
Pharmacokinetics in Pediatric Patients: Drug Distribution
 01:24
Pharmacokinetics in Pediatric Patients: Drug Metabolism
 01:26
Pharmacokinetics in Pediatric Patients: Drug Excretion