Related Experiment Video
Updated: Jul 19, 2025
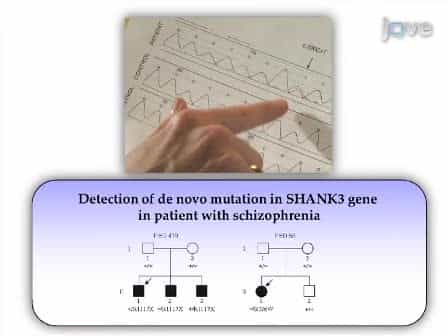

A Strategy to Identify de Novo Mutations in Common Disorders such as Autism and Schizophrenia
Published on: June 15, 2011
Use of genome sequencing to hunt for cryptic second-hit variants: analysis of 31 cases recruited to the 100 000
A Rachel Moore1,2, Jing Yu1,3, Yang Pei4
1Wellcome Centre for Human Genetics, NIHR Oxford Biomedical Research Centre, University of Oxford, Oxford, UK.

Background:
Current clinical testing methods used to uncover the genetic basis of rare disease have inherent limitations, which can lead to causative pathogenic variants being missed. Within the rare disease arm of the 100 000 Genomes Project (100kGP), families were recruited under the clinical indication 'single autosomal recessive mutation in rare disease'. These participants presented with strong clinical suspicion for a specific autosomal recessive disorder, but only one suspected pathogenic variant had been identified through standard-of-care testing. Whole genome sequencing (WGS) aimed to identify cryptic 'second-hit' variants.
Methods:
To investigate the 31 families with available data that remained unsolved following formal review within the 100kGP, SVRare was used to aggregate structural variants present in <1% of 100kGP participants. Small variants were assessed using population allele frequency data and SpliceAI. Literature searches and publicly available online tools were used for further annotation of pathogenicity.
Results:
Using these strategies, 8/31 cases were solved, increasing the overall diagnostic yield of this cohort from 10/41 (24.4%) to 18/41 (43.9%). Exemplar cases include a patient with cystic fibrosis harbouring a novel exonic LINE1 insertion in CFTR and a patient with generalised arterial calcification of infancy with complex interlinked duplications involving exons 2-6 of ENPP1. Although ambiguous by short-read WGS, the ENPP1 variant structure was resolved using optical genome mapping and RNA analysis.
Conclusion:
Systematic examination of cryptic variants across a multi-disease cohort successfully identifies additional pathogenic variants. WGS data analysis in autosomal recessive rare disease should consider complex structural and small intronic variants as potentially pathogenic second hits.

Related Concept Videos
 02:54
Evolutionary Relationships through Genome Comparisons
 01:11
Genome-wide Association Studies-GWAS
GWAS does not require the identification of the target gene involved in...
 03:00
Next-generation Sequencing
Next-Generation Sequencing Methods
Although all next-generation methods use different technologies, they all share a set of standard features....
 02:26
Comparing Copy Number Variations and SNPs
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
 01:29
Modern Molecular Taxonomy