Related Experiment Video
Updated: Jul 19, 2025

Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Computation of Absolute Binding Free Energies for Noncovalent Inhibitors with SARS-CoV-2 Main Protease
Mohammad M Ghahremanpour1, Anastasia Saar1, Julian Tirado-Rives1
1Department of Chemistry, Yale University, New Haven, Connecticut 06520-8107, United States.

Abstract:
Accurate, routine calculation of absolute binding free energies (ABFEs) for protein-ligand complexes remains a key goal of computer-aided drug design since it can enable screening and optimization of drug candidates. For development and testing of related methods, it is important to have high-quality datasets. To this end, from our own experimental studies, we have selected a set of 16 inhibitors of the SARS-CoV-2 main protease (Mpro) with structural diversity and well-distributed BFEs covering a 5 kcal/mol range. There is also minimal structural uncertainty since X-ray crystal structures have been deposited for 12 of the compounds. For methods testing, we report ABFE results from 2 μs molecular dynamics (MD) simulations using free energy perturbation (FEP) theory. The correlation of experimental and computed results is encouraging, with a Pearson's r2 of 0.58 and a Kendall τ of 0.24. The results indicate that current FEP-based ABFE calculations can be used for identification of active compounds (hits). While their accuracy for lead optimization is not yet sufficient, this activity remains addressable in separate lead series by relative BFE calculations.
Related Concept Videos
 02:18
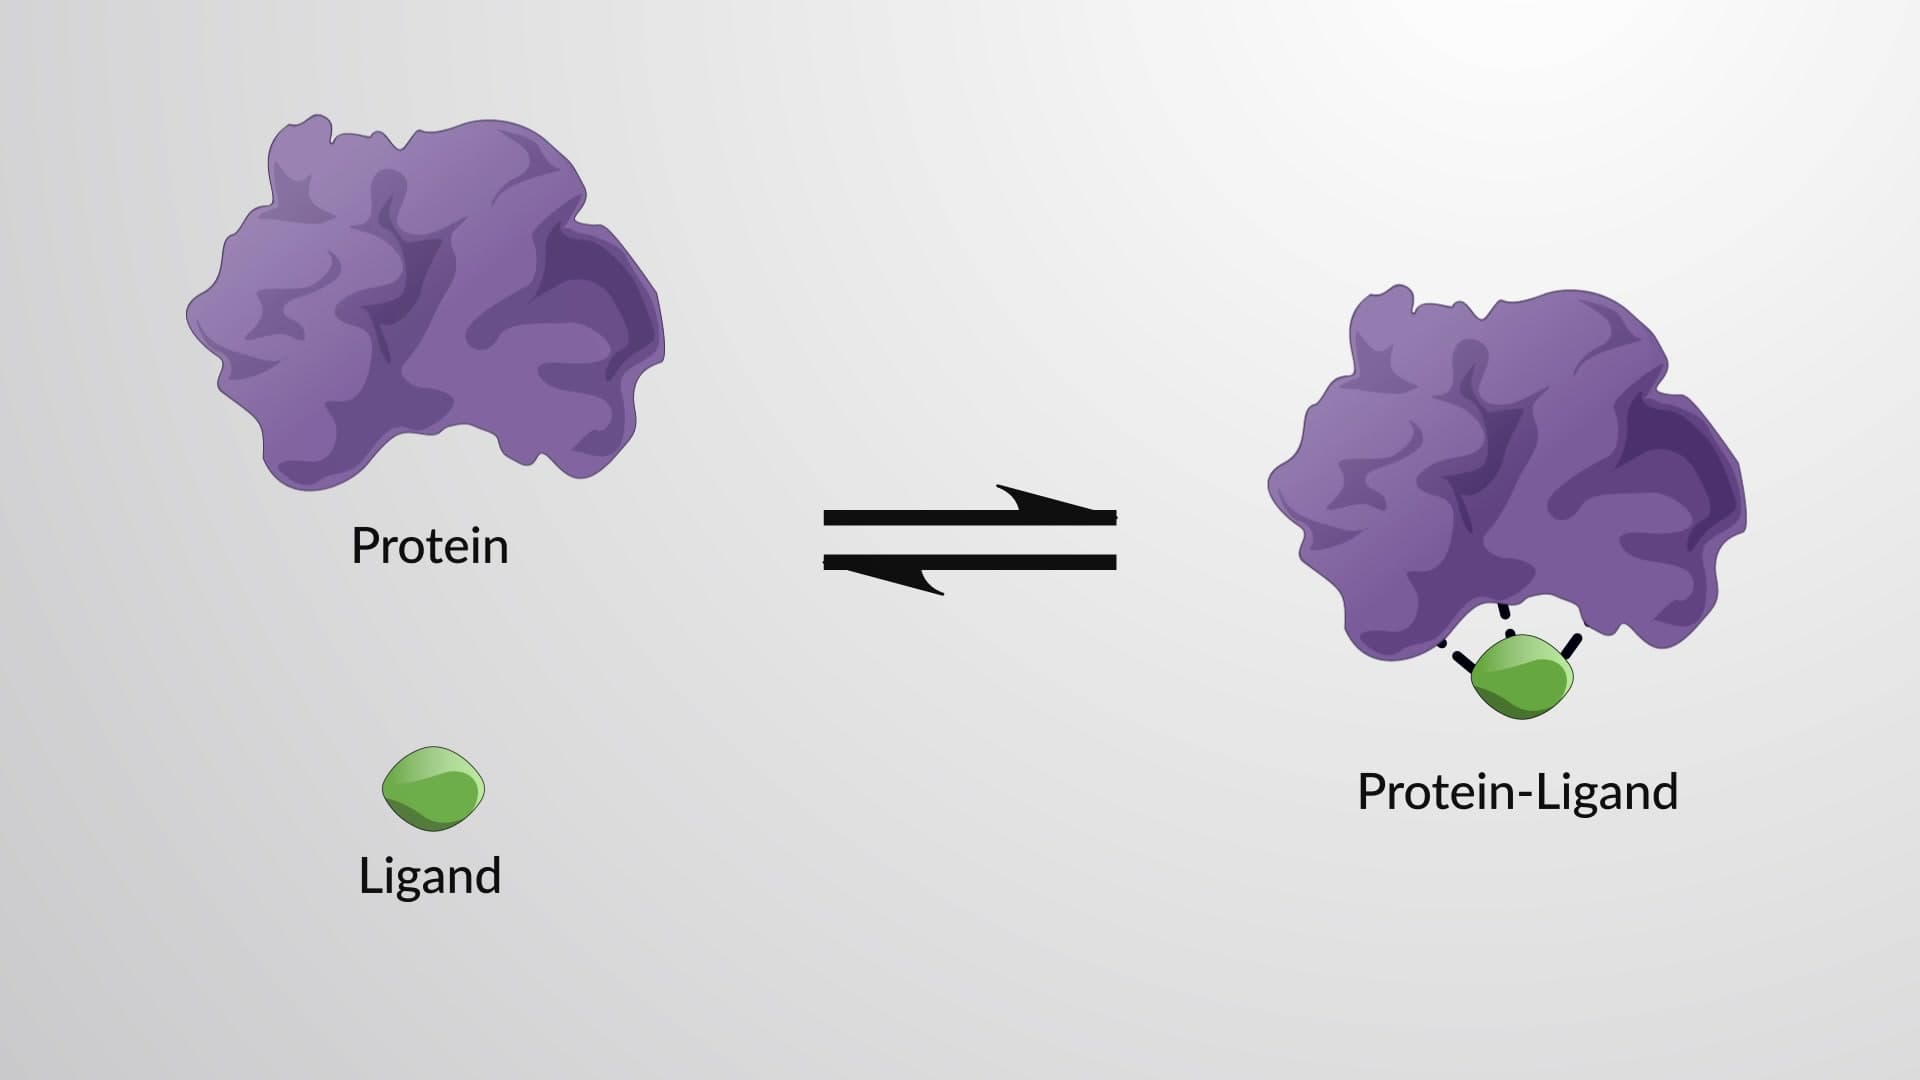
The Equilibrium Binding Constant and Binding Strength
 01:16
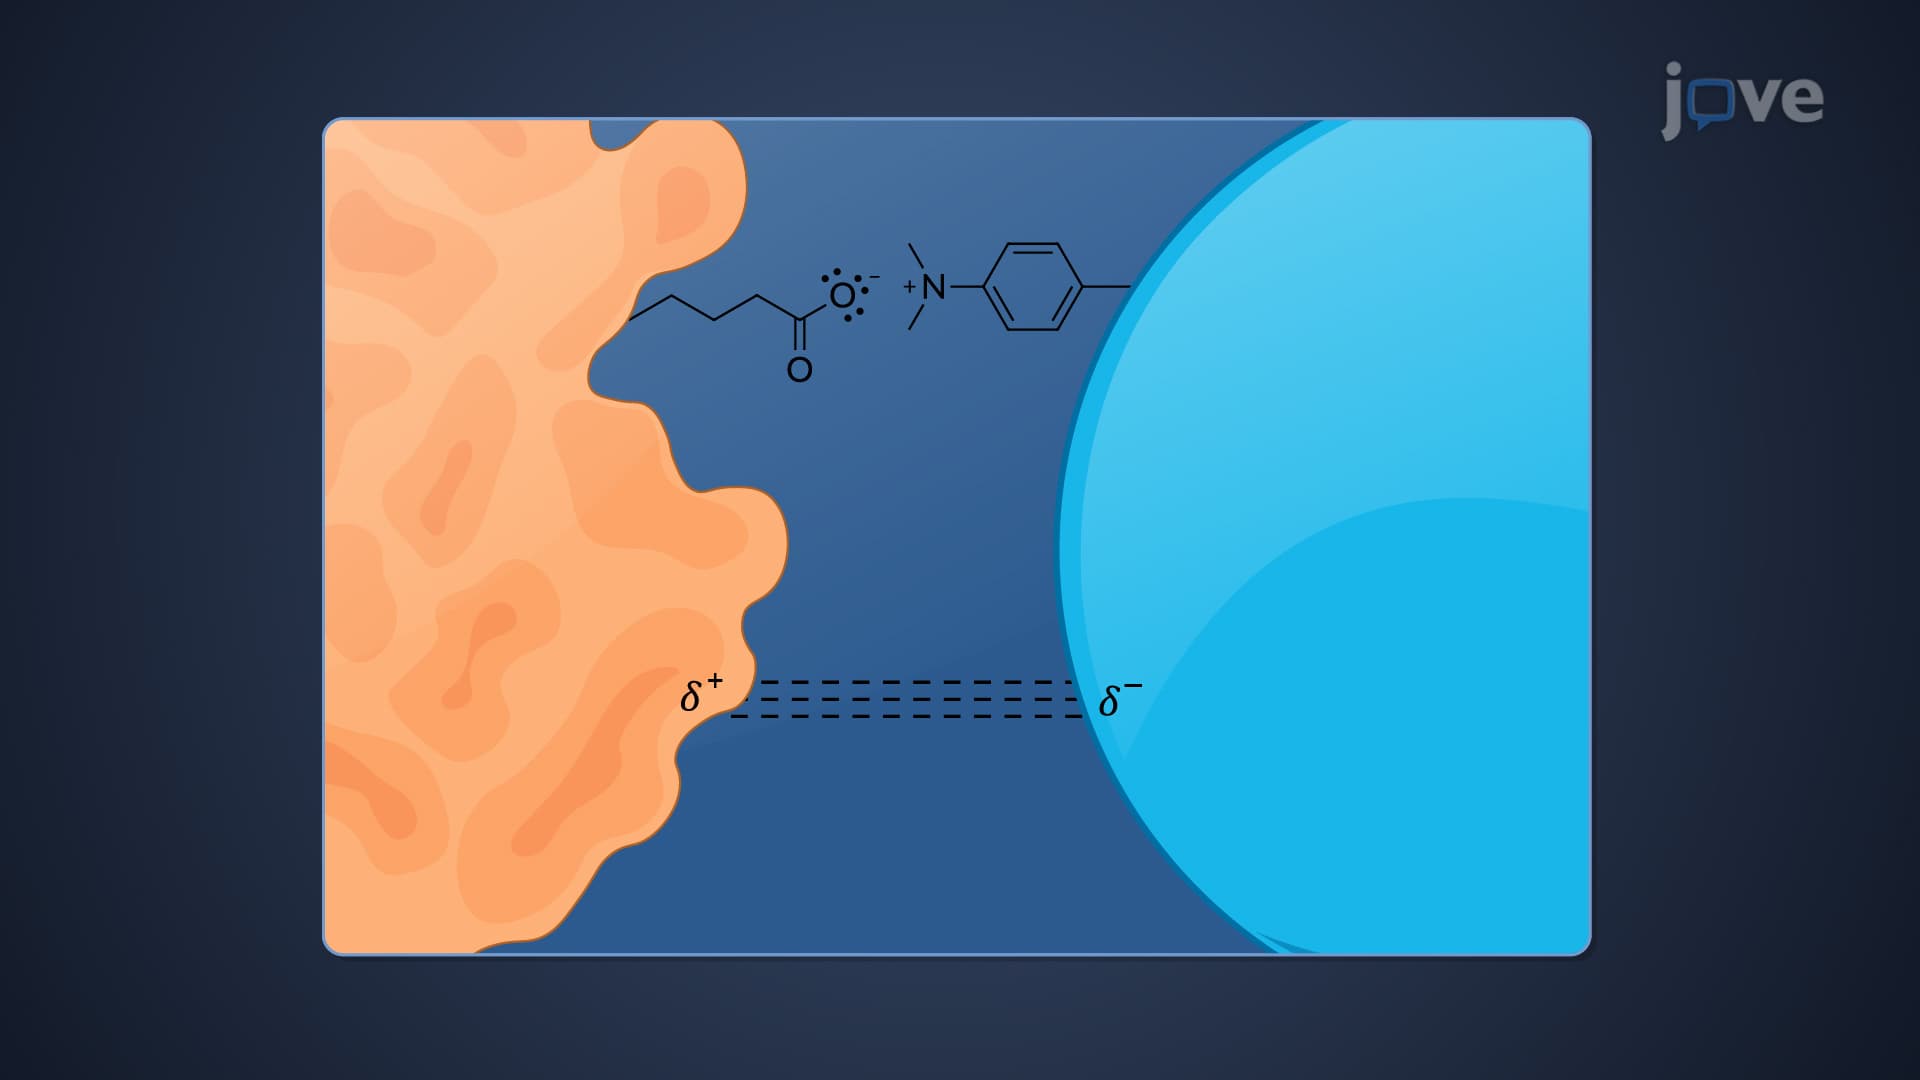
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...