Related Experiment Video
Updated: Jul 18, 2025


From Molecules to Materials: Engineering New Ionic Liquid Crystals Through Halogen Bonding
Published on: March 24, 2018
Local Chemical Clustering Enabled Ultrahigh Capacitive Energy Storage in Pb-Free Relaxors
Hui Liu1, Zheng Sun1, Ji Zhang2
1Beijing Advanced Innovation Center for Materials Genome Engineering, Department of Physical Chemistry, University of Science and Technology Beijing, Beijing 100083, China.

Abstract:
Designing Pb-free relaxors with both a high capacitive energy density (Wrec) and high storage efficiency (η) remains a remarkable challenge for cutting-edge pulsed power technologies. Local compositional heterogeneity is crucial for achieving complex polar structure in solid solution relaxors, but its role in optimizing energy storage properties is often overlooked. Here, we report that an exceptionally high Wrec of 15.2 J cm-3 along with an ultrahigh η of 91% can be achieved through designing local chemical clustering in Bi0.5Na0.5TiO3-BaTiO3-based relaxors. A three-dimensional atomistic model derived from neutron/X-ray total scattering combined with reverse Monte Carlo method reveals the presence of subnanometer scale clustering of Bi, Na, and Ba, which host differentiated polar displacements, and confirming the prediction by density functional theory calculations. This leads to a polar state with small polar clusters and strong length and direction fluctuations in unit-cell polar vectors, thus manifesting improved high-field polarizability, steadily reduced hysteresis, and high breakdown strength macroscopically. The favorable polar structure features also result in a unique field-increased η, excellent stability, and superior discharge capacity. Our work demonstrates that the hidden local chemical order exerts a significant impact on the polarization characteristic of relaxors, and can be exploited for accessing superior energy storage performance.
Related Concept Videos
 02:48
Ionic Bonding and Electron Transfer
 01:19
Complexation Equilibria: The Chelate Effect
 01:31
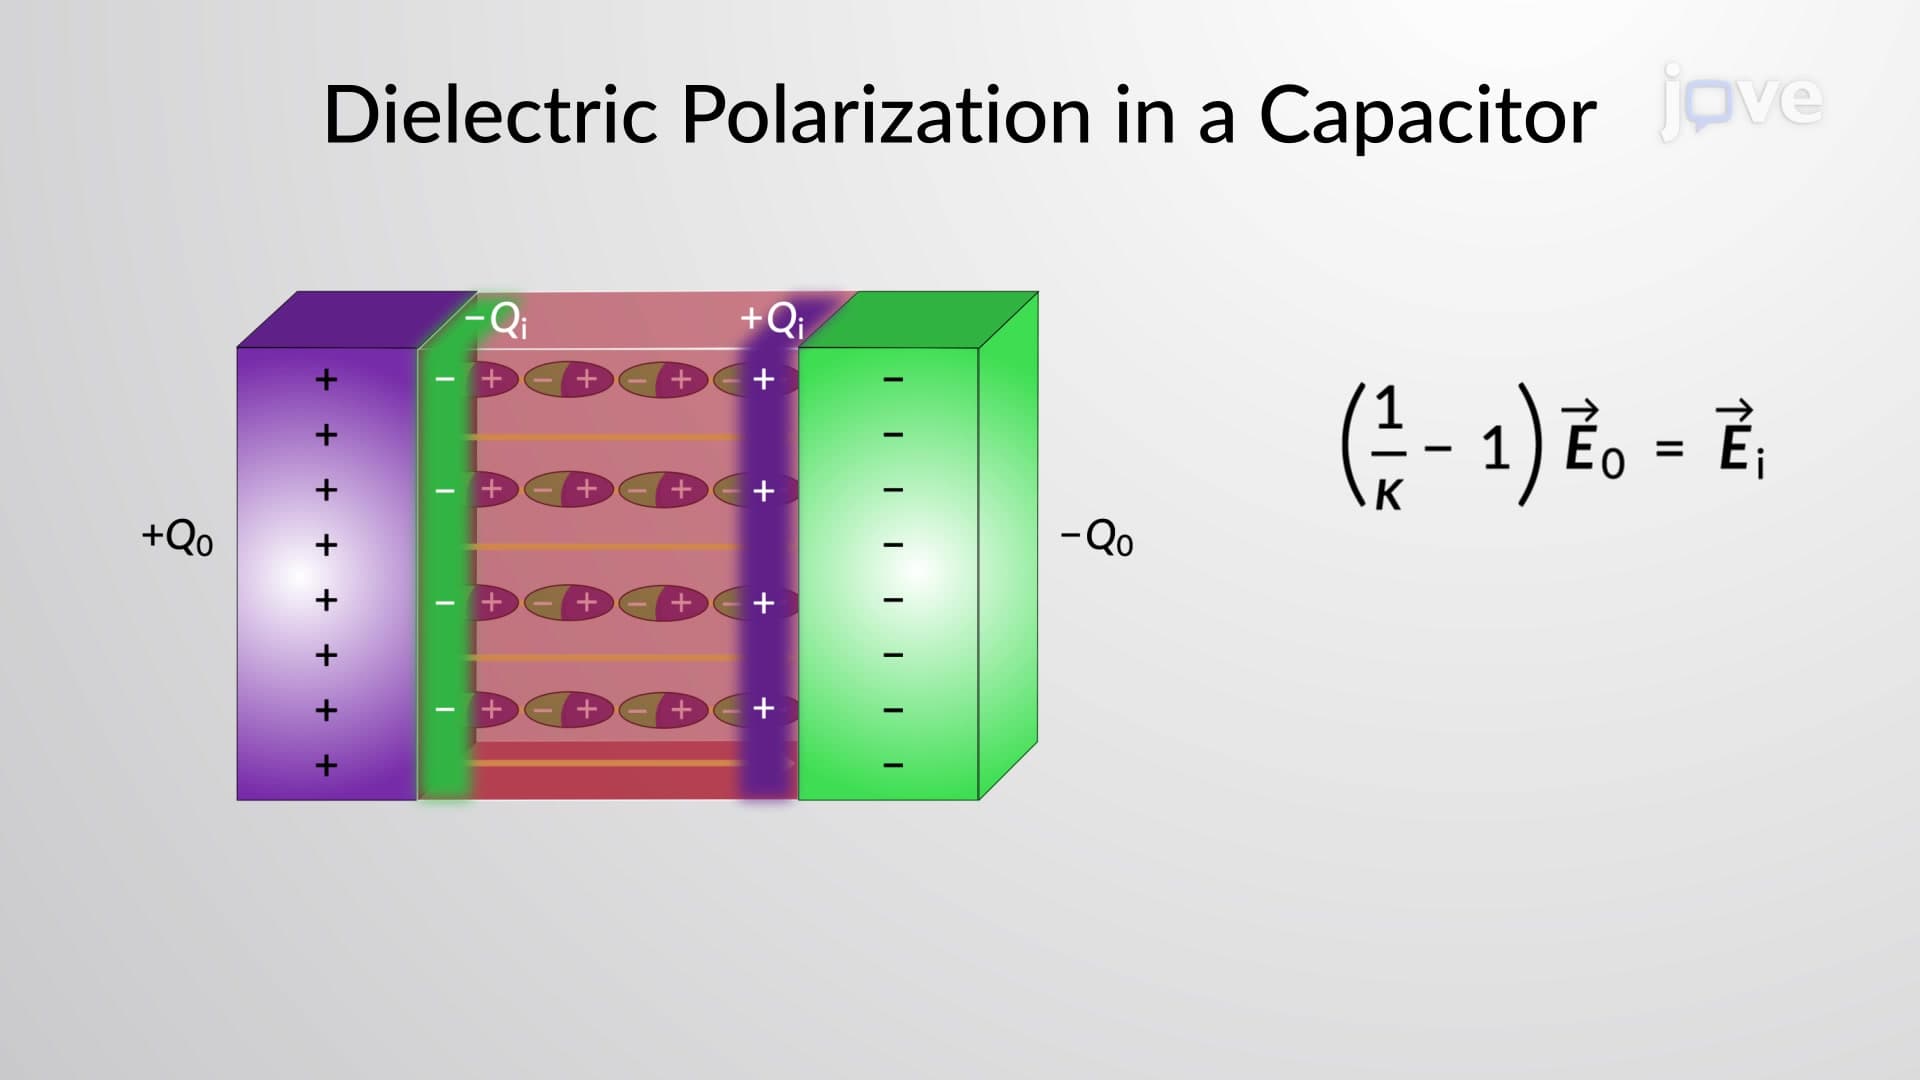
Dielectric Polarization in a Capacitor
 02:54
Trends in Lattice Energy: Ion Size and Charge
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...