Related Experiment Video
Updated: Jul 18, 2025

Author Spotlight: Streamlining Visual Dynamics to Simplify Molecular Dynamics Simulations Using Gromacs
Published on: August 9, 2024
Efficient Empirical Valence Bond Simulations with GROMACS
Gabriel Oanca1, Florian van der Ent1, Johan Åqvist1
1Department of Cell and Molecular Biology, Biomedical Center, Uppsala University, Uppsala SE-751 24, Sweden.

Abstract:
We describe a protocol to perform empirical valence bond (EVB) simulations using GROMACS software. EVB is a fast and reliable method that allows one to determine the reaction free-energy profiles in complex systems, such as enzymes, by employing classical force fields to represent a chemical reaction. Therefore, running EVB simulations is basically as fast as any classical molecular dynamics simulation, and the method uses standard free-energy calculations to map the free-energy change along a given reaction path. To exemplify and validate our EVB implementation, we replicated two cases of our earlier enzyme simulations. One of these addresses the decomposition of the activation free energy into its enthalpic and entropic components, and the other is focused on calculating the overall catalytic effect of the enzyme compared to the same reaction in water. These two examples give virtually identical results to those obtained with programs that were specifically designed for EVB simulations and show that the GROMACS implementation is robust and can be used for very large systems.

Related Concept Videos
 02:42
Valence Bond Theory
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 02:13
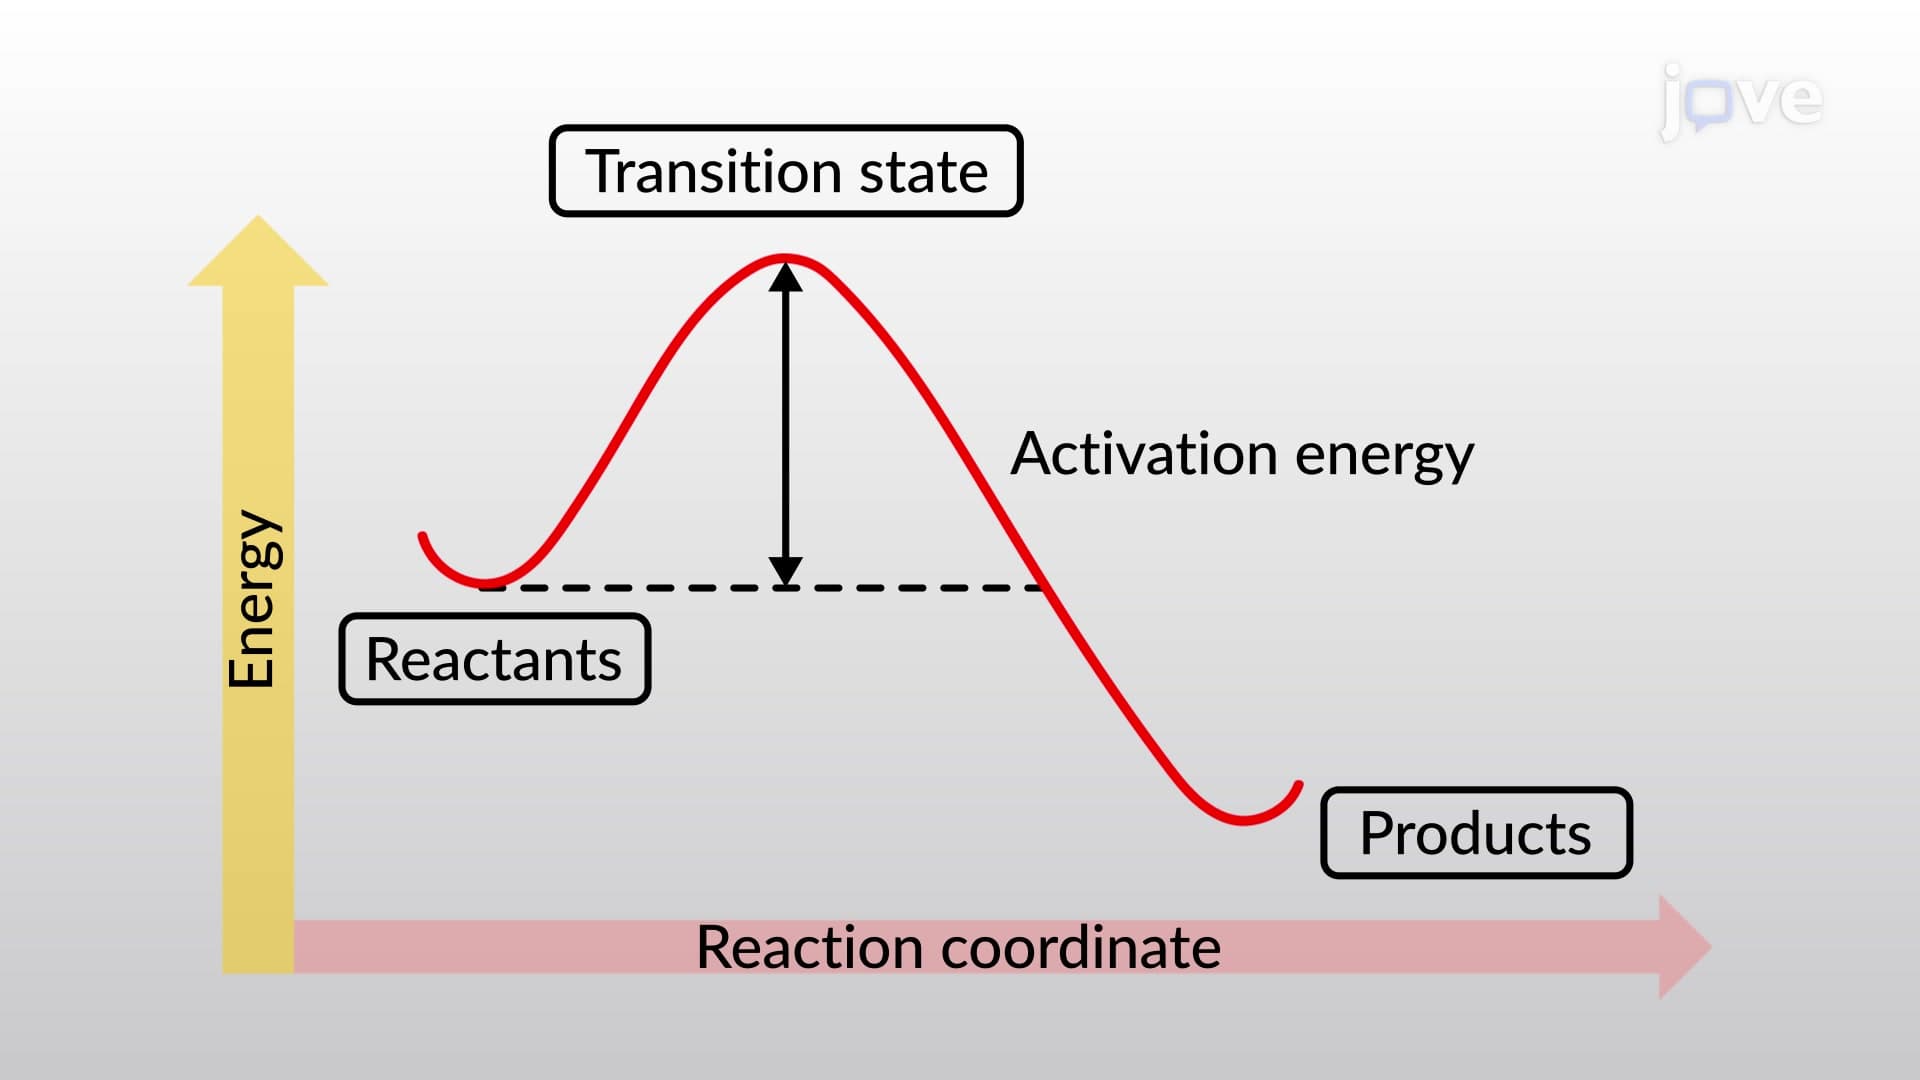
Bond Dissociation Energy and Activation Energy
 01:24
Van der Waals Interactions
 02:49
Bond Energies and Bond Lengths
 02:52
VSEPR Theory and the Basic Shapes