Related Experiment Video
Updated: Jul 17, 2025


Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
Computational methods to analyze and predict the binding mode of inhibitors targeting both human and mushroom
Federico Ricci1, Kristina Schira2, Lyna Khettabi3
1Department of Chemical, Biological, Pharmaceutical, and Environmental Sciences, University of Messina, Viale F. D'Alcontres 31, I-98166, Messina, Italy.

Abstract:
Tyrosinase, a copper-containing enzyme critical in melanin biosynthesis, is a key drug target for hyperpigmentation and melanoma in humans. Testing the inhibitory effects of compounds using tyrosinase from Agaricus bisporus (AbTYR) has been a common practice to identify potential therapeutics from synthetic and natural sources. However, structural diversity among human tyrosinase (hTYR) and AbTYR presents a challenge in developing drugs that are therapeutically effective. In this study, we combined retrospective and computational analyses with experimental data to provide insights into the development of new inhibitors targeting both hTYR and AbTYR. We observed contrasting effects of Thiamidol™ and our 4-(4-hydroxyphenyl)piperazin-1-yl-derivative (6) on both enzymes; based on this finding, we aimed to investigate their binding modes in hTYR and AbTYR to identify residues that significantly improve affinity. All the information led to the discovery of compound [4-(4-hydroxyphenyl)piperazin-1-yl](2-methoxyphenyl)methanone (MehT-3, 7), which showed comparable activity on AbTYR (IC50 = 3.52 μM) and hTYR (IC50 = 5.4 μM). Based on these achievements we propose the exploitation of our computational results to provide relevant structural information for the development of newer dual-targeting molecules, which could be preliminarily tested on AbTYR as a rapid and inexpensive screening procedure before being tested on hTYR.
More Related Videos
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 02:04
Protein-protein Interfaces
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 02:18
The Equilibrium Binding Constant and Binding Strength
 00:49
Ligand Binding and Linkage
 01:16
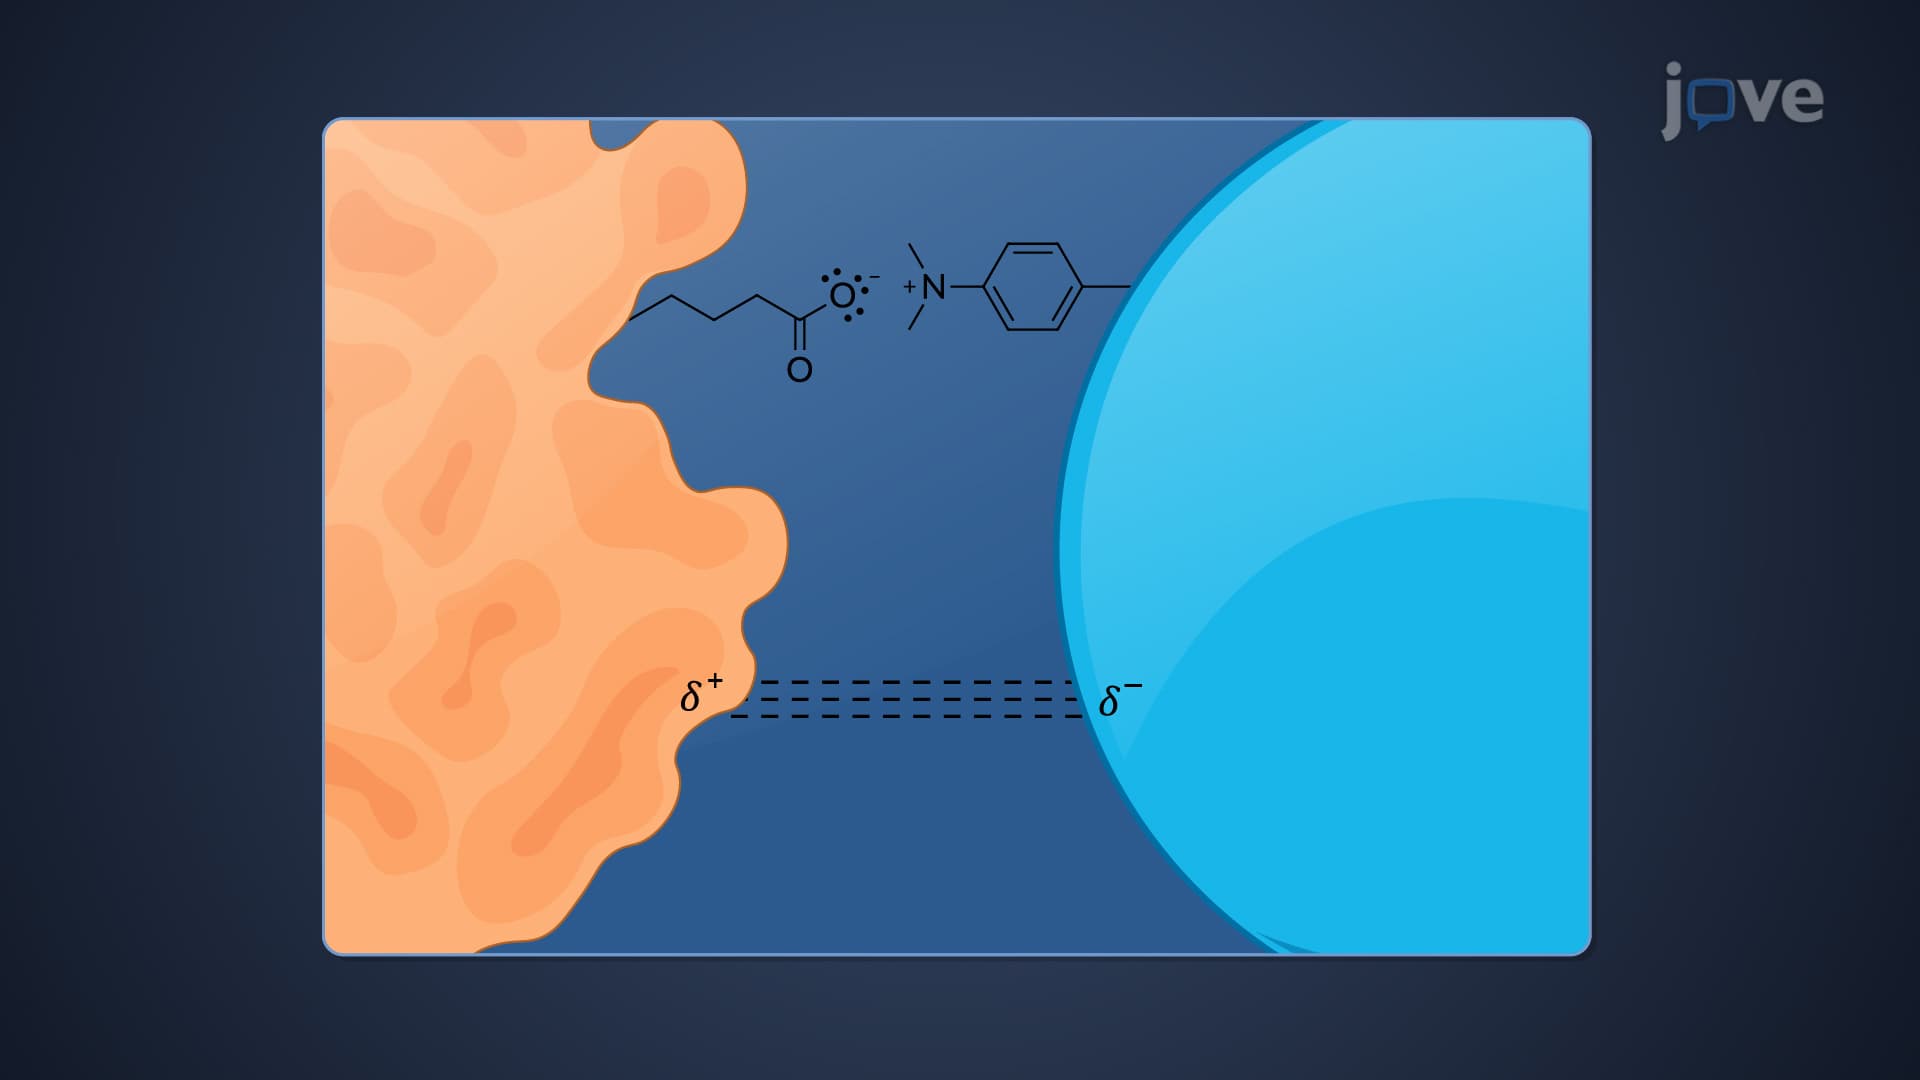
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...