Related Experiment Video
Updated: Jul 17, 2025


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Accurate and Efficient Prediction of Post-Hartree-Fock Polarizabilities of Condensed-Phase Systems
Dongbo Zhao1, Yilin Zhao2, Xin He3
1Institute of Biomedical Research, Yunnan University, Kunming 650500, P. R. China.

Abstract:
To accurately and efficiently predict the molecular response properties (such as polarizability) at post-Hartree-Fock levels for condensed-phase systems under periodic boundary conditions (PBC) is still an unaccomplished and ongoing task. We demonstrate that static isotropic polarizabilities can be cost-effectively predicted at post-Hartree-Fock levels by combining the linear-scaling generalized energy-based fragmentation (GEBF) and information-theoretic approach (ITA) quantities. In PBC-GEBF, the total molecular polarizability of an extended system is obtained as a linear combination of the corresponding quantities of a series of small embedded subsystems of several monomers. Here, we show that in the PBC-GEBF-ITA framework, one can obtain the molecular polarizabilities and establish linear relations to ITA quantities. Once these relations are established for smaller subsystems, one can predict the polarizabilities of larger subsystems directly from the molecular wavefunction (or electron density) via ITA quantities. Alternatively, one can determine the total molecular polarizability via a linear combination equation in PBC-GEBF. We have corroborated that this newly proposed PBC-GEBF-ITA protocol is much more efficient than the original PBC-GEBF approach but is not much less accurate and that this conclusion holds for both many-body perturbation theory and the coupled cluster calculations. Good efficiency and transferability of the PBC-GEBF-ITA protocol are demonstrated for periodic systems with several hundred atoms in a unit cell.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:27
Predicting Molecular Geometry
 01:08
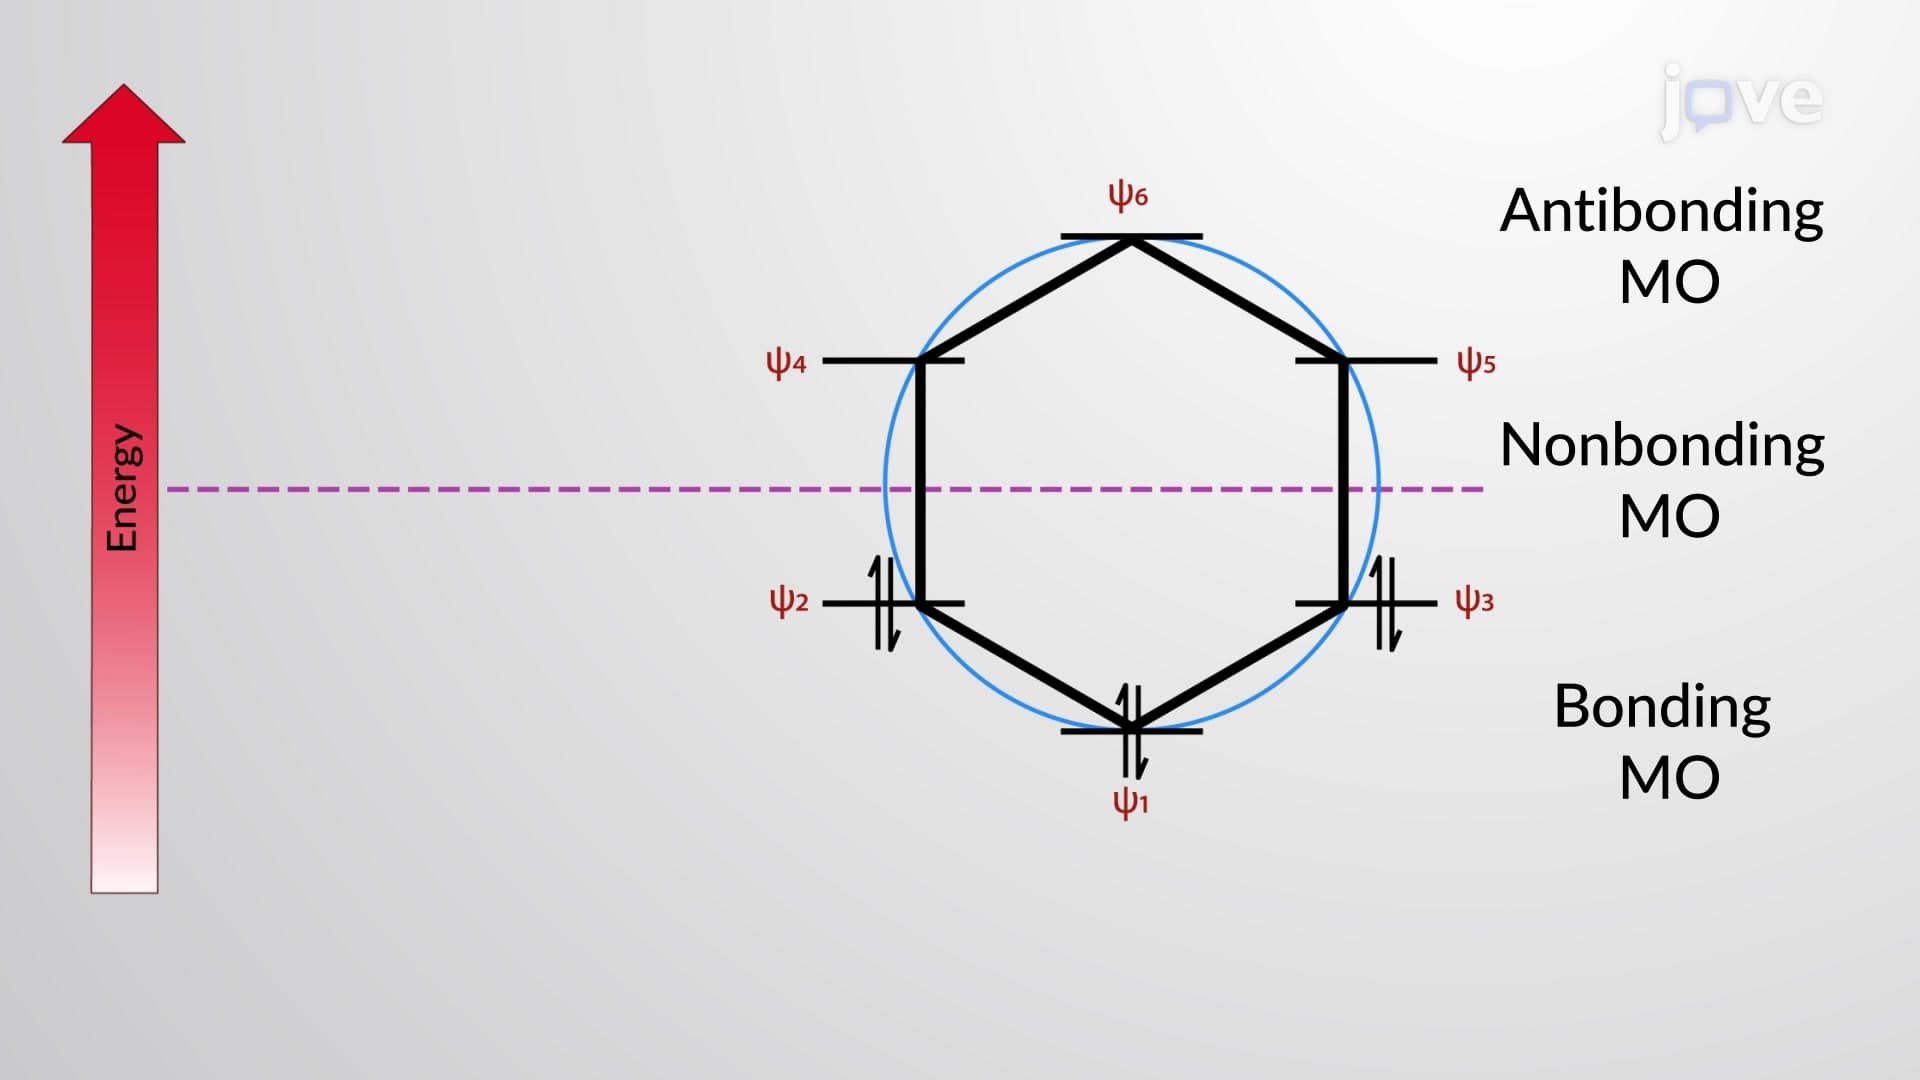
Hückel's Rule Diagram of π MOs: Frost Circle
A Frost circle is constructed by drawing a polygon whose number of edges is equal to the number of carbons of the given cyclic system, with one of the vertices pointing down. Then, a circle is drawn enclosing the polygon so...
 02:40
MO Theory and Covalent Bonding
 02:36
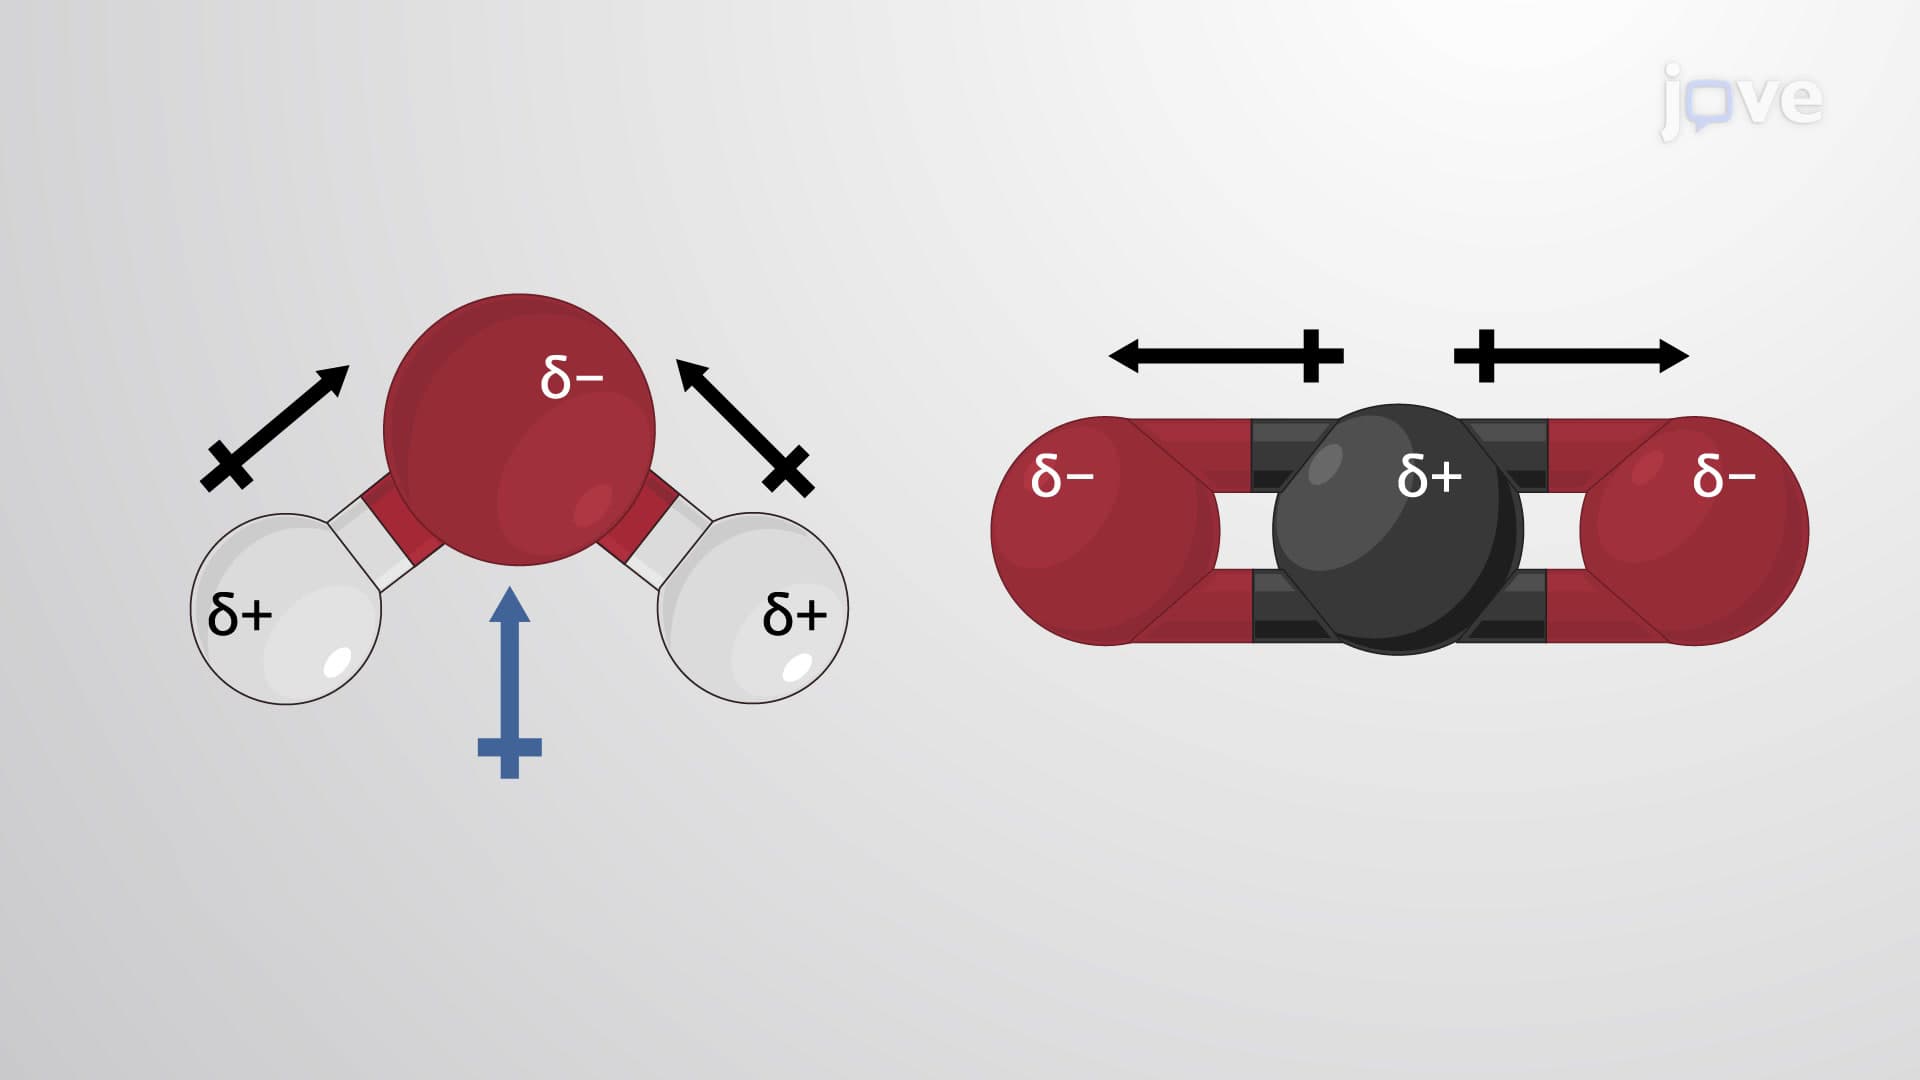
Molecular Geometry and Dipole Moments
 01:29
Potential Due to a Polarized Object
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...