Koopmans' Theorem-Compliant Long-Range Corrected (KTLC) Density Functional Mediated by Black-Box Optimization and Data-Driven Prediction for Organic Molecules
Related Experiment Video
Updated: Jul 16, 2025
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Kei Terayama1,2,3, Yamato Osaki4, Takehiro Fujita5
1Graduate School of Medical Life Science, Yokohama City University, 1-7-29 Suehiro-cho, Tsurumi-ku Kanagawa 230-0045, Japan.

Abstract:
Density functional theory (DFT) is a significant computational tool that has substantially influenced chemistry, physics, and materials science. DFT necessitates parametrized approximation for determining an expected value. Hence, to predict the properties of a given molecule using DFT, appropriate parameters of the functional should be set for each molecule. Herein, we optimize the parameters of range-separated functionals (LC-BLYP and CAM-B3LYP) via Bayesian optimization (BO) to satisfy Koopmans' theorem. Our results demonstrate the effectiveness of the BO in optimizing functional parameters. Particularly, Koopmans' theorem-compliant LC-BLYP (KTLC-BLYP) shows results comparable to the experimental UV-absorption values. Furthermore, we prepared an optimized parameter dataset of KTLC-BLYP for over 3000 molecules through BO for satisfying Koopmans' theorem. We have developed a machine learning model on this dataset to predict the parameters of the LC-BLYP functional for a given molecule. The prediction model automatically predicts the appropriate parameters for a given molecule and calculates the corresponding values. The approach in this paper would be useful to develop new functionals and to update the previously developed functionals.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:27
Predicting Molecular Geometry
 01:10
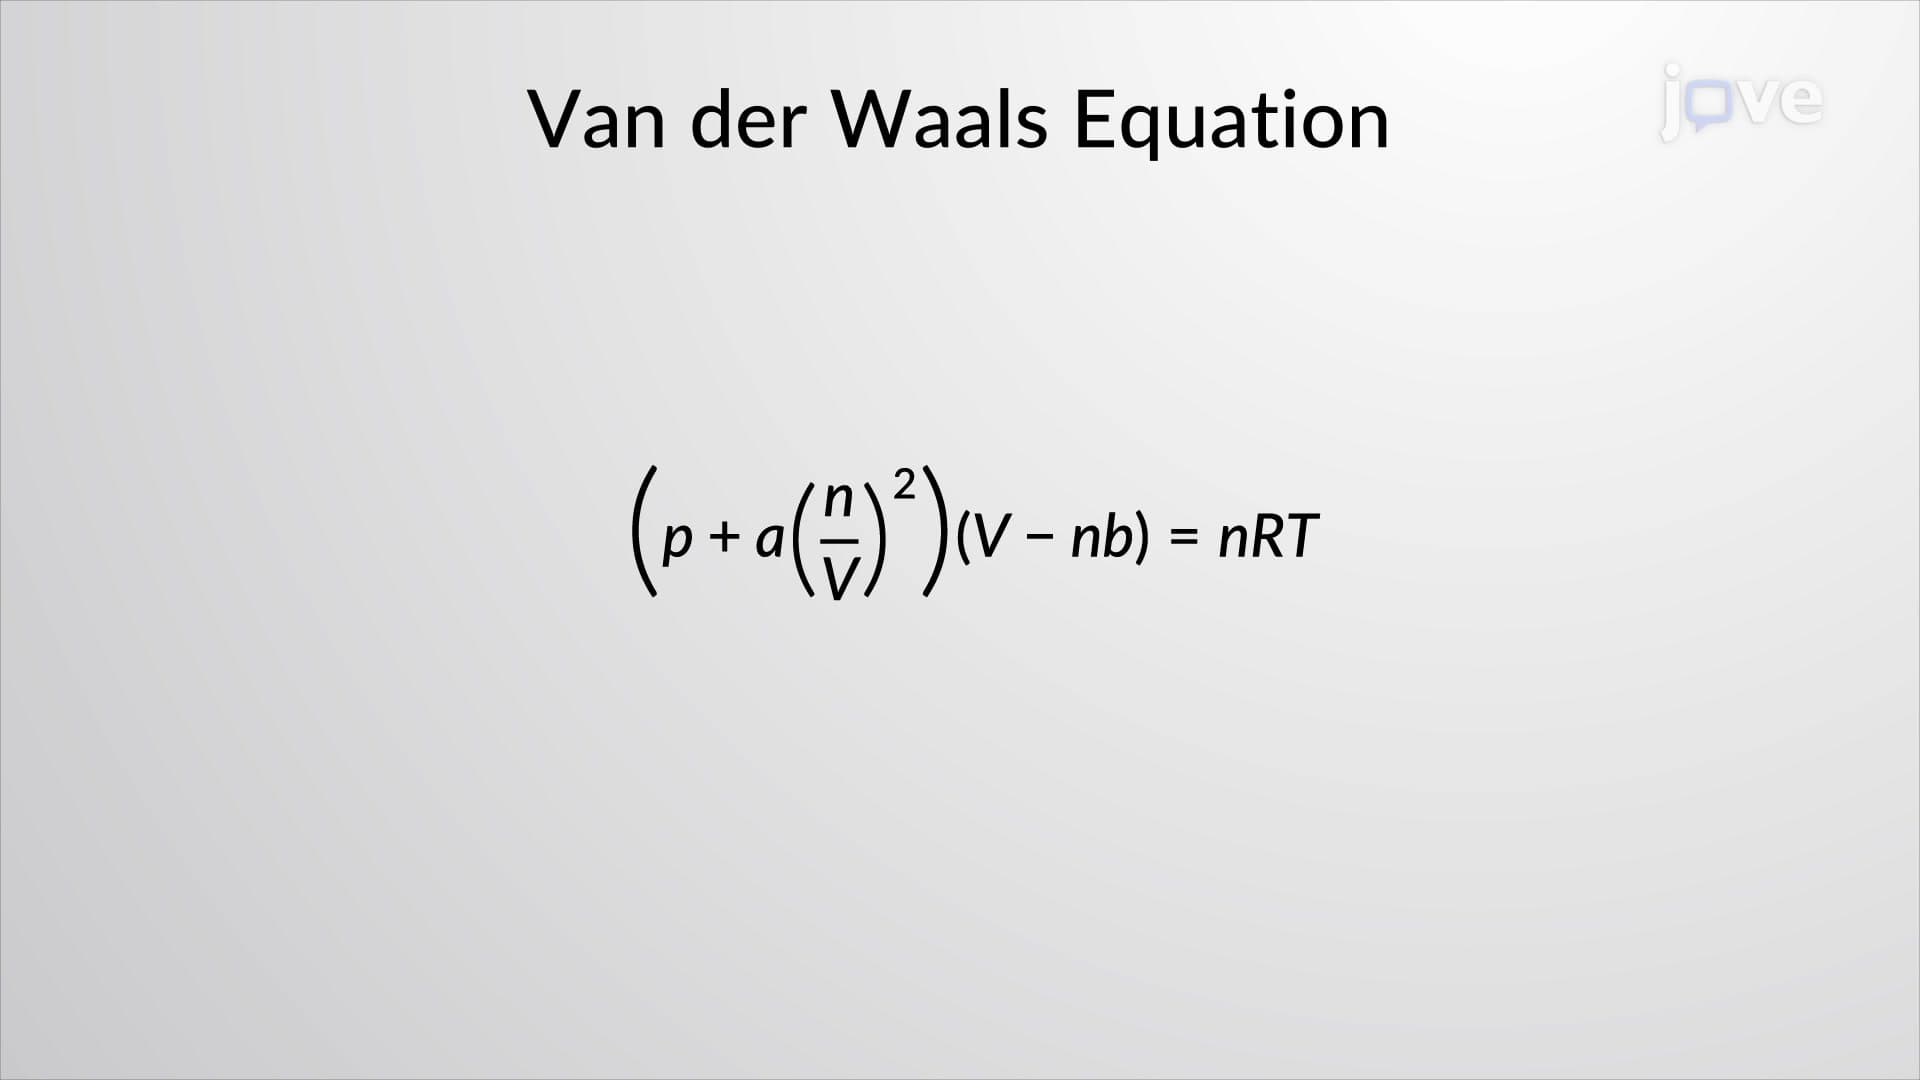
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 02:35
Molecular Orbital Theory I
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 03:51
Molecular Orbital Theory II
 02:40
MO Theory and Covalent Bonding