Related Experiment Video
Updated: Jul 14, 2025
Estimation of Urinary Nanocrystals in Humans using Calcium Fluorophore Labeling and Nanoparticle Tracking Analysis
Published on: February 9, 2021
Primary hyperoxaluria: a case series
Jawad Iqbal Rather1, Rabiya Rasheed2, Muzafar Maqsood Wani3
1Sher-I-Kashmir Institute of Medical Sciences, Srinagar, J&k, 190011, India. jawadiqbal93@gmail.com.

Background:
Primary hyperoxaluria (PH) is a rare genetic disorder characterized by the excessive production and accumulation of oxalate. We present five cases of PH, each exhibiting varying manifestations of the disorder including a case presenting as postpartum kidney failure. Notably, three of these cases involve a previously unreported mutation.
Case Presentations:
We evaluated five Indian patients who presented with varying manifestations of PH. The first case, a 30 year old woman, presented as post-partum kidney failure and was found to be having oxalate nephropathy precipitated by dietary oxalate overload in the setting of previously undiagnosed PH. Genetic analysis revealed a previously unreported mutation in the alanine-glyoxylate aminotransferase gene. The patient underwent simultaneous kidney liver transplant. The second and third cases, 26 and 28 year old women respectively, were asymptomatic siblings of the first patient, who were diagnosed through screening. The fourth case is a 12 year boy with PH type 1 presenting as nephrolithiasis and rapidly worsening kidney function requiring combined kidney liver kidney transplant. Case 5 is a 6 year old male child with type 2 PH presenting with nephrolithiasis, nephrocalcinosis and normal kidney function. All the patients were born to consanguineous parents.
Conclusions:
Due to limited clinical suspicion and inadequate diagnostic resources in certain countries with limited resources, it is possible for PH to go undiagnosed. The manifestations of the disease can range from no noticeable symptoms to severe disease. Interestingly, in some individuals with primary hyperoxaluria, the disease may not exhibit any symptoms until it is triggered by a high intake of dietary oxalate.
More Related Videos
 09:40
Isolation, Characterization, And High Throughput Extracellular Flux Analysis of Mouse Primary Renal Tubular Epithelial Cells
Published on: June 20, 2018
 09:28
Patient-derived Orthotopic Xenograft Models for Human Urothelial Cell Carcinoma and Colorectal Cancer Tumor Growth and Spontaneous Metastasis
Published on: May 12, 2019
Related Concept Videos
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 01:24
Chronic Kidney Disease II: Clinical Manifestations
 01:26
Urinary Tract Calculi II: Pathophysiology and Clinical Manifestations
 01:25
Chronic Kidney Disease I: Introduction
 01:28
Acute Pyelonephritis II: Diagnostic Studies and Management
 01:29
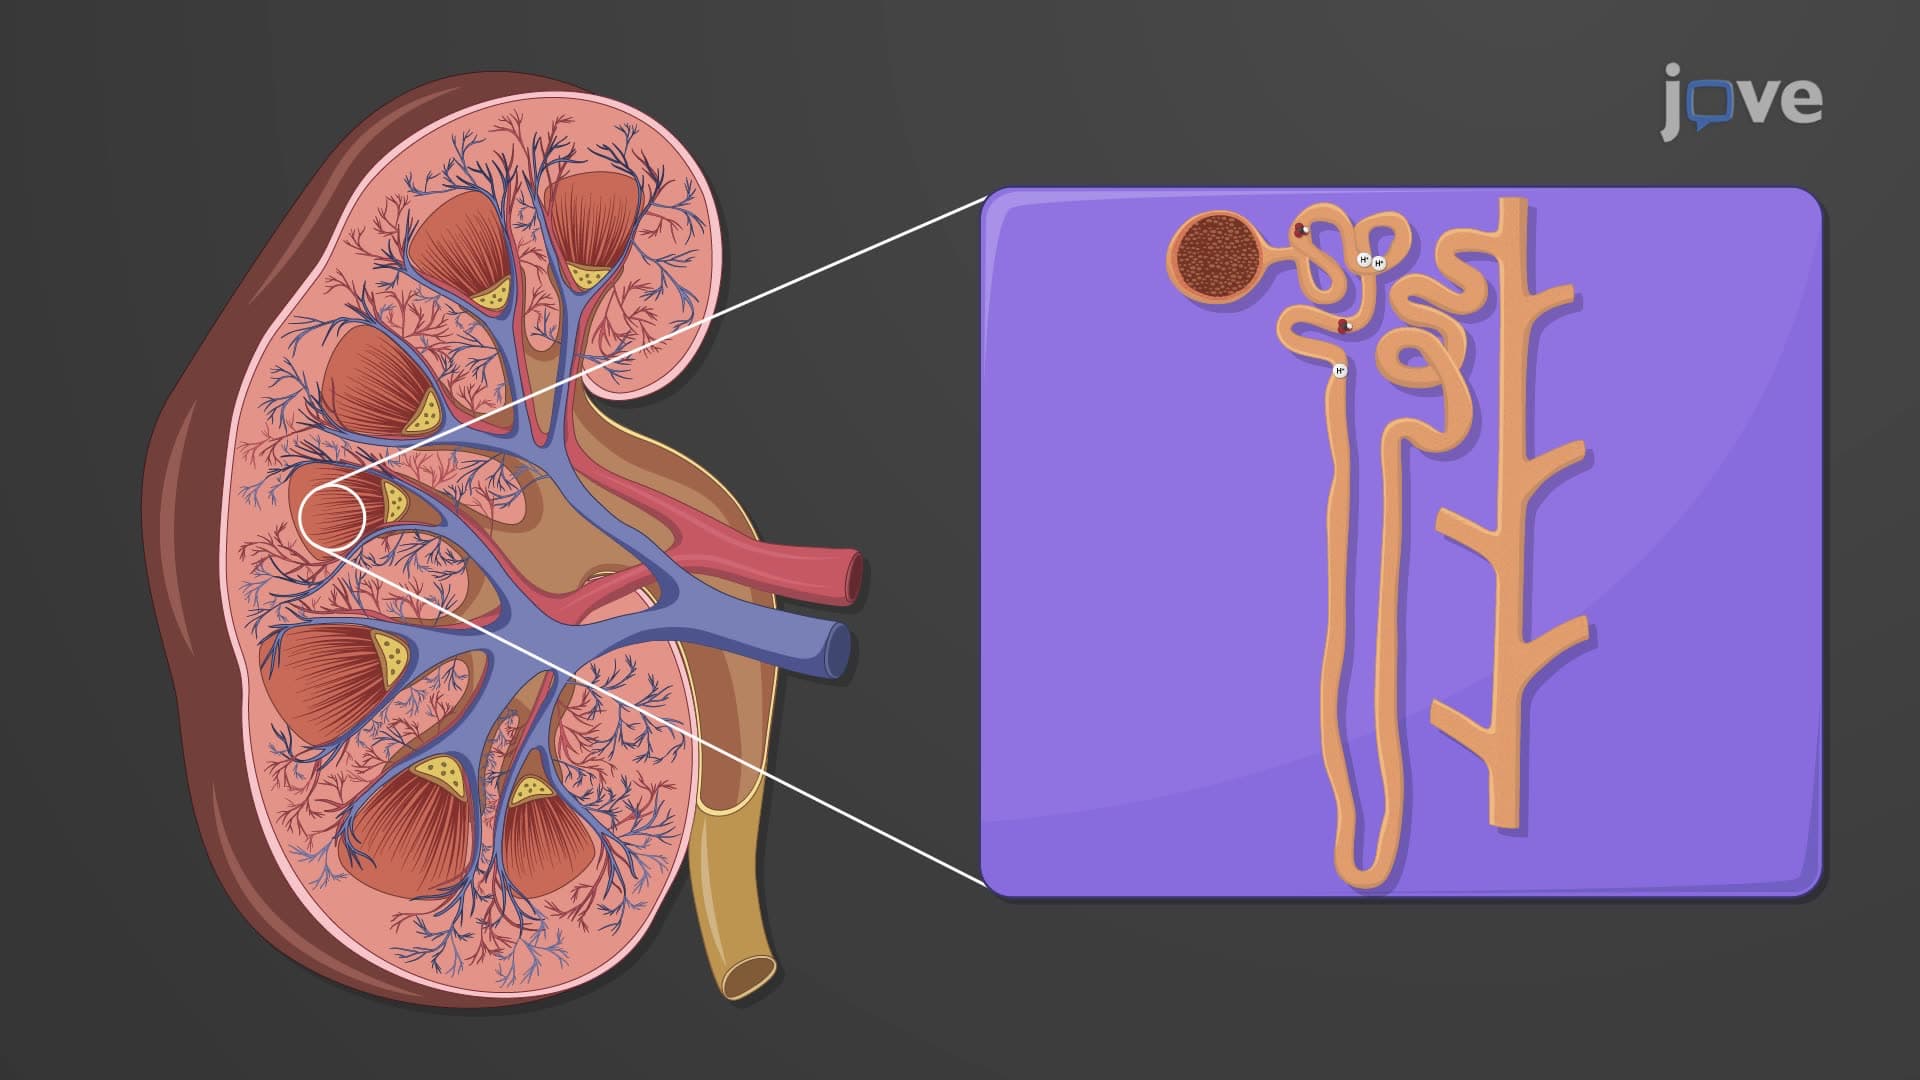
Renal Regulation of Acid-Base Balance
In the kidneys, cells within the proximal convoluted tubules (PCT) and the collecting ducts secrete hydrogen ions (H+) into the tubular fluid. Specifically, in the PCT, Na+/H+ antiporters secrete H+ while reabsorbing Na+.
However, the intercalated cells in...