Related Experiment Video
Updated: Jul 13, 2025

Author Spotlight: Decoding Mitochondrial Aging
Published on: June 30, 2023
Phenotype and natural history of mitochondrial membrane protein-associated neurodegeneration
Vassilena Iankova1, Peter Sparber2, Mohammad Rohani3
1Department of Neurology with Friedrich-Baur-Institute, University Hospital of Ludwig-Maximilians-Universität München, 80336 Munich, Germany.

Abstract:
Mitochondrial membrane protein-associated neurodegeneration (MPAN) is an ultraorphan neurogenetic disease from the group of neurodegeneration with brain iron accumulation (NBIA) disorders. Here we report cross-sectional and longitudinal data to define the phenotype, to assess disease progression and to estimate sample sizes for clinical trials. We enrolled patients with genetically confirmed MPAN from the Treat Iron-Related Childhood-Onset Neurodegeneration (TIRCON) registry and cohort study, and from additional sites. Linear mixed-effect modelling (LMEM) was used to calculate annual progression rates for the Unified Parkinson's Disease Rating Scale (UPDRS), Barry-Albright Dystonia (BAD) scale, Schwab and England Activities of Daily Living (SE-ADL) scale and the Pediatric Quality of Life Inventory (PedsQL). We investigated 85 MPAN patients cross-sectionally, with functional outcome data collected in 45. Median age at onset was 9 years and the median diagnostic delay was 5 years. The most common findings were gait disturbance (99%), pyramidal involvement (95%), dysarthria (90%), vision disturbances (82%), with all but dysarthria presenting early in the disease course. After 16 years with the disease, 50% of patients were wheelchair dependent. LMEM showed an annual progression rate of 4.5 points in total UPDRS. The total BAD scale score showed no significant progression over time. The SE-ADL scale and the patient- and parent-reported PedsQL showed a decline of 3.9%, 2.14 and 2.05 points, respectively. No patient subpopulations were identified based on longitudinal trajectories. Our cross-sectional results define the order of onset and frequency of symptoms in MPAN, which will inform the diagnostic process, help to shorten diagnostic delay and aid in counselling patients, parents and caregivers. Our longitudinal findings define the natural history of MPAN, reveal the most responsive outcomes and highlight the need for an MPAN-specific rating approach. Our sample size estimations inform the design of upcoming clinical trials.
Insights
Mitochondrial membrane protein-associated neurodegeneration (MPAN) is a rare neurological disorder. This study defines MPAN
Area of Science:
- Neurogenetics
- Neurodegenerative diseases
- Rare diseases
Background:
- Mitochondrial membrane protein-associated neurodegeneration (MPAN) is an ultra-rare neurogenetic disorder within the neurodegeneration with brain iron accumulation (NBIA) group.
- Understanding the natural history and progression of MPAN is crucial for developing effective therapeutic strategies and clinical trial designs.
- Current knowledge regarding MPAN's clinical phenotype, disease progression, and optimal outcome measures for research is limited.
Purpose of the Study:
- To define the clinical phenotype and disease progression of Mitochondrial membrane protein-associated neurodegeneration (MPAN) using cross-sectional and longitudinal data.
- To assess the rate of disease progression in MPAN patients using standardized rating scales.
- To estimate sample sizes required for future clinical trials in MPAN.
Main Methods:
- Cross-sectional and longitudinal data were collected from 85 MPAN patients enrolled in the TIRCON registry and cohort study, with functional outcome data from 45 patients.
- Linear mixed-effect modeling (LMEM) was employed to calculate annual progression rates for the Unified Parkinson's Disease Rating Scale (UPDRS), Barry-Albright Dystonia (BAD) scale, Schwab and England Activities of Daily Living (SE-ADL) scale, and the Pediatric Quality of Life Inventory (PedsQL).
- Analysis included assessment of symptom onset, frequency, and progression over time to characterize the disease's natural history.
Main Results:
- The median age of onset for MPAN was 9 years, with a median diagnostic delay of 5 years. Common early symptoms include gait disturbance (99%), pyramidal involvement (95%), and dysarthria (90%).
- After 16 years of disease, 50% of patients required wheelchair dependency. LMEM indicated an average annual progression rate of 4.5 points on the total UPDRS.
- The SE-ADL scale and PedsQL showed significant declines (3.9%, 2.14, and 2.05 points annually, respectively), while the BAD scale showed no significant progression. No distinct patient subpopulations were identified.
Conclusions:
- Cross-sectional findings clarify the order and frequency of MPAN symptoms, aiding diagnosis, shortening delays, and improving patient/caregiver counseling.
- Longitudinal data establish the natural history of MPAN, identify responsive outcomes for trials, and highlight the need for MPAN-specific assessment tools.
- Sample size estimations derived from this study will guide the design of upcoming clinical trials for MPAN.
More Related Videos
 06:07
Author Spotlight: Establishing a New Fluorescence-Based Protocol for In Vivo Mitochondrial Morphology Analysis in Parkinson's Disease
Published on: June 23, 2023
 07:32
Analyzing Mitochondrial Transport and Morphology in Human Induced Pluripotent Stem Cell-Derived Neurons in Hereditary Spastic Paraplegia
Published on: February 9, 2020
Related Concept Videos
 01:45
Mitochondrial Membranes
 01:39
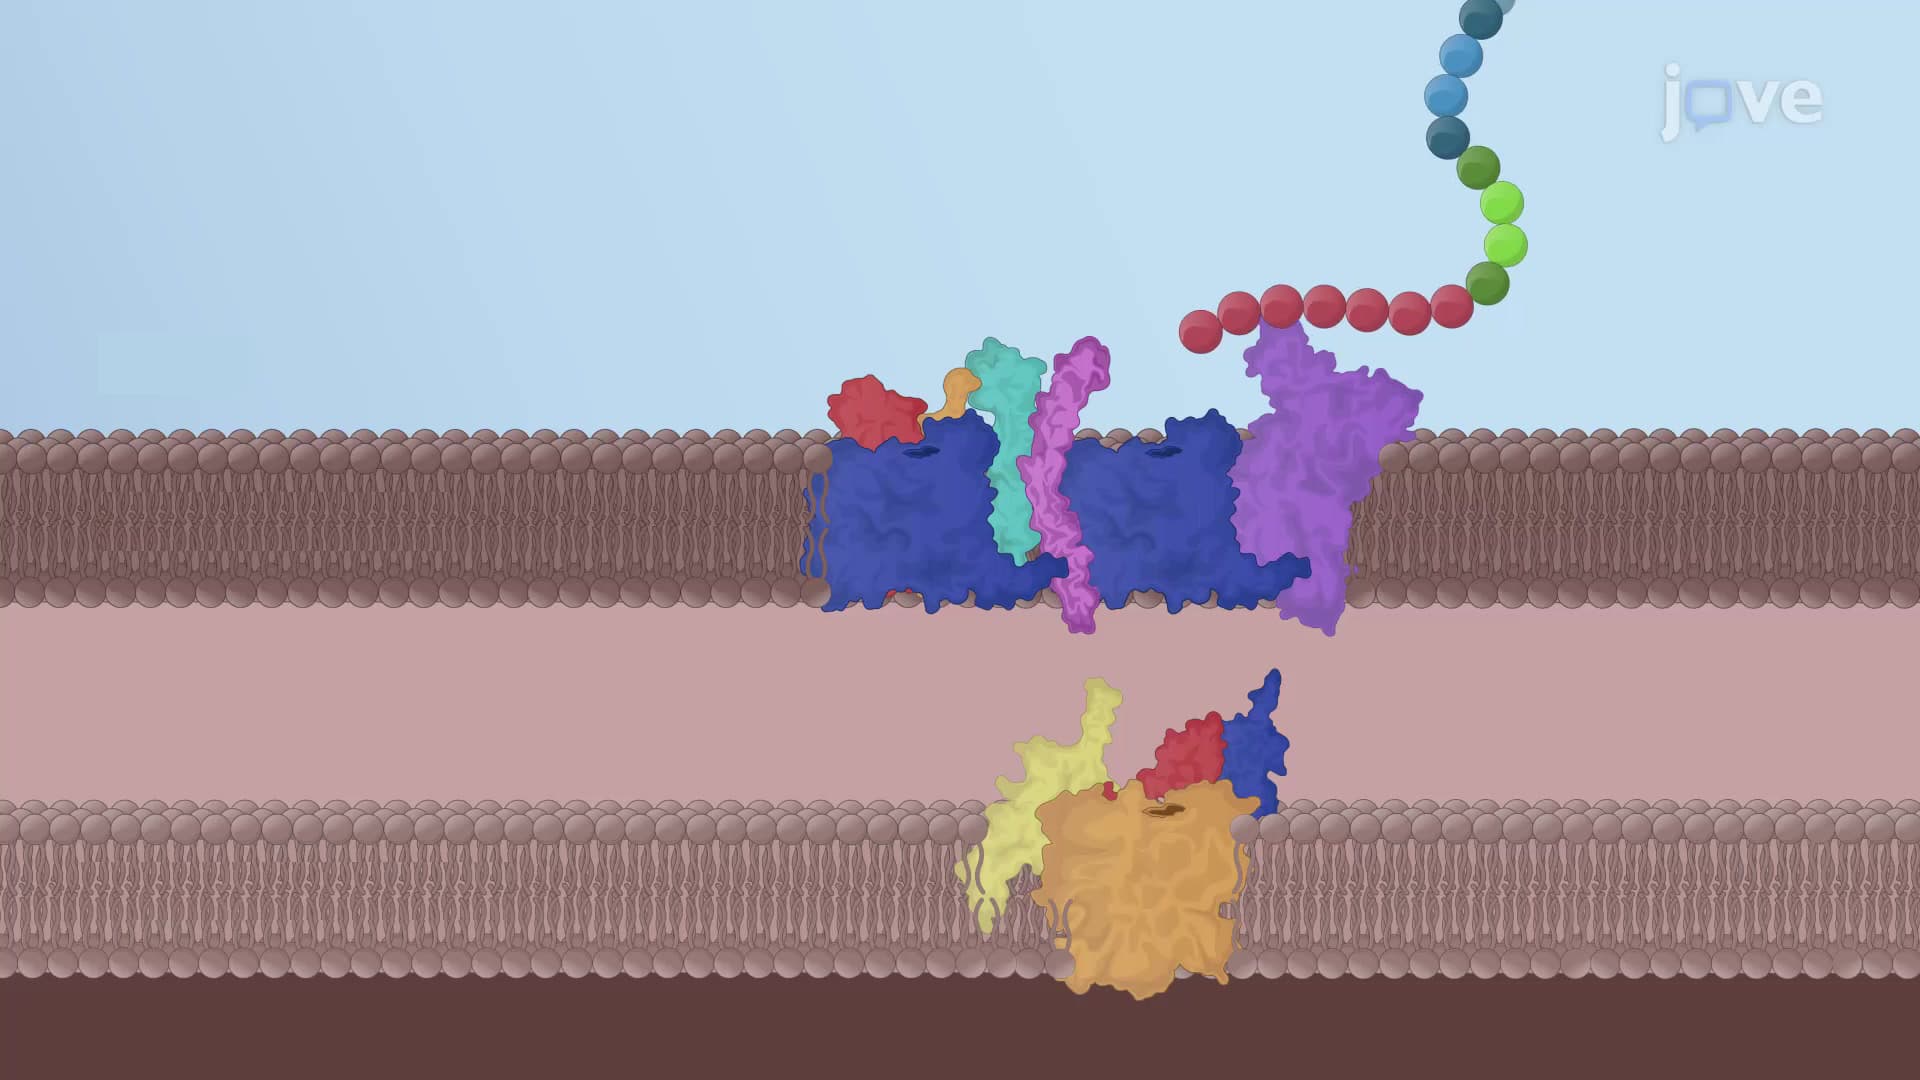
Mitochondrial Precursor Proteins
Most of the mitochondrial...
 01:48
ATP Synthase: Mechanism
 01:19
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 02:59
Animal Mitochondrial Genetics
 01:39
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...