Related Experiment Video
Updated: Jul 13, 2025

Interfacial Molecular-level Structures of Polymers and Biomacromolecules Revealed via Sum Frequency Generation Vibrational Spectroscopy
Published on: August 13, 2019
Multiphonon scattering formula of dynamic structure factors for classical Debye solids
Hikaru Kitamura1, Takaya Furukawa1
1Department of Physics, Kyoto University, Sakyo-ku, Kyoto 606-8502, Japan.

Abstract:
A semianalytic formula of the dynamic structure factor S(k,ω) for classical Debye solids over the entire wave-number (k) and frequency (ω) range is constructed by taking into account multiphonon thermal diffuse scattering up to infinite order. The formula adopts Gaussian approximations to the spatial and time decay of the multiphonon part of the displacement correlation function. Numerical illustrations for isotropic polycrystals reveal that, as k increases, sharp peaks due to one-phonon normal scattering in the hydrodynamic regime (k→0) are replaced by diffuse spectra consisting of umklapp scattering and multiphonon continuum; approach toward the ideal-gas spectra in the large-k limit is proven from analytic properties of the multiphonon term. When k coincides with a Bragg reflection point, total thermal diffuse scattering S_{TDS}(k,ω) exhibits a 1/ω divergence as ω→0, which in turn gives rise to a logarithmic enhancement of the corresponding static structure factor S_{TDS}(k). Overall accuracy of the theory is confirmed through the exact zeroth-order frequency-moment sum rule between S_{TDS}(k,ω) and S_{TDS}(k); agreement with the second-order sum rule is shown to be satisfactory except for the vicinity of the Debye cutoff region.
More Related Videos
 11:27
Studying Soft-matter and Biological Systems over a Wide Length-scale from Nanometer and Micrometer Sizes at the Small-angle Neutron Diffractometer KWS-2
Published on: December 8, 2016
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:32
The de Broglie Wavelength
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:18
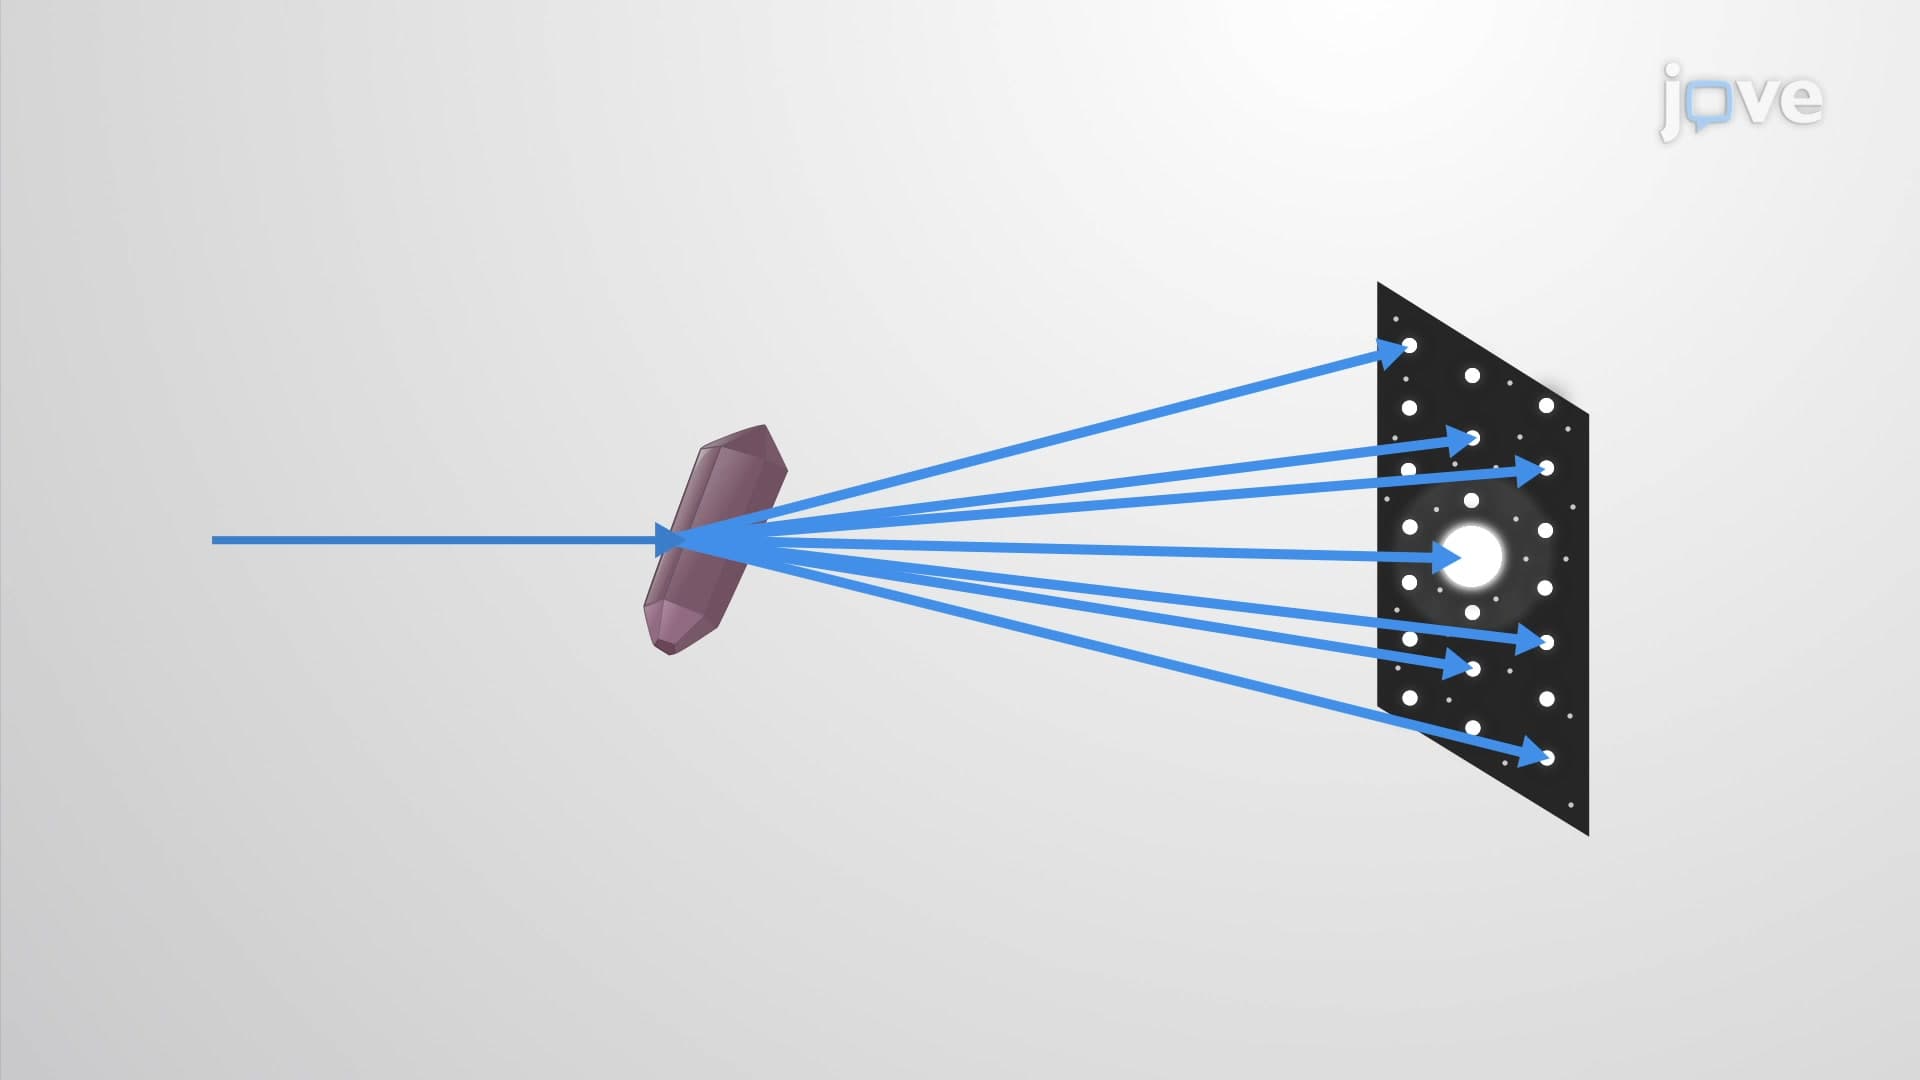
X-ray Crystallography
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 02:45
The Quantum-Mechanical Model of an Atom