Related Experiment Video
Updated: Jul 11, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Corrected density functional theory and the random phase approximation: Improved accuracy at little extra cost
1Yusuf Hamied Department of Chemistry, University of Cambridge, Cambridge, England.

Abstract:
We recently introduced an efficient methodology to perform density-corrected Hartree-Fock density functional theory [DC(HF)-DFT] calculations and an extension to it we called "corrected" HF DFT [C(HF)-DFT] [Graf and Thom, J. Chem. Theory Comput. 19 5427-5438 (2023)]. In this work, we take a further step and combine C(HF)-DFT, augmented with a straightforward orbital energy correction, with the random phase approximation (RPA). We refer to the resulting methodology as corrected HF RPA [C(HF)-RPA]. We evaluate the proposed methodology across various RPA methods: direct RPA (dRPA), RPA with an approximate exchange kernel, and RPA with second-order screened exchange. C(HF)-dRPA demonstrates very promising performance; for RPA with exchange methods, on the other hand, we often find over-corrections.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 01:10
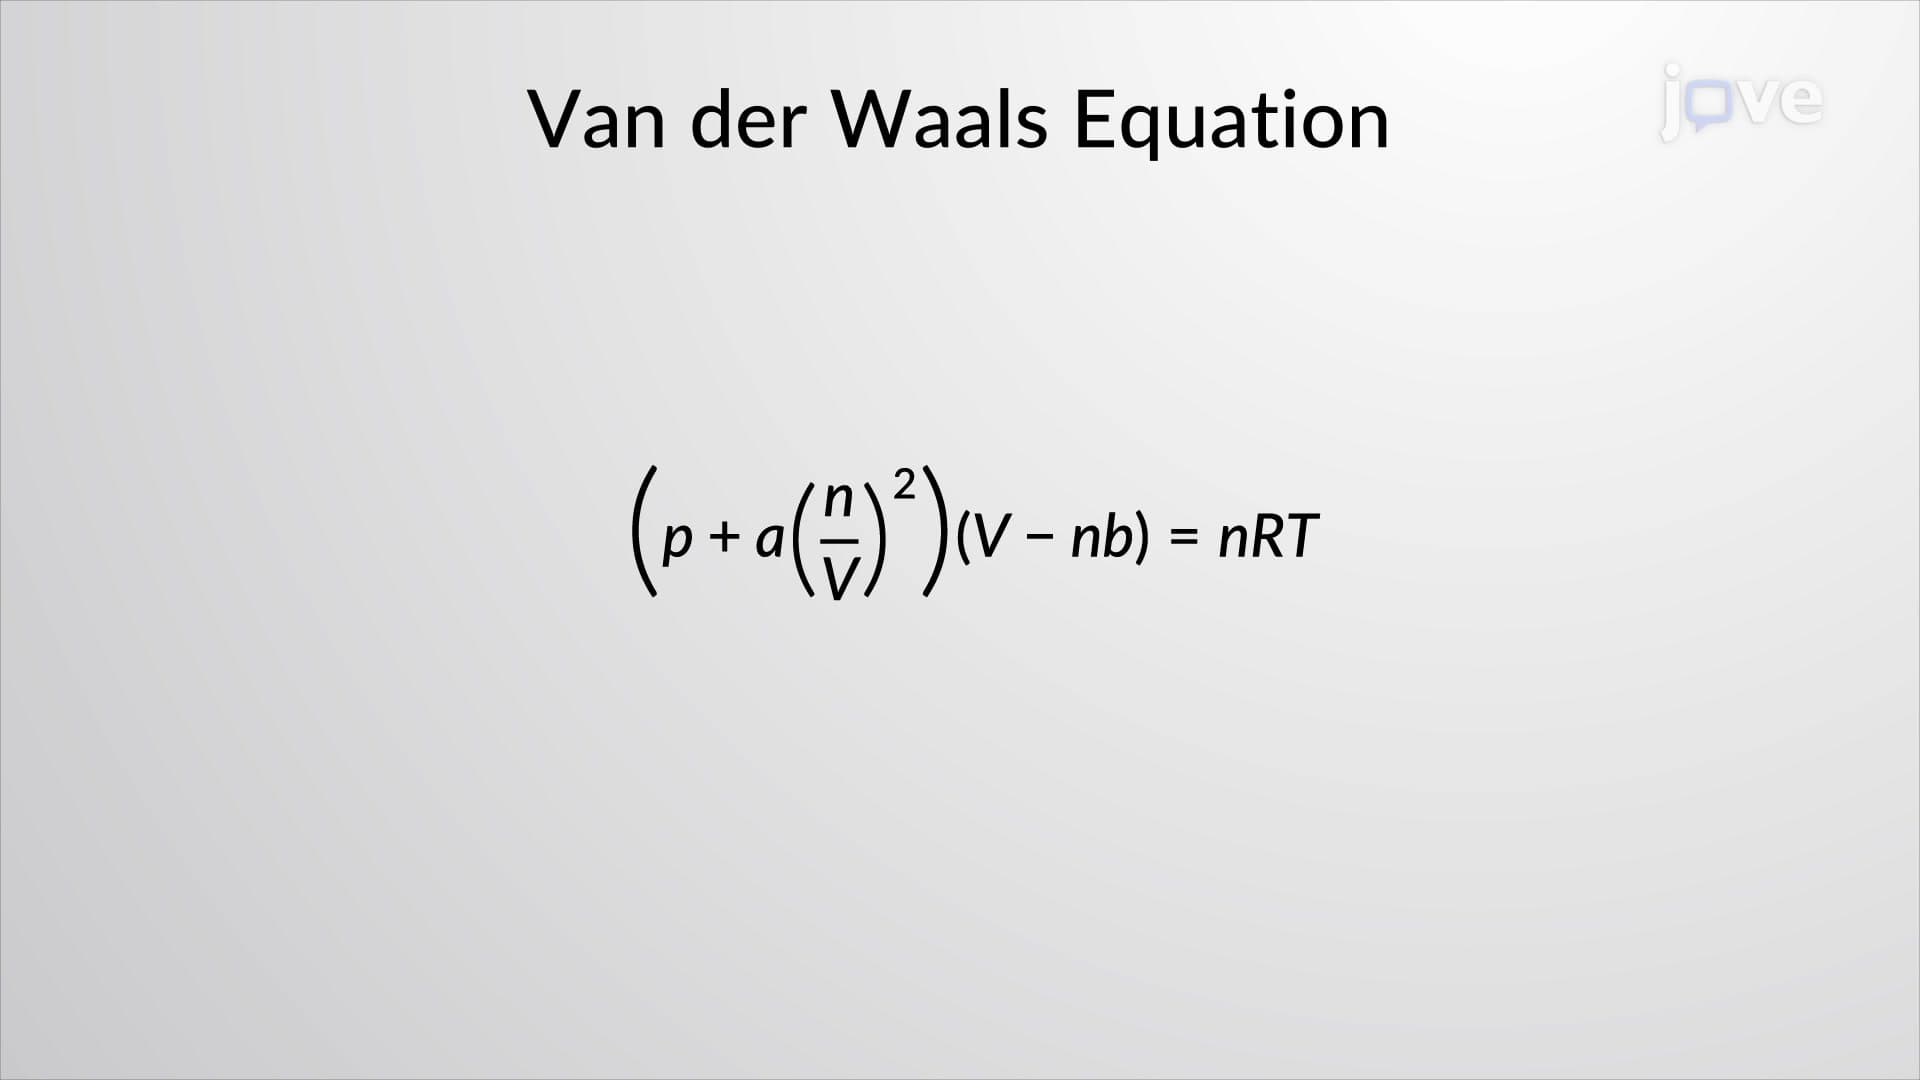
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:14
NMR Spectrometers: Resolution and Error Correction
 04:08
The Uncertainty Principle
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...