Related Experiment Video
Updated: Jul 11, 2025

Determination of Protein-ligand Interactions Using Differential Scanning Fluorimetry
Published on: September 13, 2014
Comparing Density Functional Theory Metal-Ligand Bond Dissociation Enthalpies with Experimental Solution-Phase
Edward N Brothers1, Ashfaq A Bengali2, Giovanni Scalmani1
1Gaussian, Inc., 340 Quinnipiac Street, Wallingford, Connecticut 06492, United States.

Abstract:
The predictive ability of density functional theory is fundamental to its usefulness in chemical applications. Recent work has compared solution-phase enthalpies of activation for metal-ligand bond dissociation to enthalpies of reaction for bond dissociation, and the present work continues those comparisons for 43 density functional methods. The results for ligand dissociation enthalpies of 30 metal-ligand complexes tested in this work reveal significant inadequacies of some functionals as well as challenges from the dispersion corrections to some functionals. The analysis presented here demonstrates the excellent performance of a recent density functional, M11plus, which contains nonlocal rung-3.5 correlation. We also find a good agreement between theory and experiment for some functionals without empirical dispersion corrections such as M06, r2SCAN, M06-L, and revM11, as well as good performance for some functionals with added dispersion corrections such as ωB97X-D (which always has a correction) and BLYP, B3LYP, CAM-B3LYP, and PBE0 when the optional dispersion corrections are added.
More Related Videos
 16:11
Thermochemical Studies of NiII and ZnII Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:13
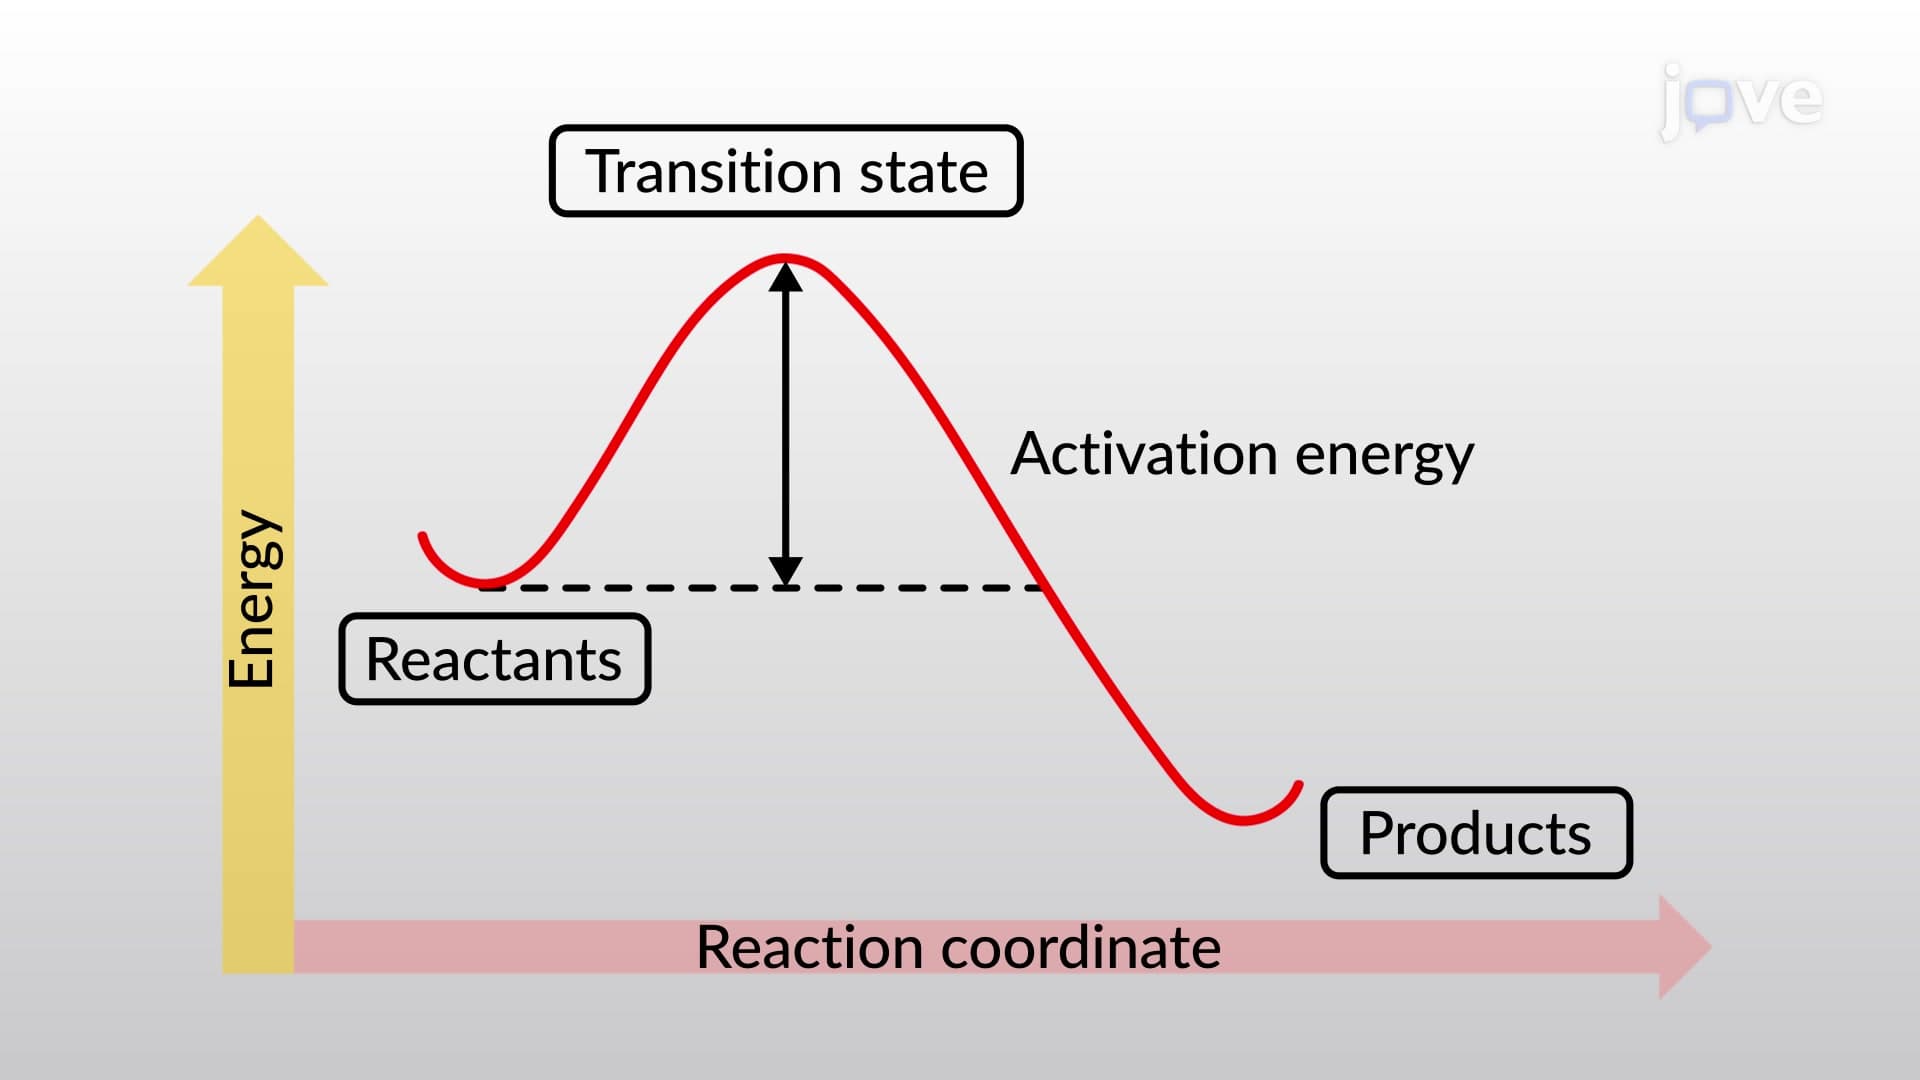
Bond Dissociation Energy and Activation Energy
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:07
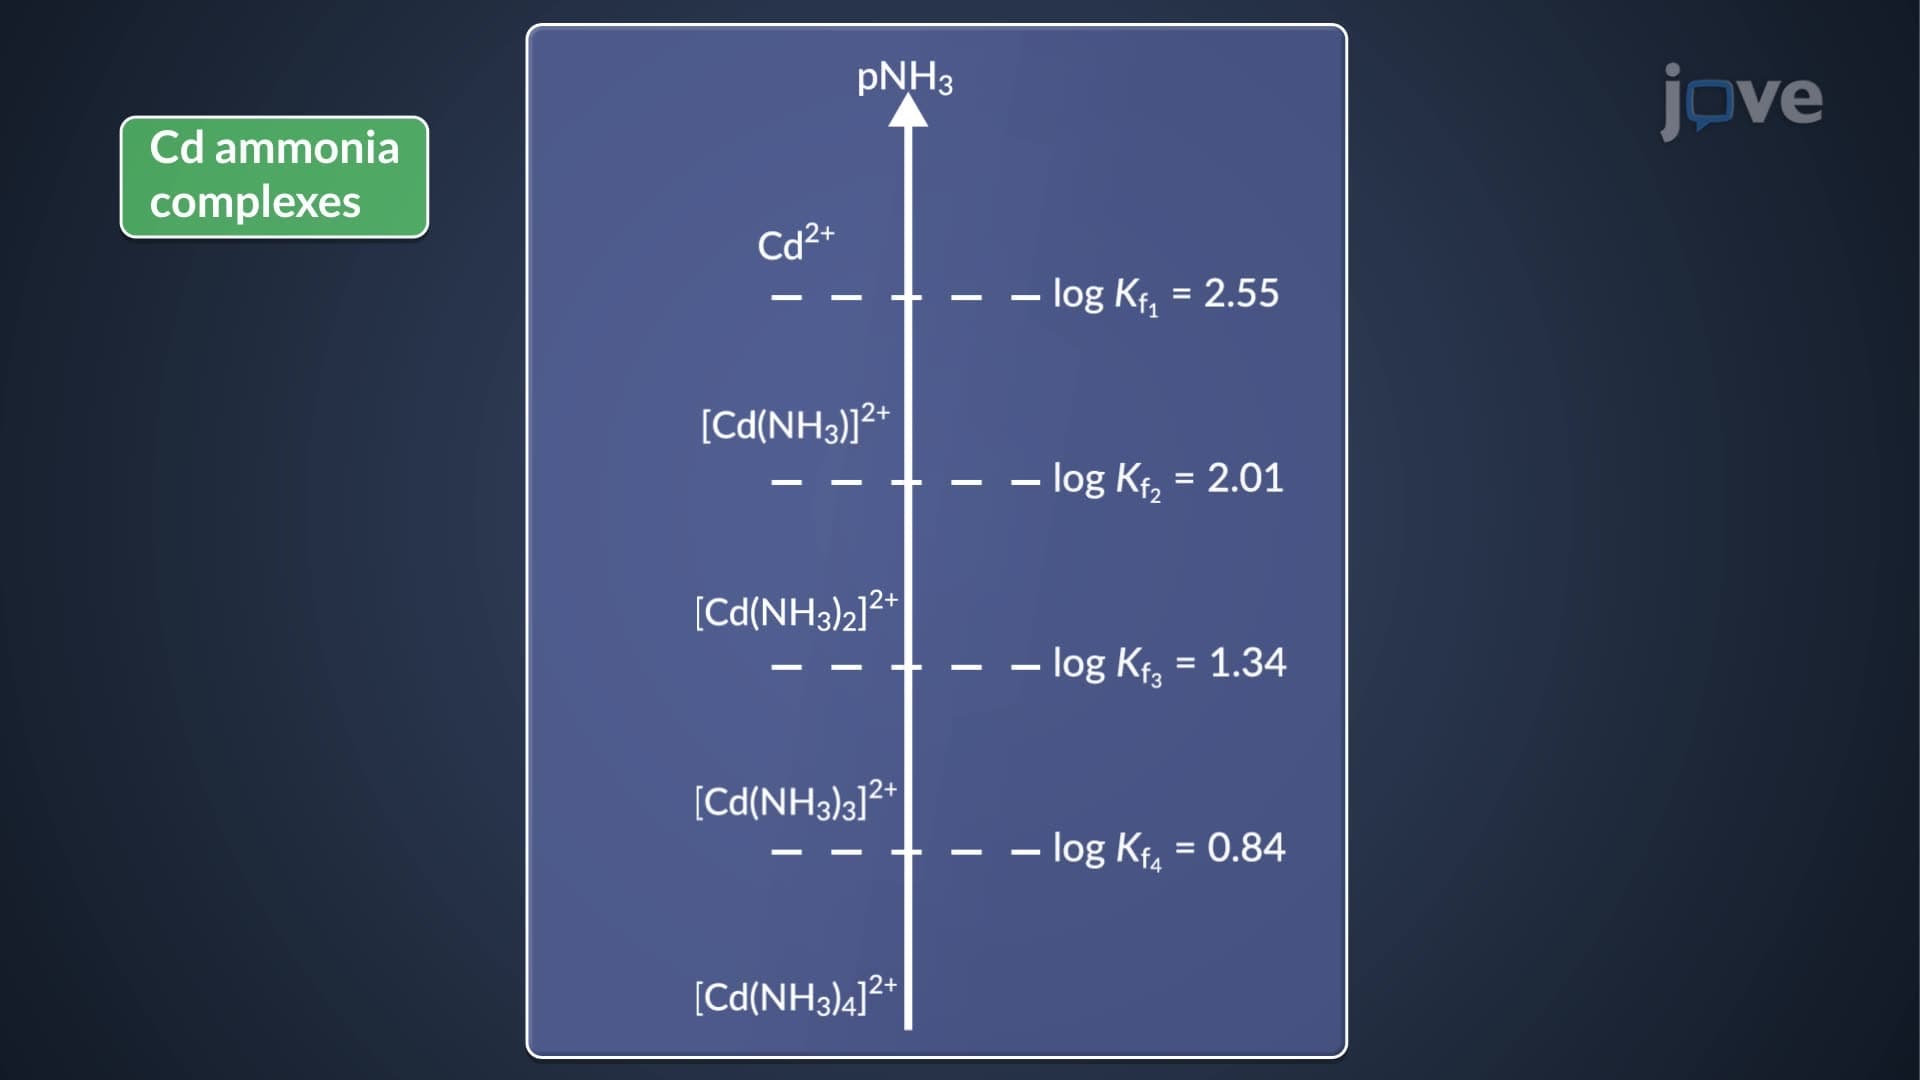
Ladder Diagrams: Complexation Equilibria
The formation constant, K1, for the formation of Cd(NH3)2+ complex from cadmium and ammonia is 3.55 × 10². Log K1 (i.e. pNH3) is 2.55, and...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:23
Complexation Equilibria: Overview
The equilibrium constant of the complexation reaction is represented as the formation constant...
 02:49
Bond Energies and Bond Lengths