Related Experiment Video
Updated: Jul 10, 2025


Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
The Impact of Data on Structure-Based Binding Affinity Predictions Using Deep Neural Networks
Pierre-Yves Libouban1, Samia Aci-Sèche1, Jose Carlos Gómez-Tamayo2
1Institute of Organic and Analytical Chemistry (ICOA), UMR7311, Université d'Orléans, CNRS, Pôle de Chimie rue de Chartres, 45067 Orléans, CEDEX 2, France.

Abstract:
Artificial intelligence (AI) has gained significant traction in the field of drug discovery, with deep learning (DL) algorithms playing a crucial role in predicting protein-ligand binding affinities. Despite advancements in neural network architectures, system representation, and training techniques, the performance of DL affinity prediction has reached a plateau, prompting the question of whether it is truly solved or if the current performance is overly optimistic and reliant on biased, easily predictable data. Like other DL-related problems, this issue seems to stem from the training and test sets used when building the models. In this work, we investigate the impact of several parameters related to the input data on the performance of neural network affinity prediction models. Notably, we identify the size of the binding pocket as a critical factor influencing the performance of our statistical models; furthermore, it is more important to train a model with as much data as possible than to restrict the training to only high-quality datasets. Finally, we also confirm the bias in the typically used current test sets. Therefore, several types of evaluation and benchmarking are required to understand models' decision-making processes and accurately compare the performance of models.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 06:50
Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:18
The Equilibrium Binding Constant and Binding Strength
 02:04
Protein-protein Interfaces
 01:22
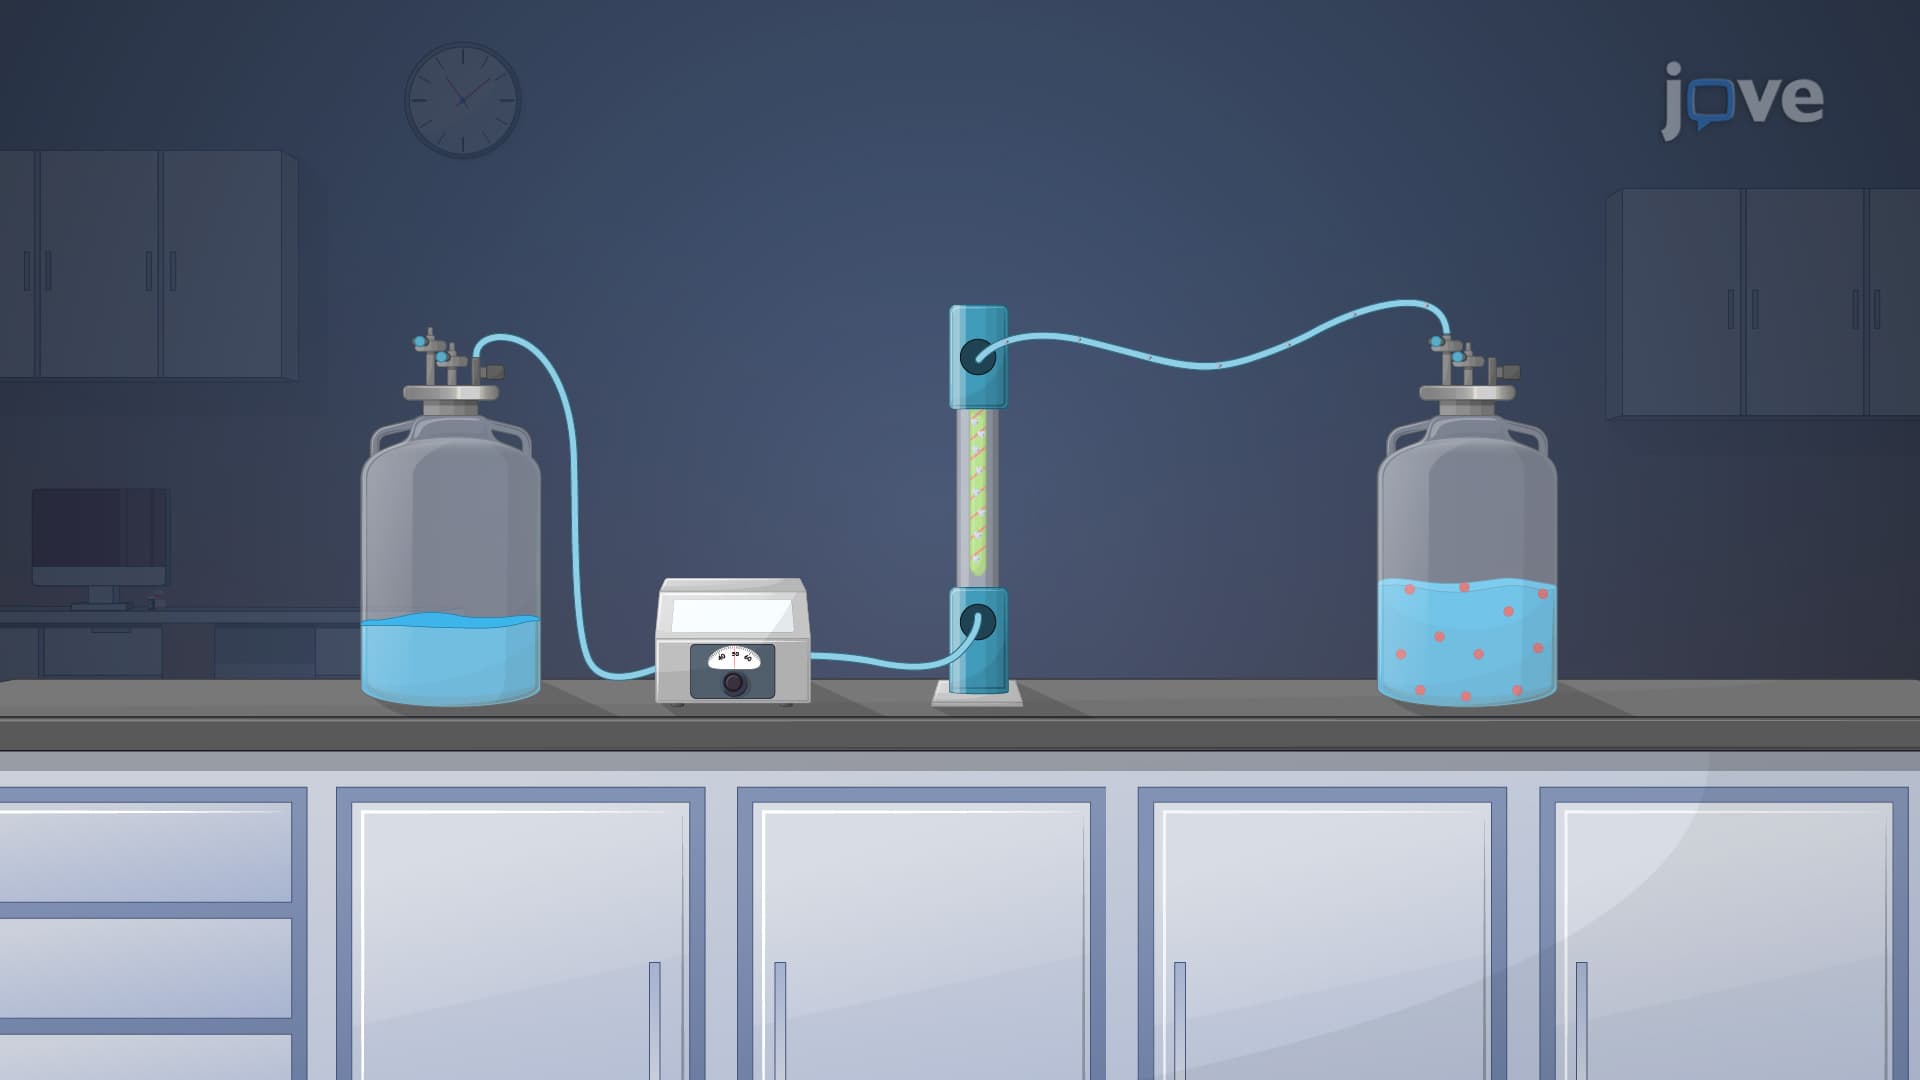
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:24
Protein Organization
The primary structure of a protein is its amino acid sequence....