

Single-chain simulation of Ising density functional theory for weak polyelectrolytes
Alejandro Gallegos1, Marcus Müller2, Jianzhong Wu1
1Department of Chemical and Environmental Engineering, University of California, Riverside, California 92521, USA.

Abstract:
Conventional theories of weak polyelectrolytes are either computationally prohibitive to account for the multidimensional inhomogeneity of polymer ionization in a liquid environment or oversimplistic in describing the coupling effects of ion-explicit electrostatic interactions and long-range intrachain correlations. To bridge this gap, we implement the Ising density functional theory (iDFT) for ionizable polymer systems using the single-chain-in-mean-field algorithm. The single-chain-in-iDFT (sc-iDFT) shows significant improvements over conventional mean-field methods in describing segment-level dissociation equilibrium, specific ion effects, and long-range intrachain correlations. With an explicit consideration of the fluctuations of polymer configurations and the position-dependent ionization of individual polymer segments, sc-iDFT provides a faithful description of the structure and thermodynamic properties of inhomogeneous weak polyelectrolyte systems across multiple length scales.
More Related Videos
 07:31
Author Spotlight: Advancing Cell Membrane Biophysics - Exploring Interactions and Challenges Through Experimental and Computational Approaches
Published on: September 1, 2023
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 01:08
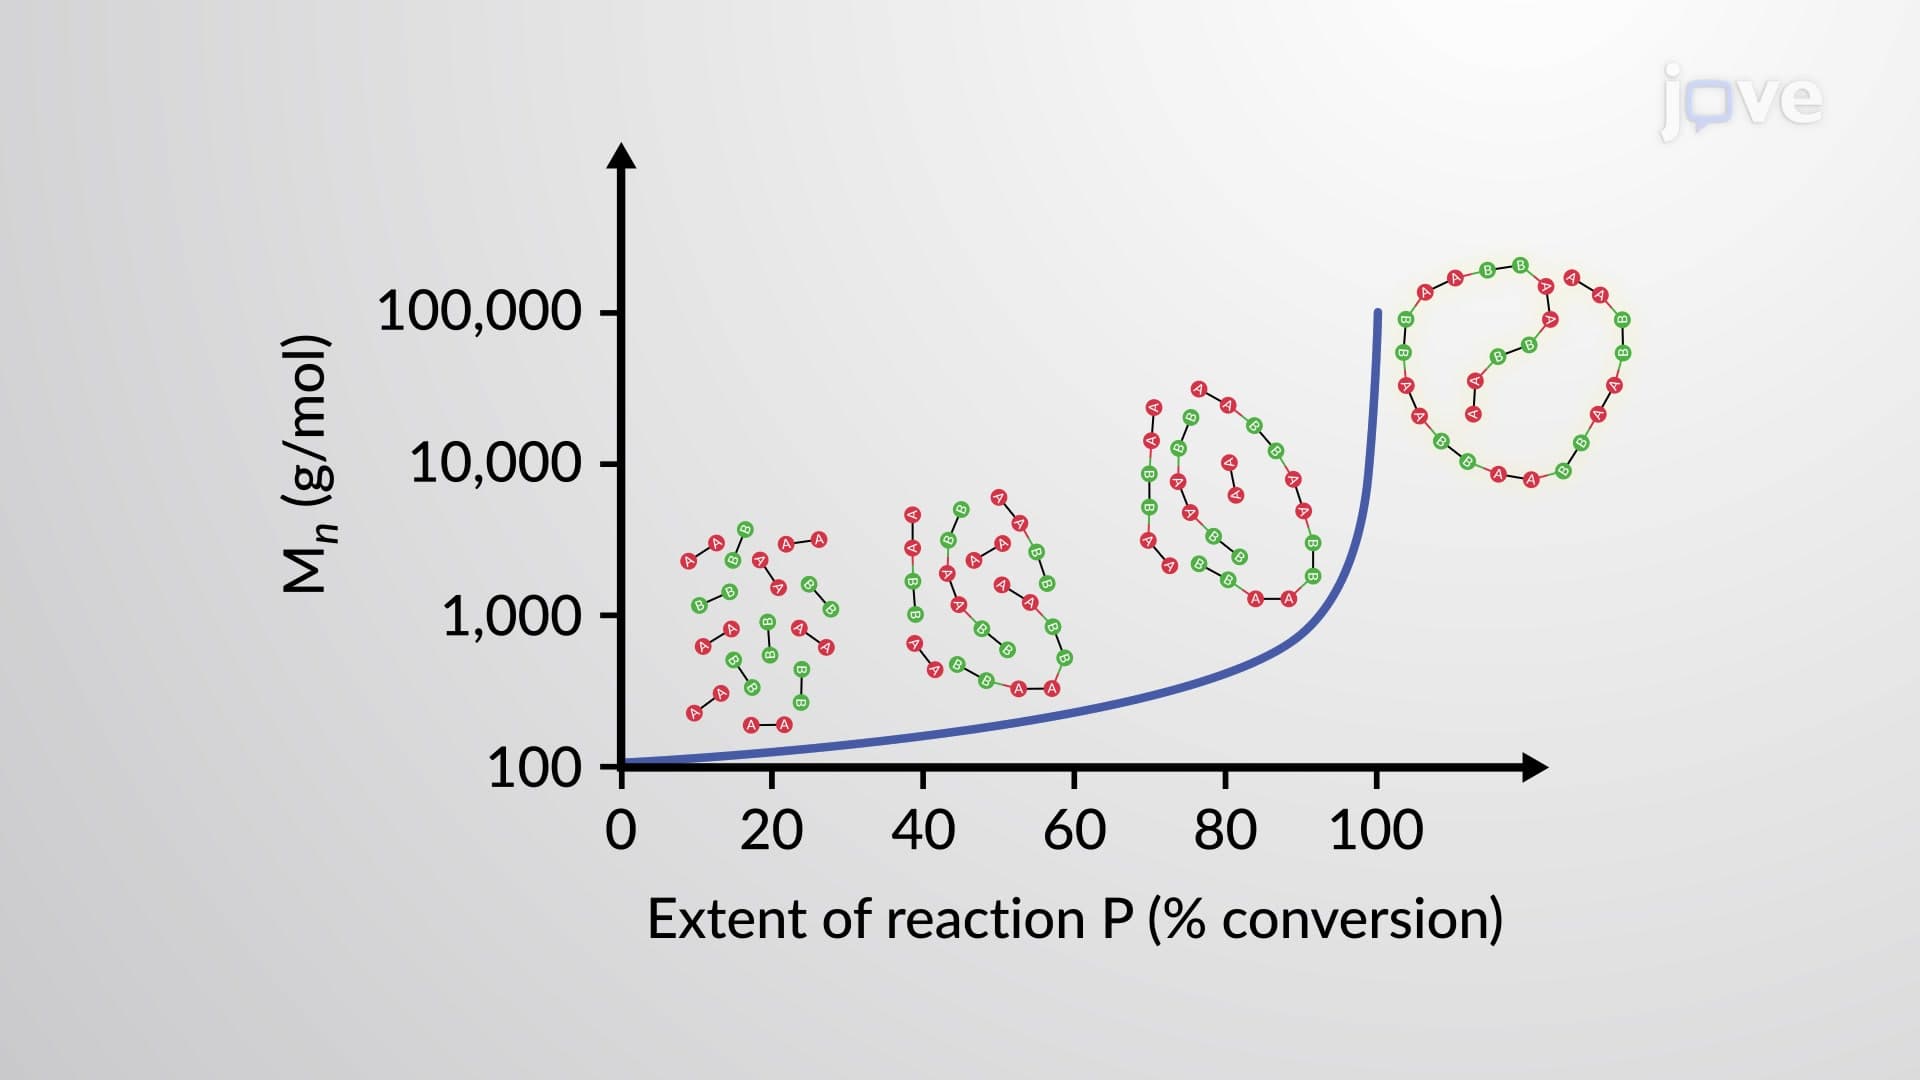
Molecular Weight of Step-Growth Polymers
As the step-growth polymerization involves step-wise condensation of monomers, the molecular weight also builds up eventually. Consequently, high molecular weight polymers are obtained at the late stages of the polymerization, where 99% of monomers have been consumed.
The extent of the...
 01:10
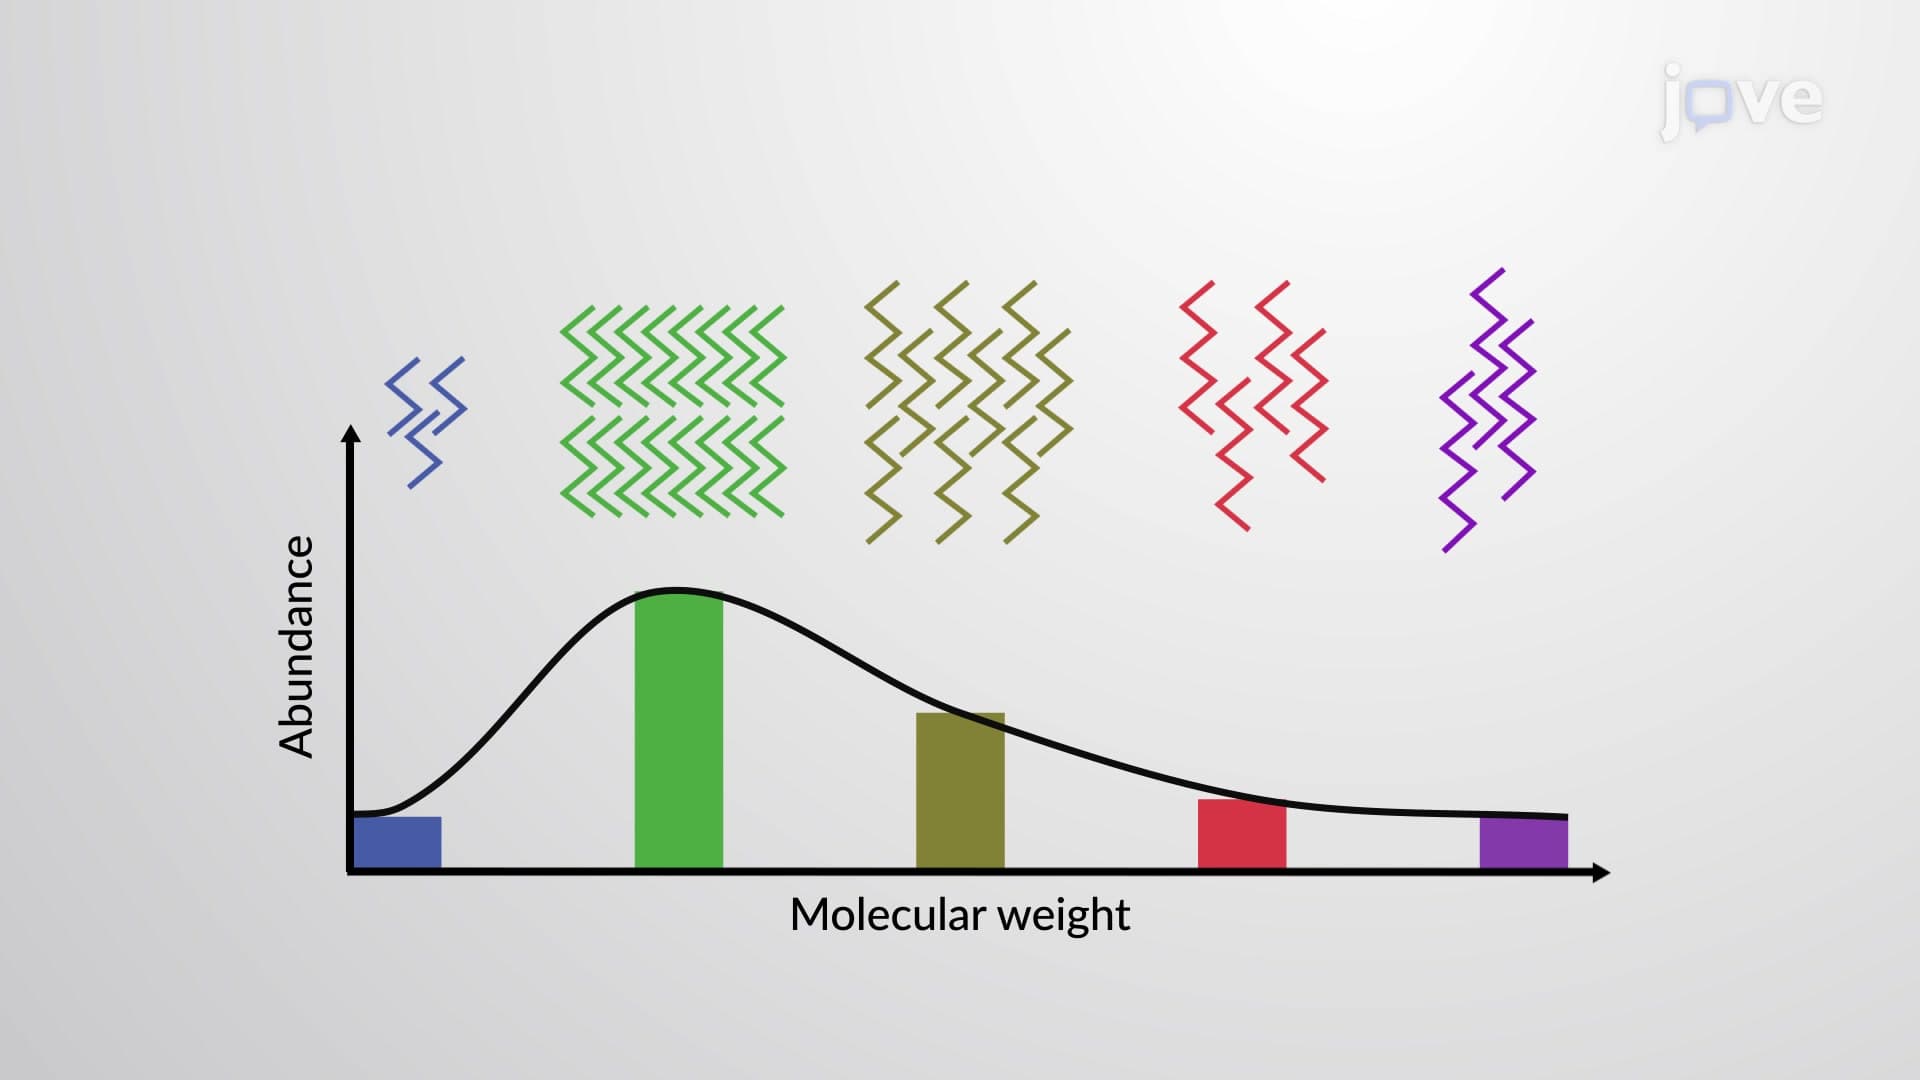
Polymers: Molecular Weight Distribution