Related Experiment Video
Updated: Jul 8, 2025
Genotyping Single Nucleotide Polymorphisms in the Mitochondrial Genome by Pyrosequencing
Published on: February 10, 2023
Genetic, metabolic and clinical delineation of an MRPS23-associated mitochondrial disorder
Chupong Ittiwut1,2, Rungnapa Ittiwut1,2, Chulaluck Kuptanon3
1Center of Excellence for Medical Genomics, Department of Pediatrics, Faculty of Medicine, Chulalongkorn University, Bangkok, Thailand.

Abstract:
MRPS23 is a nuclear gene encoding a mitochondrial ribosomal protein. A patient with a mitochondrial disorder was found to carry a variant in MRPS23. More cases are necessary to establish MRPS23 as a mitochondrial disease gene. Of 5134 exomes performed in our center, we identified five independent patients who had similar clinical manifestations and were homozygous for the same germline variant c.119C>T; p.P40L in MRPS23. Detailed clinical findings, mitochondrial enzyme activity assays from cultured skin fibroblasts, PCR-Sanger-sequencing, and variant age estimation were performed. Their available family members were also studied. Eight members homozygous for the MRPS23 p.P40L were identified. All were from Hmong hilltribe. Seven presented with alteration of consciousness and recurrent vomiting, while the eighth who was a younger brother of a proband was found pre-symptomatically. Patients showed delayed growth and development, hearing impairment, hypoglycemia, lactic acidosis, and liver dysfunction. In vitro assays of cultured fibroblasts showed combined respiratory chain complex deficiency with low activities of complexes I and IV. PCR-Sanger-sequencing confirmed the variant, which was estimated to have occurred 1550 years ago. These results establish the MRPS23-associated mitochondrial disorder inherited in an autosomal recessive pattern and provide insight into its clinical and metabolic features.
More Related Videos

Related Concept Videos
 01:20
Inborn Errors of Metabolism
 01:39
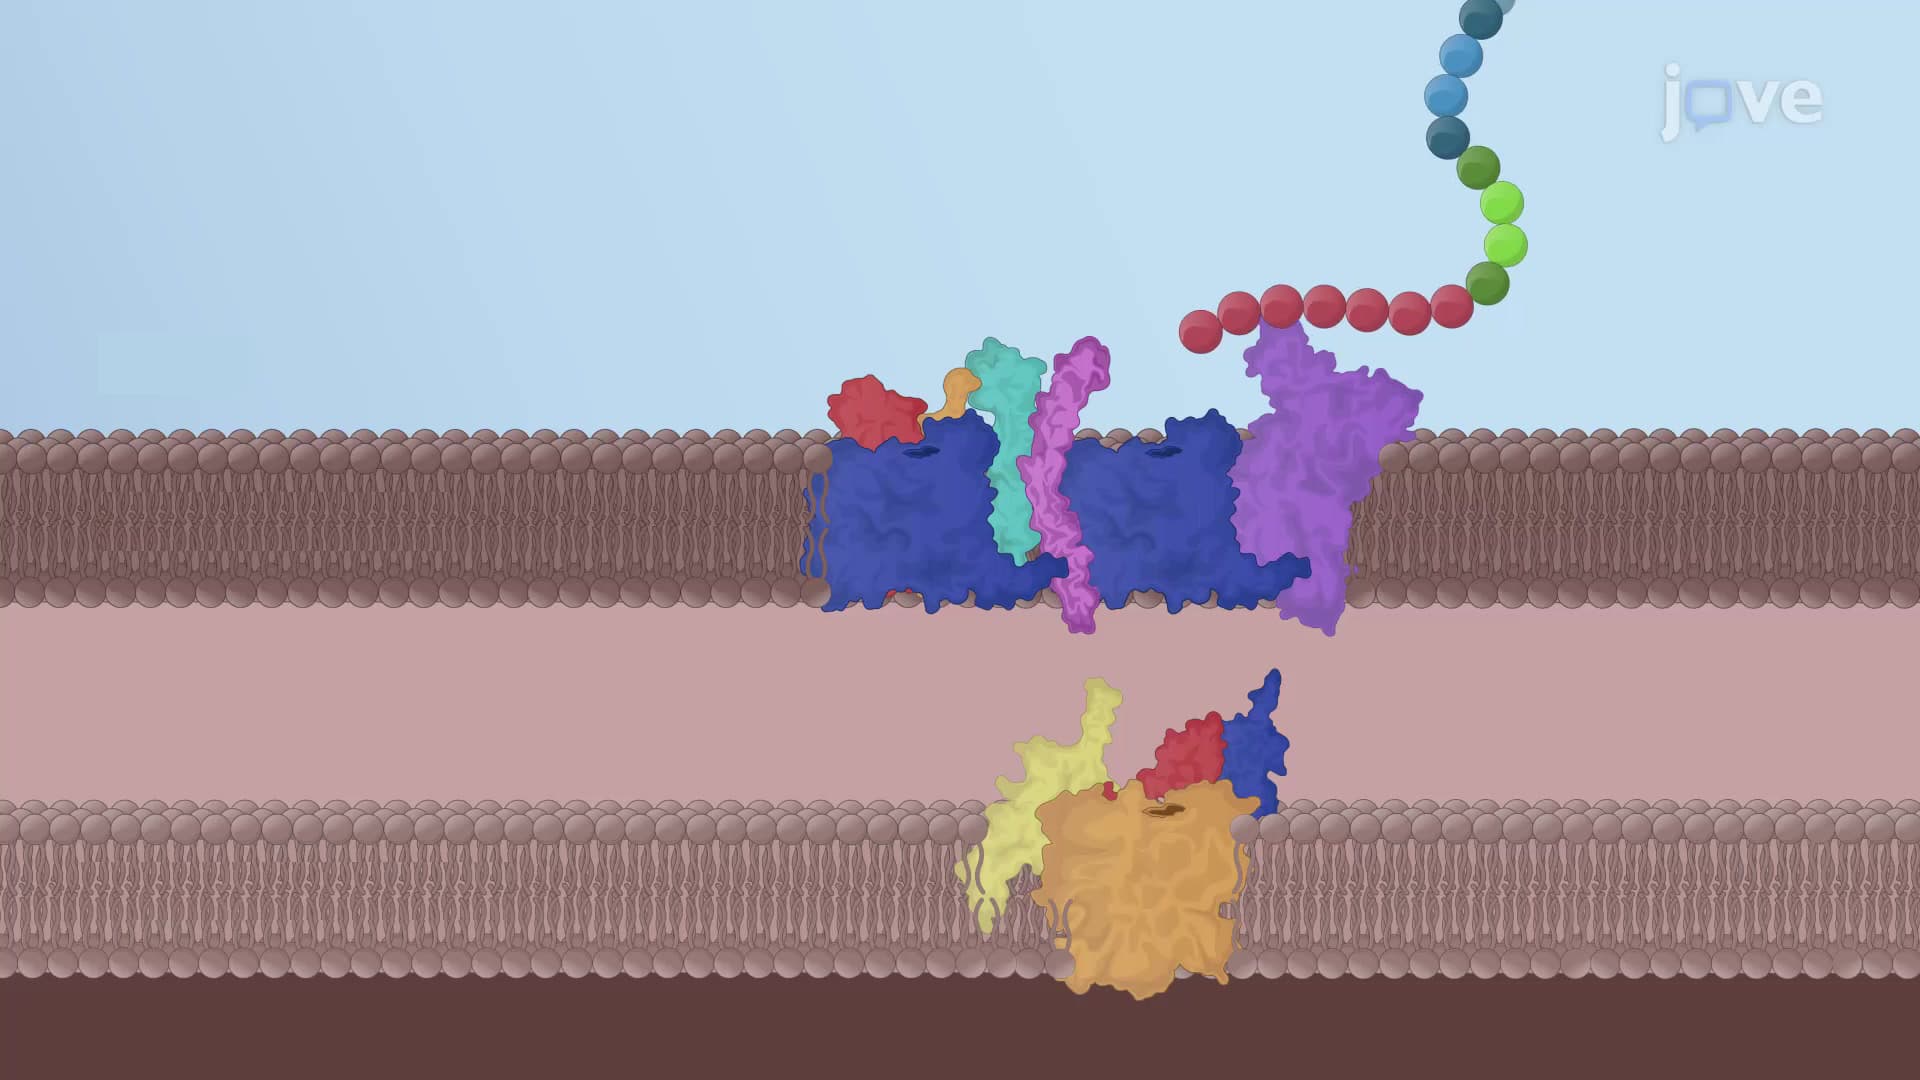
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 02:59
Animal Mitochondrial Genetics
 01:19
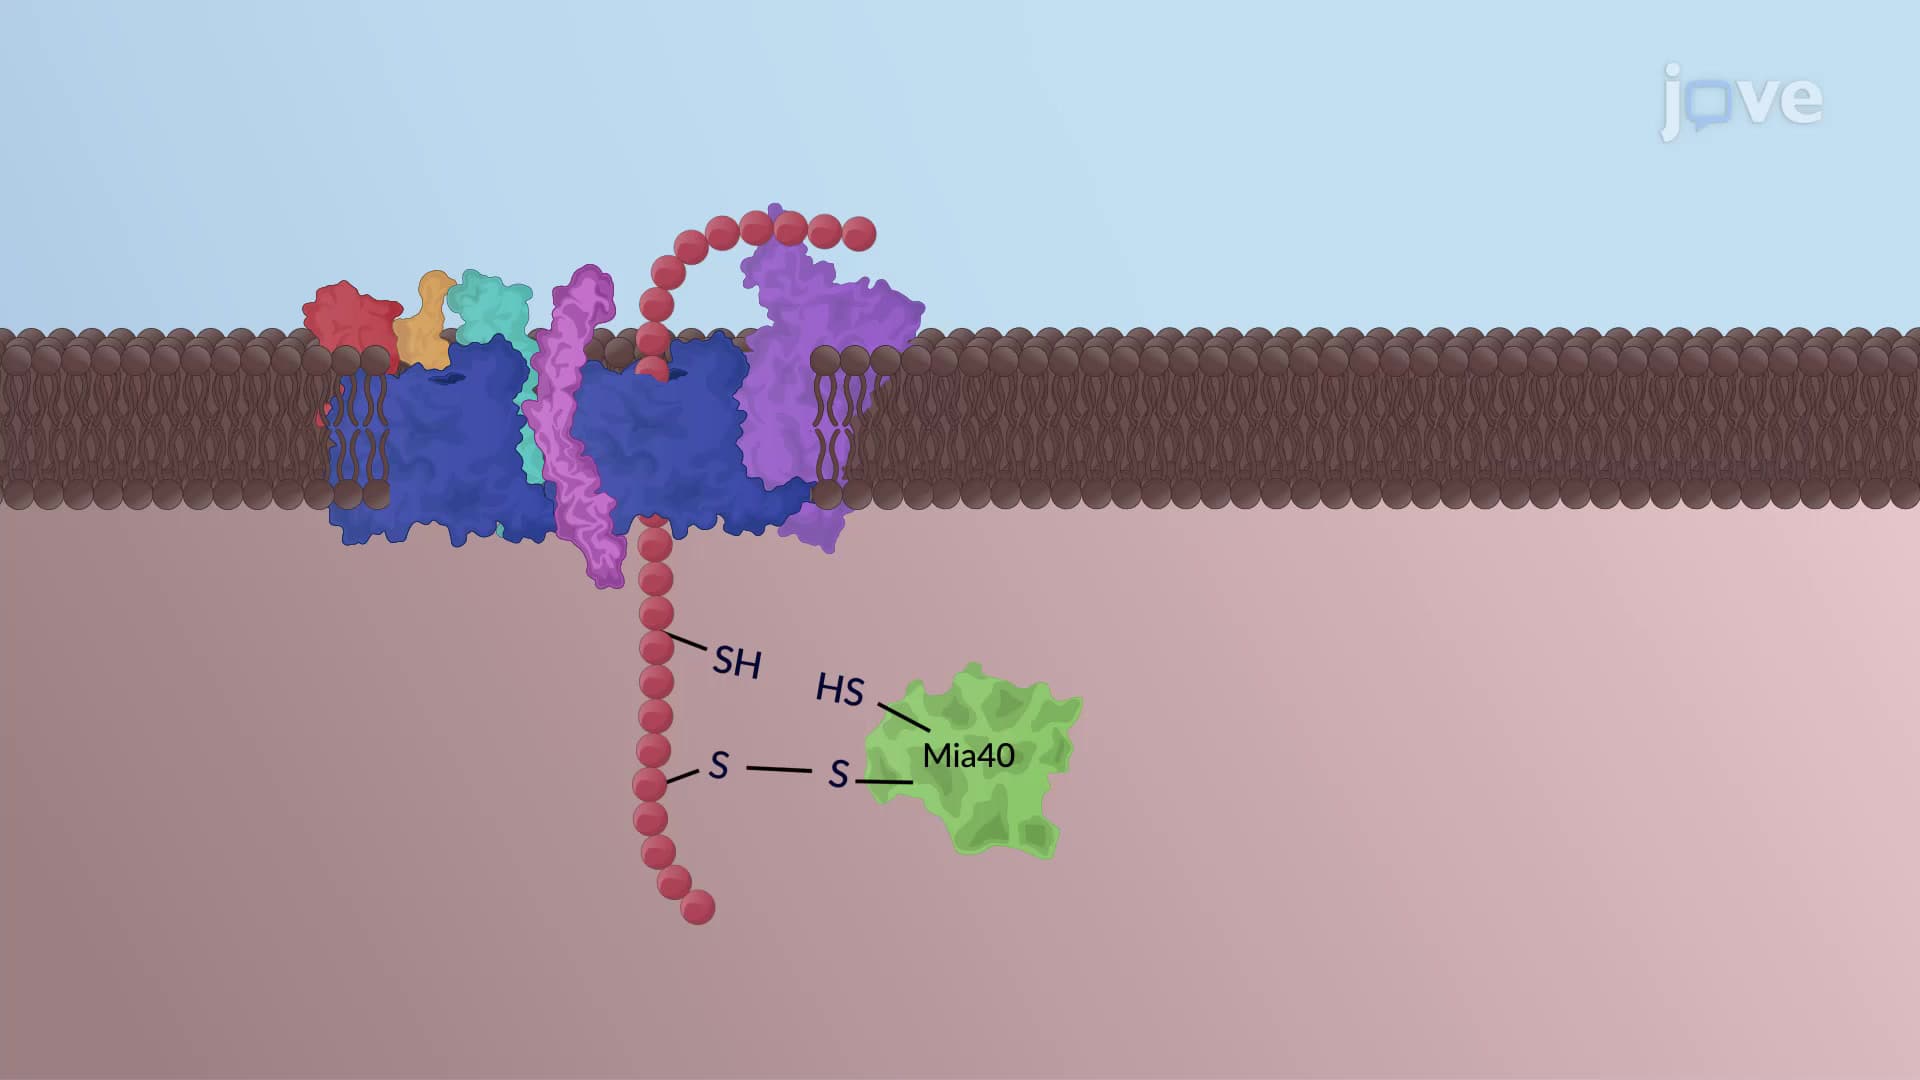
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:39
Mitochondrial Precursor Proteins
Most of the mitochondrial...