Interpreting forces as deep learning gradients improves quality of predicted protein structures

Jonathan Edward King1, David Ryan Koes2
1Joint PhD Program in Computational Biology, Carnegie Mellon University-University of Pittsburgh, Pittsburgh, Pennsylvania.

Abstract:
Protein structure predictions from deep learning models like AlphaFold2, despite their remarkable accuracy, are likely insufficient for direct use in downstream tasks like molecular docking. The functionality of such models could be improved with a combination of increased accuracy and physical intuition. We propose a new method to train deep learning protein structure prediction models using molecular dynamics force fields to work toward these goals. Our custom PyTorch loss function, OpenMM-Loss, represents the potential energy of a predicted structure. OpenMM-Loss can be applied to any all-atom representation of a protein structure capable of mapping into our software package, SidechainNet. We demonstrate our method's efficacy by finetuning OpenFold. We show that subsequently predicted protein structures, both before and after a relaxation procedure, exhibit comparable accuracy while displaying lower potential energy and improved structural quality as assessed by MolProbity metrics.


Related Concept Videos
 01:24
Protein Organization
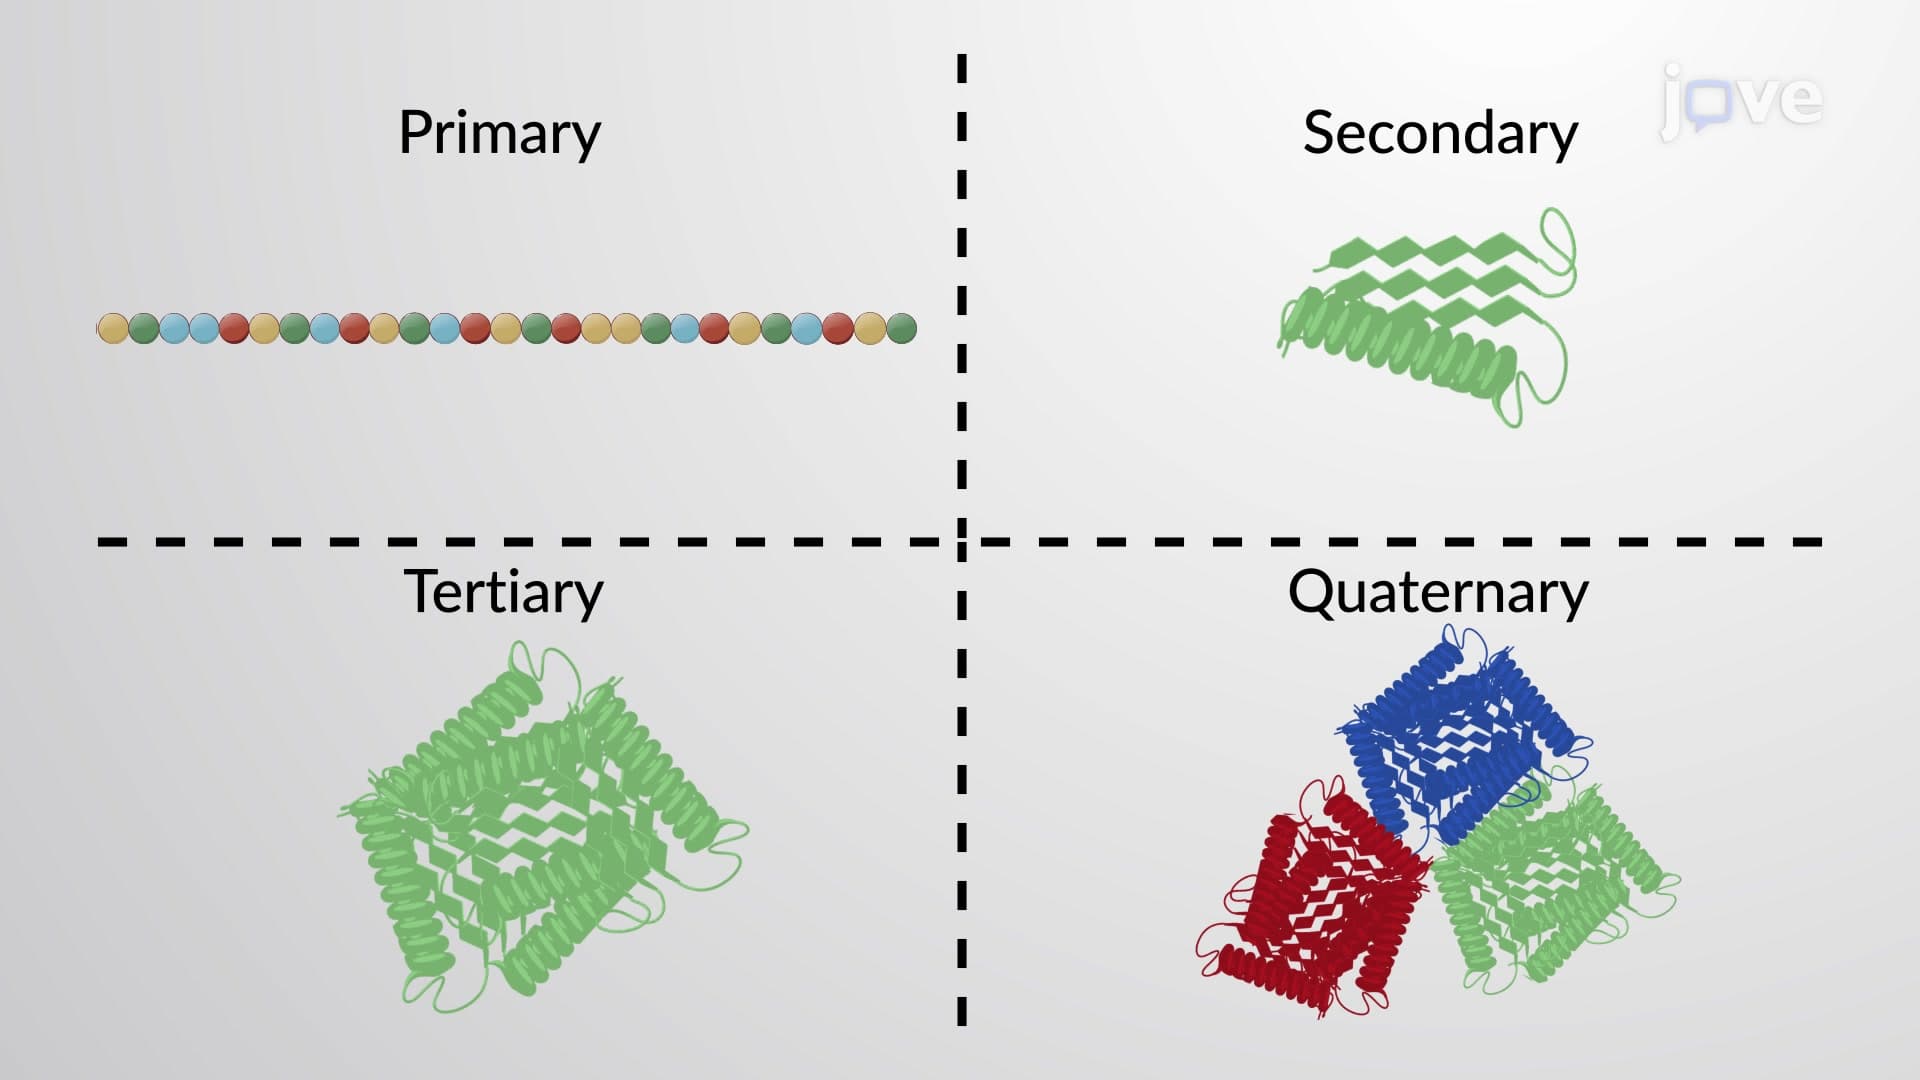
The primary structure of a protein is its amino acid sequence....
 02:15
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme...
02:15Protein and Protein Structures
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to...
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 02:04
Protein-protein Interfaces