Related Experiment Video
Updated: Jul 8, 2025


Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Improved Peptide Docking with Privileged Knowledge Distillation using Deep Learning
Zicong Zhang1, Jacob Verburgt2, Yuki Kagaya2
1Department of Computer Science, Purdue University, West Lafayette, Indiana, 47907, USA.

Abstract:
Protein-peptide interactions play a key role in biological processes. Understanding the interactions that occur within a receptor-peptide complex can help in discovering and altering their biological functions. Various computational methods for modeling the structures of receptor-peptide complexes have been developed. Recently, accurate structure prediction enabled by deep learning methods has significantly advanced the field of structural biology. AlphaFold (AF) is among the top-performing structure prediction methods and has highly accurate structure modeling performance on single-chain targets. Shortly after the release of AlphaFold, AlphaFold-Multimer (AFM) was developed in a similar fashion as AF for prediction of protein complex structures. AFM has achieved competitive performance in modeling protein-peptide interactions compared to previous computational methods; however, still further improvement is needed. Here, we present DistPepFold, which improves protein-peptide complex docking using an AFM-based architecture through a privileged knowledge distillation approach. DistPepFold leverages a teacher model that uses native interaction information during training and transfers its knowledge to a student model through a teacher-student distillation process. We evaluated DistPepFold's docking performance on two protein-peptide complex datasets and showed that DistPepFold outperforms AFM. Furthermore, we demonstrate that the student model was able to learn from the teacher model to make structural improvements based on AFM predictions.
More Related Videos
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 02:04
Protein-protein Interfaces
 01:20
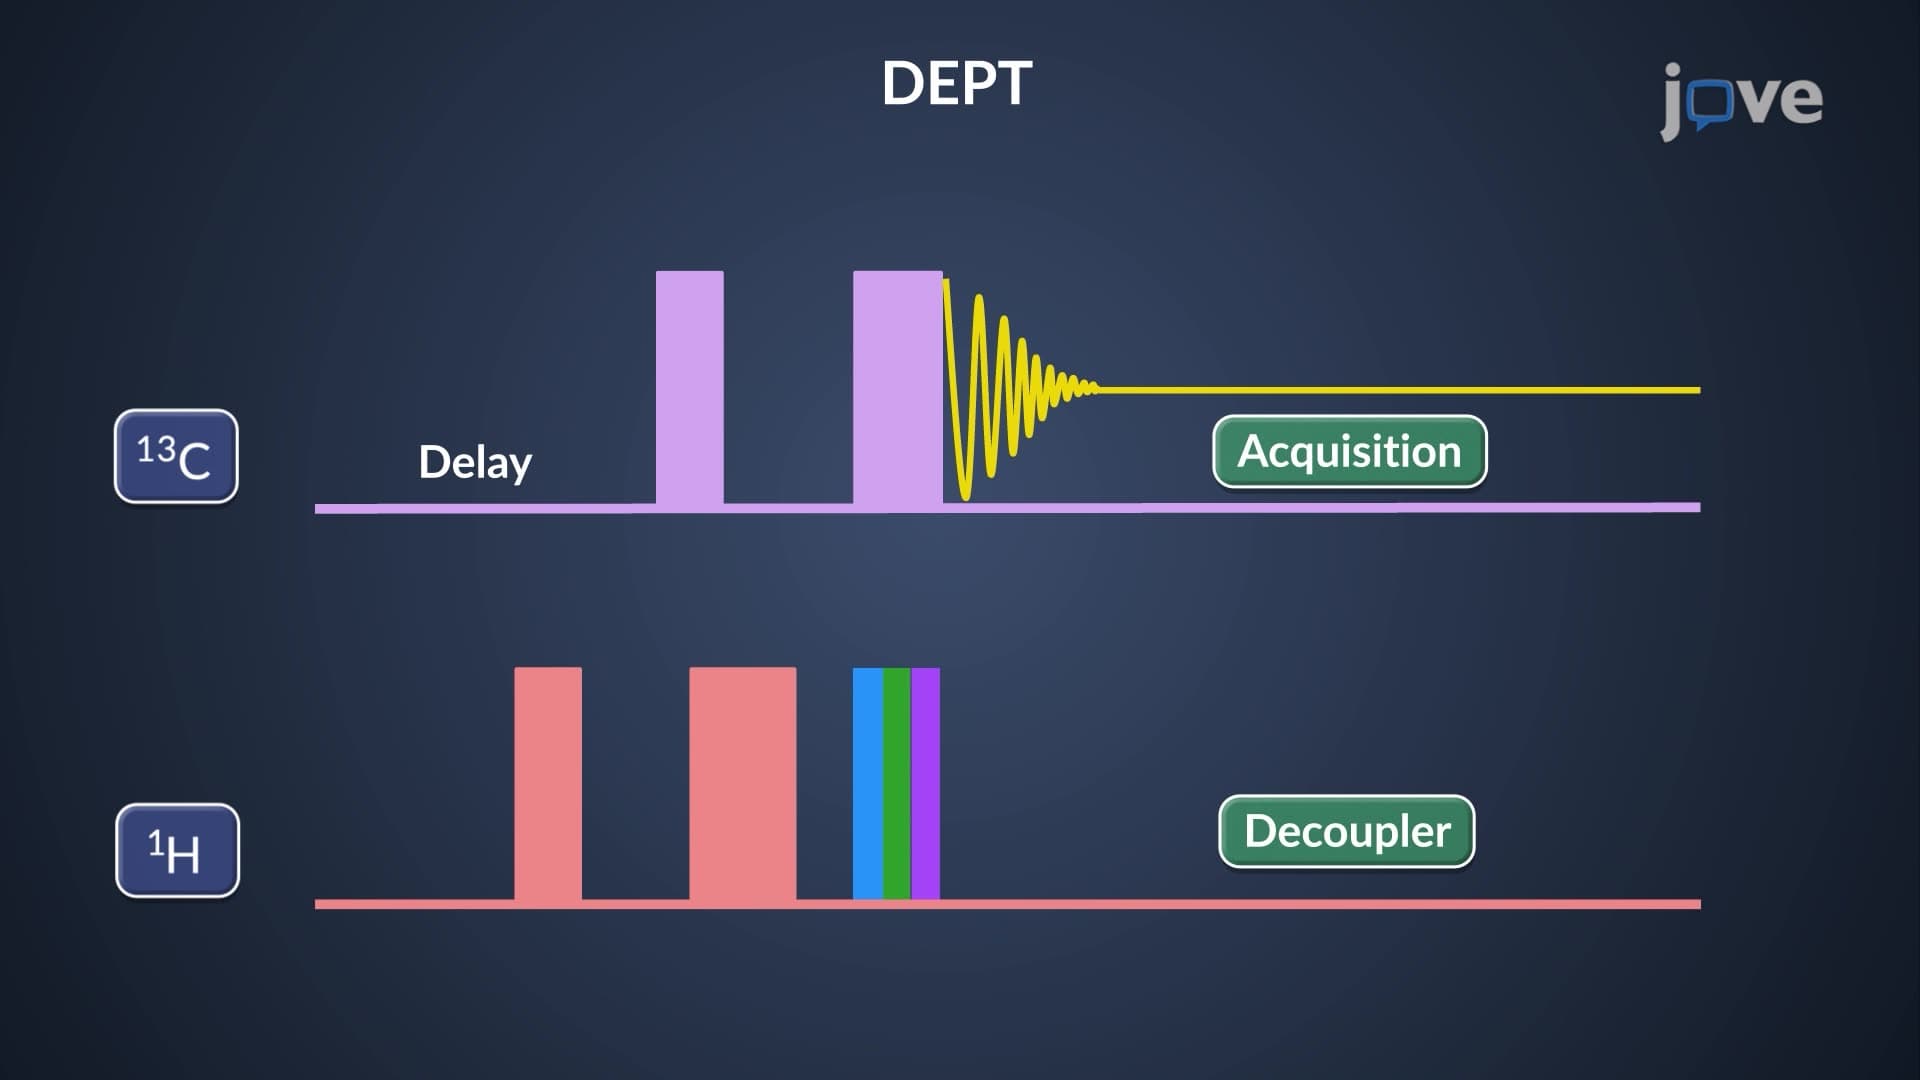
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...