Related Experiment Video
Updated: Jul 7, 2025
Determination of the Transport Rate of Xenobiotics and Nanomaterials Across the Placenta using the ex vivo Human Placental Perfusion Model
Published on: June 18, 2013
Successful Prediction of Fetal Exposure to Dual BCRP/P-gp Drug Substrates Using the Efflux Ratio-Relative Expression
Ankit Balhara1, Mengyue Yin1, Jashvant D Unadkat1
1Department of Pharmaceutics, University of Washington, Seattle, Washington, USA.

To inform fetal drug safety, it is important to determine or predict fetal drug exposure throughout pregnancy. The former is not possible in the first or second trimester. In contrast, at the time of birth, fetal drug exposure, relative to maternal exposure, can be estimated as Kp,uu (unbound fetal umbilical venous (UV) plasma area under the curve (AUC)/unbound maternal plasma (MP) AUC), provided the observed UV/MP values, spanning the dosing interval, are available from multiple maternal-fetal dyads. However, this fetal Kp,uu cannot be extrapolated to other drugs. To overcome the above limitations, we have used an efflux ratio-relative expression factor (ER-REF) approach to successfully predict the fetal Kp,uu of P-gp substrates. Because many drugs taken by pregnant people are also BCRP substrates, here, we extend this approach to drugs that are effluxed by both placental BCRP and P-gp or P-gp alone. To verify our predictions, we chose drugs for which UV/MP data were available at term: glyburide and imatinib (both BCRP and P-gp substrates) and nelfinavir (only P-gp substrate). First, the ER of the drugs was determined using Transwells and MDCKII cells expressing either BCRP or P-gp. Then, the ER was scaled using the proteomics-informed REF value to predict the fetal Kp,uu of the drug at term. The ER-REF predicted fetal Kp,uu of glyburide (0.43), imatinib (0.42), and nelfinavir (0.40) fell within two-fold of the corresponding in vivo fetal Kp,uu (0.44, 0.37, and 0.46, respectively). These data confirm that the ER-REF approach can successfully predict fetal drug exposure to BCRP/P-gp and P-gp substrates, at term.
To inform fetal drug safety, it is important to determine or predict fetal drug exposure throughout pregnancy. The former is not possible in the first or second trimester. In contrast, at the time of birth, fetal drug exposure, relative to maternal exposure, can be estimated as Kp,uu (unbound fetal umbilical venous (UV) plasma area under the curve (AUC)/unbound maternal plasma (MP) AUC), provided the observed UV/MP values, spanning the dosing interval, are available from multiple maternal-fetal dyads. However, this fetal Kp,uu cannot be extrapolated to other drugs. To overcome the above limitations, we have used an efflux ratio-relative expression factor (ER-REF) approach to successfully predict the fetal Kp,uu of P-gp substrates. Because many drugs taken by pregnant people are also BCRP substrates, here, we extend this approach to drugs that are effluxed by both placental BCRP and P-gp or P-gp alone. To verify our predictions, we chose drugs for which UV/MP data were available at term: glyburide and imatinib (both BCRP and P-gp substrates) and nelfinavir (only P-gp substrate). First, the ER of the drugs was determined using Transwells and MDCKII cells expressing either BCRP or P-gp. Then, the ER was scaled using the proteomics-informed REF value to predict the fetal Kp,uu of the drug at term. The ER-REF predicted fetal Kp,uu of glyburide (0.43), imatinib (0.42), and nelfinavir (0.40) fell within two-fold of the corresponding in vivo fetal Kp,uu (0.44, 0.37, and 0.46, respectively). These data confirm that the ER-REF approach can successfully predict fetal drug exposure to BCRP/P-gp and P-gp substrates, at term.
More Related Videos
 00:05
In Silico Modeling Method for Computational Aquatic Toxicology of Endocrine Disruptors: A Software-Based Approach Using QSAR Toolbox
Published on: August 28, 2019
 17:28
Human Pluripotent Stem Cell Based Developmental Toxicity Assays for Chemical Safety Screening and Systems Biology Data Generation
Published on: June 17, 2015
Related Concept Videos
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion,...
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:26
Pharmacokinetic Models: Comparison and Selection Criterion
Physiological models take a detailed approach by considering specific molecular processes. They can predict drug distribution, metabolism, and elimination changes, providing a comprehensive understanding of how drugs interact with the body.
 01:06
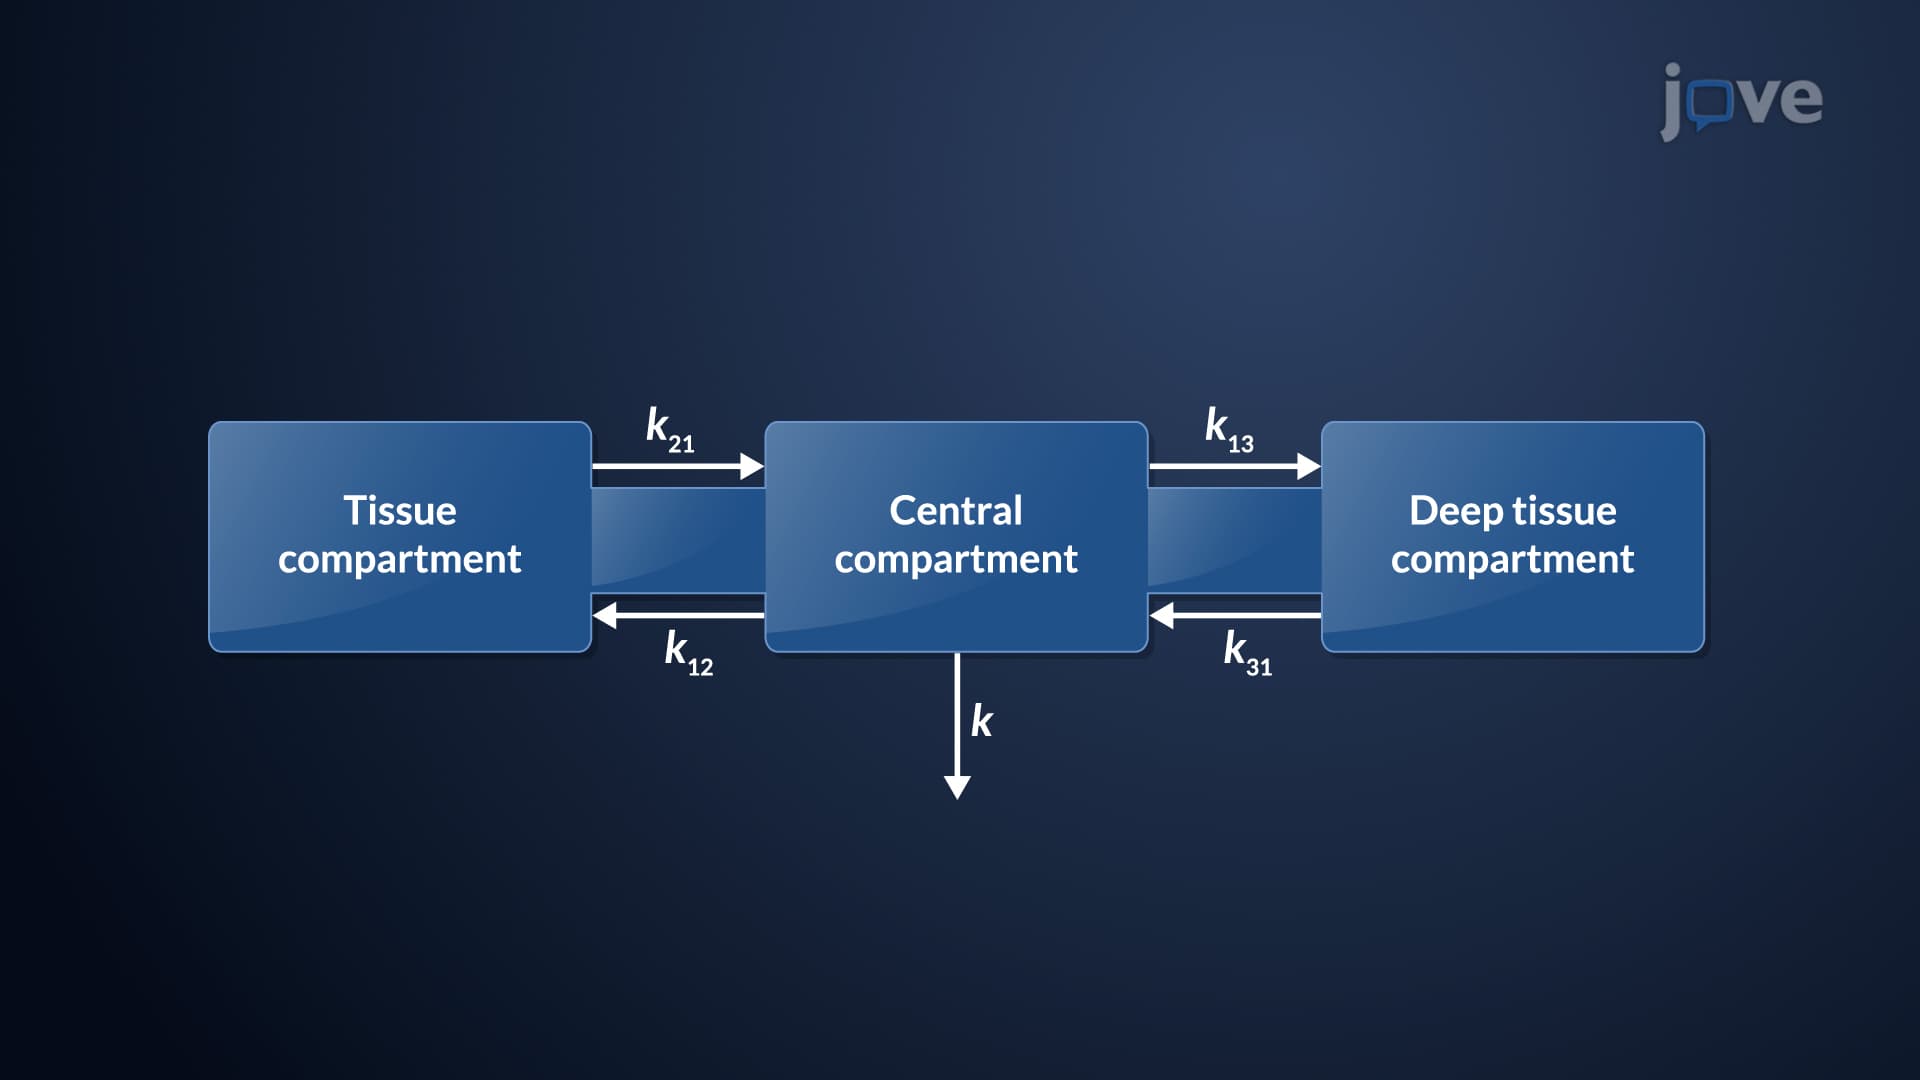
Three-Compartment Open Model
 00:57
Physiological Pharmacokinetic Models: Blood Flow-Limited Versus Diffusion-Limited Models
 01:20
Pharmacokinetic Models: Overview
There are three primary types of models: empirical, compartment, and physiological. Empirical models, with minimal...