Related Experiment Video
Updated: Jul 6, 2025

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Six-dimensional quantum dynamics study for the dissociative chemisorption of H2 on pure and alloyed AgAu surfaces
Tianhui Liu1,2, Tianze Peng2, Bina Fu2,3,4
1School of Sciences, Great Bay University, Songshan Lake International Innovation Entrepreneurship Community A5, Dongguan 523000, China.

Abstract:
The 6D time-dependent wave packet calculations were performed to explore H2 dissociation on Ag, Au, and two AgAu alloy surfaces, using four newly fitted potential energy surfaces based on the neural network fitting to density functional theory energy points. The ligand effect resulting from the Ag-Au interaction causes a reduction in the barrier height for H2+Ag/Au(111) compared to H2+Ag(111). However, the scenario is reversed for H2+Au/Ag(111) and H2+Au(111). The 6D dissociation probabilities of H2 on Ag/Au(111) surfaces are significantly higher than those on the pure Ag(111) surface, but the corresponding results for H2 on Au/Ag(111) surfaces are substantially lower than those on the pure Au(111) surface. The reactivity of H2 on Au(111) is larger than that on Ag(111), despite Ag(111) having a slightly lower static barrier height. This can be attributed to the exceptionally small dissociation probabilities at the hcp and fcc regions, which are at least 100 times smaller compared to those at the bridge or top site for H2+Ag(111). Due to the late barrier being more pronounced, the vibrational excitation of H2 on Ag(111) is more effective in promoting the reaction than on Au(111). Moreover, a high degree of alignment dependence is detected for the four reactions, where the H2 dissociation has the highest probability at the helicopter alignment, as opposed to the cartwheel alignment.
More Related Videos
 14:11
Quantification of Hydrogen Concentrations in Surface and Interface Layers and Bulk Materials through Depth Profiling with Nuclear Reaction Analysis
Published on: March 29, 2016
 07:53
Analysis of Complex Molecules and Their Reactions on Surfaces by Means of Cluster-Induced Desorption/Ionization Mass Spectrometry
Published on: March 1, 2020
Related Concept Videos
 03:35
Hybridization of Atomic Orbitals II
 03:51
Molecular Orbital Theory II
 02:38
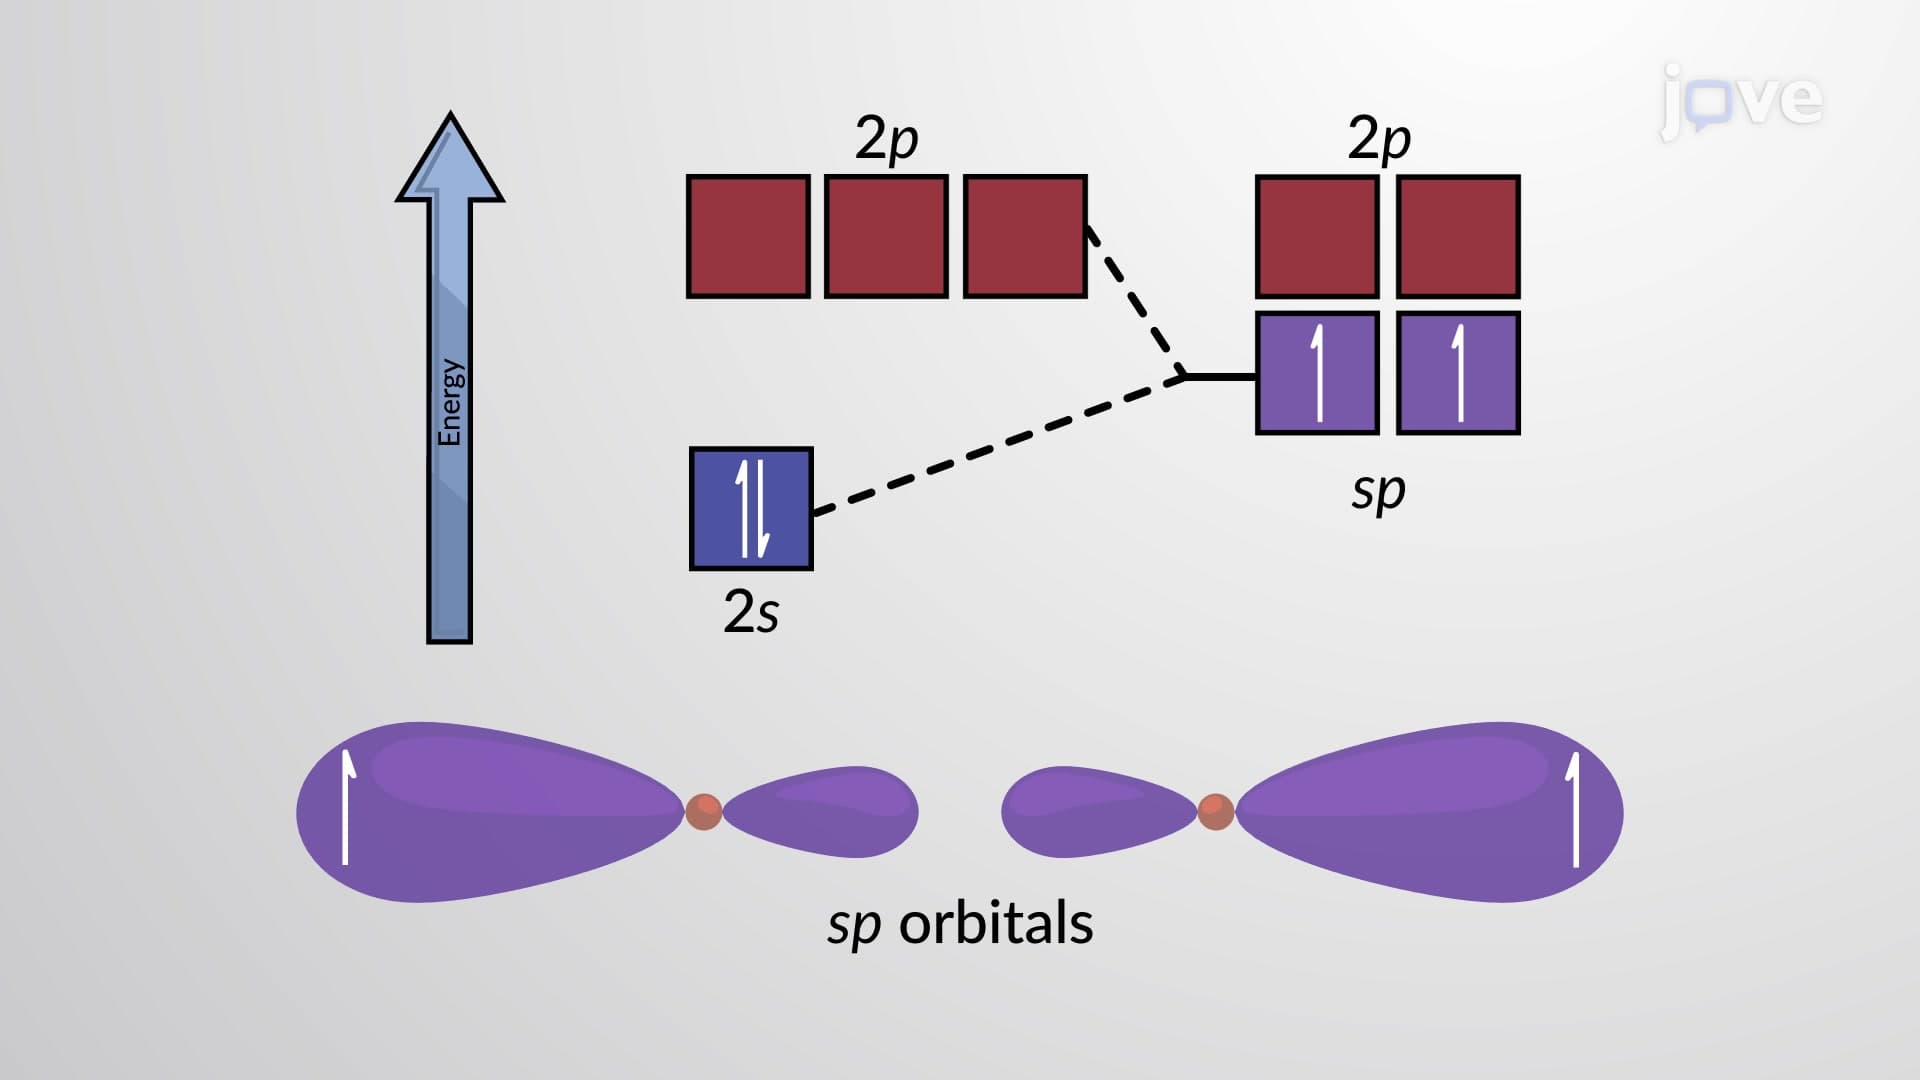
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 03:24
Hybridization of Atomic Orbitals I
 01:25
Atomic Absorption Spectroscopy: Atomization Methods
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...