
A deep learning model for predicting selected organic molecular spectra
Zihan Zou1, Yujin Zhang2, Lijun Liang3
1School of Chemistry and Chemical Engineering, Qilu University of Technology (Shandong Academy of Science), Jinan, China.

Abstract:
Accurate and efficient molecular spectra simulations are crucial for substance discovery and structure identification. However, the conventional approach of relying on the quantum chemistry is cost intensive, which hampers efficiency. Here we develop DetaNet, a deep-learning model combining E(3)-equivariance group and self-attention mechanism to predict molecular spectra with improved efficiency and accuracy. By passing high-order geometric tensorial messages, DetaNet is able to generate a wide variety of molecular properties, including scalars, vectors, and second- and third-order tensors-all at the accuracy of quantum chemistry calculations. Based on this we developed generalized modules to predict four important types of molecular spectra, namely infrared, Raman, ultraviolet-visible, and 1H and 13C nuclear magnetic resonance, taking the QM9S dataset containing 130,000 molecular species as an example. By speeding up the prediction of molecular spectra at quantum chemical accuracy, DetaNet could help progress toward real-time structural identification using spectroscopic measurements.
More Related Videos

Related Concept Videos
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:26
Spectroscopy of Carboxylic Acid Derivatives
 01:14
Molecular Spectroscopy: Absorption and Emission
 01:24
IR Spectroscopy: Molecular Vibration Overview
Stretching vibrations are vibrational motions that occur along the bond line, changing the bond length or distance between two bonded atoms. They are further distinguished as symmetric or asymmetric. In symmetric stretching, the...
 01:14
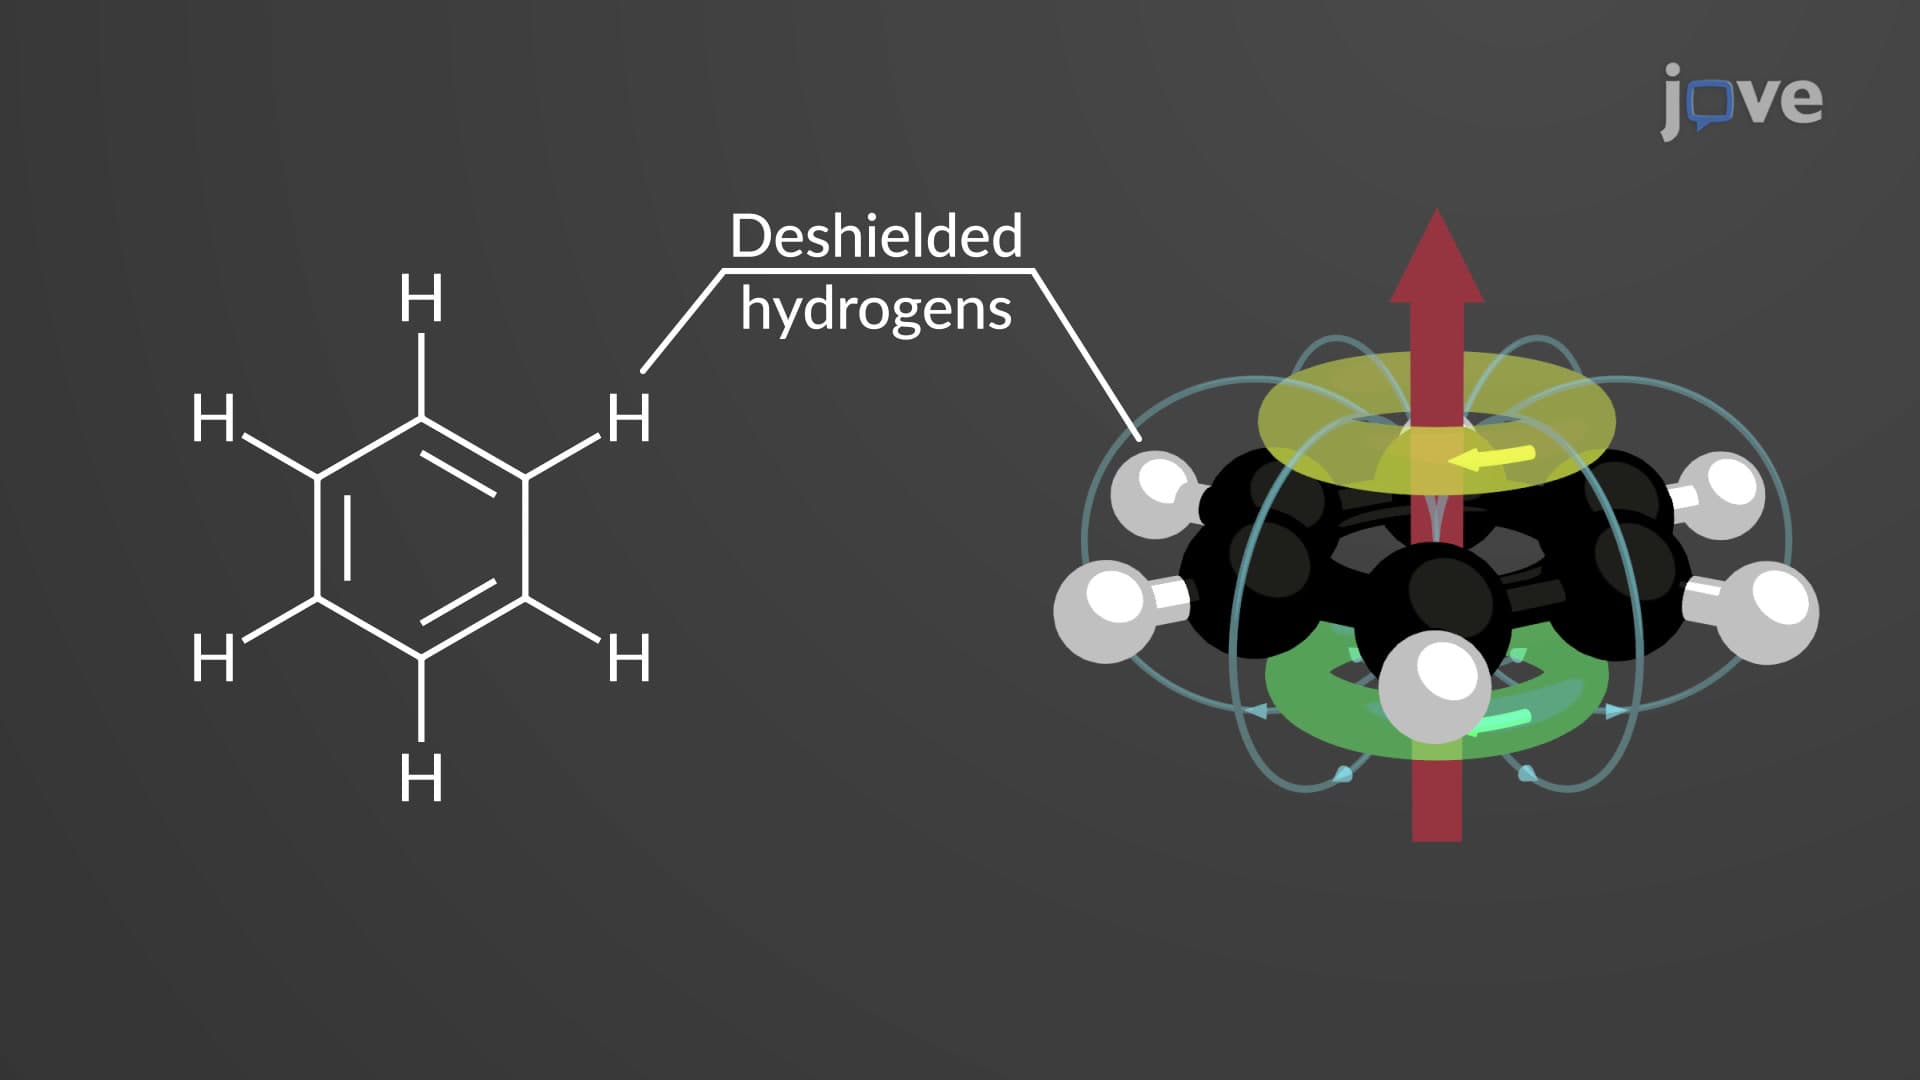
NMR Spectroscopy of Aromatic Compounds
 02:27
Predicting Molecular Geometry