Computational Procedure for Predicting Excipient Effects on Protein-Protein Affinities
Related Experiment Video
Updated: Jul 4, 2025

Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Gregory L Dignon1, Ken A Dill1,2,3
1Laufer Center for Physical and Quantitative Biology, Stony Brook University, 100 Nicolls Road, Stony Brook, New York 11794, United States.

Abstract:
Protein-protein interactions lie at the center of many biological processes and are a challenge in formulating biological drugs, such as antibodies. A key to mitigating protein association is to use small-molecule additives, i.e., excipients that can weaken protein-protein interactions. Here, we develop a computationally efficient model for predicting the viscosity-reducing effect of different excipient molecules by combining atomic-resolution MD simulations, binding polynomials, and a thermodynamic perturbation theory. In a proof of principle, this method successfully ranks the order of four types of excipients known to reduce the viscosity of solutions of a particular monoclonal antibody. This approach appears useful for predicting the effects of excipients on protein association and phase separation, as well as the effects of buffers on protein solutions.

Related Concept Videos
 02:04
Protein-protein Interfaces
 01:22
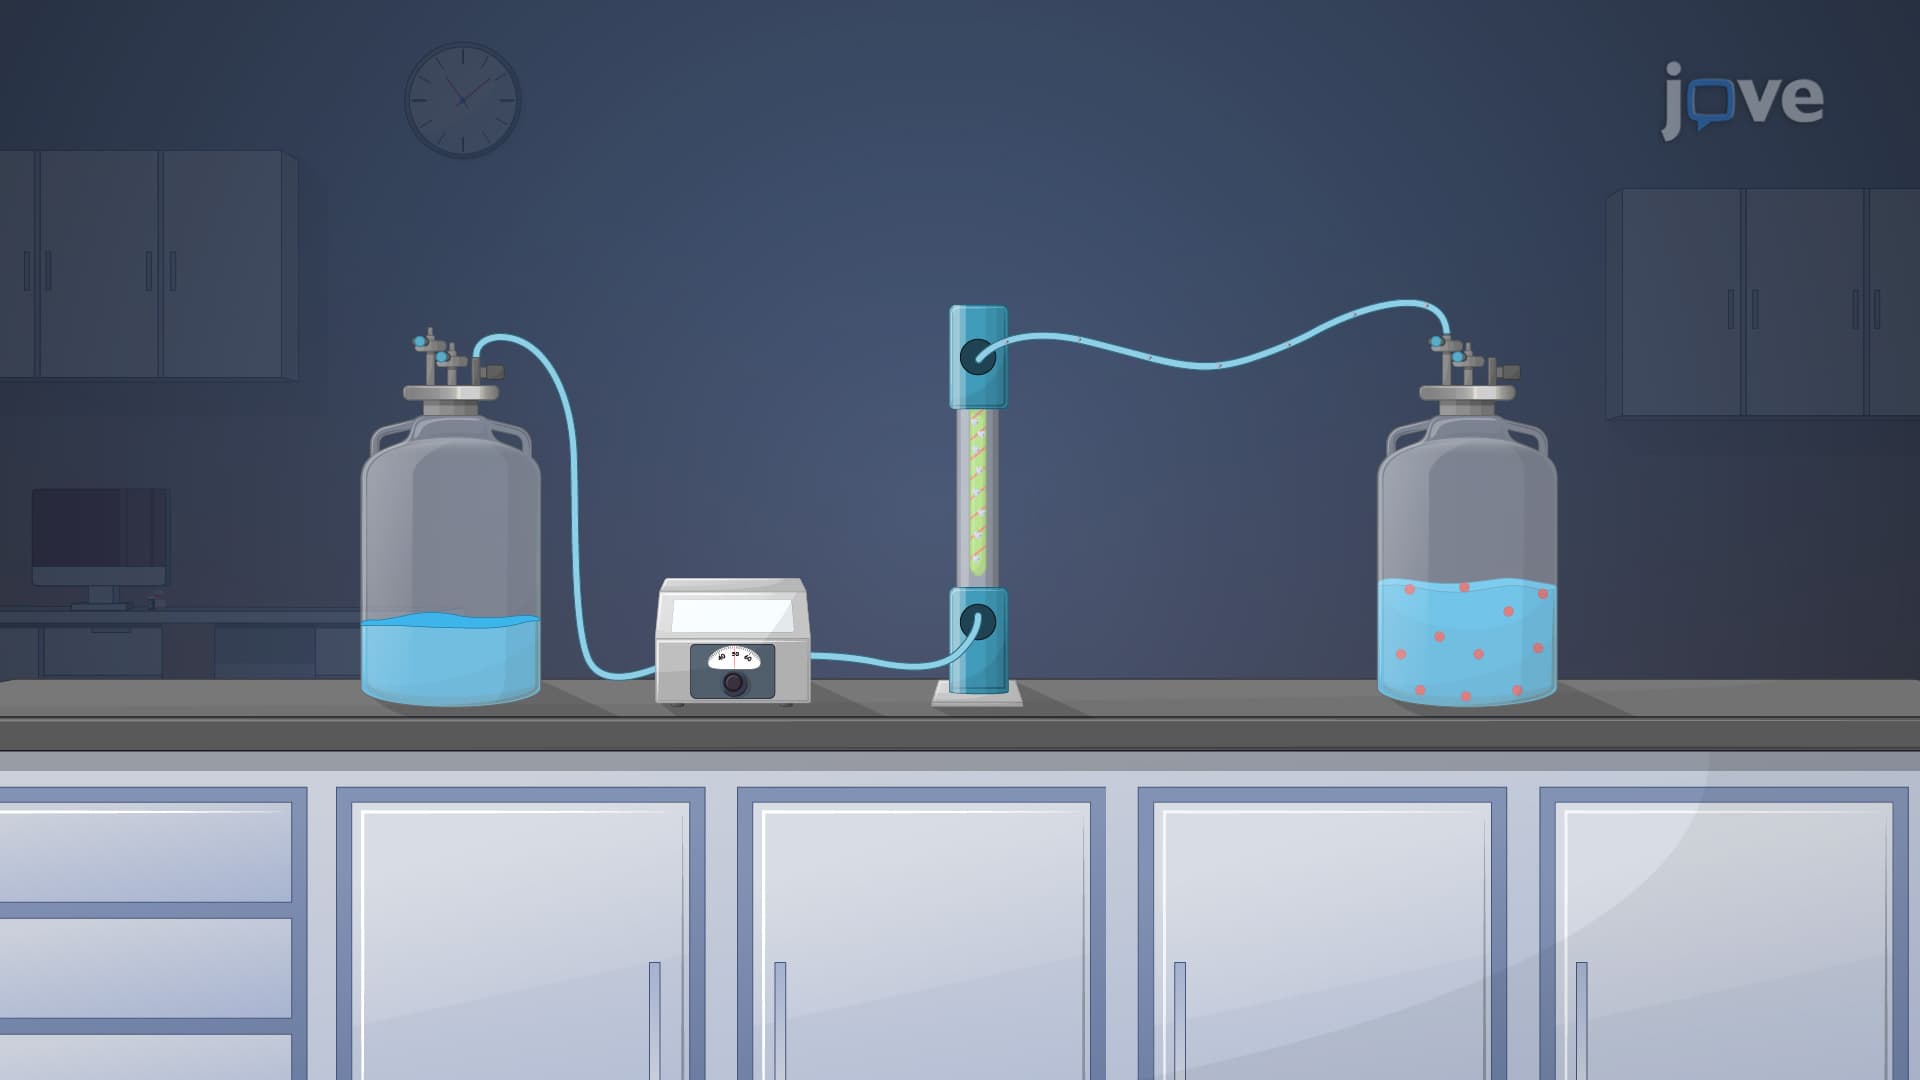
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 02:18
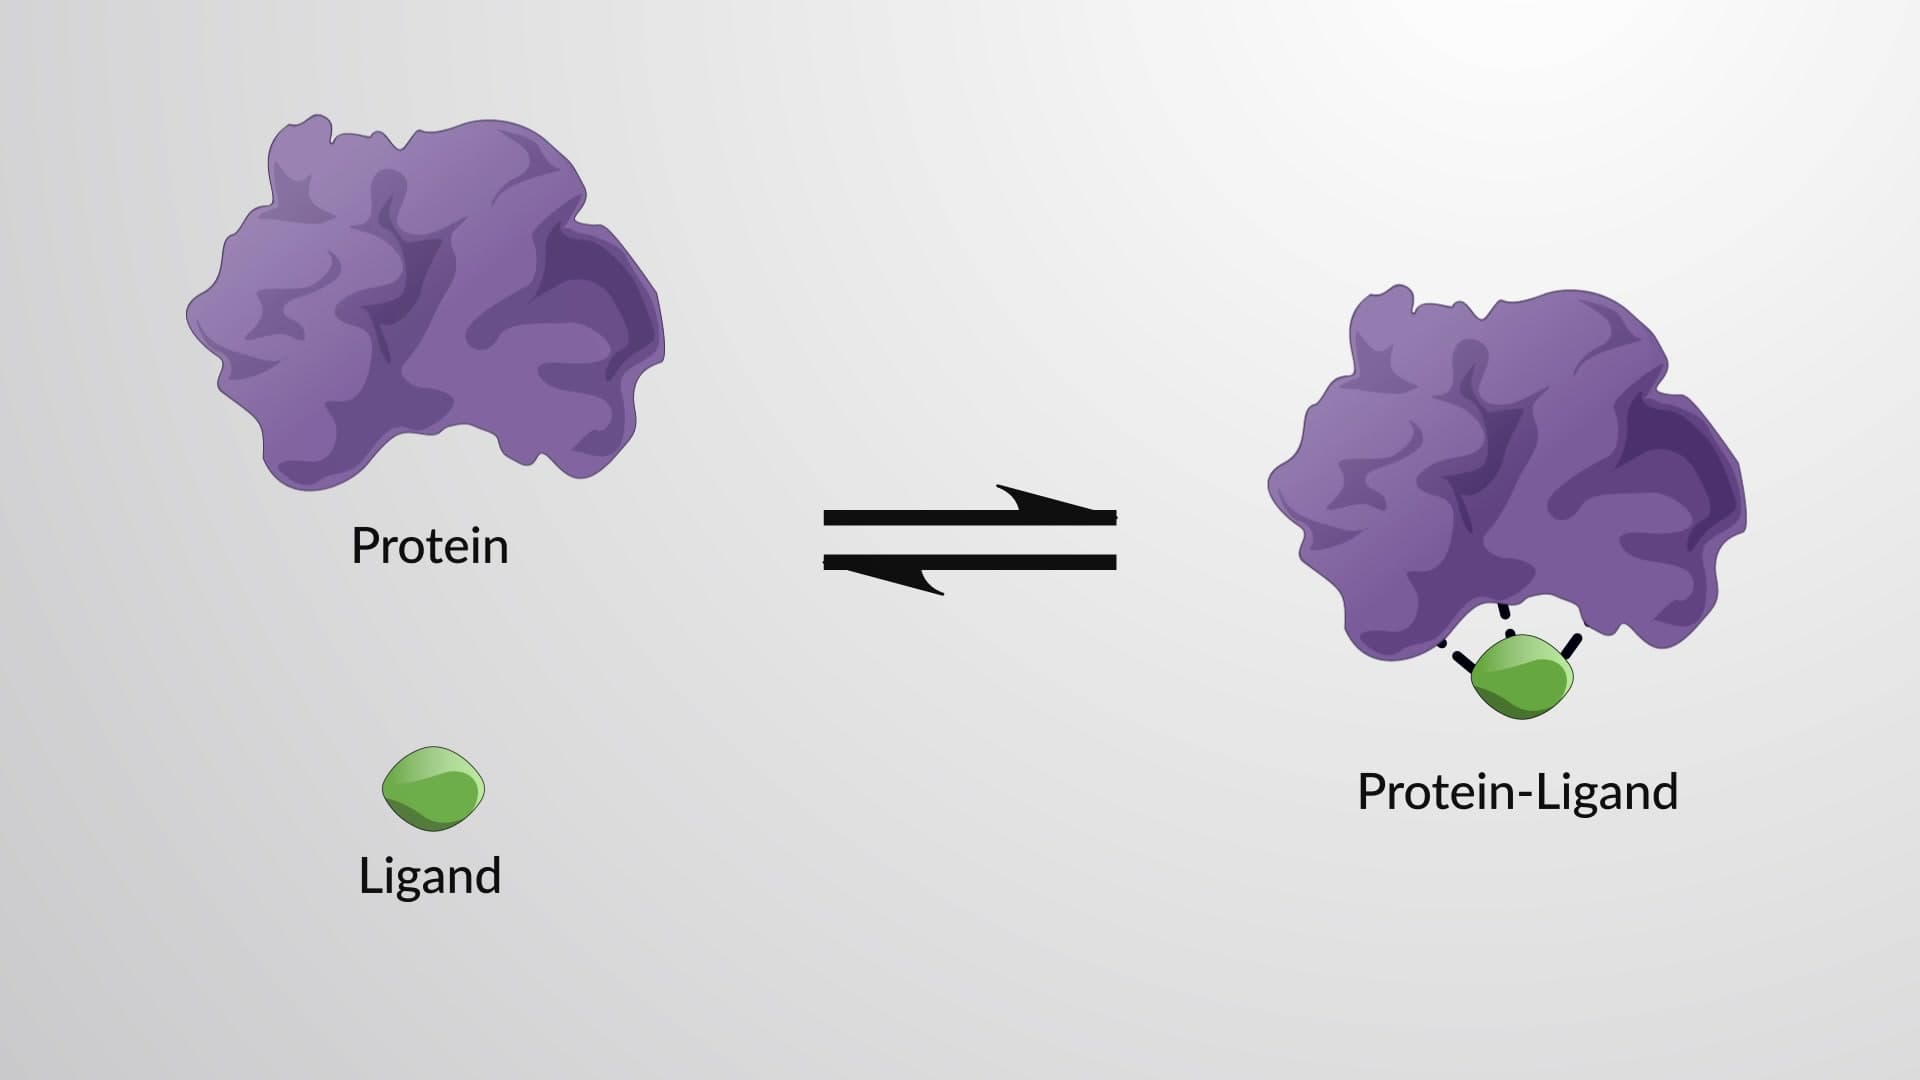
The Equilibrium Binding Constant and Binding Strength
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding