Related Experiment Video
Updated: Jul 4, 2025
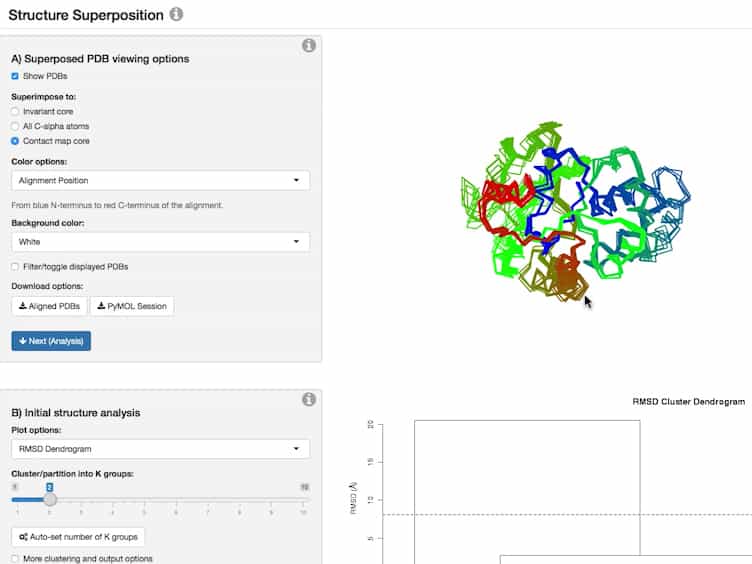

Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Sequence clustering confounds AlphaFold2
Joseph W Schafer1, Devlina Chakravarty1, Ethan A Chen1
1National Center for Biotechnology Information, National Library of Medicine, National Institutes of Health, Bethesda, MD 20894.

Abstract:
Though typically associated with a single folded state, some globular proteins remodel their secondary and/or tertiary structures in response to cellular stimuli. AlphaFold21 (AF2) readily generates one dominant protein structure for these fold-switching (a.k.a. metamorphic) proteins2, but it often fails to predict their alternative experimentally observed structures3,4. Wayment-Steele, et al. steered AF2 to predict alternative structures of a few metamorphic proteins using a method they call AF-cluster5. However, their Paper lacks some essential controls needed to assess AF-cluster's reliability. We find that these controls show AF-cluster to be a poor predictor of metamorphic proteins. First, closer examination of the Paper's results reveals that random sequence sampling outperforms sequence clustering, challenging the claim that AF-cluster works by "deconvolving conflicting sets of couplings." Further, we observe that AF-cluster mistakes some single-folding KaiB homologs for fold switchers, a critical flaw bound to mislead users. Finally, proper error analysis reveals that AF-cluster predicts many correct structures with low confidence and some experimentally unobserved conformations with confidences similar to experimentally observed ones. For these reasons, we suggest using ColabFold6-based random sequence sampling7-augmented by other predictive approaches-as a more accurate and less computationally intense alternative to AF-cluster.

Related Concept Videos
 03:21
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
 01:22
Protein Folding
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...