Related Experiment Video
Updated: Jul 4, 2025

Analysis of Complex Molecules and Their Reactions on Surfaces by Means of Cluster-Induced Desorption/Ionization Mass Spectrometry
Published on: March 1, 2020
Quantum hardware calculations of the activation and dissociation of nitrogen on iron clusters and surfaces
Georgia Christopoulou1, Cono Di Paola1, Floris Eelke Elzinga2
1Quantinuum, Terrington House, 13-15 Hills Road, CB2 1NL, Cambridge, UK. Georgia.Christopoulou@quantinuum.com.

Abstract:
Catalytic processes are the cornerstone of chemical industry, and catalytic conversion of nitrogen to ammonia remains one of the largest industrial processes implemented. Rational design of catalysts and catalytic reactions largely depends on approximate computational chemistry methods, such as density functional theory, which, however, suffer from limited accuracy, especially for strongly-correlated materials. Rigorous ab initio methods which account for static and dynamic electron correlation, while arbitrarily accurate for small systems, are generally too expensive to be applied to modelling of catalytic cycles, due to prohibitive time and space computational complexity with respect to the size of the active space. Recent advances in quantum computing give hope for enabling access to accurate ab initio methods at scale. Herein, we present a prototype hybrid quantum-classical workflow for modeling chemical reactions on surfaces, applied to proof-of-concept models of activation and dissociation of nitrogen on small Fe clusters and a single-layer (221) iron surface. First, we determined the structures of species present in the catalytic cycle at DFT level and studied their electronic structure using CASSCF. We show that it is possible to decouple the half-filled Fe-3d band from the Fe-N and N-N bond orbitals, thereby reducing the active space significantly. Subsequently, we translated the CASSCF wavefunctions into corresponding qubit quantum states, using the Adaptive Variational Quantum Eigensolver, and estimated their energies using a state vector simulator, H1-1E quantum emulator and (for selected systems) H1-1 quantum computer. We demonstrated that if a sufficiently small active orbital space is chosen, ground state energies obtained with classical methods and with the quantum computer are in reasonable agreement. We argue that once quantum computing methods are scaled up so that larger active spaces are accessible, they can offer a tremendous practical advantage to the computational catalysis community.
Related Concept Videos
 02:13
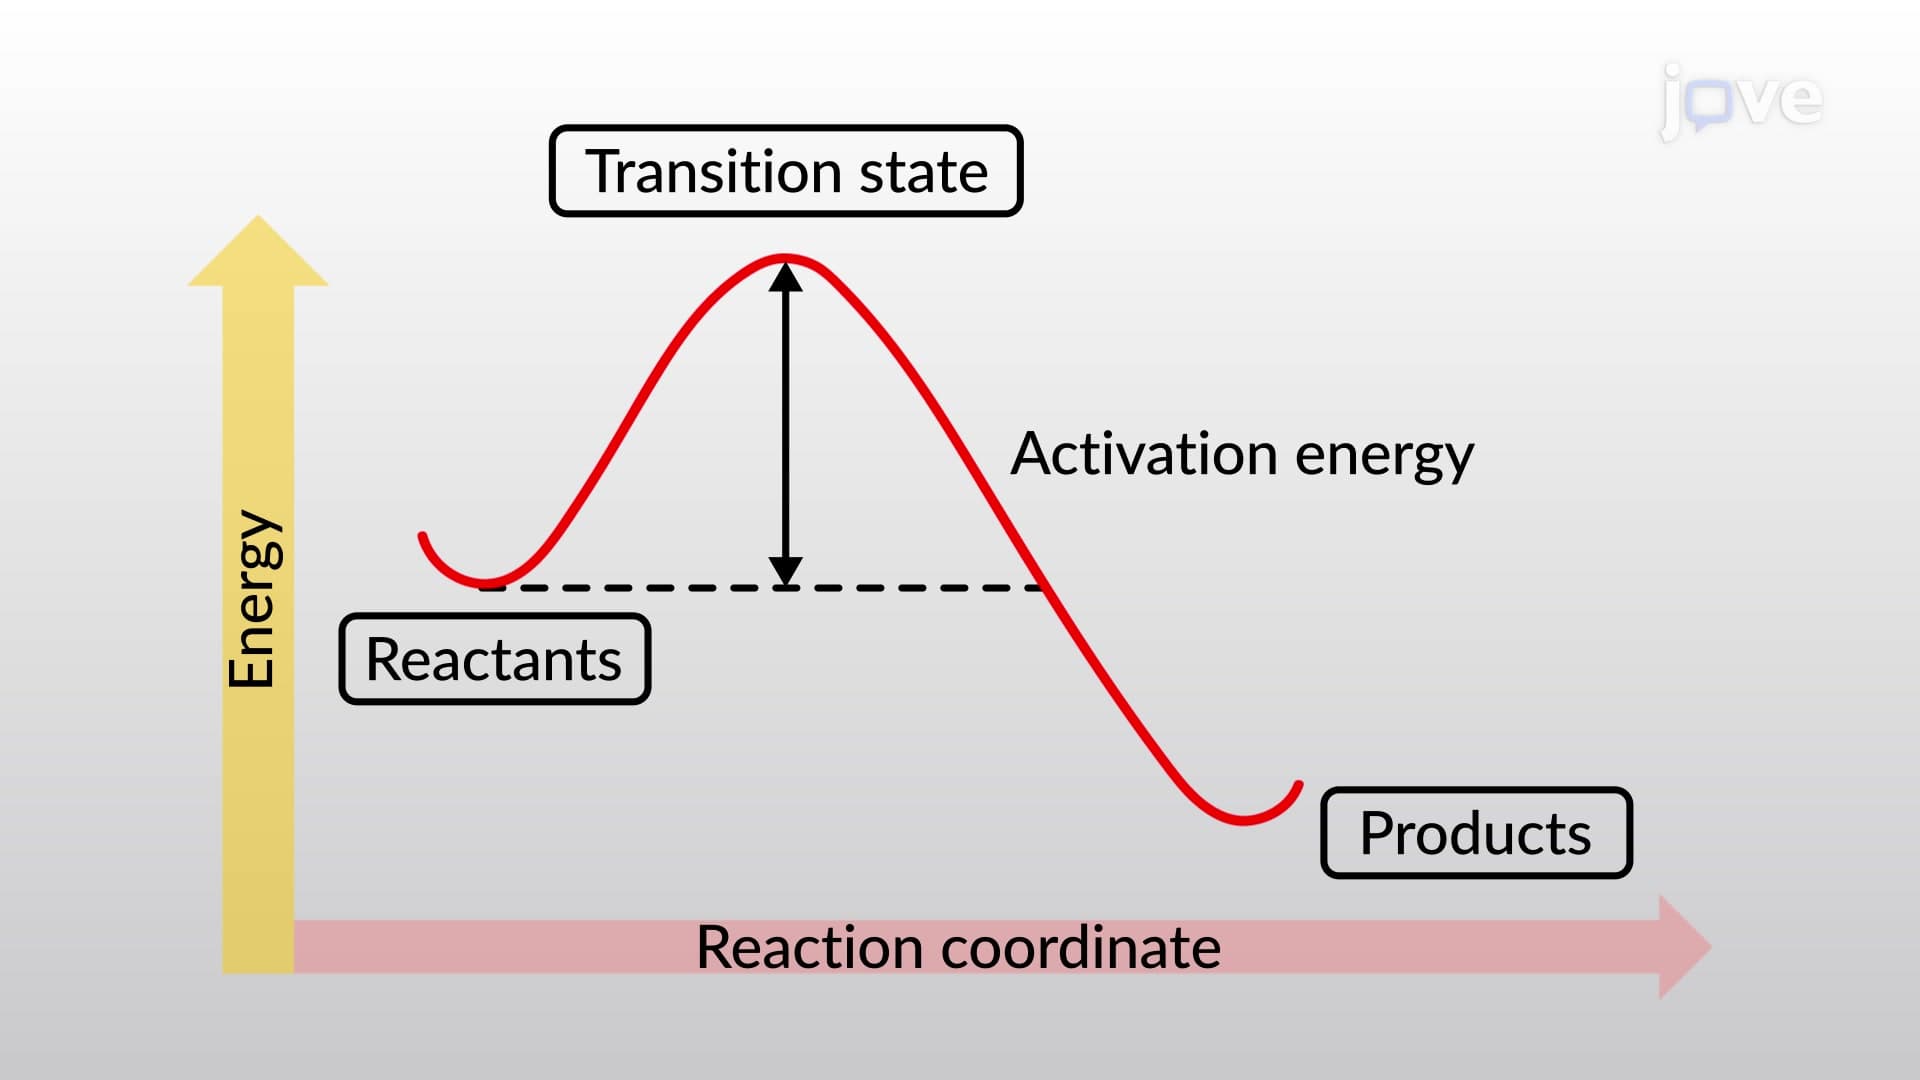
Bond Dissociation Energy and Activation Energy
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:34
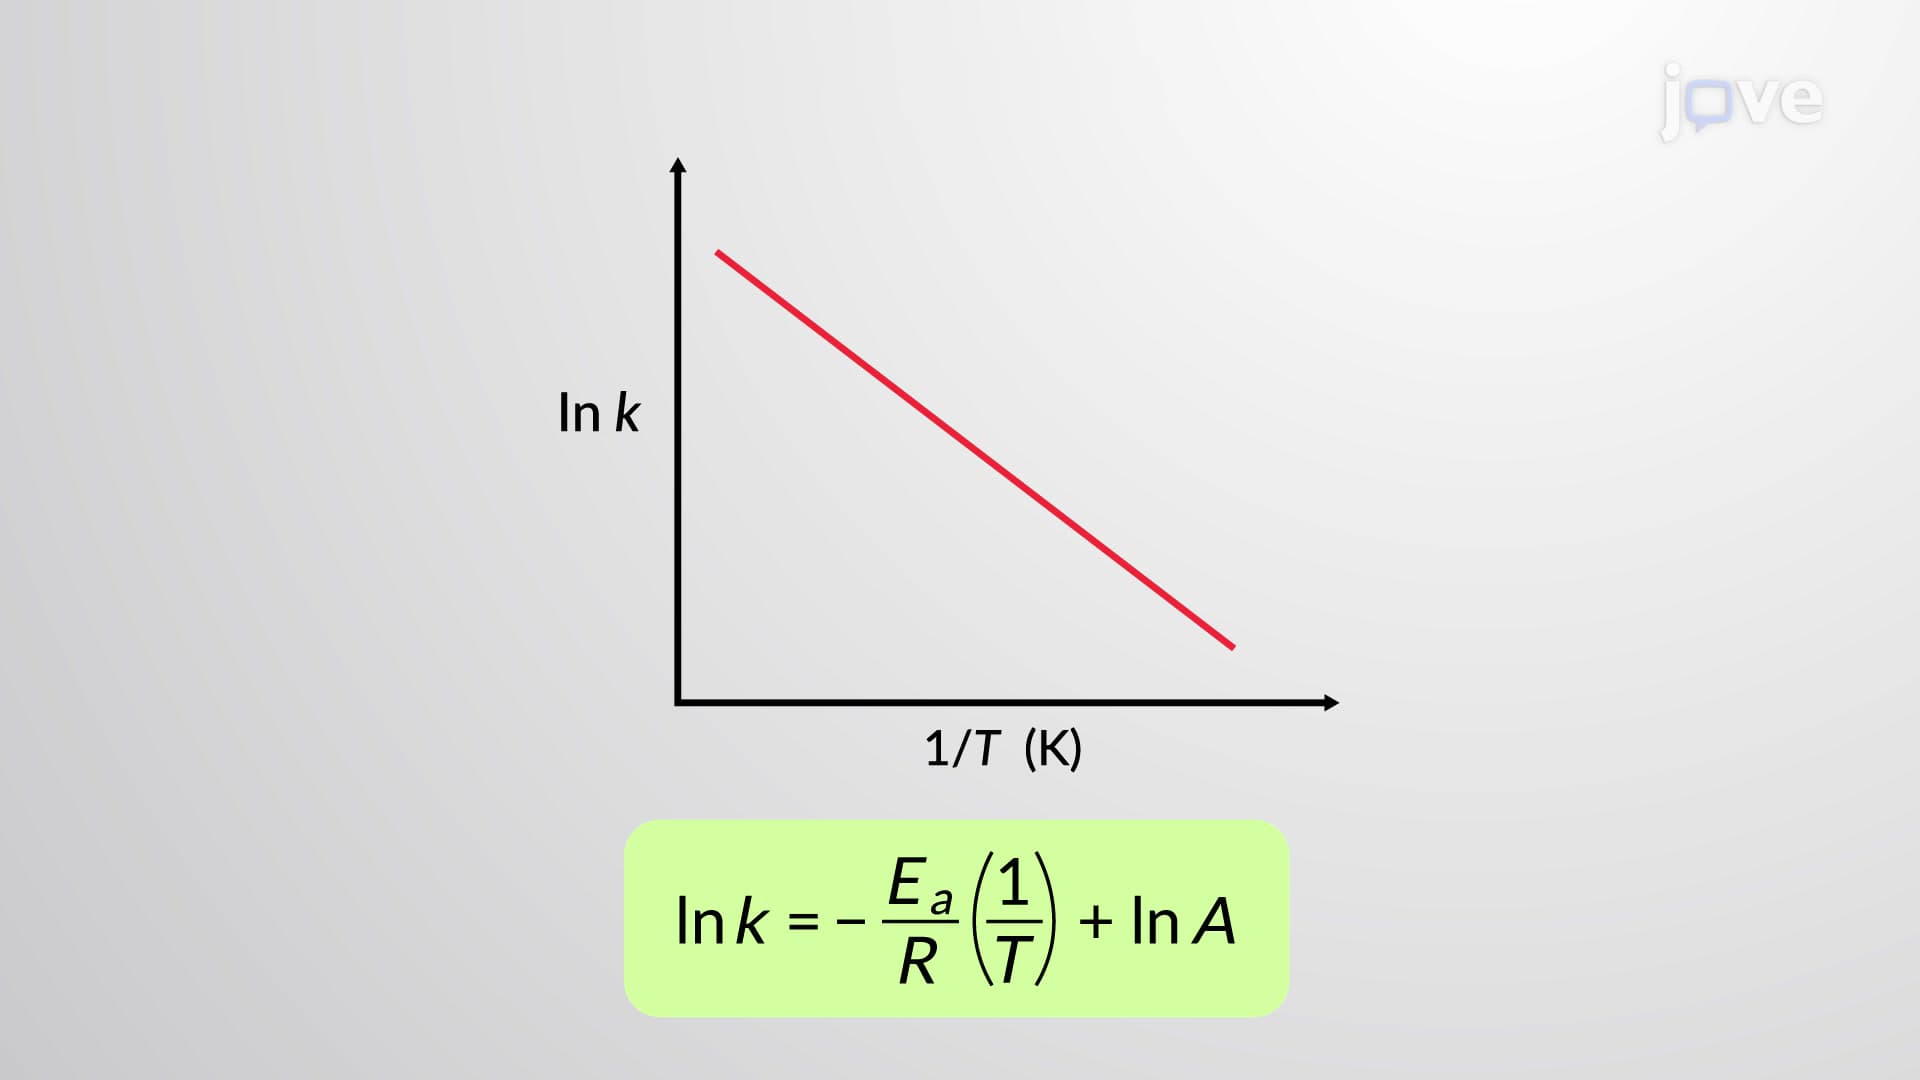
Arrhenius Plots
The Arrhenius equation can be used...
 01:25
Deactivation Processes: Jablonski Diagram
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
 02:42
Valence Bond Theory