Related Experiment Video
Updated: Jul 4, 2025

A Novel Method to Determine the Longitudinal Antibacterial Activity of Drug-Eluting Materials
Published on: March 3, 2023
Minimal impact of low-density lipoprotein apheresis on vancomycin serum concentration: A case report
Yoko Hiyama1, Takashi Tomita1, Hiroaki Matsuo2
1Department of Pharmaceutical Services, Hiroshima University Hospital, 1-2-3 Kasumi, Minami-ku, Hiroshima, 734-8551, Japan.

Low-density lipoprotein apheresis (LDL-A) is a blood purification therapy used to treat refractory ulcers in patients with arteriosclerosis obliterans. We describe a case of vancomycin treatment in a patient undergoing maintenance hemodialysis and LDL-A therapy and assess its impact on serum vancomycin concentration. The patient underwent LDL-A twice a week (Mondays and Fridays) and maintenance dialysis three times a week (Tuesdays, Thursdays, and Saturdays) for diabetic nephropathy associated with type 1 diabetes mellitus. Following the wound culture results, vancomycin was initiated with a 1.75 g administration post-dialysis. Serum vancomycin levels before and after LDL-A, measured on the subsequent day, exhibited only slight fluctuations within the intermeasurement variability range. Despite continuing vancomycin administration at the standard dose in patients undergoing hemodialysis, the serum concentration remained consistent, suggesting a minimal impact of LDL-A on vancomycin pharmacokinetics.
Low-density lipoprotein apheresis (LDL-A) is a blood purification therapy used to treat refractory ulcers in patients with arteriosclerosis obliterans. We describe a case of vancomycin treatment in a patient undergoing maintenance hemodialysis and LDL-A therapy and assess its impact on serum vancomycin concentration. The patient underwent LDL-A twice a week (Mondays and Fridays) and maintenance dialysis three times a week (Tuesdays, Thursdays, and Saturdays) for diabetic nephropathy associated with type 1 diabetes mellitus. Following the wound culture results, vancomycin was initiated with a 1.75 g administration post-dialysis. Serum vancomycin levels before and after LDL-A, measured on the subsequent day, exhibited only slight fluctuations within the intermeasurement variability range. Despite continuing vancomycin administration at the standard dose in patients undergoing hemodialysis, the serum concentration remained consistent, suggesting a minimal impact of LDL-A on vancomycin pharmacokinetics.
More Related Videos
Related Concept Videos
 01:09
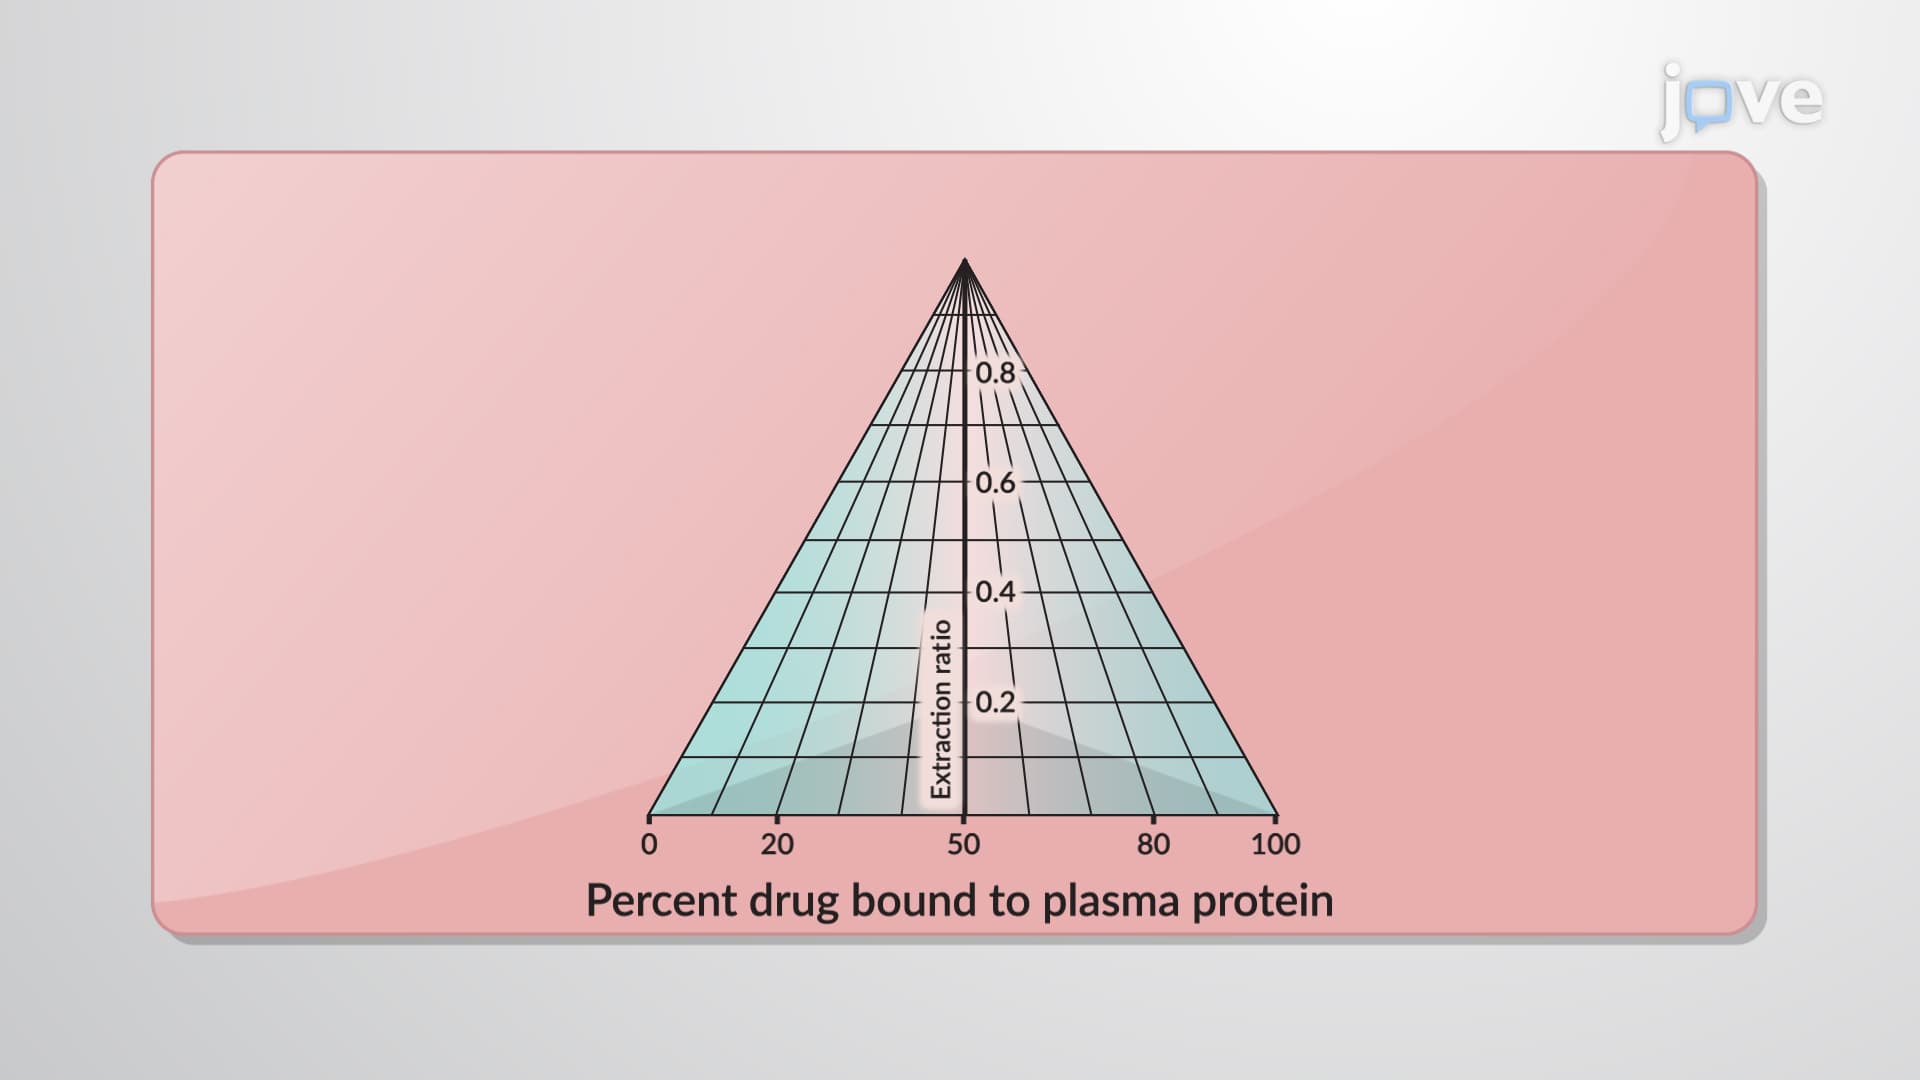
Hepatic Drug Clearance: Effect of Protein Binding
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
 01:18
Factors Affecting Protein-Drug Binding: Drug-Related Factors
One crucial factor in drug-protein binding is the drug's lipophilicity or its affinity for fat. More lipophilic drugs tend to have higher binding extents. For example, highly lipophilic drugs like cloxacillin exhibit substantial protein binding, with as much as 95% of the drug binding to proteins. In...
 01:25
Drug Distribution: Volume of Distribution
 01:29
Drug Distribution: Plasma Protein Binding
 01:14
Time Course of Drug Effect
 01:22
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
To quantify the extent of bioavailability, pharmacologists often use a parameter called .