Related Experiment Video
Updated: Jul 4, 2025

Membrane Transport Processes Analyzed by a Highly Parallel Nanopore Chip System at Single Protein Resolution
Published on: August 16, 2016
Unplugging lateral fenestrations of NALCN reveals a hidden drug binding site within the pore module

Abstract:
The sodium (Na + ) leak channel (NALCN) is a member of the four-domain voltage-gated cation channel family that includes the prototypical voltage-gated sodium and calcium channels (Na V s and Ca V s, respectively). Unlike Na V s and Ca V s, which have four lateral fenestrations that serve as routes for lipophilic compounds to enter the central cavity to modulate channel function, NALCN has bulky residues (W311, L588, M1145 and Y1436) that block these openings. Structural data suggest that oc-cluded lateral fenestrations underlie the pharmacological resistance of NALCN to lipophilic compounds, but functional evidence is lacking. To test this hypothesis, we unplugged the fenestrations of NALCN by substituting the four aforementioned resi-dues with alanine (AAAA) and compared the effects of Na V , Ca V and NALCN block-ers on both wild-type (WT) and AAAA channels. Most compounds behaved in a simi-lar manner on both channels, but phenytoin and 2-aminoethoxydiphenyl borate (2-APB) elicited additional, distinct responses on AAAA channels. Further experiments using single alanine mutants revealed that phenytoin and 2-APB enter the inner cav-ity through distinct fenestrations, implying structural specificity to their modes of ac-cess. Using a combination of computational and functional approaches, we identified amino acid residues critical for 2-APB activity, supporting the existence of drug bind-ing site(s) within the pore region. Intrigued by the activity of 2-APB and its ana-logues, we tested additional compounds containing the diphenylmethane/amine moiety on WT channels. We identified compounds from existing clinically used drugs that exhibited diverse activity, thus expanding the pharmacological toolbox for NALCN. While the low potencies of active compounds reiterate the resistance of NALCN to pharmacological targeting, our findings lay the foundation for rational drug design to develop NALCN modulators with refined properties.
Significance Statement:
The sodium leak channel (NALCN) is essential for survival: mutations cause life-threatening developmental disorders in humans. However, no treatment is currently available due to the resistance of NALCN to pharmacological targeting. One likely reason is that the lateral fenestrations, a common route for clinically used drugs to enter and block related ion channels, are occluded in NALCN. Using a combination of computational and functional approaches, we unplugged the fenestrations of NALCN which led us to the first molecularly defined drug binding site within the pore region. Besides that, we also identified additional NALCN modulators from existing clinically used therapeutics, thus expanding the pharmacological toolbox for this leak channel.
Related Concept Videos
 01:30
Ligand-Gated Ion Channel Receptor: Gating Mechanism
 01:32
Pinching-off of Coated Vesicles
 01:18
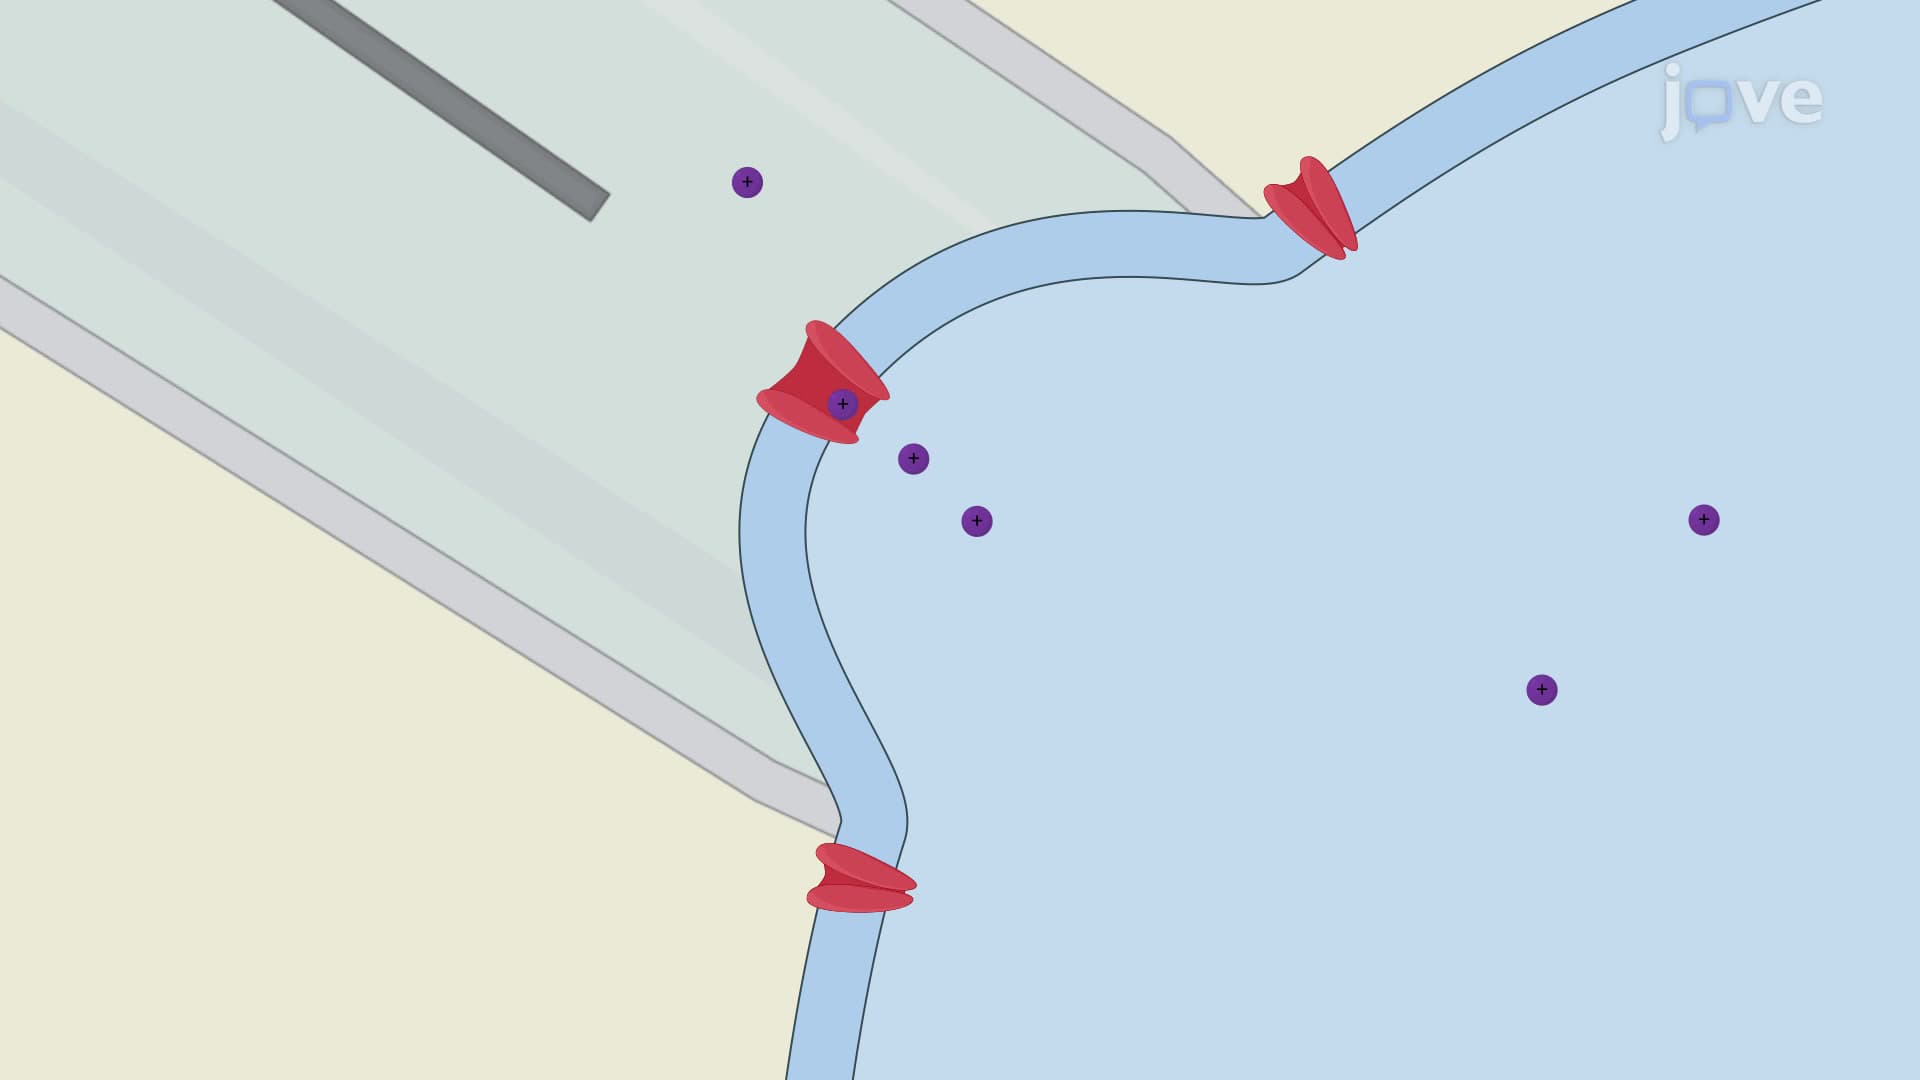
Patch Clamp
In this method, a glass micropipette containing electrolyte solution is tightly sealed against a small portion of the cell membrane. As a result, a patch of the cell...
 01:19
Ligand-gated Ion Channels
Three Subfamilies of Ligand-gated Ion Channels
Ligand-gated ion channels fall into three subfamilies. The 'Cys-loop' includes the nicotinic acetylcholine receptors, γ-aminobutyric acid (GABA), glycine, and 5-hydroxytryptamine receptors. The second one is the 'Pore-loop' channels that...