Related Experiment Video
Updated: Jul 2, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Potential energy surfaces and dynamic properties via ab initio composite and density functional approaches
Prajay Patel1,2, Joseph Chung1, Max Aksel Bowman1
1Department of Chemistry, Michigan State University, East Lansing, Michigan, USA.

Abstract:
Vibrational spectroscopy enables critical insight into the structural and dynamic properties of molecules. Presently, the majority of theoretical approaches to spectroscopy employ wavefunction-based ab initio or density functional methods that rely on the harmonic approximation. This approximation breaks down for large molecules with strongly anharmonic bonds or for molecules with large internuclear separations. An alternative to these methods involves generating molecular anharmonic potential energy surfaces (potentials) and using them to extrapolate the vibrational frequencies. This study examines the efficacy of density functional theory (DFT) and the correlation consistent Composite Approach (ccCA) in generating anharmonic frequencies from potentials of small main group molecules. Vibrational self-consistent field Theory (VSCF) and post-VSCF methods were used to calculate the fundamental frequencies of these molecules from their potentials. Functional choice, basis set selection, and mode-coupling are also examined as factors in influencing accuracy. The absolute deviations for the calculated frequencies using potentials at the ccCA level of theory were lower than the potentials at the DFT level. With DFT resulting in bending modes that are better described than those of ccCA, a multilevel DFT:ccCA approach where DFT potentials are used for single vibrational mode potentials and ccCA is used for vibrational mode-mode couplings can be utilized for larger polyatomic systems. The frequencies obtained with this multilevel approach using VCIPSI-PT2 were closer to experimental frequencies than the scaled harmonic frequencies, indicating the success of utilizing post-VSCF methods to generate more accurate representations of computed infrared spectra.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 01:26
Thermodynamic Potentials
 01:12
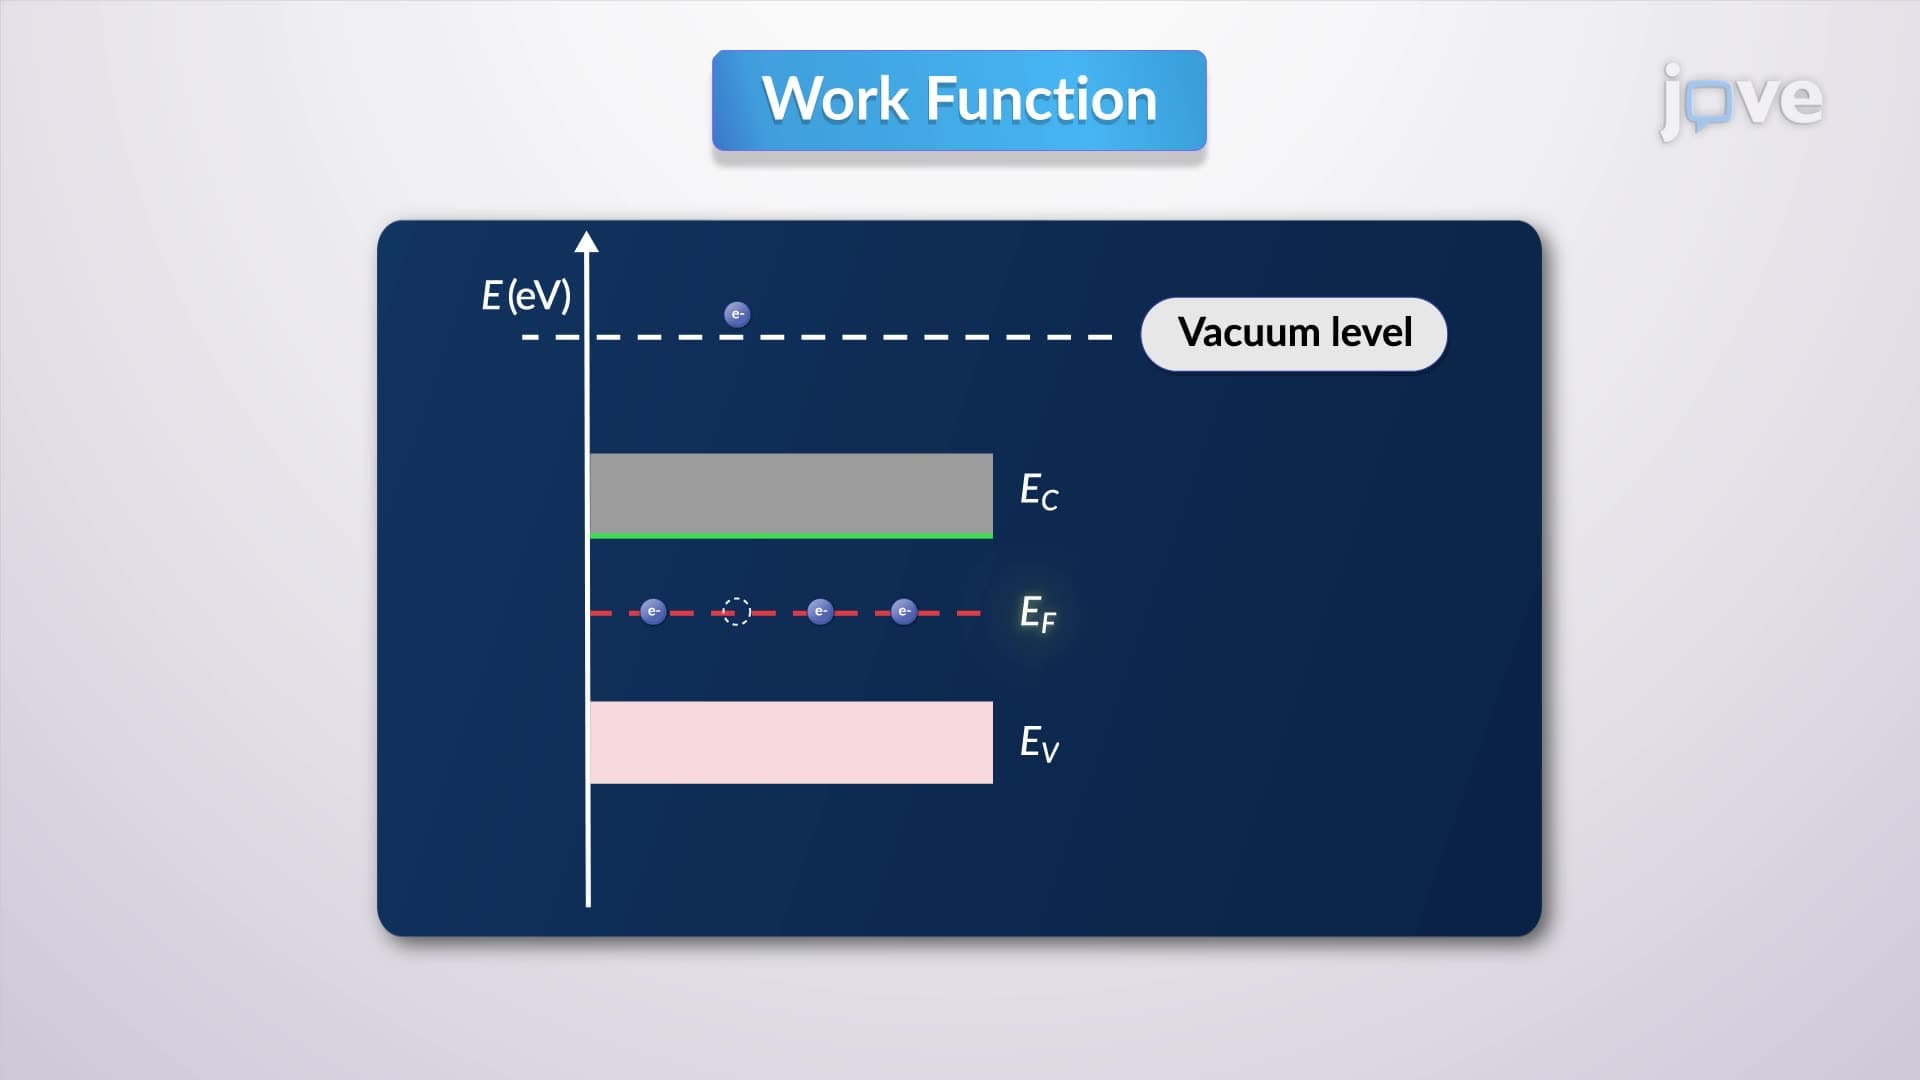
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 01:16
Potential-Energy Criterion for Equilibrium
 02:36
Molecular Geometry and Dipole Moments
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 01:13
Force and Potential Energy in One Dimension