Related Experiment Video
Updated: Jul 1, 2025


In Vitro Enzyme Measurement to Test Pharmacological Chaperone Responsiveness in Fabry and Pompe Disease
Published on: December 20, 2017
Interplay between mitochondrial dysfunction and lysosomal storage: challenges in genetic metabolic muscle diseases
Mengjiao Zhang1, Jiechao Niu1, Mengmeng Xu1
1Department of Pediatrics, The First Affiliated Hospital of Zhengzhou University, Zhengzhou, China.

Background:
Pompe disease (PD) is a rare, progressive autosomal recessive lysosomal storage disorder that directly impacts mitochondrial function, leading to structural abnormalities and potentially culminating in heart failure or cardiogenic shock. The clinical course and molecular mechanisms of the disease remain incompletely understood.
Methods:
We performed a retrospective analysis to examine the clinical manifestations, genetic traits, and the relationship between PD and mitochondrial function in a pediatric patient. This comprehensive evaluation included the use of ultrasound echocardiograms, computed tomography (CT) scans, electrocardiograms, mutagenesis analysis, and structural analysis to gain insights into the patient's condition and the underlying mechanisms of PD. For structural analysis and visualization, the structure of protein data bank ID 5KZX of human GAA was used, and VMD software was used for visualization and analysis.
Results:
The study revealed that a 5-month-old male infant was admitted due to fever, with physical examination finding abnormal cardiopulmonary function and hepatomegaly. Laboratory tests and echocardiography confirmed heart failure and hypertrophic cardiomyopathy. Despite a week of treatment, which normalized body temperature and reduced pulmonary inflammation, cardiac abnormalities did not show significant improvement. Further genetic testing identified a homozygous mutation c.2662G>T (p.E888) in the GAA gene, leading to a diagnosis of Infantile-Onset Pompe Disease (IOPD).
Conclusions:
Although enzyme replacement therapy can significantly improve the quality of life for patients with PD, enhancing mitochondrial function may represent a new therapeutic strategy for treating PD.
Related Concept Videos
 01:22
Lysosomal Hydrolases
 01:20
Inborn Errors of Metabolism
 01:19
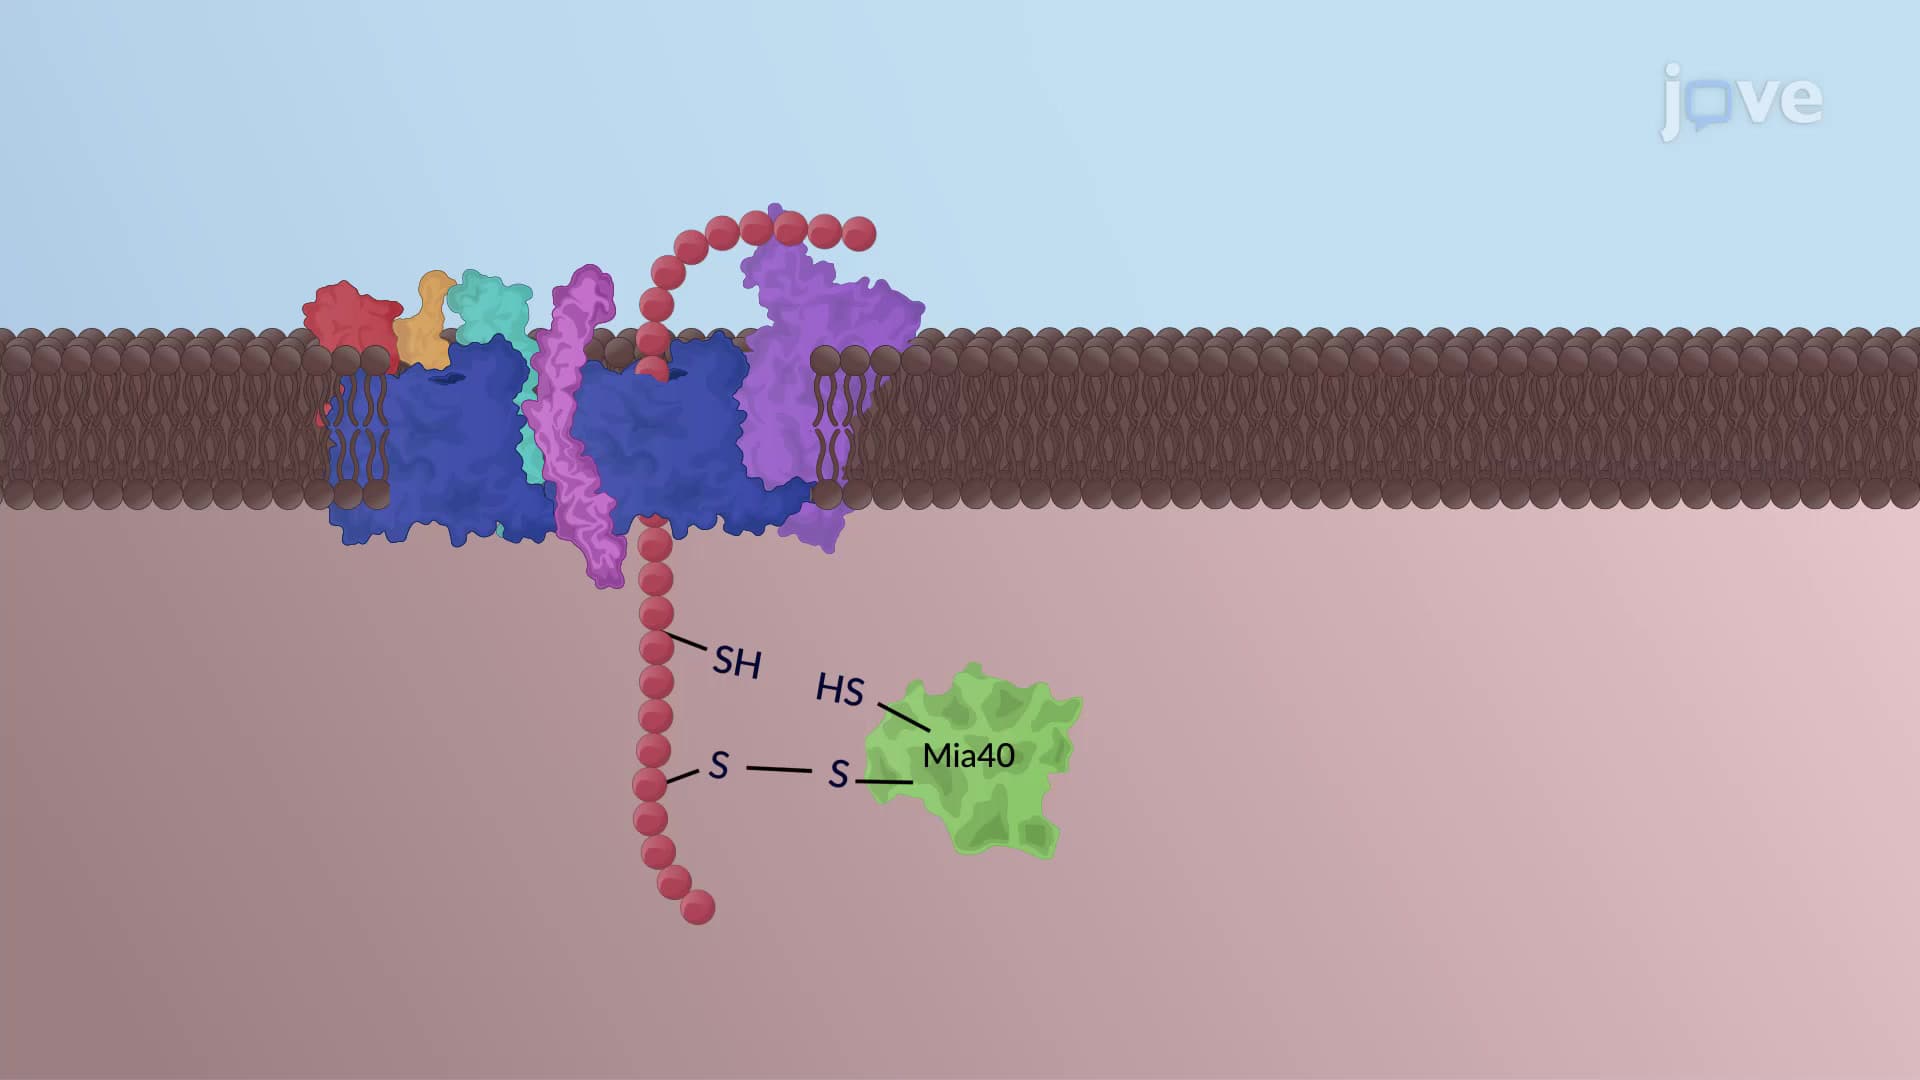
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:21
Satellite Stem Cells and Muscular Dystrophy
 01:48
ATP Synthase: Mechanism
 01:39
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...