Related Experiment Video
Updated: Jul 1, 2025

A Combined 3D Tissue Engineered In Vitro/In Silico Lung Tumor Model for Predicting Drug Effectiveness in Specific Mutational Backgrounds
Published on: April 6, 2016
Understanding CYP3A4 and P-gp mediated drug-drug interactions through PBPK modeling - Case example of pralsetinib
Christine Bowman1, Michael Dolton1, Fang Ma1
1Genentech, Inc., South San Francisco, California, USA.

Abstract:
Pralsetinib, a potent and selective inhibitor of oncogenic RET fusion and RET mutant proteins, is a substrate of the drug metabolizing enzyme CYP3A4 and a substrate of the efflux transporter P-gp based on in vitro data. Therefore, its pharmacokinetics (PKs) may be affected by co-administration of potent CYP3A4 inhibitors and inducers, P-gp inhibitors, and combined CYP3A4 and P-gp inhibitors. With the frequent overlap between CYP3A4 and P-gp substrates/inhibitors, pralsetinib is a challenging and representative example of the need to more quantitatively characterize transporter-enzyme interplay. A physiologically-based PK (PBPK) model for pralsetinib was developed to understand the victim drug-drug interaction (DDI) risk for pralsetinib. The key parameters driving the magnitude of pralsetinib DDIs, the P-gp intrinsic clearance and the fraction metabolized by CYP3A4, were determined from PBPK simulations that best captured observed DDIs from three clinical studies. Sensitivity analyses and scenario simulations were also conducted to ensure these key parameters were determined with sound mechanistic rationale based on current knowledge, including the worst-case scenarios. The verified pralsetinib PBPK model was then applied to predict the effect of other inhibitors and inducers on the PKs of pralsetinib. This work highlights the challenges in understanding DDIs when enzyme-transporter interplay occurs, and demonstrates an important strategy for differentiating enzyme/transporter contributions to enable PBPK predictions for untested scenarios and to inform labeling.
More Related Videos
 10:33
Development of Inhibitors of Protein-protein Interactions through REPLACE: Application to the Design and Development Non-ATP Competitive CDK Inhibitors
Published on: October 26, 2015
 09:17
A Robust Pneumonia Model in Immunocompetent Rodents to Evaluate Antibacterial Efficacy against S. pneumoniae, H. influenzae, K. pneumoniae, P. aeruginosa or A. baumannii
Published on: January 2, 2017
Related Concept Videos
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion,...
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:23
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance
A study on guinea pigs examined the...
 01:22
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
 01:16
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:27
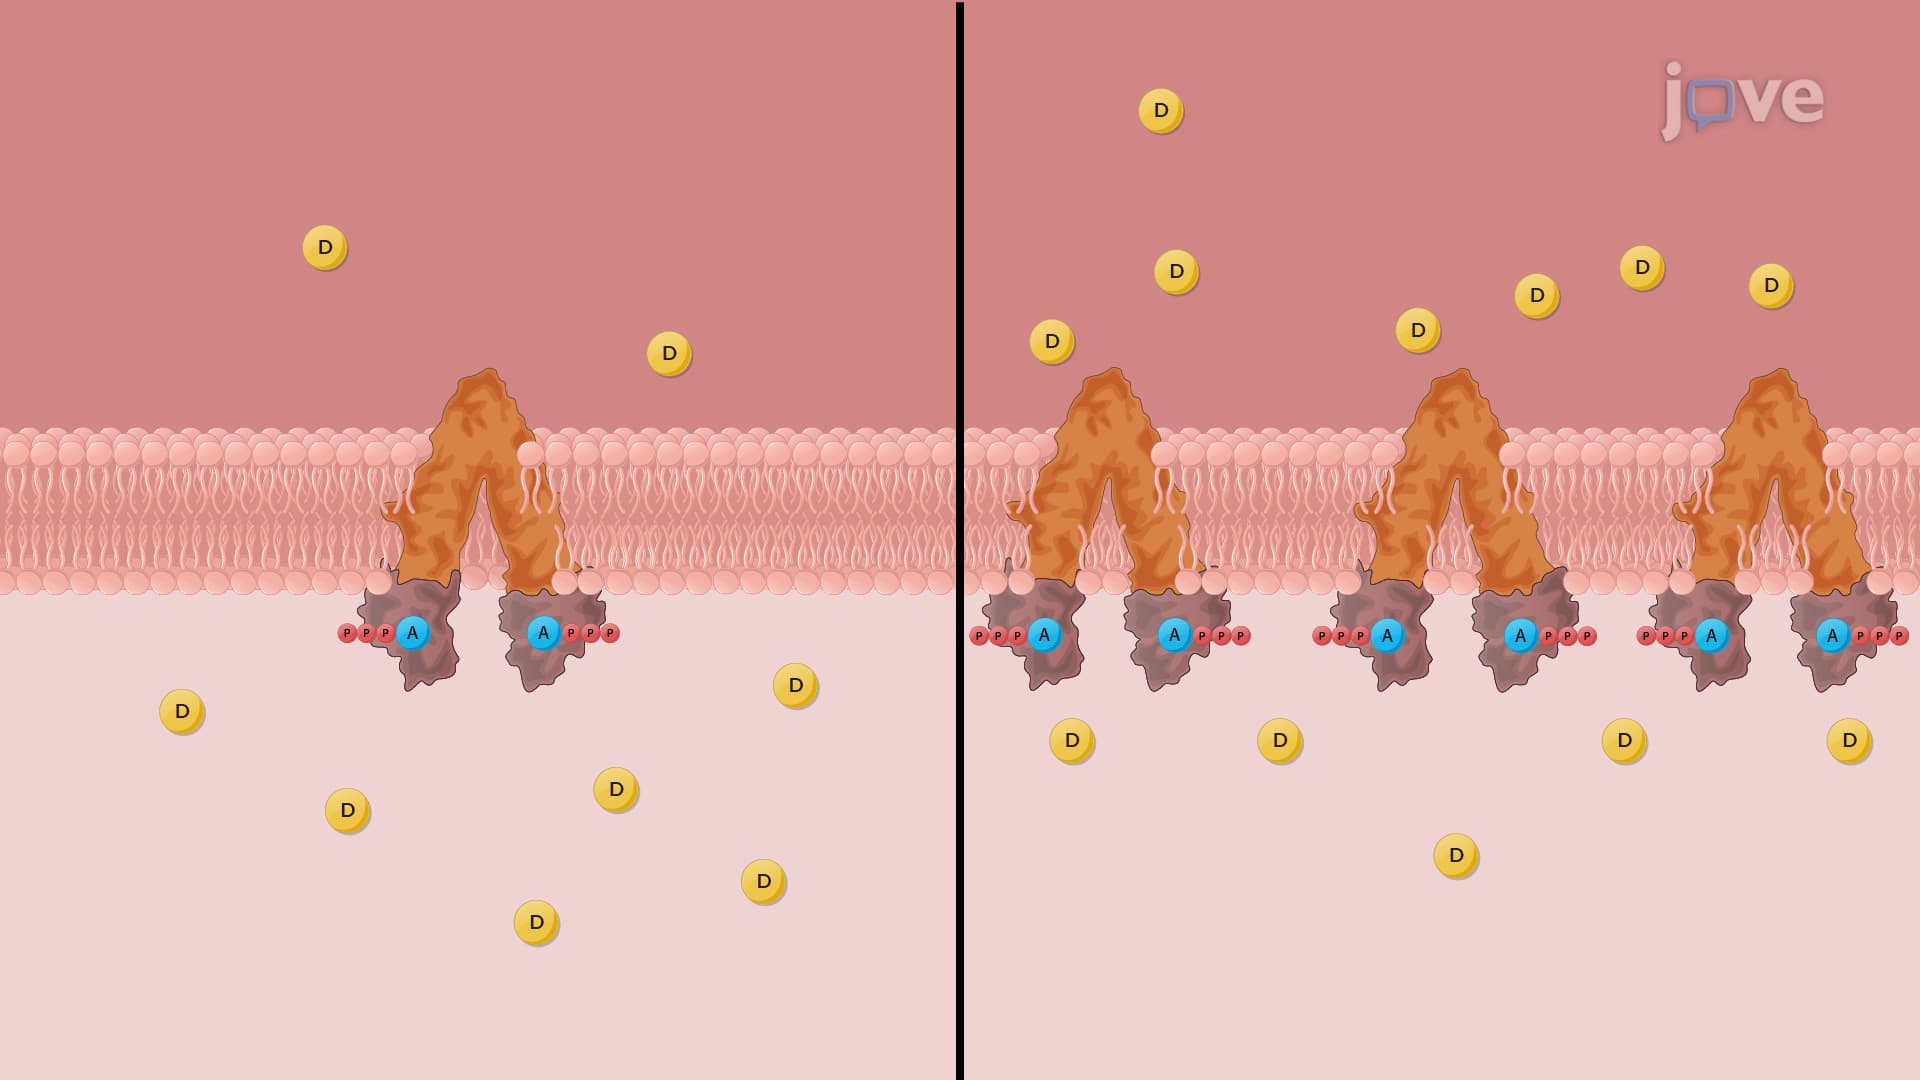
Nonlinear Pharmacokinetics: Role of Transporters
Polymorphisms occurring in drug transporters can alter...