Related Experiment Video
Updated: Jun 18, 2026

Isolation of Cortical Microglia with Preserved Immunophenotype and Functionality From Murine Neonates
Published on: January 30, 2014
Neurotoxic Microglial Activation via IFNγ-Induced Nrf2 Reduction Exacerbating Alzheimer's Disease
You Jung Kang1,2, Seung Jae Hyeon3, Amanda McQuade4,5,6,7
1Institute of Quantum Biophysics, Sungkyunkwan University, Suwon, Gyeonggi, 16419, Republic of Korea.

Abstract:
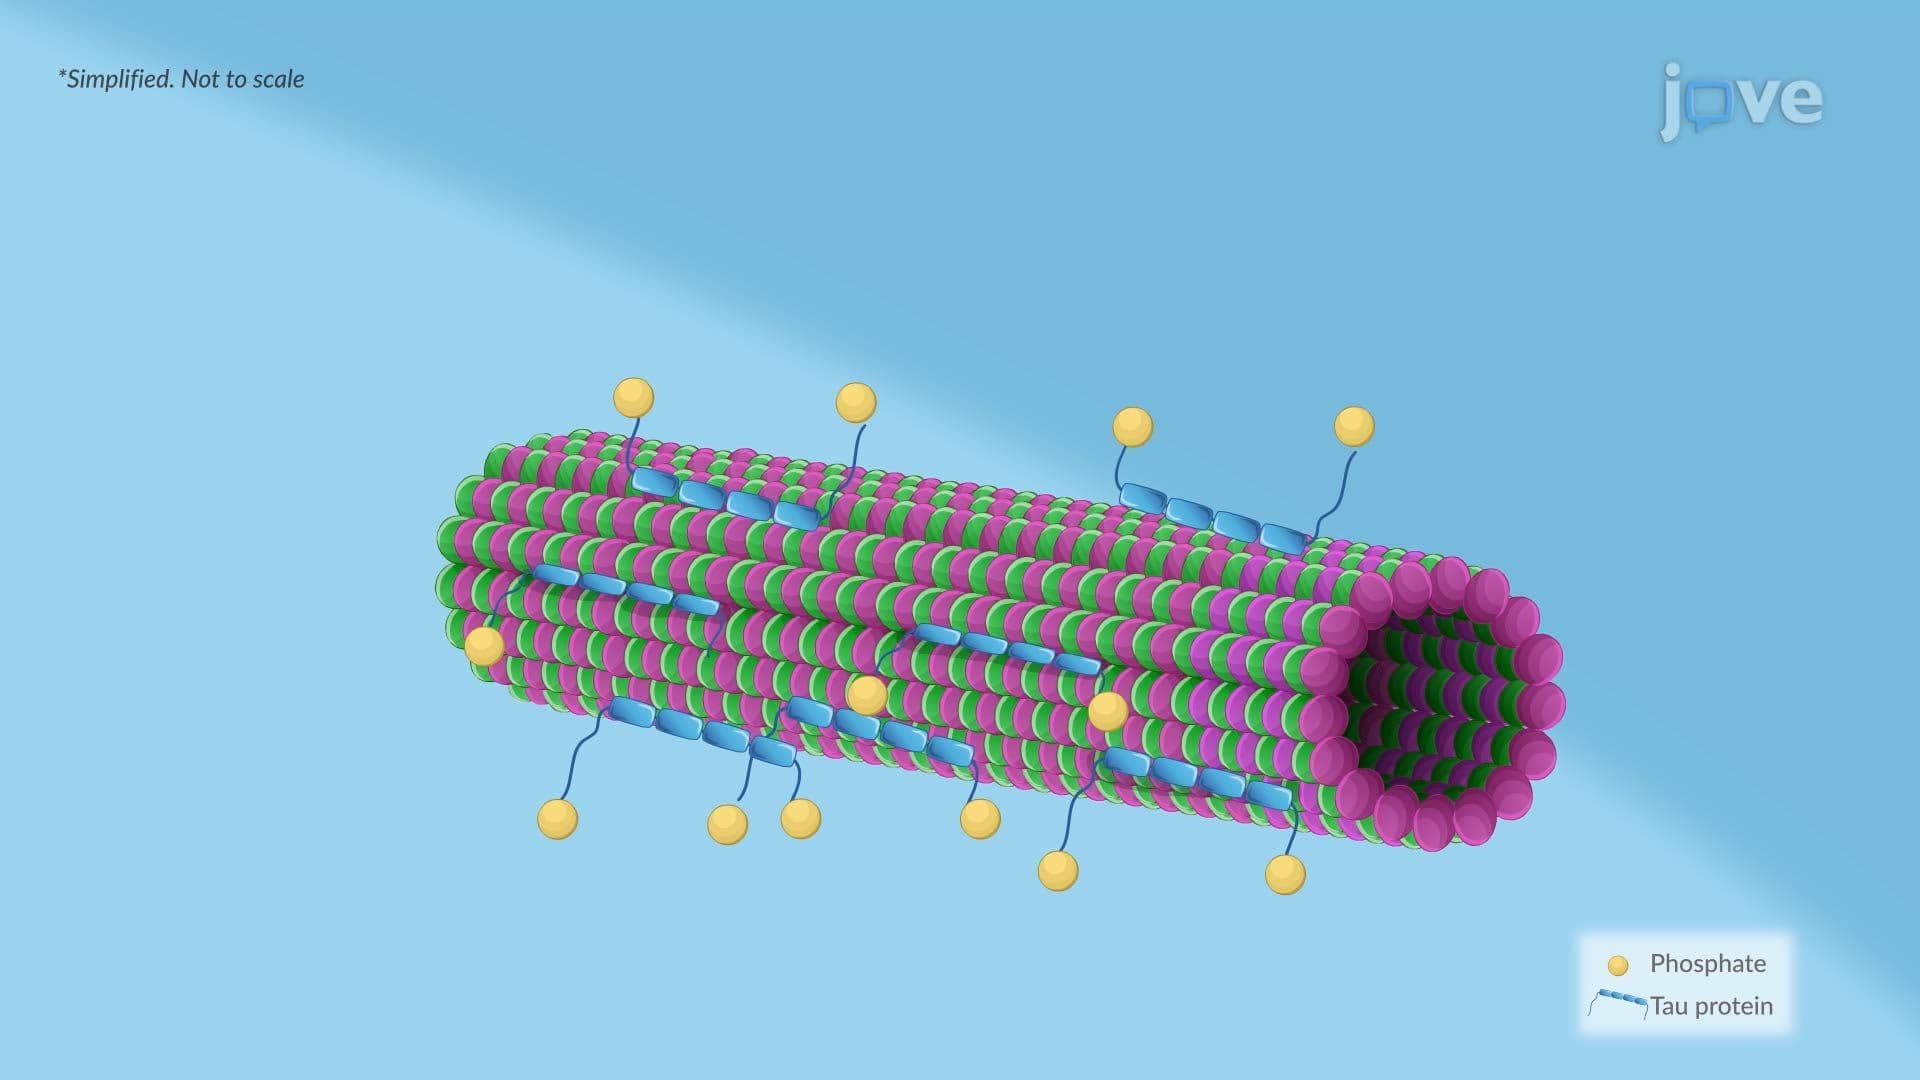
Microglial neuroinflammation appears to be neuroprotective in the early pathological stage, yet neurotoxic, which often precedes neurodegeneration in Alzheimer's disease (AD). However, it remains unclear how the microglial activities transit to the neurotoxic state during AD progression, due to complex neuron-glia interactions. Here, the mechanism of detrimental microgliosis in AD by employing 3D human AD mini-brains, brain tissues of AD patients, and 5XFAD mice is explored. In the human and animal AD models, amyloid-beta (Aβ)-overexpressing neurons and reactive astrocytes produce interferon-gamma (IFNγ) and excessive oxidative stress. IFNγ results in the downregulation of mitogen-activated protein kinase (MAPK) and the upregulation of Kelch-like ECH-associated Protein 1 (Keap1) in microglia, which inactivate nuclear factor erythroid-2-related factor 2 (Nrf2) and sensitize microglia to the oxidative stress and induces a proinflammatory microglia via nuclear factor kappa B (NFκB)-axis. The proinflammatory microglia in turn produce neurotoxic nitric oxide and proinflammatory mediators exacerbating synaptic impairment, phosphorylated-tau accumulation, and discernable neuronal loss. Interestingly, recovering Nrf2 in the microglia prevents the activation of proinflammatory microglia and significantly blocks the tauopathy in AD minibrains. Taken together, it is envisioned that IFNγ-driven Nrf2 downregulation in microglia as a key target to ameliorate AD pathology.
Insights
Alzheimer
Area of Science:
- Neuroscience
- Immunology
- Cell Biology
Background:
- Microglial neuroinflammation shifts from protective to toxic in Alzheimer's disease (AD).
- The transition mechanism from beneficial to detrimental microglial states in AD remains unclear.
- Complex neuron-glia interactions complicate understanding microglial roles in AD progression.
Purpose of the Study:
- To elucidate the mechanism of detrimental microgliosis in Alzheimer's disease.
- To investigate the role of interferon-gamma (IFNγ) and oxidative stress in microglial activation.
- To identify potential therapeutic targets for mitigating AD pathology.
Main Methods:
- Utilized 3D human AD mini-brains, post-mortem AD patient brain tissues, and 5XFAD mouse models.
- Analyzed molecular pathways including MAPK, Keap1, Nrf2, and NFκB signaling.
- Investigated the impact of Nrf2 recovery on microglial activation and AD pathology.
Main Results:
- Amyloid-beta (Aβ) and reactive astrocytes induce IFNγ and oxidative stress in AD models.
- IFNγ downregulates MAPK and upregulates Keap1 in microglia, inactivating Nrf2.
- Inactivated Nrf2 sensitizes microglia to oxidative stress, promoting a proinflammatory state via NFκB, leading to neurotoxicity and tauopathy.
- Restoring Nrf2 function in microglia blocked proinflammatory activation and significantly reduced tauopathy.
Conclusions:
- IFNγ-induced Nrf2 downregulation in microglia is a critical driver of neuroinflammation and neurodegeneration in AD.
- Targeting the IFNγ-Nrf2 axis in microglia presents a promising therapeutic strategy for Alzheimer's disease.
- Understanding these molecular mechanisms is key to developing effective AD treatments.