Related Experiment Video
Updated: Jun 30, 2025
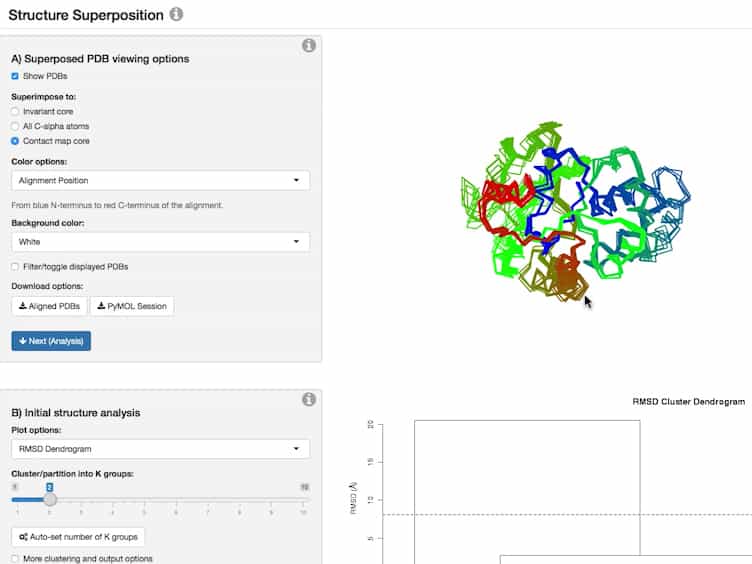

Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Classifying protein kinase conformations with machine learning
Ivan Reveguk1, Thomas Simonson1
1Laboratoire de Biologie Structurale de la Cellule (CNRS UMR7654), Ecole Polytechnique, Palaiseau, France.

Abstract:
Protein kinases are key actors of signaling networks and important drug targets. They cycle between active and inactive conformations, distinguished by a few elements within the catalytic domain. One is the activation loop, whose conserved DFG motif can occupy DFG-in, DFG-out, and some rarer conformations. Annotation and classification of the structural kinome are important, as different conformations can be targeted by different inhibitors and activators. Valuable resources exist; however, large-scale applications will benefit from increased automation and interpretability of structural annotation. Interpretable machine learning models are described for this purpose, based on ensembles of decision trees. To train them, a set of catalytic domain sequences and structures was collected, somewhat larger and more diverse than existing resources. The structures were clustered based on the DFG conformation and manually annotated. They were then used as training input. Two main models were constructed, which distinguished active/inactive and in/out/other DFG conformations. They considered initially 1692 structural variables, spanning the whole catalytic domain, then identified ("learned") a small subset that sufficed for accurate classification. The first model correctly labeled all but 3 of 3289 structures as active or inactive, while the second assigned the correct DFG label to all but 17 of 8826 structures. The most potent classifying variables were all related to well-known structural elements in or near the activation loop and their ranking gives insights into the conformational preferences. The models were used to automatically annotate 3850 kinase structures predicted recently with the Alphafold2 tool, showing that Alphafold2 reproduced the active/inactive but not the DFG-in proportions seen in the Protein Data Bank. We expect the models will be useful for understanding and engineering kinases.

Related Concept Videos
 02:54
Protein Kinases and Phosphatases
Protein kinases
Many proteins in the cell are regulated by phosphorylation, the addition of a phosphate group. A family of enzymes called kinases...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:24
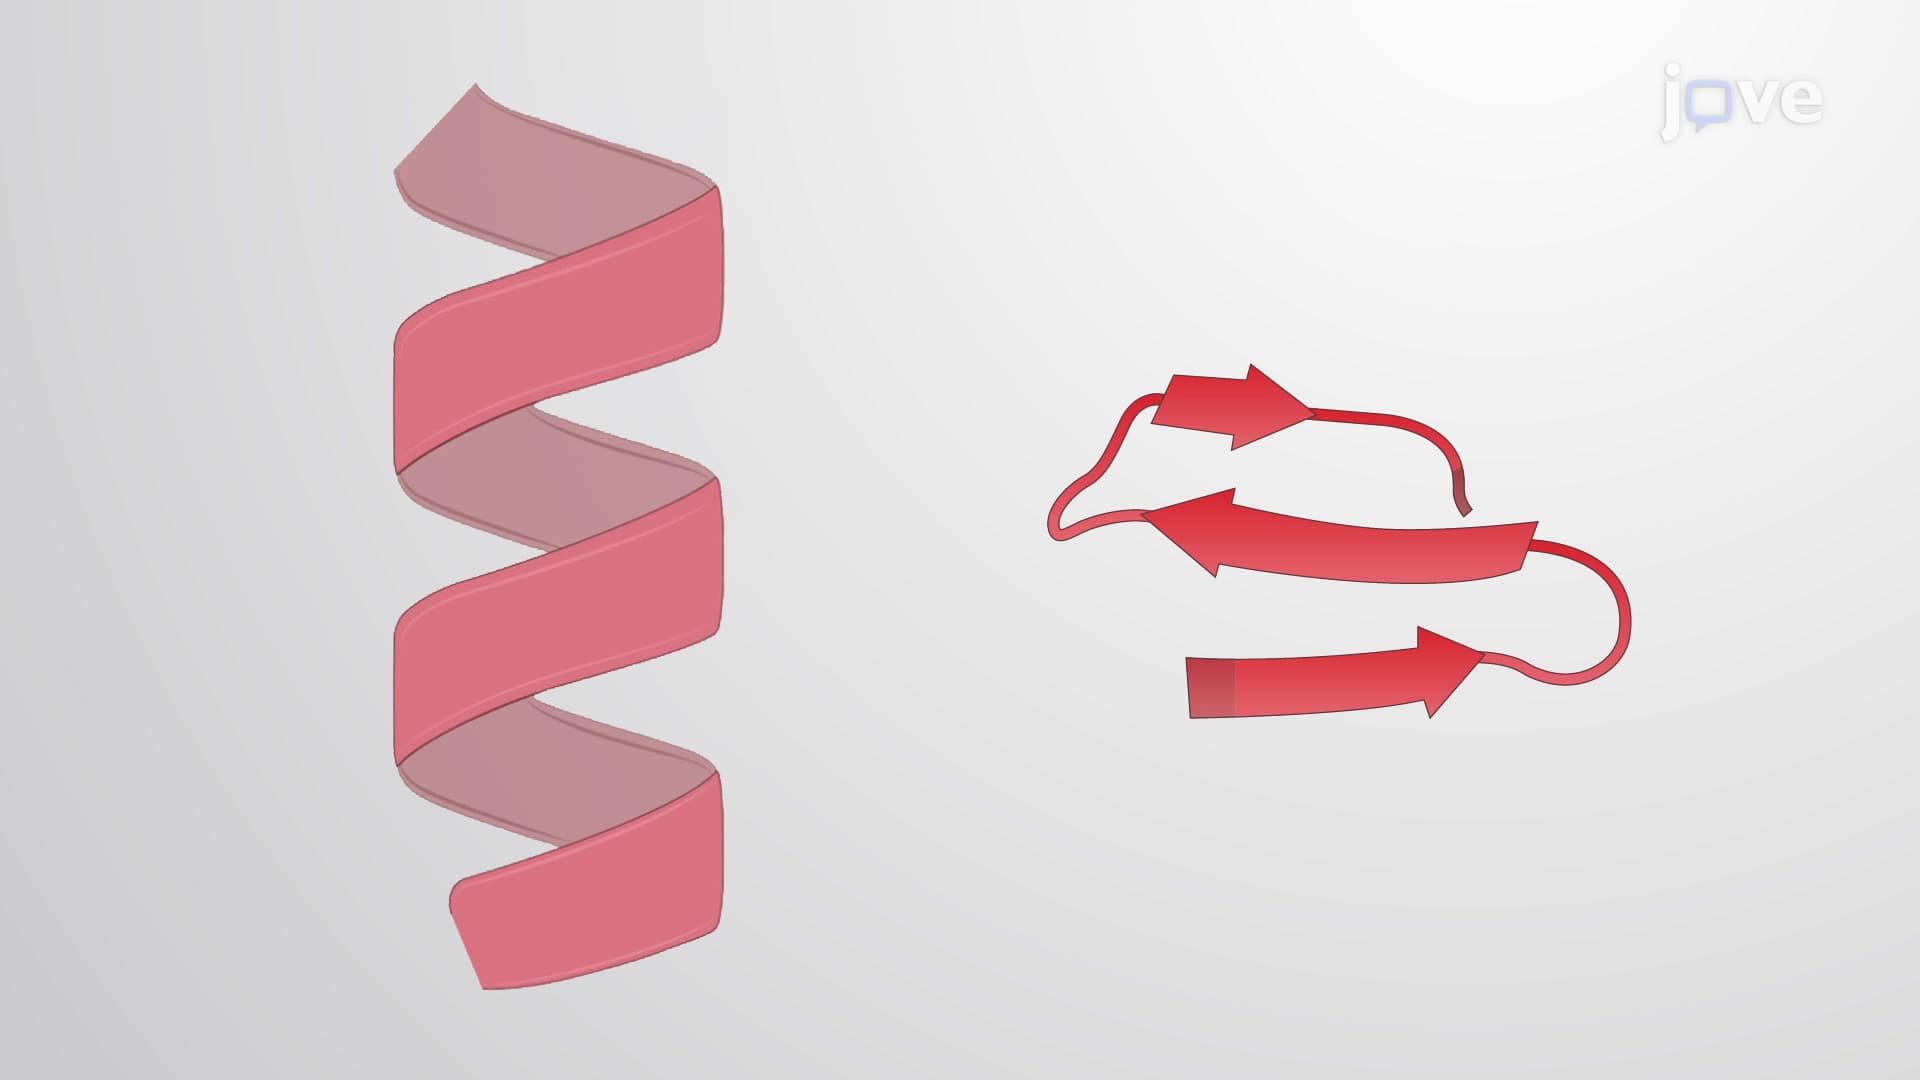
Protein Organization
The primary structure of a protein is its amino acid sequence....
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to...
 01:07
MAPK Signaling Cascades
 02:04
Protein-protein Interfaces